Preview: "Cage escape" in photosensitized of diazonium derivatives
英文原文与中文译文逐段对照 · English original with Chinese translation
Chem Catalysis
《化学催化》
Preview “Cage escape” in photosensitized diazonium derivatives
“重氮衍生物光敏化反应中的‘笼逃逸’”预览
Jianning Zhang ${}^{1}$ and Fanyang Mo ${}^{1, * }$
张建宁${}^{1}$和莫凡洋${}^{1, * }$
In this issue of Chem Catalysis, Troian-Gautier and co-workers focus on the mechanism of the photosensitized activation reaction of diazonium derivatives by studying the electron-transfer process between photosensitizers and diazonium salts. Their proposed “cage-escape” mechanistic picture is verified in the C-B bond-formation reaction of diazonium salts.
在本期《化学催化》中,特罗安 - 高蒂埃及其同事通过研究光敏剂与重氮盐之间的电子转移过程,聚焦于重氮衍生物光敏化活化反应的机理。他们提出的“笼逃逸”机理图景在重氮盐的C - B键形成反应中得到了验证。
Diazonium salts are widely used radical precursors in organic synthesis, the dyeing industry, and the modification of metallic or non-metallic material surfaces. ${}^{1 - 3}$ In organic synthesis, classical named reactions (e.g., the Sandmeyer, Pschorr, and Gomberg-Bachmann reactions) are applied in the transformation of diazonium salts to other functional organic molecules. Photosensitized activation reactions of diazonium salts have also been developed for C-C or C-heteroatom bond formation ${}^{4,5}$ (Figure 1A).
重氮盐是有机合成、染料工业以及金属或非金属材料表面改性中广泛使用的自由基前体。${}^{1 - 3}$在有机合成中,经典的命名反应(如桑德迈尔反应、普肖尔反应和贡伯格 - 巴赫曼反应)被用于将重氮盐转化为其他功能性有机分子。重氮盐的光敏化活化反应也已被开发用于形成C - C键或C - 杂原子键${}^{4,5}$(图1A)。
The article by Troian-Gautier and coworkers in this issue of Chem Catalysis focuses on the primitive reaction steps in the photosensitized aryl diazonium activation of diazonium salts, i.e., the excitation of the photocatalyst and the electron-transfer process. ${}^{6}$ Detailed experimental, spectroscopic, and theoretical results provide precise kinetic information on the interaction between photosensitizers (PSs) and diazonium salts. Further, the C-B bond formation of diazonium salts is an example that verifies that the observations and different yields are consistent with the proposed mechanism.
特罗安 - 高蒂埃及其同事在本期《化学催化》上发表的文章聚焦于重氮盐光敏化芳基重氮活化中的原始反应步骤,即光催化剂的激发和电子转移过程。${}^{6}$详细的实验、光谱和理论结果提供了关于光敏剂(PSs)与重氮盐之间相互作用的精确动力学信息。此外,重氮盐的C - B键形成是一个例子,验证了观察结果和不同产率与所提出的机理一致。
Nine commercial or synthesized PSs (including six rare-earth transition-metal PSs and three earth-abundant transition-metal PSs) and 13 quenchers (including ten aryl diazonium salts and three calix[4]arene tetradiazonium derivatives) were chosen as the studied molecules (Figure 1B). The Hammett parameters (σ) of 13 quenchers ranged from -0.83 to 0.78 according to previous work, ${}^{7}$ indicateing the different electronegativities of the diazonium salts.
选择了九种商业或合成的PSs(包括六种稀土过渡金属PSs和三种储量丰富的过渡金属PSs)以及13种猝灭剂(包括十种芳基重氮盐和三种杯[4]芳烃四重氮衍生物)作为研究分子(图1B)。根据先前的工作,13种猝灭剂的哈米特参数(σ)范围为 - 0.83至0.78,${}^{7}$表明重氮盐的不同电负性。
The authors recorded the ground-state absorption spectra and photoluminescence (PL) spectra under excitation within the visible range of solar light to show the differences among the photophysical properties of PSs. The iridium and ruthenium transition-metal PSs showed absorption features at wavelengths greater than ${400}\mathrm{\;{nm}}$ (i.e.,[Ru(bpy) ${}_{3}$ ] ${}^{2 + }$ : 450 nm), and some earth-abundant transition-metal PSs showed absorption peaks around ${500}\mathrm{\;{nm}}$ (i.e.,[Fe(phtmeimb) ${}_{2}$ ] ${}^{ + }$ : 500 nm). The PL spectra of different PSs under solar light ranged from ${472}\mathrm{\;{nm}}$ for ${\left\lbrack \operatorname{lr}\left( {\left( {\mathrm{{dFCF}}}_{3}\mathrm{{ppy}}\right) }_{2}\left( \mathrm{{dtb}}\right) \right) \right\rbrack }^{ + }$ to ${737}\mathrm{\;{nm}}$ for ${\left\lbrack \mathrm{{Os}}{\left( \mathrm{{bpy}}\right) }_{3}\right\rbrack }^{2 + }$ . Time-resolved PL recorded the excited-state lifetimes ranging from 2 ns for ${\left\lbrack \mathrm{{Fe}}{\left( \text{ ph }\text{ meim }\mathrm{b}\right) }_{2}\right\rbrack }^{ + }$ to 2,010 ns for [Ir( ${\left( {\mathrm{{dFCF}}}_{3}\mathrm{{ppy}}\right) }_{2}{\left( \mathrm{{dtb}}\right) }^{ + }$ .
作者记录了在太阳光可见光范围内激发下的基态吸收光谱和光致发光(PL)光谱,以展示PSs光物理性质的差异。铱和钌过渡金属PSs在波长大于${400}\mathrm{\;{nm}}$(即[Ru(bpy)${}_{3}$]${}^{2 + }$:450 nm)处显示吸收特征,一些储量丰富的过渡金属PSs在${500}\mathrm{\;{nm}}$左右显示吸收峰(即[Fe(phtmeimb)${}_{2}$]${}^{ + }$:500 nm)。不同PSs在太阳光下的PL光谱范围从${472}\mathrm{\;{nm}}$(对于${\left\lbrack \operatorname{lr}\left( {\left( {\mathrm{{dFCF}}}_{3}\mathrm{{ppy}}\right) }_{2}\left( \mathrm{{dtb}}\right) \right) \right\rbrack }^{ + }$)到${737}\mathrm{\;{nm}}$(对于${\left\lbrack \mathrm{{Os}}{\left( \mathrm{{bpy}}\right) }_{3}\right\rbrack }^{2 + }$)。时间分辨PL记录的激发态寿命范围从${\left\lbrack \mathrm{{Fe}}{\left( \text{ ph }\text{ meim }\mathrm{b}\right) }_{2}\right\rbrack }^{ + }$的2 ns到[Ir(${\left( {\mathrm{{dFCF}}}_{3}\mathrm{{ppy}}\right) }_{2}{\left( \mathrm{{dtb}}\right) }^{ + }$的2010 ns。
Cyclic voltammetry and differential pulse voltammetry (DPV) determined the reduction and oxidation potentials of the PSs and quenchers. The excited-state oxidation potentials of the PSs ranged from -0.63 to $- {0.78}\mathrm{\;V}$ versus normal hydrogen electrode (NHE), and the reduction potentials of the quenchers ranged from -0.51 to ${0.77}\mathrm{\;V}$ versus NHE, trending linearly with their Hammett parameters. These results suggest the thermodynamic spontaneity of the electron-transfer process from PSs to quenchers.
循环伏安法和差分脉冲伏安法(DPV)测定了光催化剂(PSs)和猝灭剂的还原和氧化电位。相对于标准氢电极(NHE),PSs的激发态氧化电位范围为-0.63至$- {0.78}\mathrm{\;V}$,猝灭剂的还原电位范围为-0.51至${0.77}\mathrm{\;V}$,与它们的哈米特参数呈线性趋势。这些结果表明了从PSs到猝灭剂的电子转移过程的热力学自发性。
The results of steady-state quenching experiments also verified the thermodynamic spontaneity. In almost all cases, an increase in quencher concentration resulted in a decrease in PS excited-state lifetime. The quenching rate constants $\left( {\mathrm{k}}_{\mathrm{q}}\right)$ increased with the driving force for electron transfer before reaching a plateau, which could have been caused by a solvent diffusion limit.
稳态猝灭实验的结果也证实了热力学自发性。在几乎所有情况下,猝灭剂浓度的增加导致PS激发态寿命的降低。猝灭速率常数$\left( {\mathrm{k}}_{\mathrm{q}}\right)$随着电子转移驱动力的增加而增加,直到达到一个平稳期,这可能是由溶剂扩散限制引起的。
The photolysis experiments confirmed that the photoreaction of PSs and quenchers is truly via excited-state electron transfer. Under continuous irradiation, the metal-to-ligand charge transfer (MLCT) transitions of PSs were gradually bleached as the reaction of PSs and quenchers proceeded. This indicates that excited-state electron transfer can occur. Moreover, reduction experiments of ascorbic acid tested the reversibility of the reaction. Nanosecond transient absorption experiments also confirmed the presence of oxidative excited-state electrons.
光解实验证实,PSs和猝灭剂的光反应确实是通过激发态电子转移进行的。在连续照射下,随着PSs和猝灭剂反应的进行,PSs的金属到配体电荷转移(MLCT)跃迁逐渐被漂白。这表明可以发生激发态电子转移。此外,抗坏血酸的还原实验测试了反应的可逆性。纳秒瞬态吸收实验也证实了氧化激发态电子的存在。
The excited PSs and quenchers form an “encounter complex” and then undergo a cage-escape process to generate products (Figure 1A). The authors estimated the cage-escape yields $\left( {\varphi }_{\mathrm{{CE}}}\right)$ , which describe the efficiency with which the oxidized and reduced species separate. The typical ${\varphi }_{\mathrm{{CE}}}$ value (i.e., methyviolo-gen as electron acceptors and anthracene derivatives as electron donors) ranges from 14% to 27%. Most oxidized and reduced species cannot escape a solvent cage. On the contrary, they regenerate the initial state via geminate charge recombination. The estimated ${\varphi }_{\mathrm{{CE}}}$ value for diazonium derivatives can reach almost 100%, which indicates the complete release of the diazonium radical. A possible reason for the high ${\varphi }_{\mathrm{{CE}}}$ value is the irreversible generation of inert ${\mathrm{N}}_{2}$ gas. The ${\mathrm{N}}_{2}$ gas extrusion impedes back electron transfer within the solvent cage, so the diazonium radical can be released efficiently after one-electron reduction.
激发的PSs和猝灭剂形成一个“遭遇复合物”,然后经历一个笼逃逸过程以产生产物(图1A)。作者估计了笼逃逸产率$\left( {\varphi }_{\mathrm{{CE}}}\right)$,它描述了氧化和还原物种分离的效率。典型的${\varphi }_{\mathrm{{CE}}}$值(即以甲基紫精作为电子受体和蒽衍生物作为电子供体)范围为14%至27%。大多数氧化和还原物种无法逃离溶剂笼。相反,它们通过双生电荷复合再生初始状态。重氮衍生物的估计${\varphi }_{\mathrm{{CE}}}$值几乎可以达到100%,这表明重氮自由基的完全释放。{{F}1}值高的一个可能原因是惰性${\mathrm{N}}_{2}$气体的不可逆生成。${\mathrm{N}}_{2}$气体的挤出阻碍了溶剂笼内的反向电子转移,因此重氮自由基在单电子还原后可以有效地释放出来。
${}^{1}$ School of Materials Science and Engineering, Peking University, Beijing 100871, China
${}^{1}$ 材料科学与工程学院,北京大学,北京100871,中国
*Correspondence:
*通讯作者:
https://doi.org/10.1016/j.checat.2023.100527
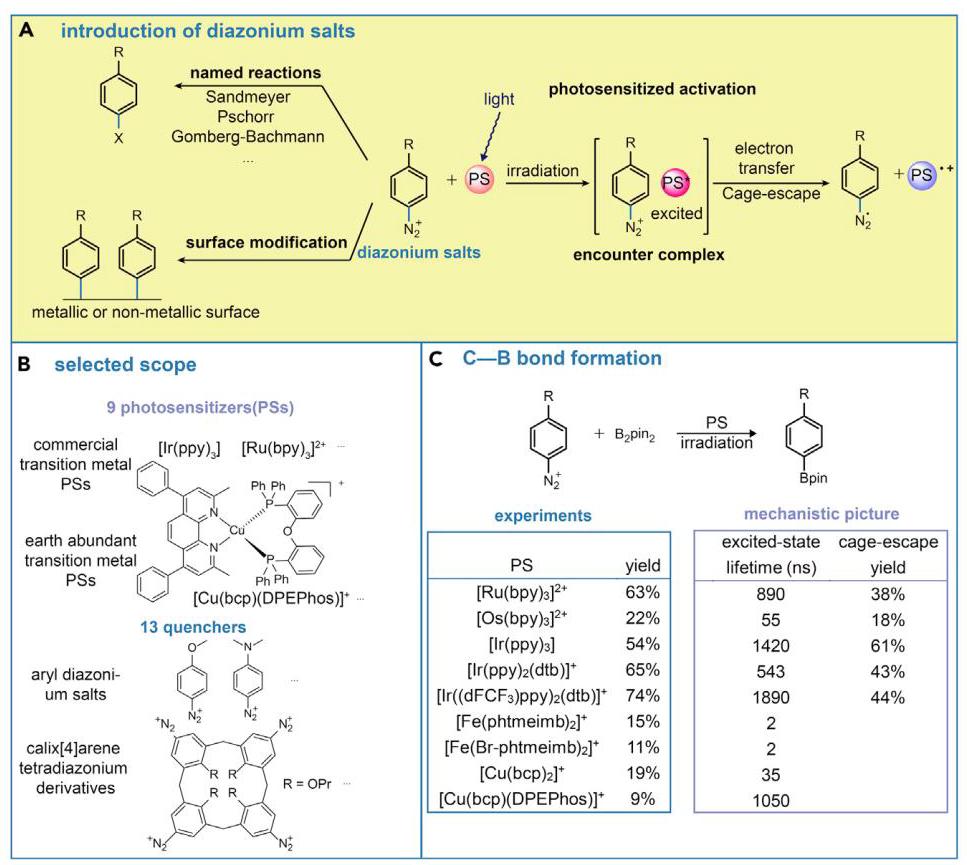
Figure 1. Mechanism insights and experimental verification of photosensitized activation reaction of diazonium derivatives
图1. 重氮衍生物光敏活化反应的机理见解和实验验证
(A) The introduction of diazonium salt transformation.
(A) 重氮盐转化的介绍。
(B) The selected substrate scope.
(B) 所选底物范围。
(C) Experimental verification of the proposed mechanism.
(C) 所提出机理的实验验证。
Furthermore, theoretical density functional theory calculation confirmed that C-N bond cleavage occurs after the electron-transfer process.
此外,理论密度泛函理论计算证实,电子转移过程后发生C-N键断裂。
Troian-Gautier and co-workers chose the C-B bond formation of diazonium salts as an actual photoredox catalysis reaction to verify the proposed mechanistic picture (Figure 1C) and measured the product yields. They found a correlation between the reaction yields and ${\varphi }_{\mathrm{{CE}}}$ , showing that differences in yields can come from differences in cage-escape yields in the transformation of diazonium salts.
Troian-Gautier及其同事选择重氮盐的C-B键形成作为实际的光氧化还原催化反应,以验证所提出的机理图(图1C)并测量产物产率。他们发现反应产率与${\varphi }_{\mathrm{{CE}}}$之间存在相关性,表明产率差异可能来自重氮盐转化过程中笼逃逸产率的差异。
Overall, this work by Troian-Gautier and colleagues investigated the electron-transfer process in the photosensitive reaction of diazonium salts. The authors estimated the cage-escape yields, and the values can explain the yield differences in actual reactions. This could inspire the design of more efficient visible-light-mediated transformations.
总体而言,Troian-Gautier及其同事的这项工作研究了重氮盐光敏反应中的电子转移过程。作者估计了笼逃逸产率,这些值可以解释实际反应中的产率差异。这可能会激发更高效的可见光介导转化的设计。
DECLARATION OF INTERESTS
利益声明
The authors declare no competing interests.
作者声明不存在利益冲突。
REFERENCES
参考文献
On a new series of bodies in which nitrogen is substituted for hydrogen. Phil. Trans. Roy. Soc. Lond. 154, 667-731.
Mo, F., Qiu, D., Zhang, L., and Wang, J. (2021). Recent development of aryl diazonium chemistry for the derivatization of aromatic compounds. Chem. Rev. 121, 5741-5829.
Bélanger, D., and Pinson, J. (2011). Electrografting: A powerful method for surface modification. Chem. Soc. Rev. 40, 3995-4048.
Cano-Yelo, H., and Deronzier, A. (1984). Photooxidation of tris(2,2’-bipyridine)ruthenium(II) by para-substituted benzene diazonium salts in acetonitrile. Two-compartment J. Chem. Soc. Faraday Trans. 1 80, 3011-3019.
Cano-Yelo, H., and Deronzier, A. (1984). Photooxidation of some carbinols by the Ru(II) polypyridyl complex-aryl diazonium salt
Ripak, A., De Kreijger, S., Sampaio, R.N., Vincent, C.A., Cauet, E., Jabin, I., Tambar, U.K., Elias, B., and Troian-Gautier, L. (2023). Photosensitized activation of diazonium derivatives for C-B bond formation. Chem Catal. 3, 100490.
Hansch, C., Leo, A., and Taft, R.W. (1991). A survey of Hammett substituent constants and resonance and field parameters. Chem. Rev.