Transition Metal Free Stannylation of Alkyl Halides: the Rapid Synthesis of Alkyltrimethylstannanes
英文原文与中文译文逐段对照 · English original with Chinese translation
Songyi Li, ${}^{\text{ § }}$ Chang Lian, ${}^{\text{ § }}$ Guanglu Yue, Jianning Zhang, Di Qiu,* and Fanyang Mo*
李松毅,${}^{\text{ § }}$ 连畅,${}^{\text{ § }}$ 岳光露,张建宁,邱迪,* 及莫凡洋*
Cite This: J. Org. Chem. 2022, 87, 4291-4297
引用本文:《有机化学杂志》2022年,第87卷,4291 - 4297页
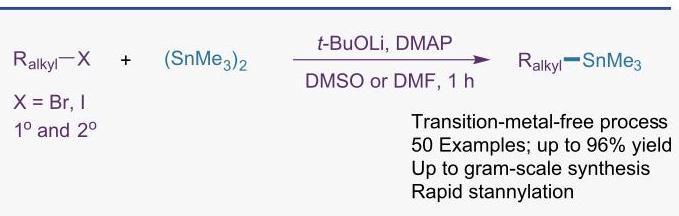
ABSTRACT: A transition metal free stannylation reaction of alkyl bromides and iodides with hexamethyldistannane has been developed. This protocol is operationally convenient and features a rapid reaction and good functional group tolerance. A wide range of functionalized primary and secondary alkyl and benzyl trimethyl stannanes are prepared in moderate to excellent yields. The success of the gram-scale procedure and tandem Stille coupling reaction has allowed this protocol to demonstrate potential for application in organic synthesis. Both experimental and theoretical studies reveal the mechanistic details of this stannylation reaction.
摘要:已开发出一种使用六甲基二锡烷对溴代烃和碘代烃进行无过渡金属参与的锡基化反应。该方法操作简便,具有反应迅速和良好的官能团耐受性的特点。以中等至优异的产率制备了多种官能化的伯、仲烷基及苄基三甲基锡烷。克级规模的反应及串联的施蒂勒偶联反应的成功使该方法在有机合成中展现出应用潜力。实验和理论研究均揭示了该锡基化反应的机理细节。
1. INTRODUCTION
1. 引言
The Stille cross-coupling reaction using organotin compounds as the coupling partners has demonstrated great applications in organic synthesis and other fields. ${}^{1 - 5}$ In particular, in the synthesis of functional materials and polymers, the Stille cross-coupling reaction features apparent advantages. ${}^{6,7}$ Apart from the widely applied arylstannanes, corresponding alkylstannanes and benzylstannanes have also been extensively explored as versatile building blocks in carbon-carbon bond-forming reactions. ${}^{8 - {14}}$ In particular, alkyl and benzyl trimethylstan-nanes have been utilized not only as important building blocks for $\mathrm{C} - \mathrm{C}$ bond formation but also in selectively constructing $\mathrm{C} - \mathrm{O}$ and $\mathrm{C} - \mathrm{X}$ bonds. ${}^{9,{15} - {19}}$ The development of the synthetic methods for various organotin derivatives has become a crucial issue. To date, compared with those of arylstannanes, the synthetic routes toward alkylstannanes are very limited. Traditional methods for accessing these reagents involve the nucleophilic substitution of alkyl metallic reagents with trialkyltin chloride. ${}^{{20} - {24}}$ Despite the synthetic utility, this method suffers from several limitations, such as rigorous reaction conditions, poor functional group tolerance, etc. Additionally, several other reactions for accessing alkylstan-nanes have been infrequently reported, including the carbene insertion of diazo compounds with ${\mathrm{{Bu}}}_{3}\mathrm{{SnH}},{}^{{25} - {28}}$ as well as the addition of alkenes with ${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}.{}^{{29} - {31}}$ Alternatively, Pd-catalyzed stannylation of organohalides with ditin compounds represents a reliable route for synthesizing stannanes (Figure 1a). ${}^{{32},{33}}$ However, these methods are effective for only primary benzyl halides, but not for general alkyl counterparts.
使用有机锡化合物作为偶联伙伴的Stille交叉偶联反应在有机合成和其他领域已展现出巨大的应用价值。${}^{1 - 5}$ 特别是在功能材料和聚合物的合成中,Stille交叉偶联反应具有明显的优势。${}^{6,7}$ 除了广泛应用的芳基锡烷外,相应的烷基锡烷和苄基锡烷也作为碳 - 碳键形成反应中的通用结构单元得到了广泛研究。${}^{8 - {14}}$ 特别是,烷基和苄基三甲基锡烷不仅被用作形成$\mathrm{C} - \mathrm{C}$键的重要结构单元,还用于选择性构建$\mathrm{C} - \mathrm{O}$和$\mathrm{C} - \mathrm{X}$键。${}^{9,{15} - {19}}$ 各种有机锡衍生物合成方法的开发已成为一个关键问题。迄今为止,与芳基锡烷相比,烷基锡烷的合成路线非常有限。传统的制备这些试剂的方法涉及用三烷基氯化锡对烷基金属试剂进行亲核取代。${}^{{20} - {24}}$ 尽管具有合成实用性,但该方法存在一些局限性,如反应条件苛刻、官能团耐受性差等。此外,很少有其他制备烷基锡烷的反应被报道,包括重氮化合物与${\mathrm{{Bu}}}_{3}\mathrm{{SnH}},{}^{{25} - {28}}$的卡宾插入反应以及烯烃与${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}.{}^{{29} - {31}}$的加成反应。另外,钯催化的有机卤化物与二锡化合物的锡化反应是合成锡烷的可靠途径(图1a)。${}^{{32},{33}}$ 然而,这些方法仅对伯苄基卤化物有效,对一般的烷基卤化物则无效。
Although some stannylation reactions have been established, it is still highly desirable to explore transition metal free approaches to access these synthetically valued building blocks. In 2017, Wang disclosed a stannylation of N-tosylhydrazones with ${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}$ under transition metal free conditions. ${}^{34}$ This protocol is mainly suitable for synthesizing benzylstannanes. On the contrary, the transition metal free alkyl borylation process has seen great progress in the past five years. ${}^{{35},{36}}$ On the basis of our recent synthetic investigations of transition metal free borylation, ${}^{37}$ a program of study was undertaken to structurally, kinetically, and computationally characterize transition metal free stannylation of a wide range of alkyl halides (Figure 1b).
尽管已经建立了一些锡化反应,但探索无过渡金属的方法来获得这些具有合成价值的结构单元仍然非常必要。2017年,Wang公开了在无过渡金属条件下用${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}$对N - 甲苯磺酰腙进行锡化反应。${}^{34}$ 该方法主要适用于合成苄基锡烷。相反,无过渡金属的烷基硼化过程在过去五年中取得了很大进展。${}^{{35},{36}}$ 基于我们最近对无过渡金属硼化反应的合成研究,${}^{37}$ 开展了一项研究计划,从结构、动力学和计算方面对多种烷基卤化物的无过渡金属锡化反应进行表征(图1b)。
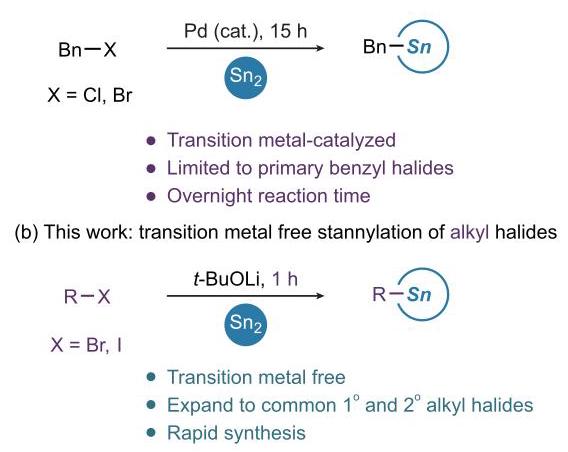
Figure 1. Stannylation of alkyl halides.
图1. 烷基卤化物的锡化反应。
2. RESULTS AND DISCUSSION
2. 结果与讨论
Development of the Reaction Conditions. At the outset, we have investigated this stannylation reaction of (3- bromopropyl)benzene SBr1 with hexamethyldistannane under transition metal free conditions (Table 1). To optimize the reaction conditions, we started by examining several representative bases (entries 1-3). Only the reaction using $t$ - BuOLi as a base afforded corresponding stannylated product 1 in 39% yield (entry 3). At the optimal reaction temperature, we found that a better yield was obtained at ${80}^{ \circ }\mathrm{C}$ (entry 4). The variation of an array of solvents indicated that DMSO as the solvent gave a yield slightly better than that with DMF (entries 6 and 7). Inspired by Jiao’s research, ${}^{38}$ as well as a suggested ${\mathrm{S}}_{\mathrm{N}}2$ mechanism (vide infra), the reaction with ${10}\mathrm{\;{mol}}\%$ DMAP added gave a superior result, providing 1 in 62% yield (entry 8). After the amount of DMAP had been optimized, the yield of 1 also increased to 71% upon addition of 0.8 equiv of DMAP (entries 8-10). Increasing the concentration did not further decrease the yield (entry 11). Furthermore, a series of differently substituted pyridines were evaluated as the additives, which proved to be less effective then DMAP (entries 12-14). Finally, the reaction with 1.4 equiv of ditin provided 1 in 92% yield (entry 15). A control experiment showed that $t$ -BuOLi was indispensable for this stannylation reaction (entry 16). For alkyl iodide substrates, we found that a prolonged reaction time at nearly room temperature is beneficial for this transformation.
反应条件的优化。首先,我们研究了在无过渡金属条件下,(3-溴丙基)苯SBr1与六甲基二锡的锡基化反应(表1)。为了优化反应条件,我们首先考察了几种代表性的碱(条目1-3)。只有使用$t$叔丁醇锂作为碱的反应以39%的产率得到了相应的锡基化产物1(条目3)。在最佳反应温度下,我们发现在${80}^{ \circ }\mathrm{C}$时产率更高(条目4)。一系列溶剂的变化表明,以二甲基亚砜为溶剂时产率略高于以N, N-二甲基甲酰胺为溶剂时(条目6和7)。受焦的研究启发,${}^{38}$以及一种推测的${\mathrm{S}}_{\mathrm{N}}2$机理(见下文),加入${10}\mathrm{\;{mol}}\%$ 4-(二甲氨基)吡啶的反应得到了更好的结果,以62%的产率得到了1(条目8)。在优化了4-(二甲氨基)吡啶的用量后,加入0.8当量的4-(二甲氨基)吡啶时,1的产率也提高到了71%(条目8-10)。提高浓度并没有进一步降低产率(条目11)。此外,评估了一系列不同取代的吡啶作为添加剂,结果证明它们的效果不如4-(二甲氨基)吡啶(条目12-14)。最后,与1.4当量的二锡反应以92%的产率得到了1(条目15)。对照实验表明,$t$叔丁醇锂对于该锡基化反应是必不可少的(条目16)。对于烷基碘底物,我们发现近室温下延长反应时间有利于这种转化。
Received: December 28, 2021
收到日期:2021年12月28日
Published: March 8, 2022
发表日期:2022年3月8日
Table 1. Optimization of the Reaction Conditions Using Alkyl Bromide SBr1 ${}^{a}$
表1. 使用烷基溴SBr1 ${}^{a}$对反应条件的优化
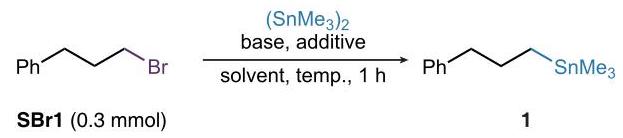
| entry | base (equiv) | additive (equiv) | solvent (mL) | temp (℃) | yield (%) |
| 1 | ${\mathrm{{Et}}}_{3}\mathrm{\;N}\left( {1.5}\right)$ | - | DMF (2) | 30 | 0 |
| 2 | $t$ -BuOK (1.5) | - | DMF (2) | 30 | 0 |
| 3 | $t$ -BuOLi (1.5) | - | DMF (2) | 30 | 39 |
| 4 | $t$ -BuOLi (1.5) | - | DMF (2) | 80 | 52 |
| 5 | $t$ -BuOLi (1.5) | - | DMF (2) | 110 | trace |
| 6 | $t$ -BuOLi (1.5) | - | DMSO (2) | 80 | 53 |
| 7 | $t$ -BuOLi (1.5) | - | MeCN (2) | 80 | trace |
| 8 | $t$ -BuOLi (1.5) | DMAP (0.1) | DMSO (2) | 80 | 62 |
| 9 | $t$ -BuOLi (1.5) | DMAP (0.8) | DMSO (2) | 80 | 71 |
| 10 | $t$ -BuOLi (1.5) | DMAP (1.0) | DMSO (2) | 80 | 60 |
| 11 | $t$ -BuOLi (1.5) | DMAP (0.8) | DMSO (1.5) | 80 | 71 |
| 12 | $t$ -BuOLi (1.5) | 4-methylpyridine (0.8) | DMSO (1.5) | 80 | 69 |
| 13 | $t$ -BuOLi (1.5) | 4-phenylpyridine (0.8) | DMSO (1.5) | 80 | 59 |
| 14 | $t$ -BuOLi (1.5) | 2,2'-bipyridine (0.8) | DMSO (1.5) | 80 | 55 |
| ${15}^{b}$ | $t$ -BuOLi (1.5) | DMAP (0.8) | DMSO (1.5) | 80 | 92 |
| 16 | - | DMAP (0.8) | DMSO (1.5) | 80 | 0 |
${}^{a}$ Reaction conditions: SBr1 (0.3 mmol), ${\mathrm{{Sn}}}_{2}{\mathrm{{Me}}}_{6}$ (0.36 mmol,1.2 equiv). DMAP [4-(dimethylamino)pyridine], DMF (N, N-dimethylformamide), DMSO (dimethyl sulfoxide). Isolated yields are reported. ${}^{b}{\mathrm{{Sn}}}_{2}{\mathrm{{Me}}}_{6}$ (0.42 mmol,1.4 equiv).
${}^{a}$反应条件:SBr1(0.3 mmol),${\mathrm{{Sn}}}_{2}{\mathrm{{Me}}}_{6}$(0.36 mmol,1.2当量)。4-(二甲氨基)吡啶(DMAP),N, N-二甲基甲酰胺(DMF),二甲基亚砜(DMSO)。报道的是分离产率。${}^{b}{\mathrm{{Sn}}}_{2}{\mathrm{{Me}}}_{6}$(0.42 mmol,1.4当量)。
With the optimized conditions in hand, we then surveyed the substrate scope of this stannylation reaction starting from a series of structurally diversified alkyl halides to obtain the corresponding alkylstannanes (Figure 2). We found that the stannylating efficiency of alkyl halides was greatly affected by the leaving properties of halogens. Phenyl propyl chloride, bromide, and iodide afforded corresponding products 1 in 23%, 92%, and 82% yields, respectively, under the reaction conditions. On this basis, we tried to use 3-phenylpropyl 4- methylbenzenesulfonate to obtain the corresponding alkyl-stannanes (1) in 59% yields. In view of the great importance of the heteroaromatic scaffold in pharmaceuticals, we synthesized trimethyl[3-(thiophen-2-yl)propyl]stannane 2 and 4-[3- (trimethylstannyl)propyl]pyridine 3 in good yields, which indicated that the heteroaromatic susbtrates were also suitable for this protocol. Continuously, a series of unactivated primary alkyl iodides/bromides were converted to the corresponding alkylstannanes in good to excellent yields (4-8). Highly volatile alkylstannanes 6 and 7 have been isolated in only discrete yields. Notably, the stannylation protocol demonstrated good functional group tolerance to various substituents, including methoxy (6), oxygen heterocycle (9), chloro (10), cyano (11), and alkenyl (12) groups. Notably, the ester group was hydrolyzed during the reaction to afford stannylative carboxylic acid derivative 13. To our delight, double stannylating product 14 starting from diiodoalkane has been obtained in 74% yield. Moreover, several alkylstannanes containing branched chains can be acquired in moderate to high yields (15 and 16). Alkylstannanes prepared from tetrahydro geranyl halides could be obtained in high yields (17). It is gratifying that this protocol has good applicability for both halides and pseudohalides. When using alkyl $p$ - toluenesulfonates as the substrates, moderate yields are afforded in some cases.
在掌握了优化条件之后,我们接着从一系列结构多样的卤代烃出发,考察了该锡基化反应的底物范围,以获得相应的烷基锡烷(图2)。我们发现,卤代烃的锡基化效率受卤素离去性质的影响很大。在反应条件下,苯丙基氯、溴和碘分别以23%、92%和82%的产率得到相应产物1。在此基础上,我们尝试使用3-苯丙基4-甲基苯磺酸酯,以59%的产率得到相应的烷基锡烷(1)。鉴于杂芳基骨架在药物中的重要性,我们以良好的产率合成了三甲基[3-(噻吩-2-基)丙基]锡烷2和4-[3-(三甲基锡基)丙基]吡啶3,这表明杂芳基底物也适用于该反应方案。接着,一系列未活化的伯烷基碘/溴以良好至优异的产率转化为相应的烷基锡烷(4-8)。高挥发性的烷基锡烷6和7仅以离散的产率分离得到。值得注意的是,该锡基化反应方案对包括甲氧基(6)、氧杂环(9)、氯(10)、氰基(11)和烯基(12)在内的各种取代基表现出良好的官能团耐受性。值得注意的是,酯基在反应过程中水解,得到锡基化羧酸衍生物13。令我们高兴的是,以二碘代烷为起始原料的双锡基化产物14的产率为74%。此外,几种含支链的烷基锡烷可以以中等至高的产率得到(15和16)。由四氢香叶基卤化物制备的烷基锡烷可以高产率得到(17)。令人欣慰的是,该反应方案对卤化物和拟卤化物均具有良好的适用性。当使用烷基$p$-甲苯磺酸酯作为底物时,在某些情况下能得到中等产率。
We then intentionally examine the functional group tolerance of the reaction. As exemplified by 18, 19, and 30, halogen atoms on the aromatic ring do not affect the reaction at all. Furthermore, we have modified the alkyl chain by using thiophenol, aniline, and phenol derivatives to obtain various alkyl halides. These heteroatom-containing alkylstannanes can be acquired in 44-85% yields (20-30). As expected, the stannylation has excellent functional group tolerance to both electron-withdrawing and electron-donating groups on the benzene ring, including phenyl (23), fluorine (24 and 30), trifluoromethyl (25), methyl sulfonate (26), acetaminomethyl (27), methylthio (28), and tert-butyl (29) groups. The stannylation reaction of natural product coniferyl alcohol derivatives was also successful, and the corresponding product 31 was isolated in 80% yield. The results for secondary halides are somewhat unsatisfactory (32 and 33), mainly due to their steric hindrance, in which corrsponding alkenes were observed as the elimination byproducts.
然后,我们特意考察了该反应的官能团耐受性。以18、19和30为例,芳环上的卤原子对反应完全没有影响。此外,我们通过使用硫酚、苯胺和苯酚衍生物对烷基链进行了修饰,以获得各种卤代烃。这些含杂原子的烷基锡烷可以以44-85%的产率得到(20-30)。正如预期的那样,该锡基化反应对苯环上的吸电子基团和供电子基团均具有优异 的官能团耐受性,包括苯基(23)、氟(24和30)、三氟甲基(25)、甲磺酸酯(26)、乙酰氨基甲基(27)、甲硫基(28)和叔丁基(29)基团。天然产物松柏醇衍生物的锡基化反应也很成功,相应产物31的分离产率为80%。仲卤化物的结果有些不尽人意(32和33),主要是由于其空间位阻,其中观察到相应的烯烃作为消除副产物。
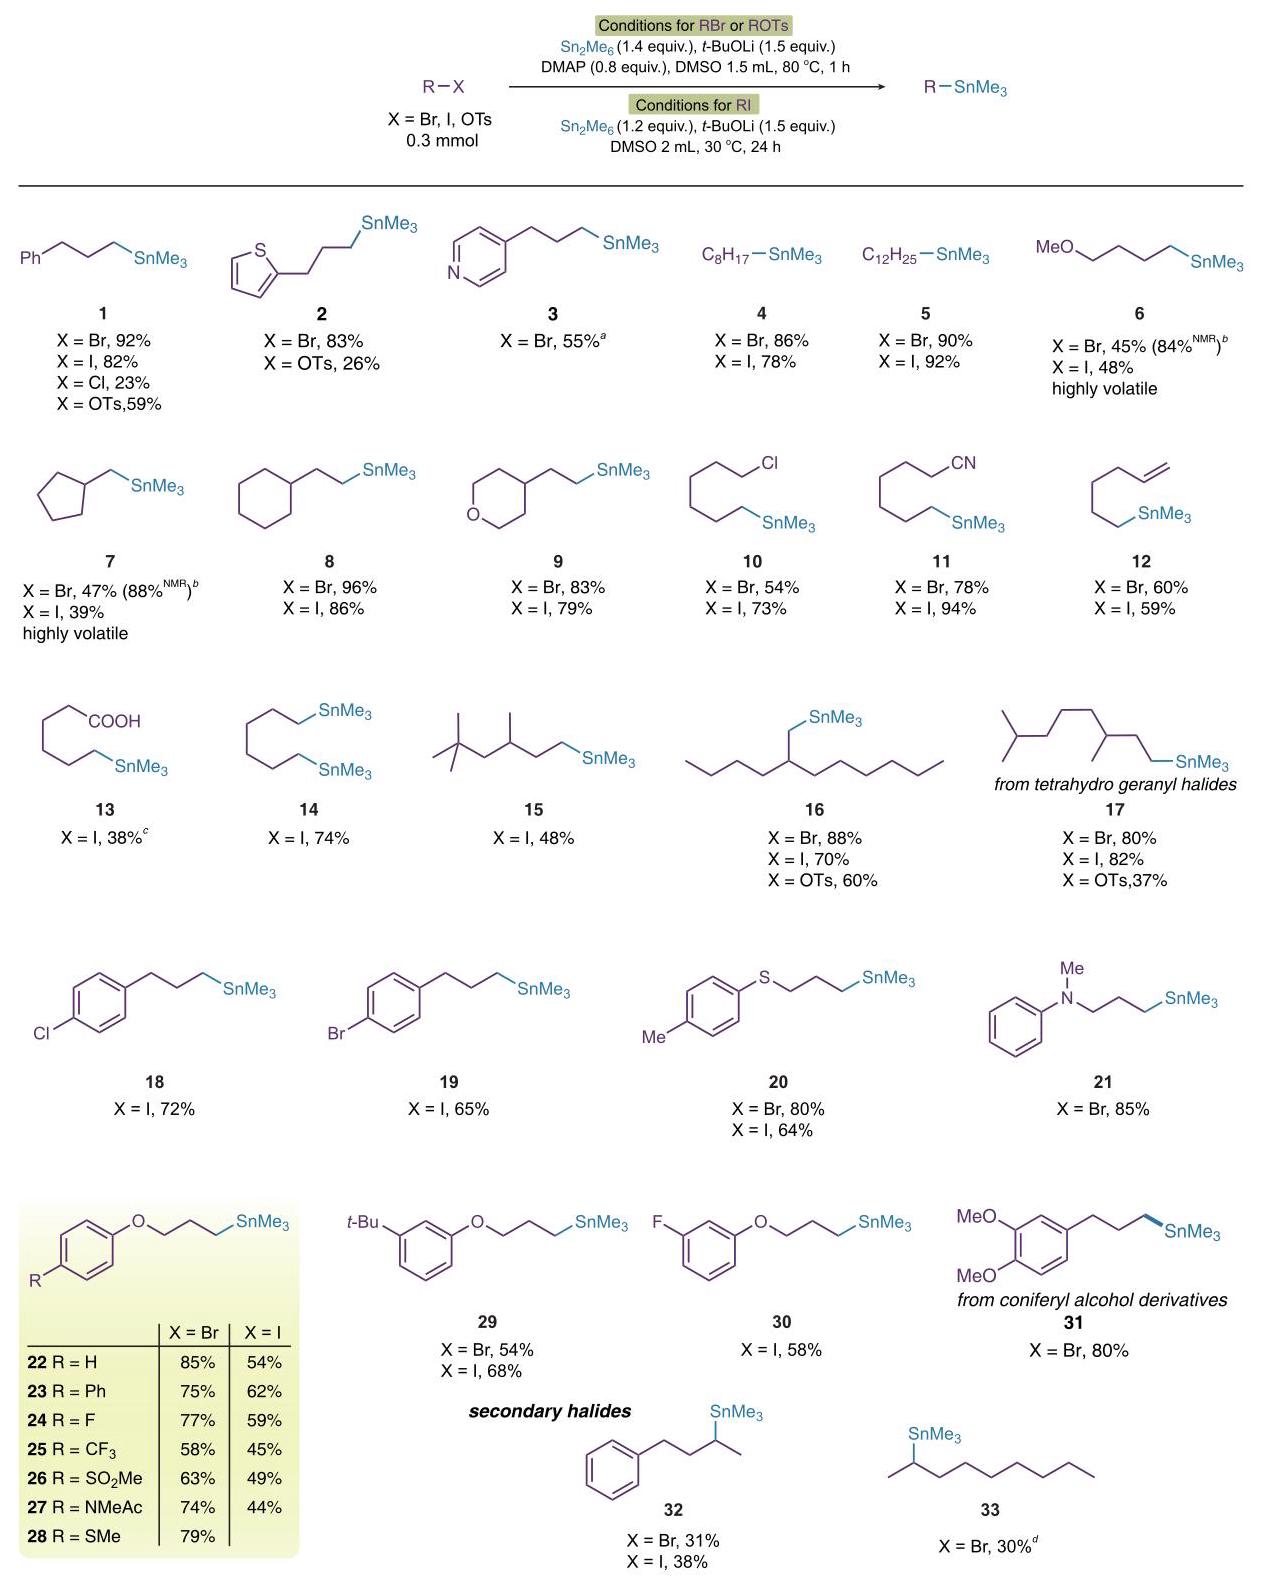
Figure 2. Substrate scope of alkyl halides. Reaction conditions for RBr or ROTs: alkyl bromide or alkyl p-toluenesulfonate (0.3 mmol, 1.0 equiv), hexamethyldistannane (1.4 equiv), t-BuOLi (1.5 equiv), DMAP (0.8 equiv), DMSO (1.5 mL), 80 ${}^{ \circ }\mathrm{C}$ , 1 h. Conditions for RI: alkyl iodide (0.3 mmol, 1.0 equiv), hexamethyldistannane (1.2 equiv), t-BuOLi (1.5 equiv), DMSO (2 mL), 30 ${}^{ \circ }\mathrm{C}$ , 24 h. Isolated yields are reported. ${}^{a}$ The reaction temperature was ${30}^{ \circ }\mathrm{C}$ . ${}^{b}\mathrm{{Te}}$ NMR yield was measured by using mesitylene as an internal standard. ${}^{\sigma }\mathrm{{Th}}$ is reaction was performed using methyl 6- iodohexanoate as the substrate. ${}^{d}$ The product was contaminated by (SnMe ${}_{3}$ ), which could not be removed by silica gel column chromatography. After silica gel column chromatography, product 33 mixed with ${\left( {\mathrm{{SnMe}}}_{3}\right) }_{2}$ was first isolated together, and then the yield of 33 was determined by ${}^{1}$ H NMR. DMAP = 4-(dimethylamino)pyridine. DMSO = dimethyl sulfoxide.
图2. 卤代烃的底物范围。RBr或ROTs的反应条件:卤代烃或对甲苯磺酸烷基酯(0.3 mmol,1.0当量),六甲基二锡烷(1.4当量),叔丁醇锂(1.5当量),4-(二甲氨基)吡啶(0.8当量),二甲基亚砜(1.5 mL),80 ${}^{ \circ }\mathrm{C}$ ,1小时。RI的反应条件:碘代烃(0.3 mmol,1.0当量),六甲基二锡烷(1.2当量),叔丁醇锂(1.5当量),二甲基亚砜(2 mL),30 ${}^{ \circ }\mathrm{C}$ ,24小时。报告的是分离产率。${}^{a}$ 反应温度为${30}^{ \circ }\mathrm{C}$ 。${}^{b}\mathrm{{Te}}$ 通过使用均三甲苯作为内标测量NMR产率。${}^{\sigma }\mathrm{{Th}}$ 该反应以6-碘己酸甲酯作为底物进行。${}^{d}$ 产物被(SnMe ${}_{3}$ )污染,无法通过硅胶柱色谱法除去。经过硅胶柱色谱法后,产物33与${\left( {\mathrm{{SnMe}}}_{3}\right) }_{2}$ 首先一起分离得到,然后通过${}^{1}$ H NMR测定33的产率。4-(二甲氨基)吡啶 = DMAP。二甲基亚砜 = DMSO。
Next, the substrate scope for benzylic bromides was investigated. Overall, various benzyl bromides were rapidly converted into the corresponding alkylstannanes in moderate to high yields at room temperature (Figure 3). A range of functional groups, such as methyl (35 and 42), tert-butyl (36), halogen (37-39, 43, and 45), boronate (40), trifluoromethoxy (41), methoxy (44), and naphthyl (46) groups, were tolerated in this protocol. By comparison, the diminished efficiency of secondary benzyl bromides was probably due to steric hindrance effects (47 and 48). It is worth mentioning that the stannylation reaction of natural product veratrol derivatives and geranyl halides was smoothly carried out to afford 49 and 50.
接下来,研究了苄基溴的底物范围。总体而言,各种苄基溴在室温下能快速转化为相应的烷基锡烷,产率适中至高(图3)。该反应体系能兼容一系列官能团,如甲基(35和42)、叔丁基(36)、卤素(37 - 39、43和45)、硼酸酯(40)、三氟甲氧基(41)、甲氧基(44)和萘基(46)。相比之下,仲苄基溴效率降低可能是由于空间位阻效应(47和48)。值得一提的是,天然产物藜芦醇衍生物和香叶基卤化物的锡化反应顺利进行,得到了49和50。
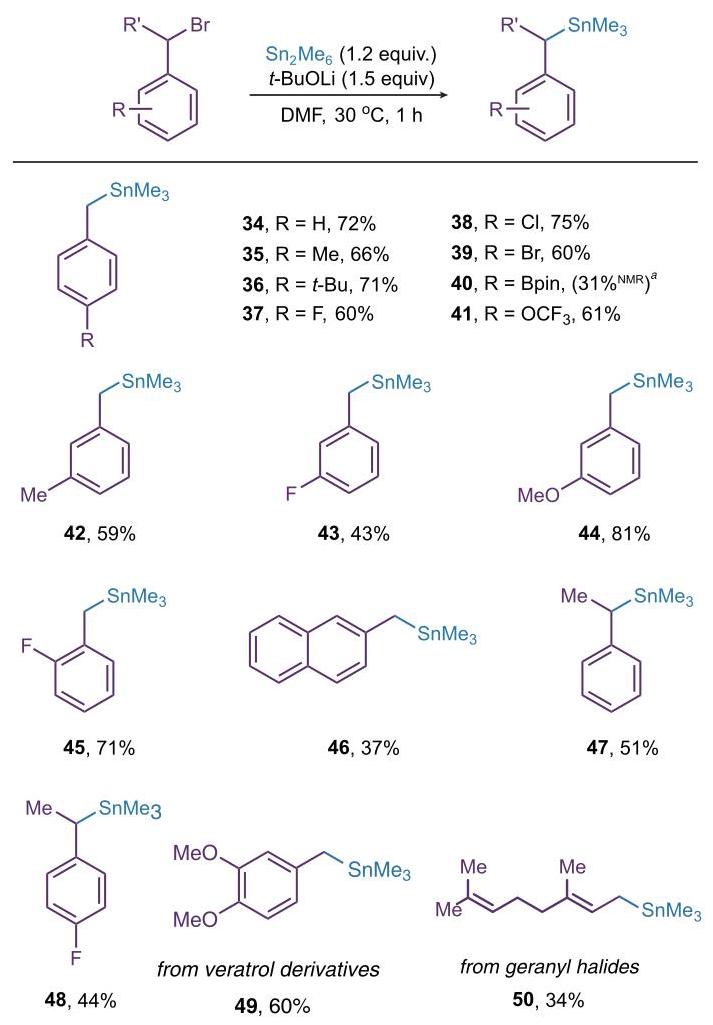
Figure 3. Substrate scope of benzyl bromides. Reaction conditions: benzyl bromide (0.3 mmol, 1.0 equiv), hexamethyldistannane (1.2 equiv), t-BuOLi (1.5 equiv), DMF (2 mL), 30 ${}^{ \circ }\mathrm{C}$ , 1 h. Isolated yields are reported. ${}^{a}$ The NMR yield was measured using mesitylene as an internal standard.
图3. 苄基溴的底物范围。反应条件:苄基溴(0.3 mmol,1.0当量),六甲基二锡烷(1.2当量),叔丁醇锂(1.5当量),N, N - 二甲基甲酰胺(2 mL),30 ${}^{ \circ }\mathrm{C}$ ,1小时。报告的是分离产率。${}^{a}$ 通过使用均三甲苯作为内标测量NMR产率。
To demonstrate the practical application of this stannylation procedure, mild reaction condition-promoted stannylations of three alkyl halides were carried out on a gram scale to form the corresponding alkylstannanes 1,11, and ${34}\left( { \leq {1.37}\mathrm{\;g}}\right)$ (Figure 4). Although the yield of 11 is moderate, the gram-scale yields of phenylpropylstannane (1) and benzylstannane (34) are comparable to those of the 0.3 mmol-scale protocol.
为了证明这种锡化反应方法的实际应用,对三种卤代烃进行了温和反应条件促进的克级规模锡化反应,以形成相应的烷基锡烷1、11和${34}\left( { \leq {1.37}\mathrm{\;g}}\right)$ (图4)。尽管11的产率适中,但苯丙基锡烷(1)和苄基锡烷(34)的克级产率与0.3 mmol规模的反应体系相当。
In the next research, we performed application studies and mechanistic investigations (Figure 5). Although the Stille coupling reactions of benzylstannanes have rarely been documented, the Pd-catalyzed Stille reaction of 34 with bromobenzene is successfully achieved to provide biaryl derivative 51 in 63% yield (Figure 5a). As for the mechanism, there are many precedents through the process of tin-based radicals (especially trialkyltin radicals) in ${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}$ -mediated transformation. ${}^{{39} - {42}}$ We therefore started looking for evidence of free radicals at first. A radical clock reaction was designed and performed with 6-iodohex-1-ene SI12 as the substrate. The result is that hex-5-en-1-yltrimethylstannane 12 was obtained as the only stannylated product, and cyclized stannylation compound ${12}^{\prime }$ was not detected at all (Figure 5b). Then, upon addition of a different amount of TEMPO or 1,1-diphenylethylene as the radical scavenger to this reaction mixture, no alkyl radical adduct was observed. In fact, the yields of 1 are not affected too much (Figure 5c). With the experimental evidence presented above, we think this stannalation might not involve radical processes. Additional experiments related to this part are described in the Supporting Information. Via a more detailed examination of the kinetics, we found that the reaction was many times faster than we expected at the beginning. The stannylation of alkyl bromides at ${80}^{ \circ }\mathrm{C}$ reached completion in $< {10}\mathrm{\;{min}}$ [10.1 (see the Supporting Information)].
在接下来的研究中,我们进行了应用研究和机理研究(图5)。尽管苄基锡烷的Stille偶联反应鲜有文献报道,但34与溴苯的钯催化Stille反应成功实现,以63%的产率得到联芳基衍生物51(图5a)。至于反应机理,在${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}$介导的转化过程中,有许多基于锡自由基(特别是三烷基锡自由基)的先例。${}^{{39} - {42}}$因此,我们首先开始寻找自由基的证据。设计并进行了以6-碘己-1-烯SI12为底物的自由基钟反应。结果是,仅得到己-5-烯-1-基三甲基锡烷12作为唯一的锡化产物,完全未检测到环化锡化化合物${12}^{\prime }$(图5b)。然后,向该反应混合物中加入不同量的TEMPO或1,1-二苯乙烯作为自由基清除剂,未观察到烷基自由基加合物。事实上,1的产率受影响不大(图5c)。基于上述实验证据,我们认为该锡化反应可能不涉及自由基过程。与这部分相关的其他实验在支持信息中进行了描述。通过对动力学更详细的研究,我们发现反应比我们最初预期的快很多倍。在${80}^{ \circ }\mathrm{C}$下,烷基溴的锡化反应在$< {10}\mathrm{\;{min}}$内完成[10.1(见支持信息)]。
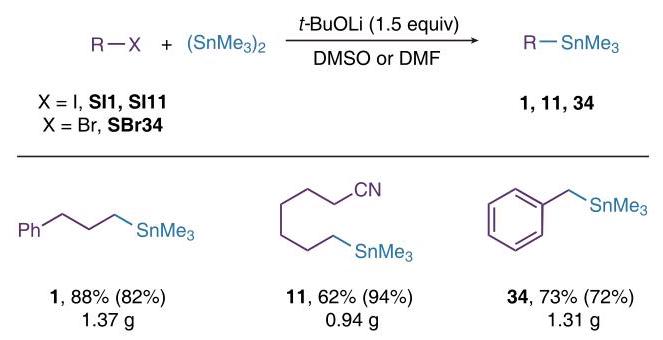
Figure 4. Gram-scale stannylation reactions. Reaction conditions: alkyl iodides SI1, SI11 (5.5 mmol, 1.0 equiv), hexamethyldistannane (1.1 equiv), t-BuOLi (1.5 equiv), DMSO (30 mL), 30 ${}^{ \circ }\mathrm{C}$ , 24 h; or alkyl bromides SBr34 (7 mmol, 1.0 equiv), hexamethyldistannane (1.05 equiv), t-BuOLi (1.5 equiv), DMF (30 mL), 30 ${}^{ \circ }\mathrm{C},4\mathrm{\;h},{\mathrm{\;N}}_{2}$ atmosphere. Isolated yield after column chromatography. The yields in parentheses refer to the reaction performed on a ${0.3}\mathrm{{mmol}}$ scale.
图4. 克级锡化反应。反应条件:烷基碘化物SI1、SI11(5.5 mmol,1.0当量),六甲基二锡烷(1.1当量),叔丁醇锂(1.5当量),二甲基亚砜(30 mL),30 ${}^{ \circ }\mathrm{C}$ ,24小时;或烷基溴化物SBr34(7 mmol,1.0当量),六甲基二锡烷(1.05当量),叔丁醇锂(1.5当量),N, N-二甲基甲酰胺(30 mL),30 ${}^{ \circ }\mathrm{C},4\mathrm{\;h},{\mathrm{\;N}}_{2}$ 气氛。柱色谱分离后的产率。括号内的产率指在${0.3}\mathrm{{mmol}}$规模上进行的反应。
(a) Pd-catalyzed Stille coupling reaction of benzylstannane with bromobenzene
(a) 苄基锡烷与溴苯的钯催化Stille偶联反应
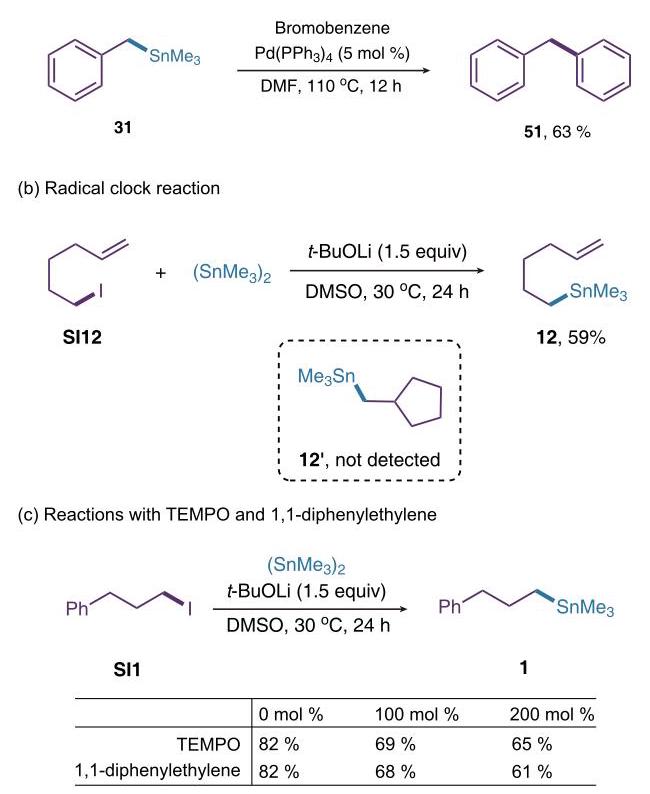
Figure 5. Derivatization of the stannylated substrate and mechanistic experiments.
图5. 锡化底物的衍生化及机理实验。
Because it is not a free radical mechanism, the nucleophilic substitution reaction is the next reasonable assumption. With the help of density functional theory (DFT) calculations, we have restored a clear and detailed reaction pathway for this stannylation reaction (Figure 6). This indicated that the heterolysis of the Sn-Sn bond was the key step for the whole transformation. Initially, we assumed the ${\mathrm{{Li}}}^{ + }$ ion would react with hexamethyldistannane to generate a ${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ + }$ ion or a $t$ - ${\mathrm{{BuO}}}^{ - }$ ion would react with hexamethyldistannane to generate a ${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ - }$ ion, but we could not find a reasonable intermediate structure of these two transformations. Then we found the Sn-Sn bond heterolysis of hexamethyldistannane was a two-step process. First, the ${\mathrm{{Li}}}^{ + }$ ion will bond with hexamethyldis-tannane to afford INT1 with a slight energy decrease (-2.47 kcal/mol). Charge distribution analysis of INT1 using an atomic dipole moment-corrected Hirshfeld population (ADCH) method ${}^{43}$ showed the charge of two Sn atoms of INT1 was slightly different (see Scheme S1). We proposed the bonding of Li and Sn will weaken the Sn-Sn bond. Second, the $t - {\mathrm{{BuO}}}^{ - }$ ion reacted with INT1 to generate INT2 and INT3 via TS1. The transformation is favored both thermodynamically (the Gibbs free energy change is -22.44 kcal/mol) and kinetically (the activation energy is 20.96 kcal/ mol). The structure and charge analysis of TS1 indicated the Sn-Sn bond is stretched to 3.46 Å and the charge of two Sn atoms is significantly different (see Scheme S2). This showed that the $\mathrm{{Sn}} - \mathrm{{Sn}}$ bond is nearly broken. The dissociation of INT3 will afford the ${\mathrm{{SnMe}}}_{3}{}^{ - }$ ion. INT3 is a widely used nucleophile that will generate the ${\mathrm{{SnMe}}}_{3}{}^{ - }$ ion. Subsequently, an ${\mathrm{S}}_{\mathrm{N}}2$ reaction of ${\mathrm{{SnMe}}}_{3}{}^{ - }$ and halide takes place to give the desired product. The Gibbs free energy change is $- {35.36}\mathrm{{kcal}}/$ mol, and the activation energy is 17.18 kcal/mol. IRC analysis of TS2 also shows the ${\mathrm{S}}_{\mathrm{N}}2$ reaction characteristics (see Scheme S3). Finally, ${\mathrm{I}}^{ - }$ would be trapped by ${\mathrm{{Li}}}^{ + }$ to form LiI.
由于这不是一个自由基机理,亲核取代反应是下一个合理的假设。借助密度泛函理论(DFT)计算,我们还原了该锡基化反应清晰而详细的反应路径(图6)。这表明Sn-Sn键的异裂是整个转化的关键步骤。最初,我们假设${\mathrm{{Li}}}^{ + }$离子会与六甲基二锡烷反应生成${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ + }$离子,或者$t$ - ${\mathrm{{BuO}}}^{ - }$离子会与六甲基二锡烷反应生成${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ - }$离子,但我们找不到这两种转化的合理中间体结构。然后我们发现六甲基二锡烷的Sn-Sn键异裂是一个两步过程。首先,${\mathrm{{Li}}}^{ + }$离子会与六甲基二锡烷结合生成INT1,能量略有降低(-2.47 kcal/mol)。使用原子偶极矩校正的Hirshfeld布居(ADCH)方法${}^{43}$对INT1进行电荷分布分析表明,INT1的两个Sn原子电荷略有不同(见方案S1)。我们认为Li与Sn的键合会削弱Sn-Sn键。其次,$t - {\mathrm{{BuO}}}^{ - }$离子与INT1反应,通过TS1生成INT2和INT3。该转化在热力学上是有利的(吉布斯自由能变化为-22.44 kcal/mol),在动力学上也是有利的(活化能为20.96 kcal/mol)。TS1的结构和电荷分析表明,Sn-Sn键被拉伸至3.46 Å,两个Sn原子的电荷有显著差异(见方案S2)。这表明$\mathrm{{Sn}} - \mathrm{{Sn}}$键几乎断裂。INT3的解离将产生${\mathrm{{SnMe}}}_{3}{}^{ - }$离子。INT3是一种广泛使用的亲核试剂,会生成${\mathrm{{SnMe}}}_{3}{}^{ - }$离子。随后,${\mathrm{{SnMe}}}_{3}{}^{ - }$与卤化物发生${\mathrm{S}}_{\mathrm{N}}2$反应,得到所需产物。吉布斯自由能变化为$- {35.36}\mathrm{{kcal}}/$ mol,活化能为17.18 kcal/mol。对TS2的IRC分析也显示了${\mathrm{S}}_{\mathrm{N}}2$反应特征(见方案S3)。最后,${\mathrm{I}}^{ - }$会被${\mathrm{{Li}}}^{ + }$捕获形成LiI。
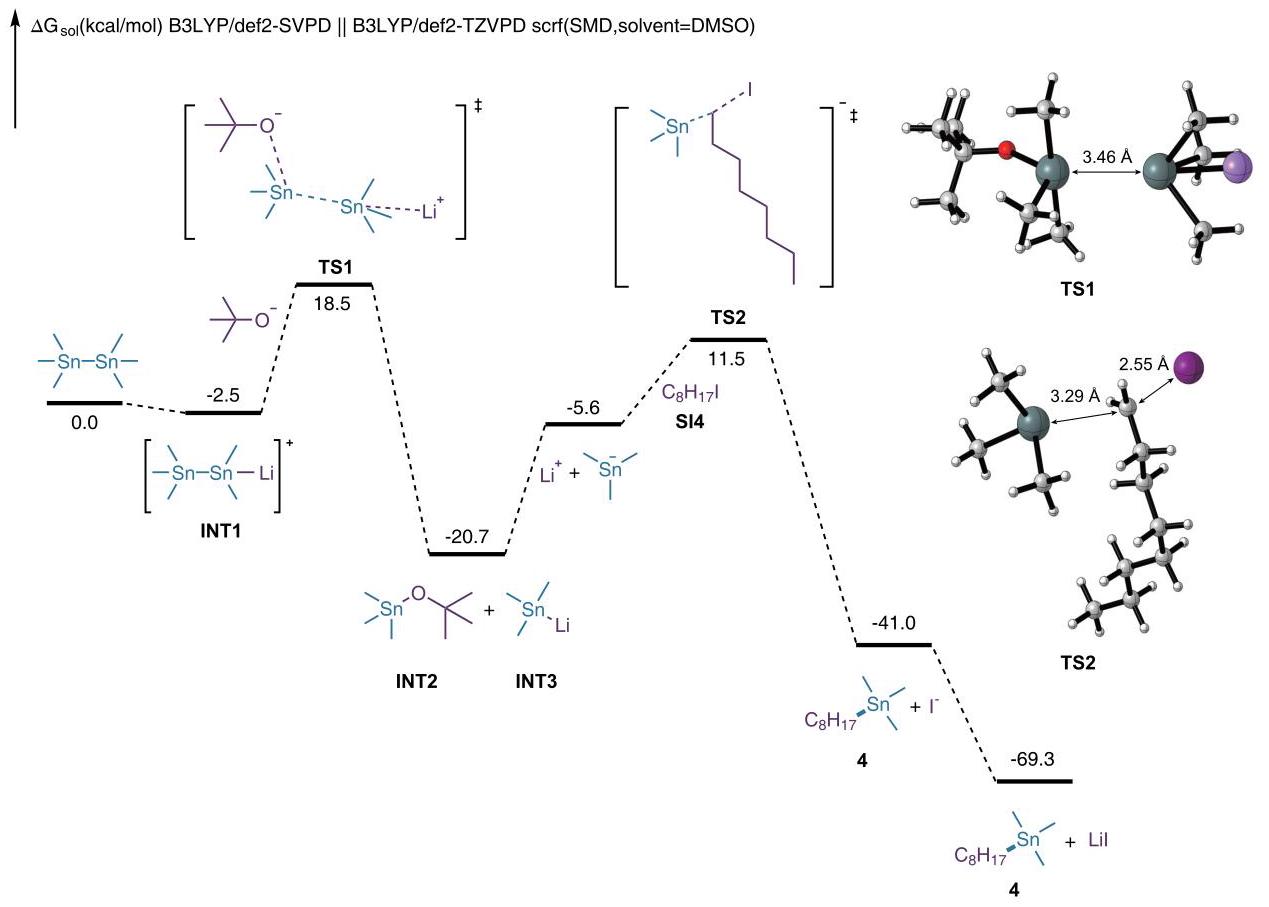
Figure 6. DFT calculations of the transition metal free stannylation reaction.
图6. 无过渡金属锡基化反应的DFT计算。
On the basis of the DFT calculation, we proposed the following mechanism (Figure 7). At first, the Sn-Sn bond of hexamethyldistannane is broken heterolytically by ${\mathrm{{Li}}}^{ + }$ and $t$ - ${\mathrm{{BuO}}}^{ - }$ to generate ${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ - }$ and ${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ + }$ simultaneously. The subsequent ${\mathrm{S}}_{\mathrm{N}}2$ reaction takes place to afford the desired product. In addition, compared with the stannylation of alkyl bromides, a small amount of DMAP is added. After a previous investigation, ${}^{38}$ we thought that the amino group in DMAP has a strong electron donating effect, which makes the pyridine nitrogen atom more nucleophilic, which can interact with the alkyl bromide involved in the reaction, thereby promoting cleavage of the carbon-bromide bond, and then complete the stannylation reaction efficiently.
基于DFT计算,我们提出了以下机理(图7)。首先,六甲基二锡烷的Sn-Sn键被${\mathrm{{Li}}}^{ + }$和$t$ - ${\mathrm{{BuO}}}^{ - }$异裂,同时生成${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ - }$和${\mathrm{{Me}}}_{3}{\mathrm{{Sn}}}^{ + }$。随后发生${\mathrm{S}}_{\mathrm{N}}2$反应,得到所需产物。此外,与烷基溴的锡基化反应相比,添加了少量的DMAP。经过之前的研究,${}^{38}$我们认为DMAP中的氨基具有很强的给电子效应,这使得吡啶氮原子更具亲核性,它可以与反应中涉及的烷基溴相互作用,从而促进碳-溴键的断裂,进而有效地完成锡基化反应。
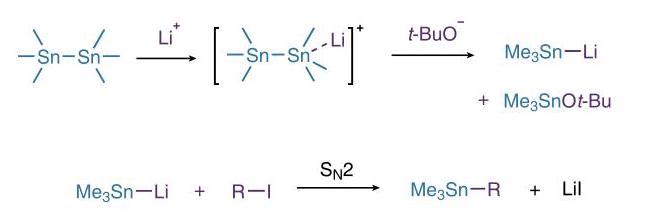
Figure 7. Proposed mechanism.
图7. 提出的机理。
3. CONCLUSIONS
3. 结论
In summary, we have disclosed a transition metal free conversion of alkyl bromides or iodides into the corresponding alkyltin compounds using hexamethyldistannane as the tin source. This reaction features a wide substrate scope, a simple, convenient, and rapid transformation, and a successful gram-scale synthesis. Preliminary experimental and theoretical investigations reveal that an in situ-formed trimethylstannyl anion might be involved during this ${\mathrm{S}}_{\mathrm{N}}2$ type of stannylation reaction. On the basis of the DFT calculations, the lithium cation is hypothesized to promote the heterolysis of the $\mathrm{{Sn}} - \mathrm{{Sn}}$ bond. This research has opened a new avenue toward the synthesis of alkylstannanes from commercially available alkyl halides and ditin compounds.
总之,我们已经公开了一种在无过渡金属条件下,使用六甲基二锡烷作为锡源,将烷基溴化物或碘化物转化为相应烷基锡化合物的方法。该反应具有底物范围广、转化简单、方便且快速,以及成功实现克级规模合成的特点。初步的实验和理论研究表明,在这种${\mathrm{S}}_{\mathrm{N}}2$类型的锡化反应过程中,可能涉及原位形成的三甲基锡阴离子。基于密度泛函理论计算,推测锂阳离子促进了$\mathrm{{Sn}} - \mathrm{{Sn}}$键的异裂。这项研究为从市售的烷基卤化物和二锡化合物合成烷基锡烷开辟了一条新途径。
ASSOCIATED CONTENT
相关内容
(c) Supporting Information
(c) 支持信息
The Supporting Information is available free of charge at https://pubs.acs.org/doi/10.1021/acs.joc.1c03135.
支持信息可在https://pubs.acs.org/doi/10.1021/acs.joc.1c03135免费获取。
General experimental methods, ${}^{1}\mathrm{H}$ NMR spectra, ${}^{13}\mathrm{C}$ NMR spetra, and coordinates and energies for the calculated structures (PDF)
一般实验方法、${}^{1}\mathrm{H}$核磁共振谱、${}^{13}\mathrm{C}$核磁共振谱以及计算结构的坐标和能量(PDF)
A UTHOR INFORMATION
作者信息
Corresponding Authors
通讯作者
Di Qiu - Tianjin Key Laboratory of Structure and Performance for Functional Molecules, College of Chemistry, Tianjin Normal University, Tianjin 300387, China;
邱迪 - 功能分子结构与性能天津市重点实验室,天津师范大学化学学院,天津 300387,中国;邮箱:
Fanyang Mo - School of Materials Science and Engineering, Peking University, Beijing 100871, China; (Corcid.org/ 0000-0002-4140-3020;
莫凡洋 - 北京大学材料科学与工程学院,北京 100871,中国;(Corcid.org/ 0000-0002-4140-3020;邮箱:
Authors
作者
Songyi Li - Tianjin Key Laboratory of Structure and Performance for Functional Molecules, College of Chemistry, Tianjin Normal University, Tianjin 300387, China
李松毅 - 功能分子结构与性能天津市重点实验室,天津师范大学化学学院,天津 300387,中国
Chang Lian - School of Materials Science and Engineering, Peking University, Beijing 100871, China
连畅 - 北京大学材料科学与工程学院,北京 100871,中国
Guanglu Yue - Tianjin Key Laboratory of Structure and Performance for Functional Molecules, College of Chemistry, Tianjin Normal University, Tianjin 300387, China
岳广禄 - 功能分子结构与性能天津市重点实验室,天津师范大学化学学院,天津 300387,中国
Jianning Zhang - School of Materials Science and Engineering, Peking University, Beijing 100871, China
张建宁 - 北京大学材料科学与工程学院,北京 100871,中国
Complete contact information is available at: https://pubs.acs.org/10.1021/acs.joc.1c03135
完整的联系信息可在:https://pubs.acs.org/10.1021/acs.joc.1c03135 查看
Author Contributions
作者贡献
${}^{\text{ § }}$ S.L. and C.L. contributed equally to this work.
${}^{\text{ § }}$ S.L.和C.L.对本工作贡献相同。
Notes
注释
The authors declare no competing financial interest.
作者声明不存在利益冲突。
ACKNOWLEDGMENTS
致谢
The authors thank the Science & Technology Development Fund of Tianjin Education Commission for Higher Education (2020KJ007) and the National Natural Science Foundation of China (Grants 22071004, 21933001 and 22150013) for financial support.
作者感谢天津市教育委员会高等教育科学技术发展基金(2020KJ007)和国家自然科学基金(项目编号22071004、21933001和22150013)的资金支持。
ERFERENCES
参考文献
(1) Neumann, W. P.; Moser, W. The organic chemistry of tin; Interscience: New York, 1970.
(2) Johansson Seechurn, C. C.; Kitching, M. O.; Colacot, T. J.; Snieckus, V. Palladium-catalyzed cross-coupling: a historical contextual perspective to the 2010 Nobel Prize. Angew. Chem., Int. Ed. 2012, 51, 5062-5085.
(3) Stille, J. K. The palladium-catalyzed cross-coupling reactions of organotin reagents with organic electrophiles [new synthetic methods (58)]. Angew. Chem., Int. Ed. 1986, 25, 508-524.
(4) Hassan, J.; Sévignon, M.; Gozzi, C.; Schulz, E.; Lemaire, M. Aryl-aryl bond formation one century after the discovery of the Ullmann reaction. Chem. Rev. 2002, 102, 1359-1470.
(5) Espinet, P.; Echavarren, A. M. The mechanisms of the Stille reaction. Angew. Chem., Int. Ed. 2004, 43, 4704-4734.
(6) Carsten, B.; He, F.; Son, H. J.; Xu, T.; Yu, L. Stille polycondensation for synthesis of functional materials. Chem. Rev. 2011, 111, 1493-1528.
(7) Murphy, A. R.; Liu, J.; Luscombe, C.; Kavulak, D.; Fréchet, J. M.; Kline, R. J.; McGehee, M. D. Synthesis, characterization, and field-effect transistor performance of carboxylate-functionalized polythio-phenes with increased air stability. Chem. Mater. 2005, 17, 4892- 4899.
(8) Kosugi, M.; Shimizu, Y.; Migita, T. Reaction of allyltin compounds: II. Facile preparation of allyl ketones via allyltins. J. Organomet. Chem. 1977, 129, C36-C38.
(9) Milstein, D.; Stille, J. K. A general, selective, and facile method for ketone synthesis from acid chlorides and organotin compounds catalyzed by palladium. J. Am. Chem. Soc. 1978, 100, 3636-3638.
(10) Kosugi, M.; Kato, Y.; Kiuchi, K.; Migita, T. Reaction of iodobenzene with trimethyltin cyanide in the presence of tetrakis (triphenylphosphine) palladium: New complex formation between the palladium and the cyanide. Chem. Lett. 1981, 10, 69-72.
(11) Kosugi, M.; Suzuki, M.; Hagiwara, I.; Goto, K.; Saitoh, K.; Migita, T. A new palladium catalyzed aromatic acetonylation by acetonyltributyltin. Chem. Lett. 1982, 11, 939-940.
(12) Labadie, J. W.; Stille, J. K. Mechanisms of the palladium-catalyzed couplings of acid chlorides with organotin reagents. J. Am. Chem. Soc. 1983, 105, 6129-6137.
(13) Falck, J.; Patel, P. K.; Bandyopadhyay, A. Stereospecific cross-coupling of $\alpha$ -(thiocarbamoyl) organostannanes with alkenyl, aryl, and heteroaryl iodides. J. Am. Chem. Soc. 2007, 129, 790-793.
(14) Li, L.; Wang, C.-Y.; Huang, R.; Biscoe, M. R. Stereoretentive Pd-catalysed Stille cross-coupling reactions of secondary alkyl azastannatranes and aryl halides. Nat. Chem. 2013, 5, 607-612.
(15) Andrianome, M.; Häberle, K.; Delmond, B. Allyl-and benzylstannanes, new reagents in terpenic synthesis. Tetrahedron 1989, 45, 1079-1088.
(16) Deline, J. E.; Miller, R. A facile preparation of benzyltrimethyl-stannanes and their utility in the synthesis of 1-benzyl-1,2- dihydroisoquinolines. Tetrahedron Lett. 1998, 39, 1721-1724.
(17) Baciocchi, E.; Ioele, M. Oxidation of benzyltrialkylstannanes by iodosylbenzene catalyzed by iron tetrakis(pentafluorophenyl)- porphyrin chloride. possible role of benzyloxy radicals. J. Org. Chem. 1995, 60, 5504-5508.
(18) Tipparaju, S. K.; Joyasawal, S.; Forrester, S.; Mulhearn, D. C.; Pegan, S.; Johnson, M. E.; Mesecar, A. D.; Kozikowski, A. P. Design and synthesis of 2-pyridones as novel inhibitors of the Bacillus anthracis enoyl-ACP reductase. Bioorg. Med. Chem. Lett. 2008, 18, 3565-3569.
(19) Cheng, Z.; Skouta, R.; Vazquez, H.; Widawsky, J. R.; Schneebeli, S.; Chen, W.; Hybertsen, M. S.; Breslow, R.; Venkataraman, L. In situ formation of highly conducting covalent Au-C contacts for single-molecule junctions. Nat. Nanotechnol. 2011, 6, 353-357.
(20) Davies, A. G.; Roberts, B. P.; Smith, J. M. Homolytic organometallic reactions. Part VIII. Kinetics of the homolytic reaction between N-halogenosuccinimides and tetra-alkyltins. J. Chem. Soc., Perkin Trans. 2 1972, 2, 2221-2224.
(21) Marton, D.; Russo, U.; Stivanello, D.; Tagliavini, G.; Ganis, P.; Valle, G. C. Preparation of benzylstannanes by zinc-mediated coupling of benzyl bromides with organotin derivatives. Physicochemical characterization and crystal structures. Organometallics 1996, 15, 1645-1650.
(22) Smith, G. F.; Kuivila, H. G.; Simon, R.; Sultan, L. Electron-transfer, halogen-metal exchange, and direct processes in formal nucleophilic substitutions on alkyl halides by trimethyltinsodium. J. Am. Chem. Soc. 1981, 103, 833-839.
(23) Lamandé-Langle, S.; Abarbri, M.; Thibonnet, J.; Duchêne, A. A novel mode of access to polyfunctional organotin compounds and their reactivity in Stille cross-coupling reaction. J. Organomet. Chem. 2009, 694, 2368-2374.
(24) Jiang, J.; Crabtree, R. H.; Brudvig, G. W. One-step trimethylstannylation of benzyl and alkyl halides. J. Org. Chem. 2016, 81, 9483-9488.
(25) Doyle, M. P.; Taunton, J.; Oon, S.-M.; Liu, M. T.; Soundararajan, N.; Platz, M. S.; Jackson, J. E. Reactivity and selectivity in intermolecular insertion reactions of chlorophenylcar-bene. Tetrahedron Lett. 1988, 29, 5863-5866.
(26) Jackson, J. E.; Soundararajan, N.; Platz, M. S.; Doyle, M. P.; Liu, M. T. Activation parameters for the reaction of phenylchloro carbene with pyridine, tri-n-butyltin hydride, and triethylsilane; evidence against the need to invoke reversibly formed complexes in the reaction of this carbene with olefins. Tetrahedron Lett. 1989, 30, 1335-1338.
(27) Andrey, O.; Landais, Y.; Planchenault, D.; Weber, V. Synthesis of $\alpha$ -(Alkoxysilyl) acetic esters. A route to 1,2 diols. Tetrahedron 1995, 51, 12083-12096.
(28) Liu, Z.; Li, Q.; Yang, Y.; Bi, X. Silver (I)-promoted insertion into X-H (X= Si, Sn, and Ge) bonds with N-nosylhydrazones. Chem. Commun. 2017, 53, 2503-2506.
(29) Leusink, A.; Noltes, J. G. Mechanism of the hydrostannation of acrylonitrile. Tetrahedron Lett. 1966, 7, 335-340.
(30) Fleming, I.; Urch, C. J. Stereospecific cyclopropane synthesis from $\gamma$ -stannyl alcohols. J. Organomet. Chem. 1985, 285, 173-191.
(31) Smith, N. D.; Mancuso, J.; Lautens, M. Metal-catalyzed hydrostannations. Chem. Rev. 2000, 100, 3257-3282.
(32) Azizian, H.; Eaborn, C.; Pidcock, A. Synthesis of organo-trialkylstannanes. The reaction between organic halides and hexaalkyldistannanes in the presence of palladium complexes. J. Organomet. Chem. 1981, 215, 49-58.
(33) Kashin, A. N.; Bumagina, I. G.; Bumagin, N. A.; Bakunin, V. N.; Beletskaya, I. P. The reactions of organometallic compounds, catalyzed by complexes of transition-metals 0.2 . the formation of tin-carbon bond in hexamethyldistannane reactions with organic halides, catalyzed by palladium complexes. Russ. J. Org. Chem. 1981, 17, 789-794.
(34) Qiu, D.; Wang, S.; Meng, H.; Tang, S.; Zhang, Y.; Wang, J. Synthesis of benzyltributylstannanes by the reaction of N-tosylhy-drazones with ${\mathrm{{Bu}}}_{3}\mathrm{{SnH}}$ . J. Org. Chem. 2017, 82, 624-632.
(35) Friese, F. W.; Studer, A. New avenues for C-B bond formation via radical intermediates. Chem. Sci. 2019, 10, 8503-8518.
(36) Liu, Q.; Zhang, L.; Mo, F. Organic borylation reactions via radical mechanism. Acta Chim. Sin. 2020, 78, 1297-1308.
(37) Liu, Q.; Hong, J.; Sun, B.; Bai, G.; Li, F.; Liu, G.; Yang, Y.; Mo, F. Transition-metal-free borylation of alkyl iodides via a radical mechanism. Org. Lett. 2019, 21, 6597-6602.
(38) Zhang, L.; Jiao, L. Visible-light-induced organocatalytic borylation of aryl chlorides. J. Am. Chem. Soc. 2019, 141, 9124-9128.
(39) Kuivila, H. G. Organotin hydrides and organic free radicals. Acc. Chem. Res. 1968, 1, 299-305.
(40) Barton, D. H. R.; Dowlatshahi, H. A.; Motherwell, W. B.; Villemin, D. A new radical decarboxylation reaction for the conversion of carboxylic acids into hydrocarbons. J. Chem. Soc., Chem. Commun. 1980, 732-733.
(41) Ingold, K. U.; Bowry, V. W. Why are organotin hydride reductions of organic halides so frequently retarded? kinetic studies, analyses, and a few remedies. J. Org. Chem. 2015, 80, 1321-1331.
(42) Juliá, F.; Constantin, T.; Leonori, D. Applications of halogen-atom transfer (XAT) for the generation of carbon radicals in synthetic photochemistry and photocatalysis. Chem. Rev. 2022, 122, 2292- 2352.
(43) Lu, T.; Chen, F. Atomic dipole moment corrected Hirshfeld population method. J. Theor. Comput. Chem. 2012, 11, 163-183.
CAS BIOFINDER DISCOVERY PLATFORM™
CAS生物发现者发现平台™
ELIMINATE DATA SILOS. FIND WHAT YOU NEED, WHEN YOU NEED IT.
消除数据孤岛。在您需要时找到您需要的东西。
A single platform for relevant, high-quality biological and toxicology research
一个用于相关高质量生物学和毒理学研究的单一平台
Streamline your R&D
简化您的研发流程
CAS