Photoinduced Decarbonylative Borylation of Alkyl Aldehyde through 4-Alkyl-1,4-Dihydropyridines
中文 / Bilingual · English version
ORGANIC CH
Check for updates
Cite this: Org. Chem. Front., 2024, 11, 1140
Received 22nd July 2023, Accepted 19th December 2023
DOI: 10.1039/d3qo01139a
DOI: 10.1039/d3qo01139a
rsc.li/frontiers-organic
rsc.li/frontiers-organic
Photoinduced decarbonylative borylation of alkyl aldehydes through 4-alkyl-1,4-dihydropyridinest
Chang Lian, ${}^{a, b}$ Jianning Zhang ${}^{a, b}$ and Fanyang Mo D $* {}^{a}$
We propose a novel approach for the facile decarbonylative borylation of aliphatic aldehydes. In this strategy, 1,4-dihydropyridines (DHPs) are synthesized in a single step from the corresponding aldehydes and subsequently reacted with bis(catecholato)diboron to form an alkyl C-B bond through a photoinduced process. Remarkably, this reaction can be conducted under mild conditions and demonstrates compatibility with a wide range of substrates. The experimental findings, in conjunction with density functional theory (DFT) calculations, reveal the involvement of an electron transfer process between the photocatalyst and DHPs. Notably, the appropriate redox properties of DHPs facilitate the generation of alkyl radicals under these conditions, eliminating the need for any external oxidant.
1 Introduction
Alkyl boronates possess significant importance in organic synthesis owing to their medicinal properties and versatile reactivities. The formation and manipulation of the C-B bond have been extensively explored in various fields, including pharmaceuticals, agrochemicals, and materials science. ${}^{1 - 5}$ Conventional approaches for the synthesis of boronic ester compounds predominantly rely on petrochemical-based starting materials such as olefins, alkyl halides (via organolithiums and Grignard reagents), and alkanes. ${}^{1}$ Nonetheless, these traditional methods, although widely employed, suffer from drawbacks such as harsh reaction conditions, regioselectivity, and the use of hazardous reagents. Our pioneering work on radical borylation ${}^{6}$ contributed to a more versatile and environmentally friendly strategy for C-B bond construction, which demonstrates broad applicability, simplicity in operation, cleanliness, and compatibility with late-stage transformations. ${}^{7}$ The development of novel borylation methodologies has sparked considerable interest among researchers.
The strategy of transforming readily available and abundant chemical compounds, such as carboxylic acids, ${}^{8 - {13}}$ alcohols, ${}^{{14},{15}}$ and amines, ${}^{{16} - {18}}$ into valuable, structurally diversified alkyl boronates is highly appealing. In 2017, Aggarwal and colleagues reported decarboxylative borylation, wherein carboxylic acids were converted into $N$ -hydroxyphthalimide (NHPI) esters, ${}^{8}$ and in 2019, the deoxygenative borylation of aliphatic alcohols was achieved by preactivating alcohol groups to xanthates or oxalates, as demonstrated by Studer’s group (Fig. 1a). ${}^{14}$ Moreover, our previous work successfully accomplished a photoinduced ring-opening borylation reaction of hemiacetals by transforming them into optically active derivatives. ${}^{19}$ These borylation reactions proceed via radical mechanisms, with the intermediates involved (such as xanthates and NHPI esters) serving as effective precursors of alkyl radicals. The strategy of pre-activation using simple starting materials has generated our interest in exploring boryla-tion reactions initiated by other radical precursors.
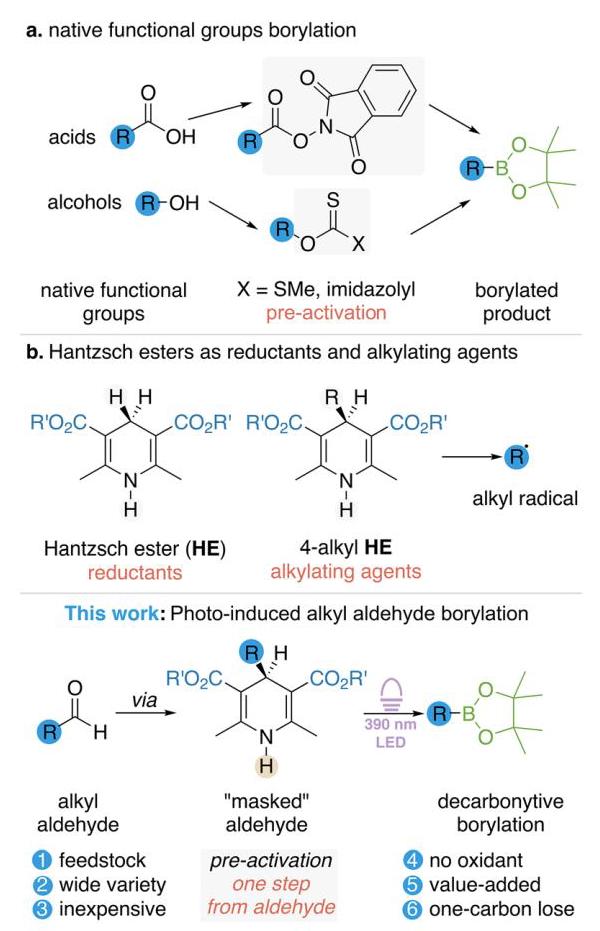
Fig. 1 Reaction design.
${}^{a}$ School of Materials Science and Engineering, Peking University, Beijing, PR China.
${}^{b}$ College of Engineering, Peking University, Beijing, PR China
† Electronic supplementary information (ESI) available. See DOI: https://doi.org/ 10.1039/d3qo01139a
Hantzsch esters are widely used organic reducing agents in a diverse range of organic transformations. ${}^{{20},{21}}$ The 4-substituted Hantzsch ester (4-alkyl-1,4-dihydropyridines), synthesized from aldehydes, serves as a valuable precursor for alkyl radicals under photochemical or electrochemical conditions in alkylation reactions (Fig. 1b). ${}^{22}$ There is no efficient method for the conversion of alkyl aldehydes to alkyl boro-nates. Building upon previous investigations on radical boryla-tion, we have chosen 4-alkyl-substituted Hantzsch esters as radical precursors and report a visible light-induced borylation of these esters (Fig. 1c).
2 Results and discussion
The well-tolerated radical precursors for borylation predominantly encompass oxidative radical precursors, such as alkyl halides or diazonium salts. ${}^{23}$ This implies that these precursors undergo an initial reduction step, leading to the formation of alkyl radicals. However, the challenge persists when attempting borylation with reductive radical precursors, such as alkanes, due to the limited availability of oxidants that exhibit compatibility with reductive diboron reagents. A commonly employed approach involves the conversion of reductive radical precursors into oxidative ones. For instance, the direct decarboxylation of alkyl carboxylic acids to generate alkyl radicals constitutes an oxidative reaction, known to be a formidable task. NHPI ester, synthesized from carboxylic acids, serves as an oxidative radical precursor, facilitating a more facile bor-ylation reaction.
The 4-substituted Hantzsch ester represents a category of reductive radical precursors that typically require the addition of oxidants to the reaction system in order to generate alkyl radicals. ${}^{22}$ This inherent necessity for oxidants often poses a challenge in the context of borylation reactions. However, it is noteworthy that we have observed the borylation of 4-substituted Hantzsch esters occurring smoothly and efficiently, even in the absence of added oxidants.
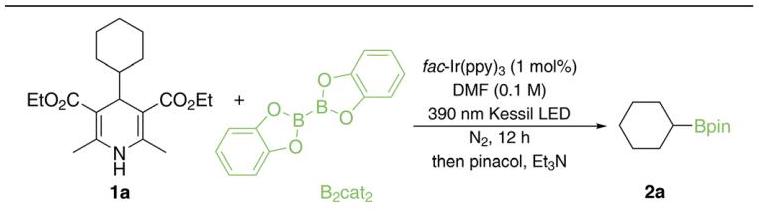
Our investigation of photocatalytic alkyl radical borylation commenced by subjecting diethyl 4-cyclohexyl-2,6-dimethyl- 1,4-dihydropyridine-3,5-dicarboxylate (1a) and bis(catecholato) diboron $\left( {{\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}}\right)$ to irradiation, and the desired borylation product 2a was obtained by transforming the initial catechol boronic ester with pinacol. The reaction was conducted using 1 mol% of fac-Ir(ppy) ${}_{3}$ as the photocatalyst in $N, N$ -dimethyl-formamide (DMF) under irradiation with a Kessil 390 nm LED lamp at room temperature for 12 hours. The obtained product exhibited a 91% GC-FID yield and an 84% isolated yield (Table 1, entry 1).
Table 1 The optimization of reaction conditions ${}^{a}$
| Entry | Deviations from standard condition | 2a ${}^{b}\left( \% \right)$ |
| 1 | None | 92% |
| 2 | DMF (0.16 M) | 75% |
| 3 | DMF (0.06 M) | 83% |
| 4 | DMAc instead of DMF | 86% |
| 5 | DCM instead of DMF | Trace |
| 6 | ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ (2 eq.) | 60% |
| 7 | ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ (5 eq.) | 92% |
| 8 | 2 h instead of overnight | 65% |
| 9 | ${\mathrm{B}}_{2}{\mathrm{{pin}}}_{2}$ instead of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ | N.D. |
| 10 | $\left\lbrack {\text{ Ir }{\left( \mathrm{{ppy}}\right) }_{2}\mathrm{{dtbbpy}}}\right\rbrack \left\lbrack {\mathrm{{PF}}}_{6}\right\rbrack$ | 67% |
| 11 | 456 nm light | 46% |
| 12 | No fac-Ir(ppy) | 9% |
| 13 | No light | N.D. |
${}^{a}$ Standard conditions: Hantzsch ester 1a ( ${0.3}\mathrm{{mmol}},{1.0}$ equiv.), ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ (1.2 mmol, 4.0 equiv.) and fac-Ir(ppy) ${}_{3}$ (1 mol%) in DMF (0.1 M, 3.0 mL), Kessil 390 nm LED lamp, ${\mathrm{N}}_{2}$ atmosphere, room temperature, 12 h. ${}^{b}$ GC-FID yields were given using $n$ -decane as internal standard. N.D., not detected.
During the exploration of reaction conditions, it was observed that maintaining the proper concentration of substrates resulted in a high yield of the product, while deviating from the standard value led to a slight decrease in yield (entry 2 and entry 3). Replacing DMF with $N, N$ -dimethylacetamide (DMAc) as the solvent provided a good yield of product 2a due to the similar properties between DMAc and DMF (entry 4). However, when DMF was replaced with another solvent such as dichloromethane (DCM), only a minimal quantity of product was observed (entry 5). This observation may be attributed to the Lewis base difference of DCM and DMF. Furthermore, the reduction of the boron reagent to 2 equivalents resulted in a significant decrease in yield, as depicted in entry 6. This observation can be attributed to the occurrence of side reactions involving ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ under light irradiation. ${}^{24}$ Increasing the amount of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ to 5 equivalents did not significantly enhance the yield (entry 7). Notably, the reaction proceeded rapidly in the first 2 hours, yielding 65% of the bor-ylation product (entry 8).
Alternative boron reagents such as bis(pinacolato)diboron $\left( {{\mathrm{B}}_{2}{\mathrm{{pin}}}_{2}}\right)$ did not exhibit reactivity in this reaction (entry 9). However, it is noteworthy that in this case, we observed the partial consumption of reagent 1a. One possible reason for the lack of borylation product formation is that the energy barrier of the reaction between ${\mathrm{B}}_{2}{\mathrm{{pin}}}_{2}$ and alkyl radicals is much higher than that of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ (see ESI Scheme S1†). Moreover, the commonly used photocatalyst $\left\lbrack {\mathrm{{Ir}}{\left( \mathrm{{ppy}}\right) }_{2}\mathrm{{dtbbpy}}}\right\rbrack \left\lbrack {\mathrm{{PF}}}_{6}\right\rbrack$ did not display comparable catalytic activity to fac-Ir(ppy) ${}_{3}$ (entry 10), which can be attributed to the differing redox abilities of these photocatalysts. Additionally, the use of a ${456}\mathrm{\;{nm}}$ LED light resulted in a lower yield of 46% compared to the ${390}\mathrm{\;{nm}}$ LED light (entry 11). Control experiments confirmed the essential roles of the photocatalyst and visible light irradiation in facilitating the borylation reaction (entries 12 and 13).
Subsequently, we investigated the substrate range using various Hantzsch esters (1) under the optimized conditions (Fig. 2). According to previous literature, the 4-alkyl-substituted Hantzsch esters can generate alkyl radicals under light irradiation. ${}^{21}$ We found that Hantzsch esters capable of forming secondary alkyl radicals were excellent substrates for this borylation reaction, resulting in the formation of the corresponding borylation products with good to excellent yields (2a-2g). It is worth mentioning that traditional methods often involve the addition of diboron reagents to double bonds, leading to the formation of 1,2-diboron compounds. ${}^{25}$ However, our borylation reaction, benefiting from its neutral reaction conditions, exhibited good tolerance towards alkenes (2h-2j). Additionally, our method displayed favorable diastereoselectivity, as demonstrated by the conversion of the norbornane DHP substrate into the titexo-borylation product 2k with moderate yield. Substrates containing benzene rings (2l-2n) were also amenable to the reaction, yielding boronates with yields ranging from 86% to 90%. Notably, functional groups such as the N-tert-butyloxycarbonyl group (20), hydroxyl (2p), fluorine (2q), and chlorine (2r) were well tolerated under the standard reaction conditions.
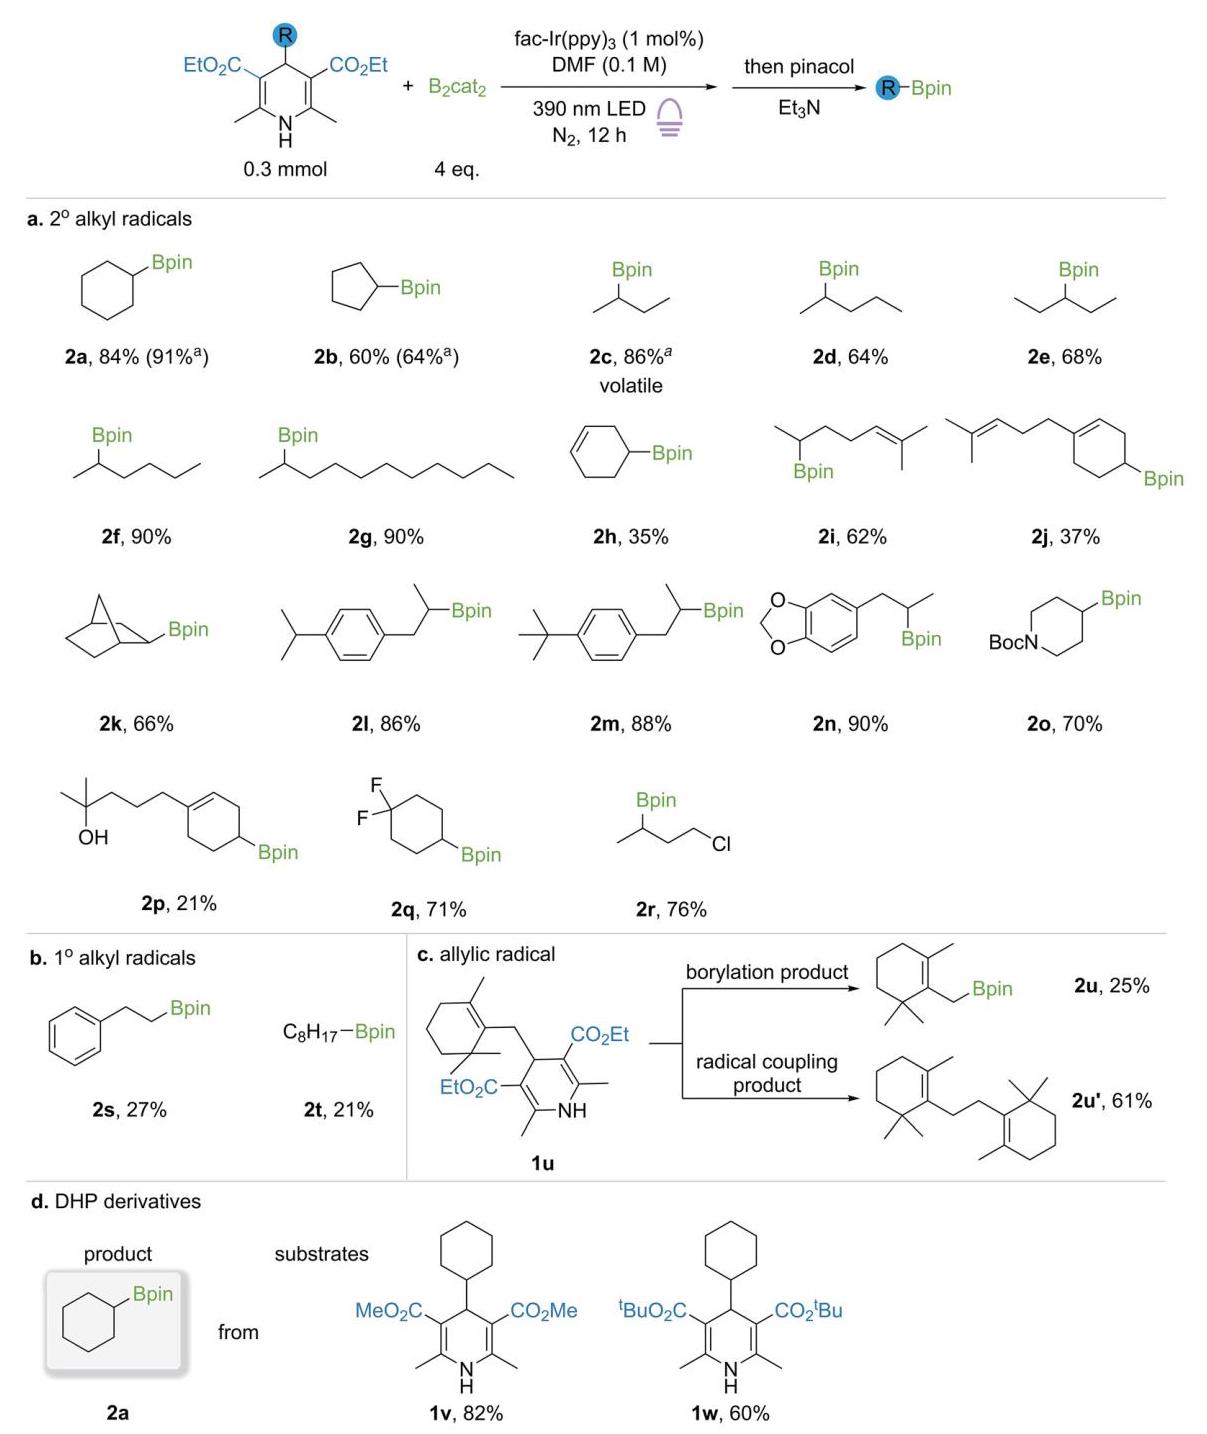
Fig. 2 Scope exploration of 4-alkyl-substituted Hantstzsch esters borylation reaction. Reaction conditions: Hantzsch ester 1 (0.3 mmol, 1.0 equiv.), ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ (1.2 mmol, 4.0 equiv.) and fac-lr(ppy) ${}_{3}$ (1 mol%) in DMF (0.1 M, 3.0 mL), Kessil 390 nm LED lamp, ${\mathrm{N}}_{2}$ atmosphere, room temperature. Isolated yield. ${}^{a}$ Yields were determined by GC-FID using $n$ -decane as internal standard.
In contrast to secondary alkyl radical precursors, primary alkyl radical precursors exhibited lower reactivity in this reaction. The borylation products of phenylpropyl and $n$ -octyl were obtained in low yields (2s, 2t). This disparity can be attributed to the relative stability of alkyl radicals, as secondary radicals are generally considered more stable than primary radicals. However, highly stable radicals are less favorable for boryla-tion. For instance, the highly stable allylic radical produced the borylation product $2\mathbf{u}$ with only a ${25}\%$ yield, while the radical coupling product $2{\mathbf{u}}^{\prime }$ was concurrently isolated in a 61% yield. We further explored the scope of Hantzsch ester derivatives (1v, 1w), and the results revealed that substituents with significant steric hindrance, such as tert-butyl, resulted in a notable decrease in borylation yield.
To demonstrate the utility of this borylation reaction, we explored various synthetic applications. Firstly, we conducted the large-scale synthesis of 2a, which yielded a satisfactory 65% yield (Fig. 3a). Furthermore, we investigated the pre- and post-conversion of Hantzsch ester substrates and borylation products. For instance, cyclohexylmethyl alcohols could be readily oxidized to cyclohexanecarboxaldehyde. By employing our proposed synthesis, we obtained the borylation product 2a, which could be further oxidized using hydrogen peroxide to yield cyclohexanol 3a. This entire reaction sequence exemplifies a “one-carbon-lose” strategy, where a primary alcohol is converted into a secondary alcohol (Fig. 3b).
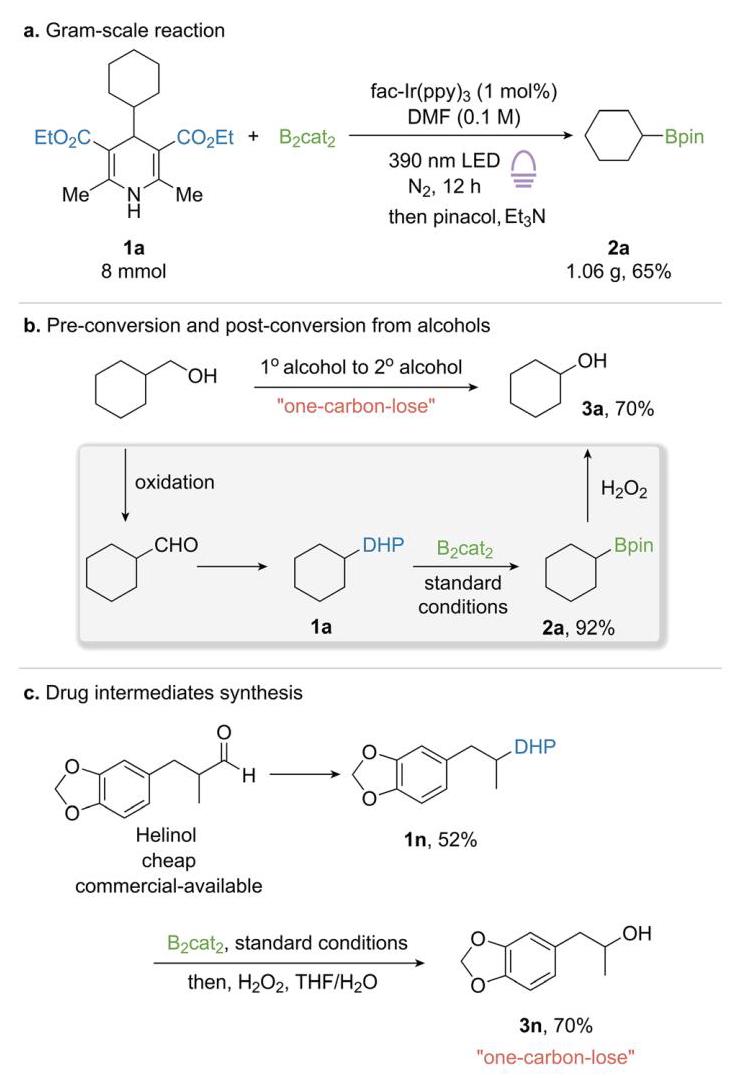
Fig. 3 Synthetic applications.
Additionally, the versatility of aldehydes in synthetic methodologies allowed for the synthesis of intriguing borylation products. For instance, acids and esters could be selectively reduced to aldehydes. We successfully accomplished the transformation of esters to the corresponding borylation product $2\mathbf{q}$ by first reducing the ester to the aldehyde (see ESI Scheme S2a†). This approach provides an alternative for bory-lation of acid derivatives. Furthermore, the reduction of lactones using diisobutylaluminum hydride (DIBAL-H) led to ring-opening aldehyde products. For example, treatment of 3-methyltetrahydro-2-furanone with DIBAL-H yielded 4-hydroxy-2-methylbutyraldehyde, which contains both hydroxyl and aldehyde groups. The transformation from aldehyde to boron ester and from hydroxyl to other functional groups, such as halogens, can be achieved as demonstrated in the case of $2\mathbf{r}$ (Scheme S2b†). This strategy allows for the synthesis of boron esters with multiple functional groups, which have been scarcely reported in the literature.
Moreover, 3n serves as an alcohol intermediate in drug synthesis and is typically obtained via a three-step synthetic route from the costly starting material piperonal. ${}^{26}$ However, by following the procedures outlined in this work (Fig. 3c), we successfully obtained the alcohol $3\mathbf{n}$ from $1\mathbf{n}$ , which can be readily prepared in a single step using the inexpensive and commercially available compound helinol. These transformations illustrate the potential applications of our borylation reaction.
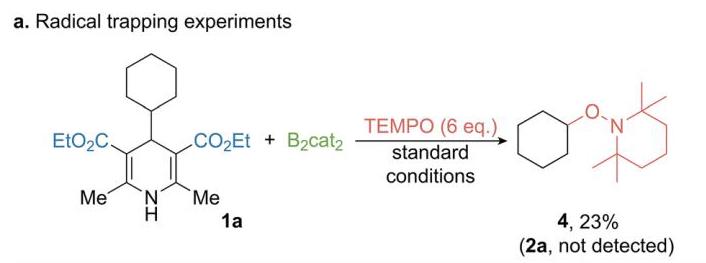
To elucidate the mechanism of this borylation reaction, additional investigations were conducted, as depicted in Fig. 4. Previous literature on the photochemical transformations of Hantzsch esters has demonstrated that alkyl radicals can be generated under light irradiation conditions. ${}^{21}$ The formation of the coupling product $2{\mathbf{u}}^{\prime }$ (Fig. 2c) supports the involvement of a radical mechanism in this reaction. To confirm the radical mechanism, we introduced 6 equivalents of TEMPO as a radical scavenger into the standard reaction system. We observed that the borylation was completely blocked, and the TEMPO scavenged product 4 was isolated in a yield of 23% (Fig. 4a).
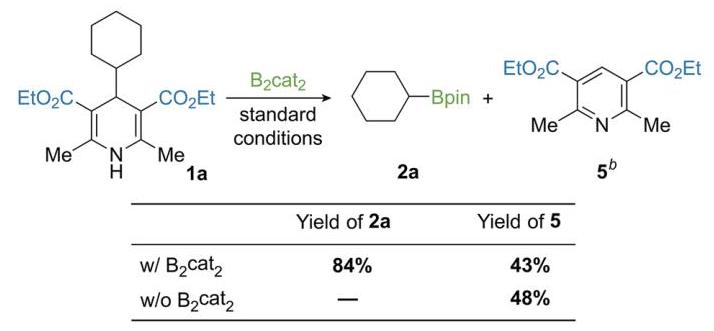
Furthermore, we isolated the by-product observed in this borylation reaction to aid in the analysis of the mechanism. Under the standard reaction conditions, the DHP group in the substrates was transformed into the pyridine product 5 , which could be easily recovered with a yield of 42% (Fig. 4b). A control experiment demonstrated that even without the addition of the boron source $\left( {{\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}}\right)$ , the pyridine product 5 could still be obtained with a yield of 50%.
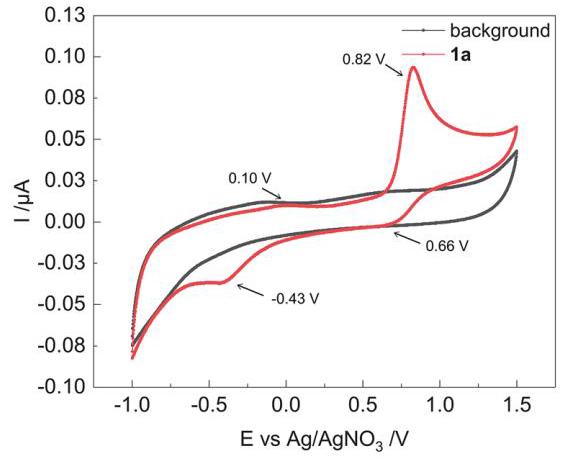
Moreover, cyclic voltammetry (CV) experiments were performed to determine the redox properties of 1a (Fig. 4c). The resulting CV curve exhibited two main redox peaks $\left( {{E}_{1/2} = }\right.$ -0.16 V and ${0.74}\mathrm{\;V}{vs}.\mathrm{{Ag}}/{\mathrm{{AgNO}}}_{3}$ reference electrode), and we attributed the wave around $- {0.16}\mathrm{\;V}$ to the oxidation of $1\mathrm{a}$ .
c. CV experiments ${}^{c}$
d. UV and FL absorption spectra ${}^{d}$
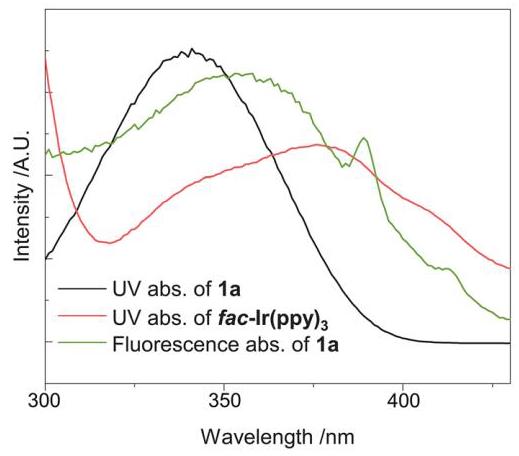
Fig. 4 Mechanism studies. ${}^{a}$ Control reaction conditions: Hantzsch ester 1a (0.3 mmol,1.0 equiv.) and fac-Ir(ppy) ${}_{3}$ (1 mol%) in DMF (0.1 M, 3.0 mL), Kessil 390 nm LED lamp, ${\mathrm{N}}_{2}$ atmosphere, room temperature. Isolated yield. ${}^{b}$ Yields were determined by GC-FID using $n$ -decane as internal standard. ${}^{c}$ Reaction conditions: solvent, MeCN; electrodes, platinum wire; reference electrode, $\mathrm{{Ag}}/{\mathrm{{AgNO}}}_{3}$ (0.01 M in $\mathrm{{MeCN}}$ ); scan rate ${0.3}\mathrm{\;V}{\mathrm{\;s}}^{-1}$ ; background, ${0.1}\mathrm{M}{\mathrm{{LiClO}}}_{4}$ in MeCN. ${}^{d}$ solvent, DMF.
Regarding the optical properties, we measured the UV-vis and fluorescence absorption spectra of substrate 1a and fac-Ir (ppy) ${}_{3}$ (Fig. 4d). The UV-vis spectra revealed that fac-Ir(ppy) ${}_{3}$ exhibited a maximum absorption wavelength around ${380}\mathrm{\;{nm}}$ , which closely corresponds to the wavelength of the lamp used in our experiments. The maximum UV-vis absorption wavelength of 1a was approximately ${340}\mathrm{\;{nm}}$ , but the fluorescence absorption spectra indicated that it also exhibited fluorescence absorption at ${390}\mathrm{\;{nm}}$ . Therefore, we proposed that under our reaction conditions, both $\mathbf{{1a}}$ and ${fac}$ - $\operatorname{Ir}{\left( \mathrm{{ppy}}\right) }_{3}$ could undergo excitation.
Based on the above observations, several intriguing questions regarding the mechanism arise in this study: (1) How does the oxidative process occur for the conversion of substrate 1a to product 5 in the absence of an external oxidant? (2) How does the photocatalyst fac-Ir(ppy) ${}_{3}$ facilitate the reaction? To address these questions, extensive investigations were conducted and will be discussed in the subsequent sections.
To analyze the reason behind the oxidation process, we conducted nuclear magnetic resonance (NMR) spectroscopy analysis on the reaction system to examine the structures of certain stable intermediates (Fig. 5). In this analysis, we replaced the solvent in the standard conditions with fully deuterated DMF and collected samples at regular intervals for NMR proton spectroscopy detection. Prior to the NMR analysis, we performed a reaction time control experiment to determine the optimal reaction duration (see ESI Scheme S3†). We observed that the borylation product could be obtained in a yield of 65% within 2 hours and 83% within 8 hours.
During NMR analysis, we observed several distinctive signals of compound 5. Specifically, signals corresponding to the methyl hydrogen at position 2 and the methylene group within the ester moiety gradually intensified as the reaction progressed, as depicted in the grey region in Fig. 5. However, it is worth noting that the signal of the hydrogen at position 4 (represented by the red dot) did not appear. Additionally, the hydrogen signals associated with the DHP group of reagent 1a were not entirely transformed into signals corresponding to product 5 (indicated by the blue and green dots) after approximately 8 hours, even though the borylation reaction was nearly completed (as illustrated in ESI Fig. S2†). This suggests that pyridine 5 may not be the sole product derived from the DHP group of $\mathbf{{1a}}$ ; there may be other products originating from the DHP group.
To investigate further, we conducted mass spectrometry analysis on the reaction system, which revealed the presence of a molecular ion peak corresponding to the dimerized product resulting from two DHP radicals (ESI Scheme S5†). Notably, the formation of this dimerized product from substrate 1a was a redox-neutral reaction.
Our previous findings have already substantiated that the yield of compound 5 is less than ${50}\%$ (as depicted in Fig. 4b). Remarkably, even in the absence of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ , the reagent $1\mathbf{a}$ can undergo transformation into compound 5. Consequently, we contend that the conversion of the DHP group assumes a pivotal role in the successful execution of these chemical reactions, rather than involving some boron-containing species. The DHP group exhibits a propensity to readily convert into its dimerized product, subsequently giving rise to pyridine 5, with other products. This mechanism obviates the need for the introduction of oxidants. Consequently, the diboron reagent can maintain its efficacy without requiring the addition of oxidants, thereby facilitating the accomplishment of the boryla-tion reaction. A similar mechanism has been proposed in previous studies. ${}^{27}$
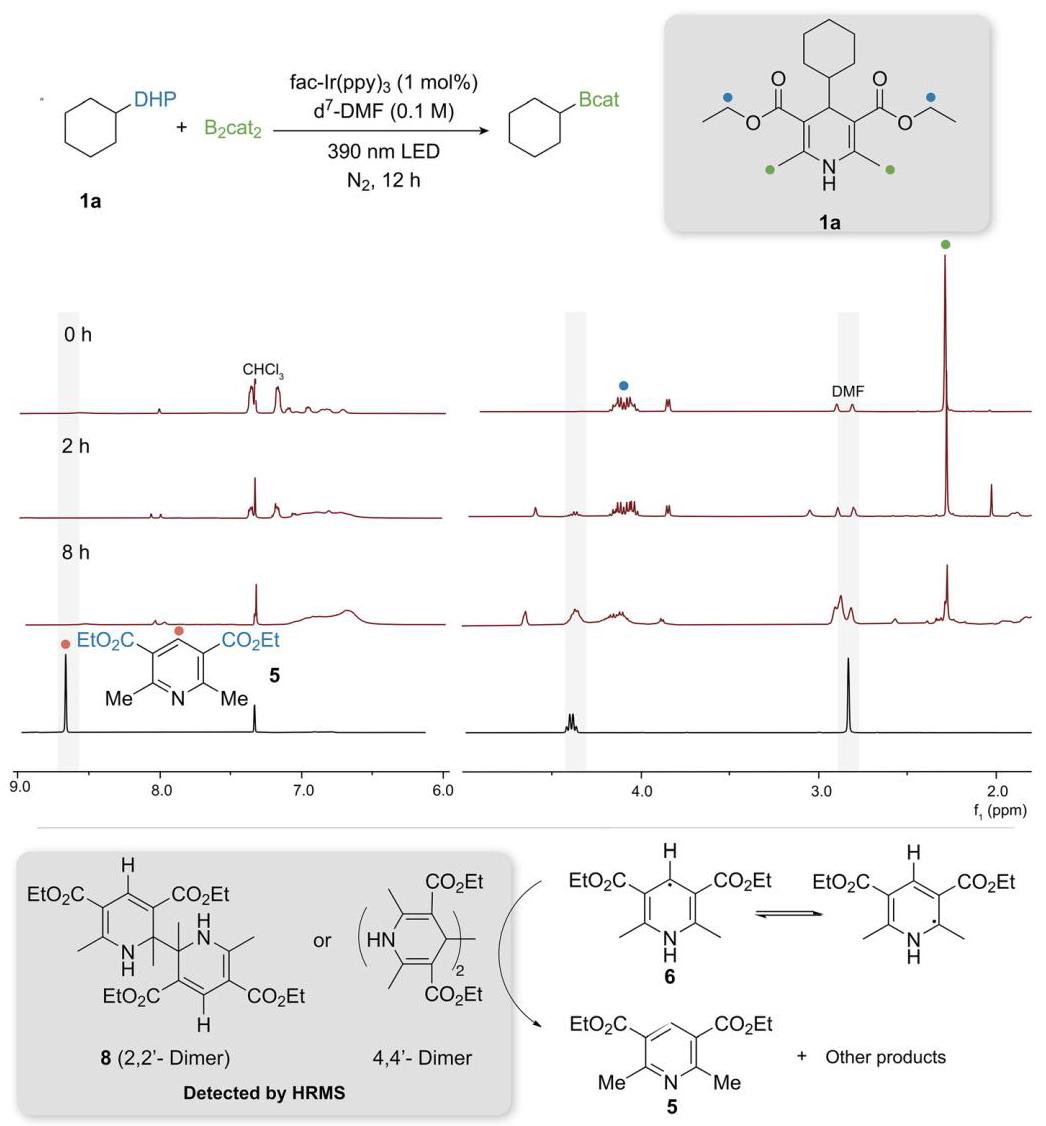
Fig. 5 NMR experiments. Reaction conditions: Hantzsch ester 1a (0.3 mmol,1.0 equiv.), B2cat (1.2 mmol, 4.0 equiv.) and fac-Ir(ppy) ${}_{3}$ (1 mol%) in ${\mathrm{d}}^{7}$ -DMF (0.1 M, 3.0 mL), Kessil 390 nm LED lamp, ${\mathrm{N}}_{2}$ atmosphere, room temperature, reaction time: 0 h, 2 h, 8 h.
To investigate the interactions between fac-Ir(ppy) ${}_{3}$ and 1a, we conducted density functional theory (DFT) calculations to determine the energy changes associated with one-electron oxidation or reduction. Through these calculations, we obtained the Gibbs free energy of substrate 1a along with its oxidized species 1a’+ and reduced species 1a’- (see ESI Scheme S4†). The redox potentials were derived from the free energy change $\left( {\Delta G}\right)$ using computational methods. ${}^{{28},{29}}$ Our calculations also involved fac-Ir(ppy) ${}_{3}$ and other intermediates.
Experimental studies have demonstrated that the excited state of fac-Ir(ppy) ${}_{3}$ (Ir(III)) can oxidize substrate $1\mathrm{a}$ to form 1a’+ while simultaneously reducing itself to fac-Ir(ppy) ${}_{3}{}^{ - }$ (Ir (II)). ${}^{{30},{31}}$ Additionally, ${}^{1}$ a* + can generate an alkyl radical along with the cationic species 7. The above DFT results confirmed that the oxidation of $\mathbf{{1a}}$ by $\operatorname{Ir}{\left( \mathrm{{III}}\right) }^{ * }$ occurred spontaneously. Furthermore, it indicated that the intermediate Ir(π) possessed a sufficiently reduced potential to facilitate the reduction of cationic species 7, resulting in the production of radical 6 and the regeneration of Ir(iii).
In our experimental results, we previously determined that the oxidation potential of 1a was approximately -0.16 V vs. Ag/ ${\mathrm{{AgNO}}}_{3}$ reference electrode (equivalent to ${0.64}\mathrm{\;V}\nu \mathrm{s}$ . standard hydrogen electrode, SHE). The reduction potential of Ir(III)* was found to be ${0.31}\mathrm{\;V}{vs}$ . saturated calomel electrode (SCE), which corresponds to ${0.56}\mathrm{\;V}{vs}$ . SHE. Both the experimental and DFT-calculated redox potentials indicated that Ir(III)* could oxidize 1a. However, it is important to note that the redox potential difference between them was relatively small, suggesting that the driving force for the oxidation process was not significant. Moreover, considering the strong reducibility of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ , a competitive process involving the reduction of Ir (III)* by ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ could also occur within the system. Due to the optical activity of the Hantzsch ester, the excited state of the Hantzsch ester is present in the reaction system. The oxidation of the excited Hantzsch ester by Ir(шi)* is thermodynamically more favorable. This may be a key factor in the reaction, as both the Hantzsch ester and fac-Ir(ppy) ${}_{3}$ are in the excited state, and the reducibility of the excited Hantzsch ester is greater than that of ${\mathrm{B}}_{2}{\mathrm{{cat}}}_{2}$ , promoting the electron transfer process between Hantzsch ester and fac-Ir(ppy) ${}_{3}$ .
The results obtained so far confirm the occurrence of electron transfer between 1a and fac-Ir(ppy) ${}_{3}$ . In addition, we sought to investigate whether simultaneous energy transfer (EnT) occurs between the Hantzsch ester and the catalyst. To explore this, we performed time-dependent density functional theory (TD-DFT) calculations to analyze the optical properties of $1\mathrm{a}$ and fac-Ir(ppy) ${}_{3}$ (Fig. 6a). The calculations revealed that the excitation wavelength for $1\mathrm{a}$ to the first excited state (S1) was ${296}\mathrm{\;{nm}}$ , while the excitation wavelength for ${fac}$ -Ir(ppy) ${}_{3}$ was ${411}\mathrm{\;{nm}}$ . This indicates that the homo-lumo gap of $1\mathrm{a}$ is larger than that of fac-Ir(ppy) ${}_{3}$ , suggesting that the energy transfer process from fac-Ir(ppy) ${}_{3}$ to $1\mathrm{a}$ is not thermodynamically favorable. Therefore, we concluded that the photocatalyst primarily governs the reaction through single electron transfer (SET), wherein the excited state of fac-Ir(ppy) ${}_{3}$ oxidizes 1a (or the excited state of 1a) to generate an alkyl radical and the cationic intermediate 7. Detailed DFT calculations were performed to validate this proposed mechanism (Fig. 6b). The results demonstrated that the Hantzsch ester substrate can indeed be oxidized by the excited state of fac-Ir(ppy) ${}_{3}$ (Ir(III), leading to the cleavage of the C-C bond in 1a and the generation of the alkyl radical and cation 7. The borylation process of the alkyl radical can proceed smoothly from both thermodynamic and kinetic perspectives, while 7 can be reduced to the radical 6, thereby regenerating Ir(III) from its reduced species Ir(II) due to the strong reducibility of the intermediate Ir(π). The subsequent transformation of 6 remains uncertain. However, an analysis of its spin density distribution indicates that the single electron of this radical is distributed with nearly equal probability at positions 2 and 4 (see ESI Scheme S6†). Therefore, we propose that 6 undergoes a dimerization process to form the 2,2’-dimeric product 8, and the DFT results confirm that this process is thermodynamically favorable.
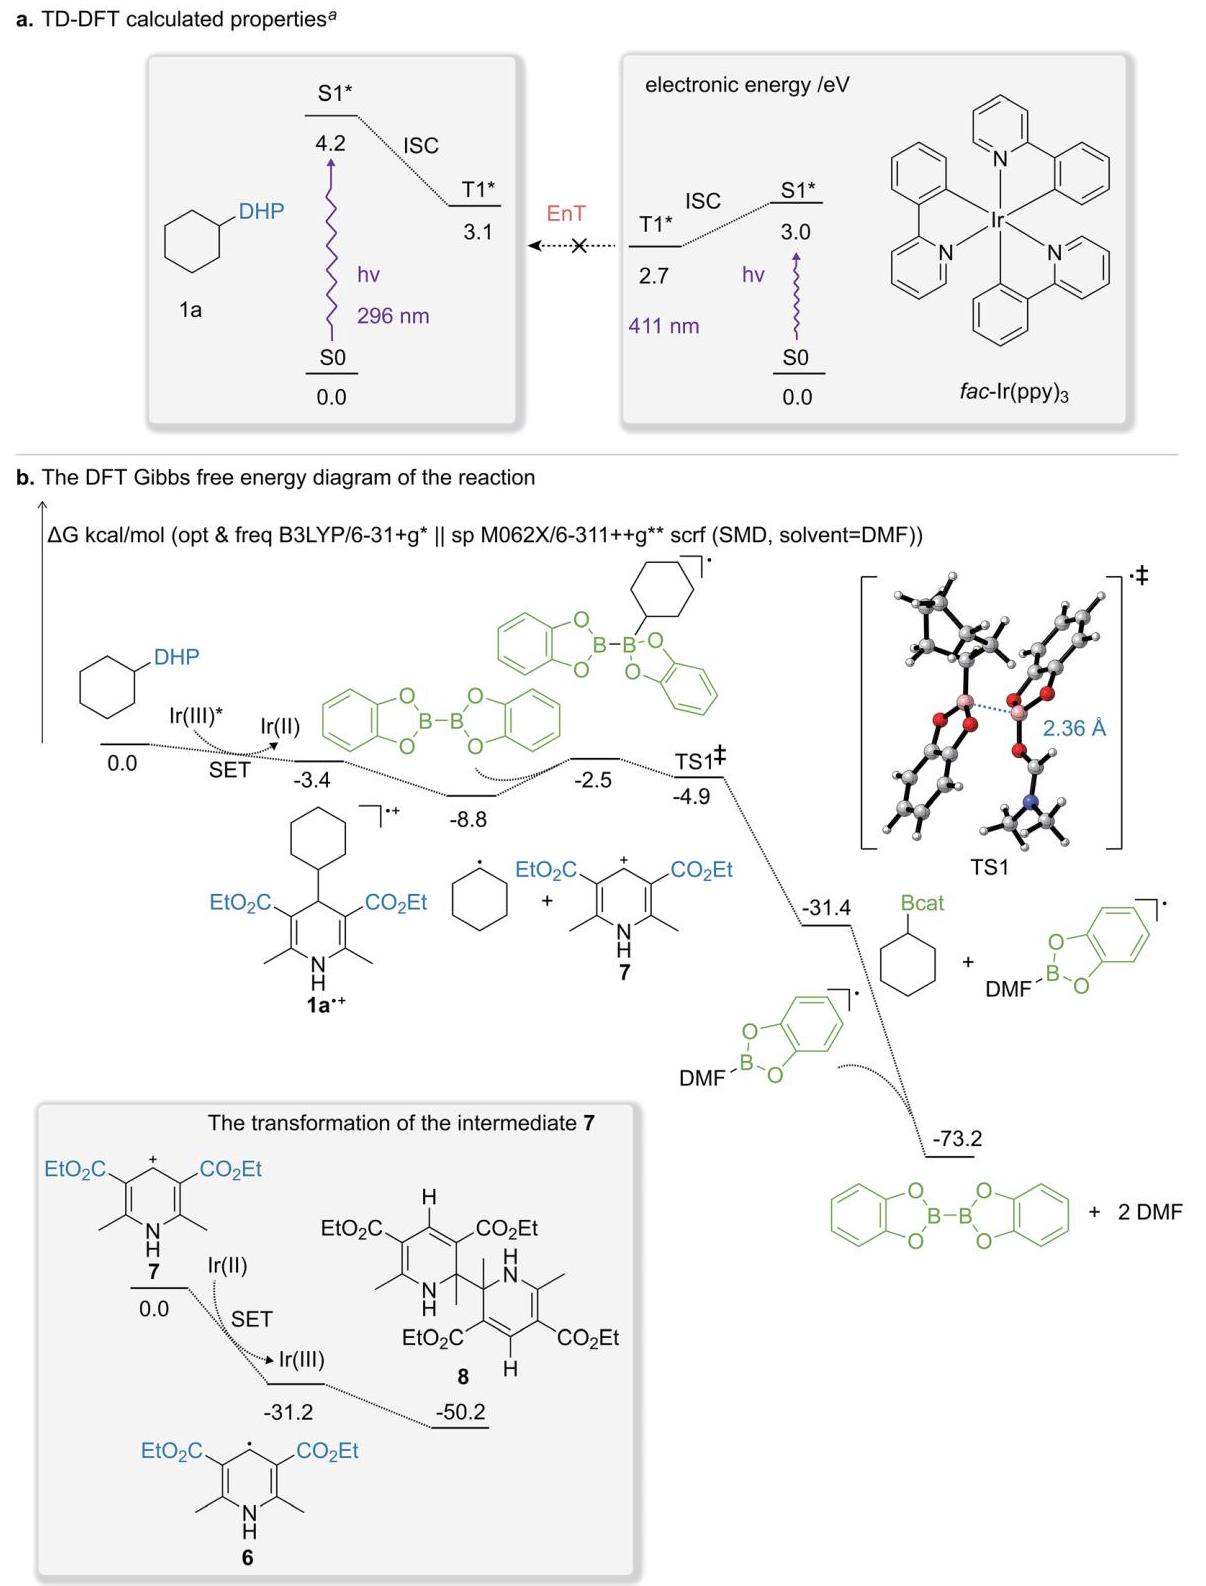
Fig. 6 DFT calculations. The S1* and T1* energies were calculated by TD-DFT under B3LYP/6-311++g** level with an implicit solvation model SMD (solvent = DMF).
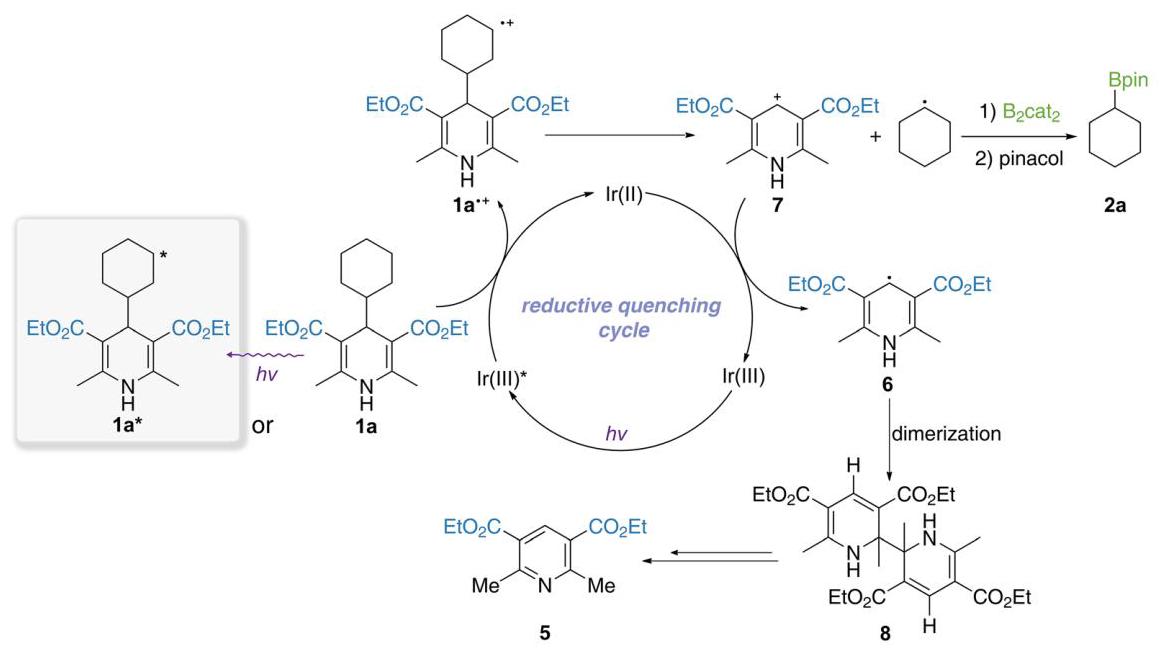
Fig. 7 Plausible mechanism.
Based on the experimental and theoretical evidence presented, along with the knowledge from previously developed radical borylation reactions, ${}^{9,{10},{17},{32}}$ we propose a plausible mechanism outlined in Fig. 7. The excitation of fac-Ir(ppy) ${}_{3}$ initiates a reductive quenching cycle in which the excited Ir(III) * species oxidizes 1a (or excited 1a) to generate the radical cation intermediate 1a*+. This intermediate then produces an alkyl radical along with the cation 7, which react with the boron reagent to form the desired borylation product. The intermediate 7 assists in the regeneration of Ir(III) to complete the quenching cycle. The product radical 6 subsequently undergoes a dimerization reaction to form the intermediate 8. Upon isolation, the dimerization products transform into the pyridine product 5. This mechanism provides an explanation for why the reaction proceeds without the need for external oxidants.
3 Conclusions
In conclusion, we have successfully developed a mild and efficient method for the radical borylation of aldehydes using 4-substituted Hantzsch esters as radical precursors. This photoredox-catalyzed reaction, conducted under visible light irradiation at room temperature, enables the transformation of aldehydes into alkyl borylation products in a “one-carbon-less” manner. The methodology exhibits excellent functional group compatibility and a broad substrate scope. By the oxidation of 4-substituted Hantzsch esters, alkyl radicals are generated and efficiently captured by the diboron reagent to afford a diverse range of alkyl borylation products. We have provided theoretical and experimental evidence confirming the formation of alkyl radicals. Furthermore, the scalability of the reaction and post-conversion of the products highlight the practical utility of this method. Through a combination of mechanistic experiments and density functional theory calculations, we have elucidated the electron transfer process between the photocatalyst and substrate, and proposed a plausible mechanism that is consistent with our experimental observations. Notably, this reaction proceeds efficiently without the need for external oxidants, offering an environmentally friendly and straightforward approach for the synthesis of alkyl boronates.
Conflicts of interest
There are no conflicts to declare.
Acknowledgements
The author thanks the National Natural Science Foundation of China (Grant No. 22071004, 21933001 and 22150013). The author also thanks the High Performance Computing Platform of Peking University and the Shenzhen Cloud Computing Center for DFT calculations support.
References
1 D. G. Hall, Boronic Acids: Preparation and Applications in Organic Synthesis, Medicine and Materials, John Wiley & Sons, 2012.
2 S. D. Bull, M. G. Davidson, J. M. Van den Elsen, J. S. Fossey, A. T. A. Jenkins, Y.-B. Jiang, Y. Kubo, F. Marken, K. Sakurai, J. Zhao, et al., Exploiting the Reversible Covalent Bonding of Boronic Acids: Recognition, Sensing, and Assembly, Acc. Chem. Res., 2013, 46, 312-326.
3 W. L. Brooks and B. S. Sumerlin, Synthesis and Applications of Boronic Acid-containing Polymers: from Materials to Medicine, Chem. Rev., 2016, 116, 1375-1397.
4 D. B. Diaz and A. K. Yudin, The Versatility of Boron in Biological Target Engagement, Nat. Chem., 2017, 9, 731- 742.
5 R. F. Barth, P. Mi and W. Yang, Boron Delivery Agents for Neutron Capture Therapy of Cancer, Cancer Commun., 2018, 38, 1-15.
6 F. Mo, Y. Jiang, D. Qiu, Y. Zhang and J. Wang, Direct Conversion of Arylamines to Pinacol Boronates: a Metal-free Borylation Process, Angew. Chem., Int. Ed., 2010, 49, 1846-1849.
7 F. W. Friese and A. Studer, New Avenues for C-B Bond Formation via Radical Intermediates, Chem. Sci., 2019, 10, 8503-8518.
8 A. Fawcett, J. Pradeilles, Y. Wang, T. Mutsuga, E. L. Myers and V. K. Aggarwal, Photoinduced Decarboxylative Borylation of Carboxylic Acids, Science, 2017, 357, 283-286.
9 D. Hu, L. Wang and P. Li, Decarboxylative Borylation of Aliphatic Esters under Visible-light Photoredox Conditions, Org. Lett., 2017, 19, 2770-2773.
10 C. Li, J. Wang, L. M. Barton, S. Yu, M. Tian, D. S. Peters, M. Kumar, A. W. Yu, K. A. Johnson, A. K. Chatterjee, et al., Decarboxylative Borylation, Science, 2017, 356, eaam7355.
11 J. Wang, M. Shang, H. Lundberg, K. S. Feu, S. J. Hecker, T. Qin, D. G. Blackmond and P. S. Baran, Cu-catalyzed Decarboxylative Borylation, ACS Catal., 2018, 8, 9537-9542.
12 D. Wei, T.-M. Liu, B. Zhou and B. Han, Decarboxylative Borylation of m CPBA-activated Aliphatic Acids, Org. Lett., 2019, 22, 234-238.
13 Q. Wei, Y. Lee, W. Liang, X. Chen, B.-s. Mu, X.-Y. Cui, W. Wu, S. Bai and Z. Liu, Photocatalytic Direct Borylation of Carboxylic Acids, Nat. Commun., 2022, 13, 7112.
14 F. W. Friese and A. Studer, Deoxygenative Borylation of Secondary and Tertiary Alcohols, Angew. Chem., Int. Ed., 2019, 58, 9561-9564.
15 J. Wu, R. M. Bär, L. Guo, A. Noble and V. K. Aggarwal, Photoinduced Deoxygenative Borylations of Aliphatic Alcohols, Angew. Chem., Int. Ed., 2019, 58, 18830- 18834.
16 J. Hu, G. Wang, S. Li and Z. Shi, Selective C- N Borylation of Alkyl Amines Promoted by Lewis base, Angew. Chem., Int. Ed., 2018, 57, 15227-15231.
17 F. Sandfort, F. Strieth-Kalthoff, F. J. Klauck, M. J. James and F. Glorius, Deaminative Borylation of Aliphatic Amines Enabled by Visible Light Excitation of an Electron Donor-acceptor Complex, Chem. - Eur. J., 2018, 24, 17210-17214.
18 J. Wu, L. He, A. Noble and V. K. Aggarwal, Photoinduced Deaminative Borylation of Alkylamines, J. Am. Chem. Soc., 2018, 140, 10700-10704.
19 Q. Liu, J. Zhang, L. Zhang and F. Mo, Photo-Induced Radical Borylation of Hemiacetals via C-C Bond Cleavage, Tetrahedron, 2021, 80, 131867.
20 A. M. F. Phillips and A. J. L. Pombeiro, Applications of Hantzsch Esters in Organocatalytic Enantioselective Synthesis, Catalysts, 2023, 13, 419.
21 P.-Z. Wang, J.-R. Chen and W.-J. Xiao, Hantzsch Esters: an Emerging Versatile Class of Reagents in Photoredox Catalyzed Organic Synthesis, Org. Biomol. Chem., 2019, 17, 6936-6951.
22 S. Ye and J. Wu, 4-Substituted Hantzsch Esters as Alkylation Reagents in Organic Synthesis, Acta Chim. Sin., 2019, 77, 814-831.
23 Z.-H. Shang, J. Pan, Z. Wang, Z.-X. Zhang and J. Wu, Transition-Metal-Free Radical Borylation Reactions, Eur. J. Org. Chem., 2023, 26, e202201379.
24 B. Wang, X. Zhang, Y. Cao, L. Zou, X. Qi and Q. Lu, Electrooxidative Activation of B-B Bond in B2cat2: Access to gem-Diborylalkanes via Paired Electrolysis, Angew. Chem., Int. Ed., 2023, 62, e202218179.
25 Y. Wen, C. Deng, J. Xie and X. Kang, Recent Synthesis Developments of Organoboron Compounds via Metal-free Catalytic Borylation of Alkynes and Alkenes, Molecules, 2018, 24, 101.
26 S. Easwar and N. P. Argade, Amano PS-catalysed Enantioselective Acylation of (±)-α-methyl-1, 3-benzodiox-ole-5-ethanol: an Efficient Resolution of Chiral Intermediates of the Remarkable Antiepileptic Drug Candidate,(-)-talampanel, Tetrahedron: Asymmetry, 2003, 14, 333-337.
Organic Chemistry Frontiers
27 K. Kano and T. Matsuo, Photoinduced One-electron Reduction of 1-benzyl-3-carbamoylpyridinium chloride and 3, 5-bis (ethoxycarbonyl)-2, 6-dimethylpyridine, Bull. Chem. Soc. Jpn., 1976, 49, 3269-3273.
28 A. V. Marenich, J. Ho, M. L. Coote, C. J. Cramer and D. G. Truhlar, Computational Electrochemistry: Prediction of Liquid-phase Reduction Potentials, Phys. Chem. Chem. Phys., 2014, 16, 15068-15106.
29 P. Winget, E. J. Weber, C. J. Cramer and D. G. Truhlar, Computational Electrochemistry: Aqueous One-electron Oxidation Potentials for Substituted Anilines, Phys. Chem. Chem. Phys., 2000, 2, 1231-1239.
30 K. Nakajima, S. Nojima, K. Sakata and Y. Nishibayashi, Visible-light-mediated Aromatic Substitution Reactions of Cyanoarenes with 4-Alkyl-1, 4-dihydropyridines through Double Carbon-Carbon Bond Cleavage, ChemCatChem, 2016, 8, 1028-1032.
31 C.-M. Wang, D. Song, P.-J. Xia, J. Wang, H.-Y. Xiang and H. Yang, Visible-light-promoted Synthesis of 1, 4-dicarbo-nyl Compounds via Conjugate Addition of Aroyl Chlorides, Chem. - Asian J., 2018, 13, 271-274.
32 B. Sun, S. Zheng and F. Mo, Transition Metal- and Light-free Radical Borylation of Alkyl Bromides and Iodides Using Silane, Chem. Commun., 2021, 57, 5674-5677.