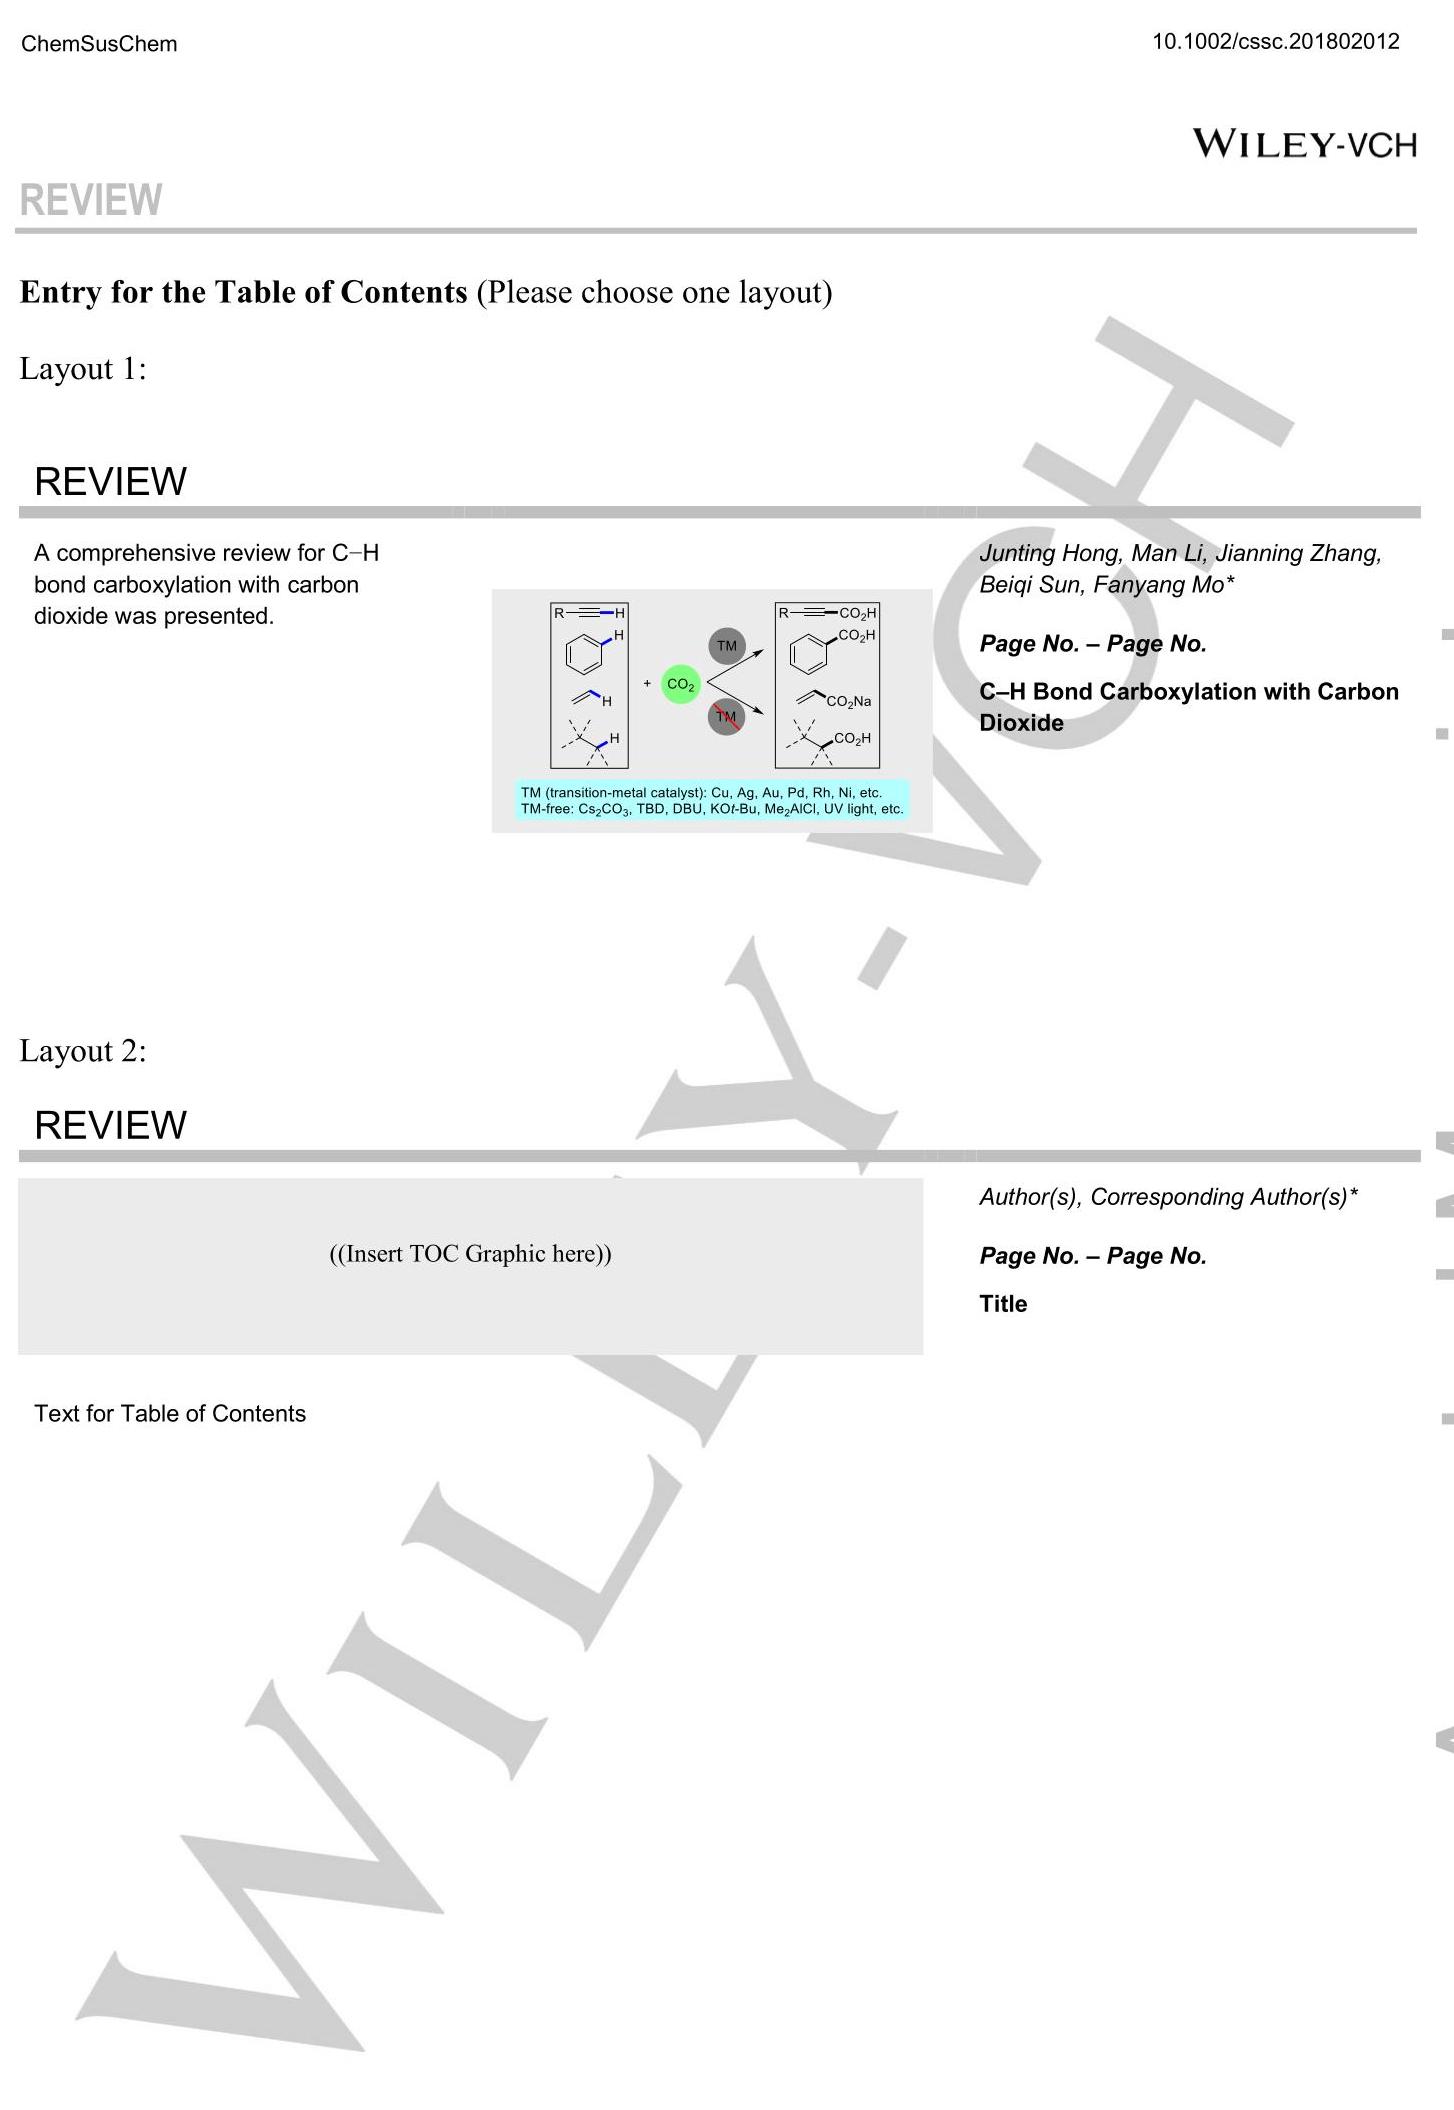
C-H Bond Carboxylation with Carbon Dioxide
中文 / Bilingual · English version
CHEMISTRY & SUSTAINABILITY
CHIEMSUSCHIM
ENERGY & MATERIALS
Accepted Article
Title: C-H Bond Carboxylation with Carbon Dioxide
Authors: Junting Hong, Man Li, Jianning Zhang, Beiqi Sun, and Fanyang Mo
This manuscript has been accepted after peer review and appears as an Accepted Article online prior to editing, proofing, and formal publication of the final Version of Record (VoR). This work is currently citable by using the Digital Object Identifier (DOI) given below. The VoR will be published online in Early View as soon as possible and may be different to this Accepted Article as a result of editing. Readers should obtain the VoR from the journal website shown below when it is published to ensure accuracy of information. The authors are responsible for the content of this Accepted Article.
To be cited as: ChemSusChem 10.1002/cssc.201802012
Link to VoR: http://dx.doi.org/10.1002/cssc.201802012
C-H Bond Carboxylation with Carbon Dioxide
Junting Hong, Man Li, Jianning Zhang, Beiqi Sun, and Fanyang Mo*
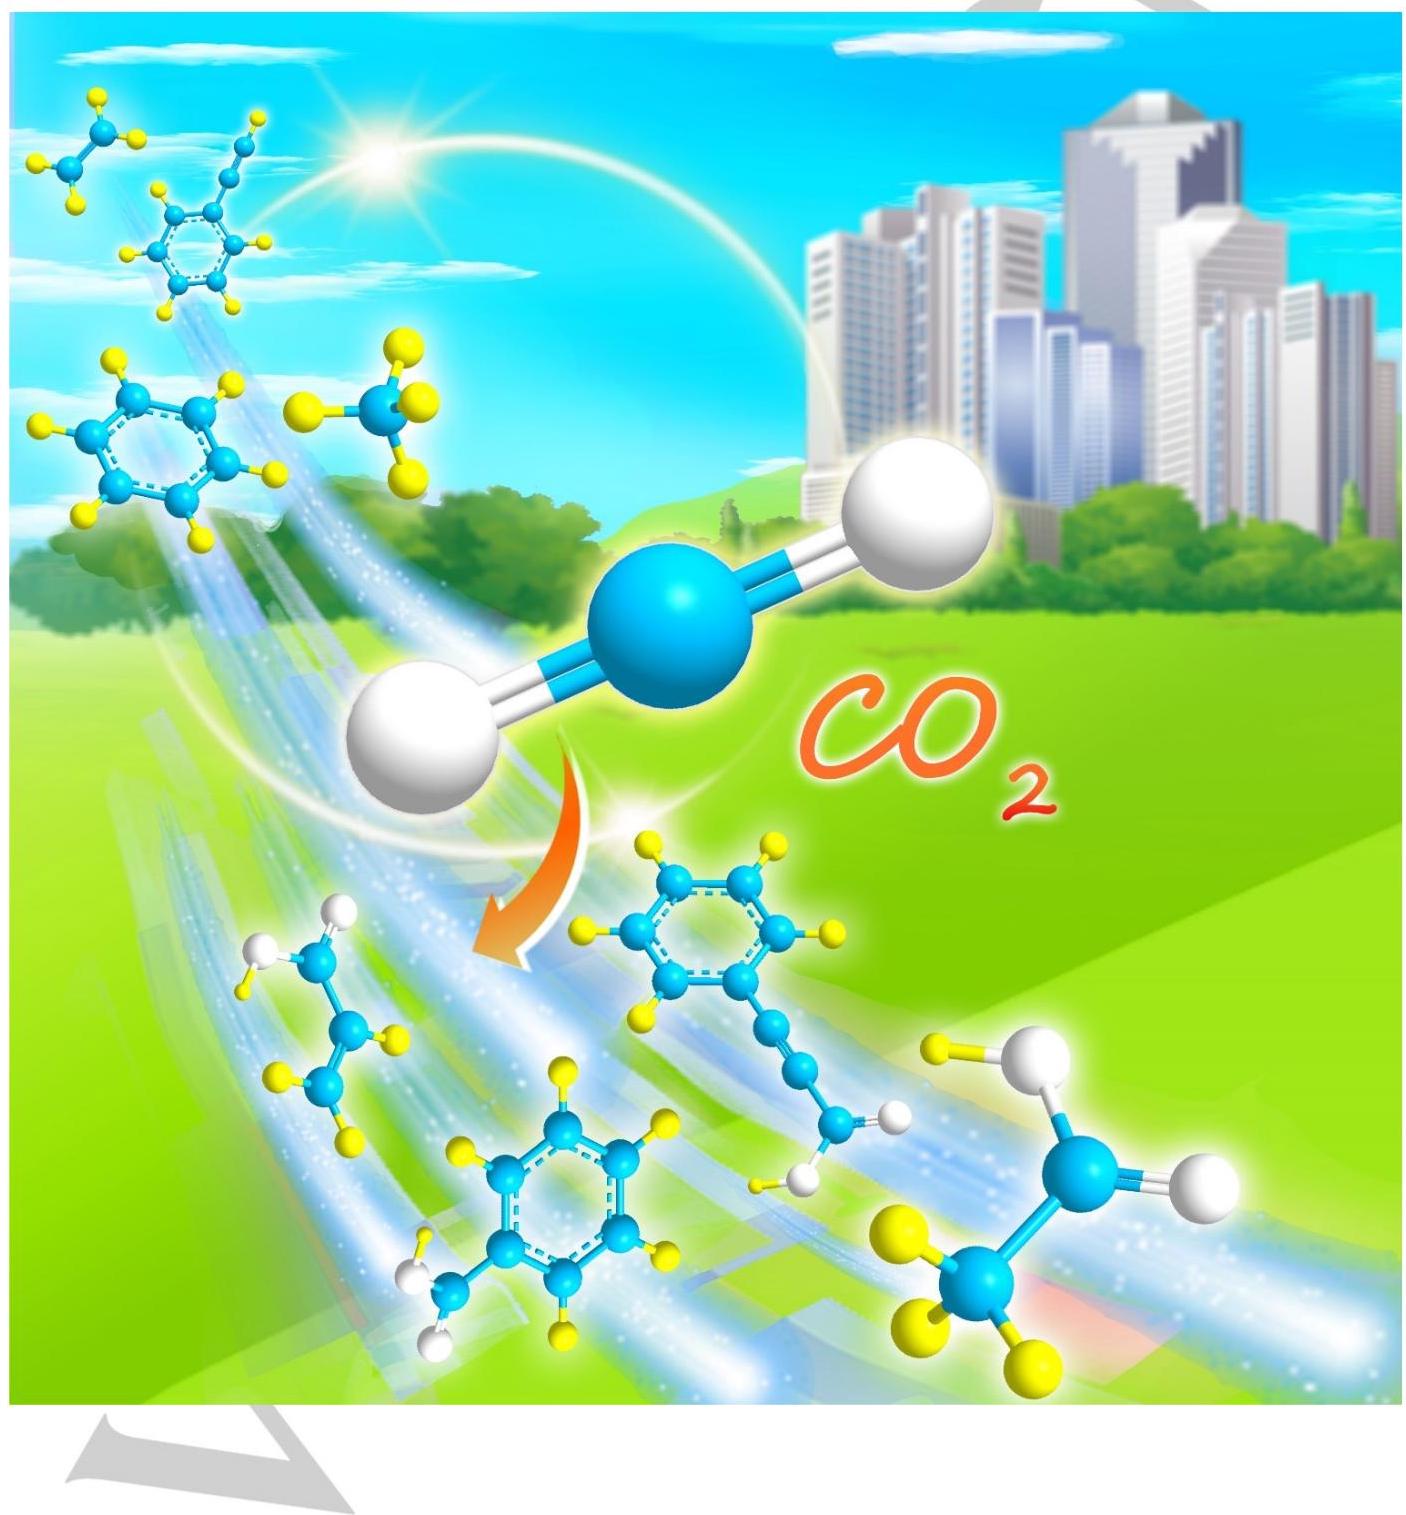
Abstract: Carbon dioxide $\left( {\mathrm{{CO}}}_{2}\right)$ is a nontoxic, renewable and abundant one-carbon (C1) source, while C-H bond functionalization represents one of the most important approaches to the construction of carbon-carbon bonds and carbon-heteroatom bonds in an atom- and step-economical manner. Merging the chemical transformation of ${\mathrm{{CO}}}_{2}$ with $\mathrm{C} - \mathrm{H}$ bond functionalization is of great importance in the synthesis of carboxylic acids and their derivatives. The contents of this review are organized according to the type of $\mathrm{C} - \mathrm{H}$ bond involved in carboxylation. The primary types of $\mathrm{C} - \mathrm{H}$ bonds are as follows: C(sp)-H bonds of terminal alkynes, C(sp2)-H bonds of (hetero)arenes, vinylic C(sp2)-H bonds, the diazo group ipso- $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds, aldehyde $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds, $\alpha - \mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds of carbonyl group, $\gamma - \mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds of carbonyl group, $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) -$ H bonds adjacent to nitrogen or oxygen atoms, $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds of $o$ -alkylphenyl ketones, allylic $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds, $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds of methane, and $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds of halogenated aliphatic hydrocarbons. In addition, multicomponent reactions, tandem reactions and key theoretical studies related to the carboxylation of C-H bonds are briefly summarized. Transition-metal-free, organocatalytic, electrochemical and light-driven methods are highlighted.
1. Introduction
With increasing attention to climate change, fixation and transformation of carbon dioxide $\left( {\mathrm{{CO}}}_{2}\right)$ , a greenhouse gas, has become a hot topic for chemical researchers. ${}^{\left\lbrack 1 - 6\right\rbrack }{\mathrm{{CO}}}_{2}$ is a one-carbon (C1) source that is non-toxic, nonflammable, renewable, abundant, and inexpensive. The use of ${\mathrm{{CO}}}_{2}$ instead of toxic phosgene or carbon monoxide in organic synthesis has high value in research and development. ${}^{\left\lbrack 7 - {13}\right\rbrack }$ Unfortunately, the high thermodynamic stability and kinetic inertness of ${\mathrm{{CO}}}_{2}$ make its chemical transformation very difficult. To solve this problem, the most classical method is to trap it with highly reactive compounds, such as Grignard or organolithium reagents. However, this method has poor functional group compatibility and requires strict anhydrous and anaerobic conditions. To address these problems, ${\mathrm{{CO}}}_{2}$ fixation processes with less reactive organometallic reagents (such as boron or zinc reagents) or halogenated hydrocarbons have been developed in recent years. ${}^{\left\lbrack {14}\right\rbrack }$ However, these methods require prefunctionalization of the substrate, and also produce needless waste, which fail to meet the requirements of green and sustainable chemistry. Although transition metal-catalyzed reductive carboxylation of unsaturated hydrocarbons partially solves the problem of the substrate prefunctionalization, these reactions still generally require stoichiometric or even excessive amounts of reductants, such as organozinc reagents, aluminum reagents, or hydrosilanes, etc. ${}^{\left\lbrack {15} - {16}\right\rbrack }$ On the other hand, the functionalization of carbon-hydrogen (C-H) bonds is an important method for constructing carbon-carbon bonds and carbon-heteroatom bonds in organic chemistry. ${}^{\left\lbrack {17} - {21}\right\rbrack }$ In light of its high atom economy and step economy, C-H bond functionalization demonstrates a wide range of application in the syntheses of pharmaceuticals, natural products, and organic materials.[22-24] Using ubiquitous $\mathrm{C} - \mathrm{H}$ bond to react with ${\mathrm{{CO}}}_{2}$ is an ideal way to produce high value-added chemicals. Key to achieving this goal is to combine the selective activation and functionalization of inert $\mathrm{C} - \mathrm{H}$ bonds with the activation of ${\mathrm{{CO}}}_{2}$ . Organized according to the types of $\mathrm{C} - \mathrm{H}$ bond involved in the reaction, this review summarizes recent progress in the functionalization of $\mathrm{C} - \mathrm{H}$ bond using ${\mathrm{{CO}}}_{2}$ as a carboxylative reagent. We noted that some published reviews cover part of contents in this paper. For example, $\mathrm{C} - \mathrm{H}$ carboxylation of aromatic compounds through ${\mathrm{{CO}}}_{2}$ fixation has been reviewed recently. ${}^{\left\lbrack 3\right\rbrack }$ The direct carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ and $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ by transition-metal-catalyzed and base-mediated reactions has also been surveyed. ${}^{\left\lbrack {25}\right\rbrack }$ However, some important aspects including several specific types of C-H carboxylation reactions, transition-metal-free reactions (including Lewis acid-mediated reactions), multicomponent reactions, tandem reactions and key theoretical studies related to the carboxylation of C-H bonds are overlooked. To this end, it’s highly desirable to thoroughly summarize the field of C-H carboxylation with ${\mathrm{{CO}}}_{2}$ . By doing so, we can find many interesting points which will enhance our understanding of this exciting field. For example, many transition-metal-catalyzed C-H carboxylation reactions with ${\mathrm{{CO}}}_{2}$ (including C(sp)-H bonds and $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds) can develop their transition-metal-free version (see section 2.2.4., 3.1.1.2., and Scheme 56); copper can catalyze all types of $\mathrm{C} - \mathrm{H}$ carboxylation reactions with ${\mathrm{{CO}}}_{2}$ (see section 2.1.1, 2.2.1, 2.3, 3.1.1.1, Scheme 71, Scheme 78, etc.); base can mediate all types of $\mathrm{C} - \mathrm{H}$ carboxylation reactions for substrates with relatively acidic C-H bonds (see section 2.2.4, 3.1.1.2, 3.3, Scheme 68, Scheme 73, etc.); all types of C-H carboxylation reactions can be incorporated into tandem or sequential reactions (see section 2.3, Scheme 45, 55 and 70, etc.).
Junting Hong received her B.S. degrees from Beijing Institute of Technology in 2017. She is currently a Ph.D. candidate in Professor Fanyang Mo’s group in Peking University. Her research interest is radical borylation of aryl and alkyl halides.
Junting Hong, Man Li, Jianning Zhang, Beigi Sun, Prof. Dr. Fanyang Mo
Department of Energy and Resources Engineering, College of Engineering, Peking University
No. 5 Yiheyuan Road, Haidian Distric, Beijing 100871, China
REVIEW
Man Li received his B.S. degree from Henan Normal University in 2010, and obtained his M.Sc. degree from Lanzhou Institute of Chemical Physics (LICP), Chinese Academy of Sciences (CAS) in 2014. He has been a Research Assistant at LICP, CAS, Shanghai Institute of Organic Chemistry, CAS and Peking University (with Assistant Professor Fanyang Mo). His research interests include chemical transformations of carbon dioxide and acid catalysis.
Jianning Zhang is a fourth year undergraduate student of Peking University. He is now studying in Professor Mo’s group.
Beiqi Sun obtained his B.E degree from
Peking University in 2018. He is
currently a Ph.D student in Dr. Fanyang
Mo’s group. His research interests are organic electrochemistry and ${\mathrm{{CO}}}_{2}$ utilization in organic synthesis.
Fanyang Mo obtained his undergraduate
degree and master degree in applied
chemistry in 2004 and 2006 at Beijing
Institute of Technology under the
supervision of Professor Zhiming Zhou. He then moved to Peking University and pursued his Ph. D study under the supervision of Professor Jianbo Wang. After receiving his PhD in 2010, he went to the US and worked with Professor Qinghai Zhang at The Scripps Research Institute and Professor Guangbin Dong at The University of Texas at Austin from 2010 to 2015. He joined Peking University as an Assistant Professor in April 2015. Current research interests of the group include ${\mathrm{{CO}}}_{2}$ utilization in organic synthesis, organic electrochemistry, and modification of semiconductor oxides.
2. Carboxylation of C(sp)-H bonds with ${\mathrm{{CO}}}_{2}$
2.1. Carboxylative coupling reaction of terminal alkynes, alkyl halides, and CO2
2.1.1. Cu(I)-catalyzed carboxylative coupling reaction of terminal alkynes, alkyl halides, and CO2
In 1994, Inoue and coworkers[26] found that Cu(I) salts (CuI/CuBr) or Ag(I) salts (AgI/AgNO ${}_{3}$ ) can catalyze the carboxylative coupling of terminal alkynes, alkyl bromides, and ${\mathrm{{CO}}}_{2}$ (Scheme 1). The choice of solvent and base has a significant effect on the reaction. Polar aprotic solvents such as DMF, DMAc and N-methylpyrrolidone are suitable for this reaction. Employment of organic bases such as triethylamine or DBU suffers from alkylation of the base with alkyl halide. Thus, an inorganic salt like ${\mathrm{K}}_{2}{\mathrm{{CO}}}_{3}$ is used as the base. A three-step reaction mechanism is proposed: (i) copper acetylide formation,(ii) ${\mathrm{{CO}}}_{2}$ insertion into the carbon-copper bond, and (iii) esterification with bromoalkane with regeneration of the copper salt catalyst (Scheme 1). Based on the above work, Oi and Inoue and coworkers reported the Cul-catalyzed polymerization of diyne, ${\mathrm{{CO}}}_{2}$ and dihaloalkane in 1996.[27] The use of 1,10-phenanthroline as a ligand accelerates the polymerization.
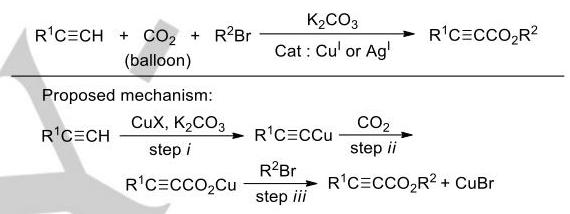
Scheme 1. Carboxylative coupling reaction of terminal alkynes catalyzed by copper(I) or silver(I) salt ${}^{\left\lbrack {26}\right\rbrack }$ In 2010, Lu and coworkers ${}^{\left\lbrack {28}\right\rbrack }$ reported the carboxylative coupling of terminal alkynes, allylic chlorides, and ${\mathrm{{CO}}}_{2}$ catalyzed by $\mathrm{N}$ - heterocyclic carbene (NHC) copper(I) complex (IPr)CuCl (Scheme 2). They found that the use of NHC ligands significantly increases catalytic activity and selectivity. Besides allylic chlorides, propargylic chlorides, benzyl chlorides, and a-chloro carbonyl compounds can all participate in the reaction. Catalyst recovery can be achieved by column chromatography. In 2014, Yuan and Lin[29] conducted a density functional theory (DFT) study on (IPr)CuCl-catalyzed carboxylative coupling reaction of terminal alkynes, allyl chloride and ${\mathrm{{CO}}}_{2}$ reported by Lu. ${}^{\left\lbrack {28}\right\rbrack }$ It was found that both the reactions of (IPr)Cu-C≡CPh and (IPr)CuOCOC≡CPh with an allylic chloride occur through an ${\mathrm{S}}_{\mathrm{N}}2$ substitution pathway. They also revealed how the reaction conditions affect the preference of one pathway over the other (carboxylative coupling versus cross coupling). In 2012, Inamoto, Kondo and coworkers ${}^{\left\lbrack {30}\right\rbrack }$ found that the Cul/PEt ${}_{3}$ system can catalyze carboxylative coupling reactions of terminal alkynes, haloalkane, and ${\mathrm{{CO}}}_{2}$ under mild conditions (Scheme 3). For some relatively inert terminal alkynes such as 4- cyanophenylacetylene, higher yields can be obtained by using the Cul/2,2’-bipyridine catalyst system while heating up to ${50}{}^{ \circ }\mathrm{C}$ . For substrates such as 4-nitrophenylacetylene, the reaction results are unsatisfactory. In 2015, He and coworkers[31] reported a Cu(I)-based ionic liquid-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ (Scheme 4). For terminal alkynes with electron-withdrawing groups, raising the reaction temperature to ${40}^{ \circ }\mathrm{C}$ can significantly improve the yield.
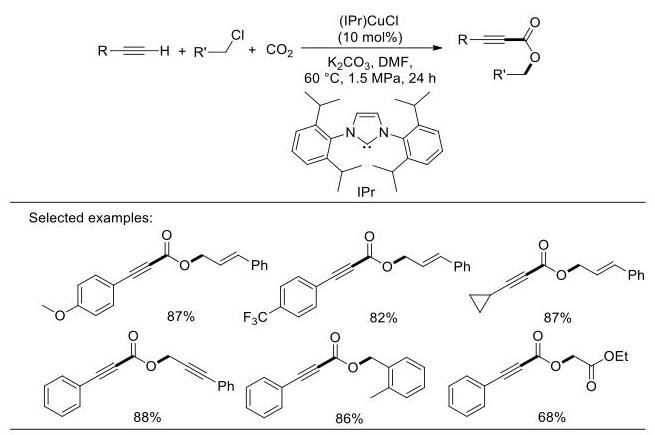
Scheme 2. (IPr)CuCl-catalyzed carboxylative coupling reaction of terminal alkynes[28]
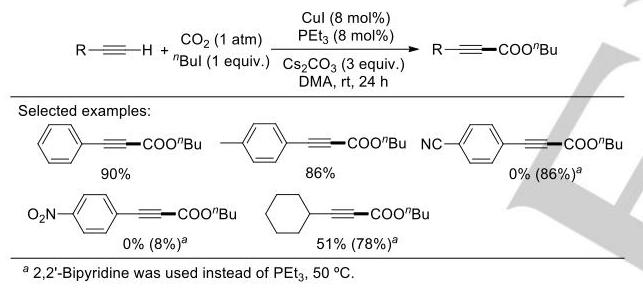
Scheme 3. Cul/PEt3-catalyzed carboxylative coupling reaction of terminal alkynes[30]
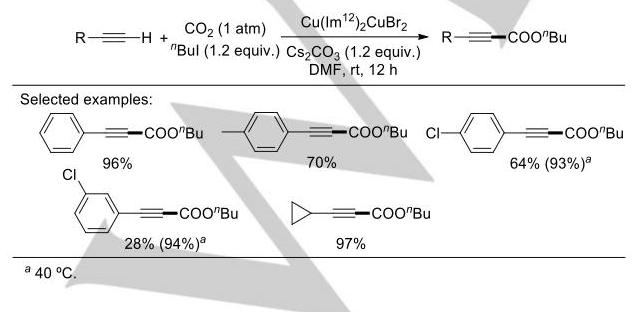
Scheme 4. Copper(I)-based ionic liquid-catalyzed carboxylative coupling reaction of terminal alkynes ${}^{\left\lbrack {31}\right\rbrack }$
The most commonly used solvents in the carboxylative coupling reactions of terminal alkynes, haloalkanes and ${\mathrm{{CO}}}_{2}$ are DMF and DMA. However, these solvents are often environmentally unfriendly. In 2013, He and coworkers[32] found that ethylene carbonate was an ideal solvent for the carboxylative coupling reaction of terminal alkynes (Scheme 5a). Ethylene carbonate has the advantages of strong solubility and low toxicity. DFT calculations show that the use of ethylene carbonate as solvent can greatly reduce the energy barrier for ${\mathrm{{CO}}}_{2}$ insertion into the Cu-C(sp) bond. Ethylene carbonate was thought to act as a potential ligand, as the reaction does not require any external ligands. Based on this work, He and coworkers[33] further developed a carboxylative coupling reaction of terminal alkynes catalyzed by activated carbon-supported CuBr (Scheme 5b). This reaction also used ethylene carbonate as solvent. Hot filtration experiment demonstrates that this reaction most likely occurs on the surface of a heterogeneous catalyst which can be reused at least five times without substantial inactivation. Notably, the catalyst on the second and third reuse performed slightly better than the freshly prepared catalyst. A possible reason was proposed according to X-ray photoelectron spectroscopy (XPS) characterization: Divalent copper species on the surface of the catalyst may be reduced to monovalent copper under the reaction conditions.
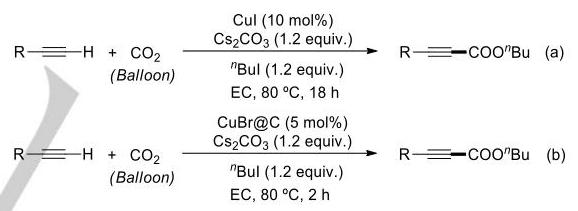
Scheme 5. Cu(I)-catalyzed carboxylative coupling reaction of terminal alkynes in ethylene carbonate[32-33]
2.1.2. Ag(I)-catalyzed carboxylative coupling reaction of terminal alkynes, alkyl halides, and CO2
In 2012, based on their previous work[34], Lu and coworkers also reported carboxylative coupling reactions of terminal alkynes, chlorides, and ${\mathrm{{CO}}}_{2}$ catalyzed by Agl (Scheme 6a). ${}^{\left\lbrack {35}\right\rbrack }$ The amount of Agl can be as low as ${0.1}\mathrm{\;{mol}}\%$ . Kinetic studies demonstrated that the activity of the ${0.1}\mathrm{\;{mol}}\% \mathrm{{AgI}}/{\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ catalytic system was significantly better than the previously reported 10 mol% (IPr) ${\mathrm{{CuCl}/K}}_{2}{\mathrm{{CO}}}_{3}$ system. Furthermore, in 2013, Lu and coworkers realized the sequential AgI-catalyzed carboxylative coupling/Ru-catalyzed cross-metathesis reactions for the synthesis of functionalized 2-alkynoates (Scheme 6b). ${}^{\left\lbrack {36}\right\rbrack }$
REVIEW
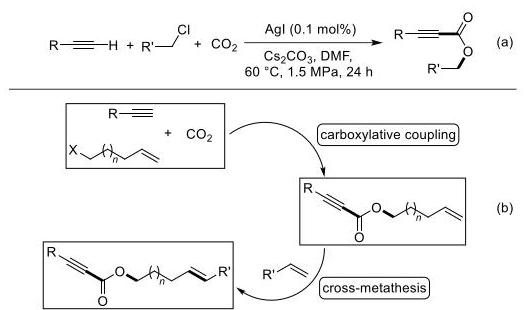
Scheme 6. Agl-catalyzed carboxylative coupling reaction of terminal alkynes ${}^{\left\lbrack {35}\right\rbrack }$ and related sequential reaction ${}^{\left\lbrack {36}\right\rbrack }$
In 2015, He and coworkers[37] reported the carboxylative coupling reaction of terminal alkyne, $n$ -butyl iodide and ${\mathrm{{CO}}}_{2}$ catalyzed by ${\mathrm{{Ag}}}_{2}{\mathrm{{WO}}}_{4}$ (Scheme 7a). The reaction can be carried out under very mild conditions: room temperature,1 atm of ${\mathrm{{CO}}}_{2}$ , without using any external ligands. They believe the catalyst is bifunctional: silver(I) activates C≡C triple bond, while tungstate anion activates ${\mathrm{{CO}}}_{2}$ , as verified by ${}^{13}\mathrm{C}$ NMR. In addition, they also successfully achieved the one-pot sequential phenylacetylene carboxylative coupling and hydroamination reaction (Scheme 7b). There are some other Ag-catalyzed carboxylative coupling reaction of terminal alkynes developed in recent years. ${}^{\left\lbrack {38} - {39}\right\rbrack }$
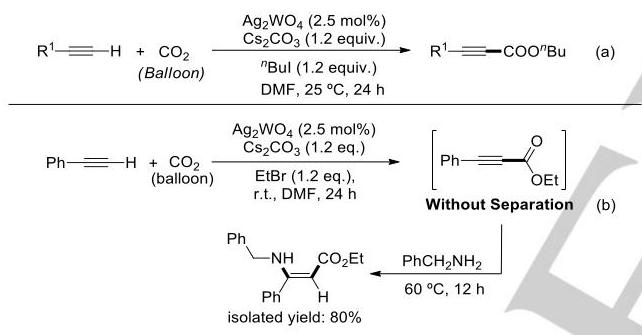
Scheme 7. ${\mathrm{{Ag}}}_{2}{\mathrm{{WO}}}_{4}$ -catalyzed carboxylative coupling reaction of terminal alkynes and related sequential reaction ${}^{\left\lbrack {37}\right\rbrack }$
2.1.3. Au(I)-catalyzed carboxylative coupling reaction of terminal alkynes, alkyl halides, and CO2
In 2008, Anastas and coworkers[40] found that Aul can catalyze the carboxylative coupling of phenylacetylene, 1-bromo-2- pentyne and ${\mathrm{{CO}}}_{2}$ to give 1,6-diyne (Scheme 8).
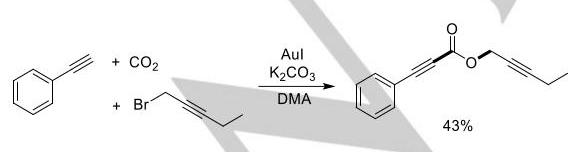
Scheme 8. Aul-catalyzed carboxylative coupling reaction of phenylacetylene ${}^{\left\lbrack {40}\right\rbrack }$
2.2. Carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$
2.2.1. Cu(I)-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$
In 2010, Gooßen ${}^{\left\lbrack {41}\right\rbrack }$ and Zhang ${}^{\left\lbrack {42}\right\rbrack }$ almost simultaneously reported the catalytic carboxylation of terminal alkynes catalyzed by $\mathrm{{Cu}}\left( \mathrm{I}\right)$ complexes. In Gooßen’s report, it is necessary to use different catalytic systems for aromatic and aliphatic alkynes (Scheme 9). Ethers, halogens, trifluoromethyl group and alkenyl group are all compatible, while unprotected amino group and hydroxyl group are not. In Zhang’s report, the choice of ligand, base and temperature has a large influence on the reaction outcome (Scheme 10). Interestingly, they found that by using the CuCl-TMEDA system, the reaction proceeds in the absence of any basic additives. For most terminal alkynes, good yields can be obtained with the CuCl-TMEDA system. For the very inert 4- nitrophenylacetylene, the use of CuCl-TMEDA system gives lower yields, while using poly-N-heterocyclic carbene (PNHC) as ligands gives good results. The scope of substrates for the reaction is wide and the conditions are mild (room temperature, 1 atm of ${\mathrm{{CO}}}_{2}$ ). It is worth mentioning that when the ratio of $\mathrm{{CuCl}}$ and PNHC ligands is 1:2, the performance of catalyst $\mathrm{P}{\left( \mathrm{{NHC}}\right) }_{0.5}{\left( \mathrm{{NHC}} - \mathrm{{Cu}}\right) }_{0.5}$ is better than that of the catalyst at 3:2 ratio. This result shows that the free carbene species in the catalyst plays an important role in the reaction. It is proposed that the key to extremely high catalytic activity is that organic catalysts (NHC) and organometallic catalysts operate synergistically in one system. In 2014, Wang, Zhang and coworkers[43] calculated the possible mechanism of the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by $\mathrm{{Cu}}\left( \mathrm{I}\right) - \mathrm{{NHC}}$ complexes and the role of $\mathrm{{NHC}}$ through DFT calculations. In 2015, Trivedi and coworkers[44] reported the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by ferrocene-based bisphosphine ligand and Cul Complex C (Scheme 11). The structure of catalyst C was confirmed by X-ray crystal diffraction. In 2017, Beletskaya and coworkers ${}^{\left\lbrack {45}\right\rbrack }$ first used ${\mathrm{{Al}}}_{2}{\mathrm{O}}_{3}$ -supported copper particles to catalyze the direct carboxylation of various terminal alkynes with ${\mathrm{{CO}}}_{2}$ . The catalyst is heterogeneous, highly active, easily available, and recyclable.
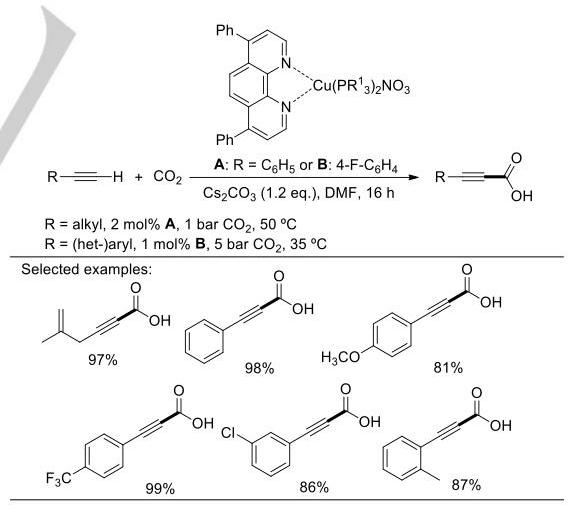
Scheme 9. Copper(I) complexes-catalyzed carboxylation of terminal alkynes reported by Gooßen and co-workers[41]
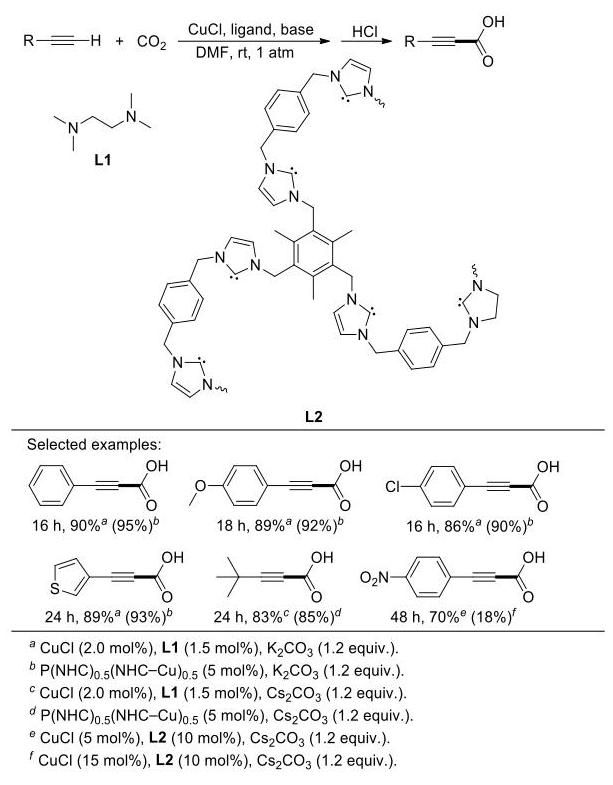
Scheme 10. Copper(I) complexes-catalyzed carboxylation of terminal alkynes reported by Zhang and co-workers[42]
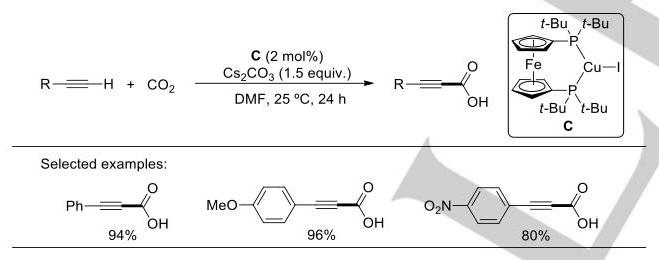
Scheme 11. Complex C-catalyzed carboxylation of terminal alkynes[44]
In 2018, Gooßen and coworkers[46] reported the salt-free copper-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ (Scheme 12). The reaction utilizes a copper(I) catalyst which enables the insertion of carbon dioxide into alkyne $\mathrm{C} - \mathrm{H}$ bonds by using 2,2,6,6-tetramethylpiperidine (TMP) as the base. The ammonium alkynecarboxylate can be hydrogenated to the primary alcohol and water by a rhodium/molybdenum catalyst, regenerating the amine base.
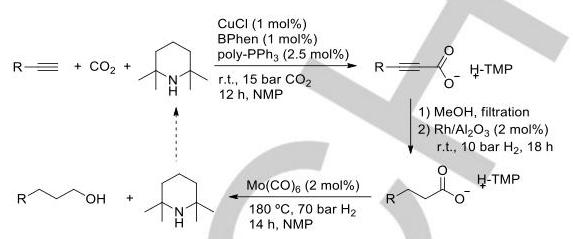
Scheme 12. The salt-free copper-catalysed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {46}\right\rbrack }$
2.2.2. Ag(I)-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$
In 2011, Lu and coworkers[34] reported an Agl-catalyzed direct carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ (Scheme 13a). The optimal amount of Agl is 1 mol%, while increasing the amount of catalyst to 20 mol% decreases the yield. It was reasoned that the catalyst can promote both carboxylation and decarboxylation reactions. This reaction does not require any external ligands which will reduce product yield instead. In 2014, Luo and Zhang and coworkers[47] studied the mechanism of silver-catalyzed carboxylation ${}^{\left\lbrack {34}\right\rbrack }$ of phenylacetylene with ${\mathrm{{CO}}}_{2}$ through DFT calculations. The calculated results show the true catalytically active species is a ${\mathrm{{CsCO}}}_{3}$ -coordinated Ag complex rather than neutral alkynyl silver species. Through theoretical calculations, they also explained the experimentally observed inhibition of the reaction by organic ligands. In 2012, Gooßen and coworkers[48] reported a catalytic system analogous to Lu and coworkers (Scheme 13b). In their report, the amount of Ag(I) catalyst used can be as low as 500 ppm. In 2014, Hong and coworkers[49] found that combusted exhaust gas can be used for organic reactions related to ${\mathrm{{CO}}}_{2}$ after capturing ${\mathrm{{CO}}}_{2}$ in aqueous ethanolamine solution. Successfully combining the capture and utilization of ${\mathrm{{CO}}}_{2}$ , this strategy was applied to the direct carboxylation of terminal alkynes catalyzed by Agl (Scheme 13c). Similar results were obtained using ${\mathrm{{CO}}}_{2}$ in the combusted exhaust gas and directly using hyper-pure ${\mathrm{{CO}}}_{2}$ . Notably, even though the aqueous ethanolamine solution was reused 55 times, the yield of the reaction was still not significantly reduced.
In 2014, Jover and Maseras[50] used DFT calculations to study the mechanism of the copper-, silver- and gold-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ . They found the catalytic systems of these three metals all experienced similar reaction pathways: coordination of alkyne and metal, deprotonation with an external base, and nucleophilic attack on the ${\mathrm{{CO}}}_{2}$ molecule by the metal-acetylide complex.
In 2016, Fang and Zhou and coworkers ${}^{\left\lbrack {51}\right\rbrack }$ reported the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by the $\mathrm{{Ag}}\left( \mathrm{I}\right) -$ NHC complex (Scheme 14). In 2012, Zhang and coworkers developed a PNHC-supported silver nanoparticle catalyst system ${}^{\left\lbrack {52}\right\rbrack }$ based on their previous ${\text{ work }}^{\left\lbrack {42}\right\rbrack }$ . Similar to the Cu(I)-PNHC system, this system can also efficiently catalyze the carboxylation of terminal alkynes at room temperature under 1 atm of ${\mathrm{{CO}}}_{2}$ . A mechanism involving similar synergistic effects of organic catalyst (NHC) and organometallic catalyst was proposed. As a heterogeneous catalytic system, the catalyst can be easily recovered and reused. Later they developed NHC-Ag/Pd-catalyzed reductive carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ and ${\mathrm{H}}_{2}.{}^{\left\lbrack {53}\right\rbrack }$ In 2014, Finashina and coworkers ${}^{\left\lbrack {54}\right\rbrack }$ reported an Ag/F-Al ${}_{2}{\mathrm{O}}_{3}$ -catalyzed carboxylation of phenylacetylene with ${\mathrm{{CO}}}_{2}$ in 62.1% yield.
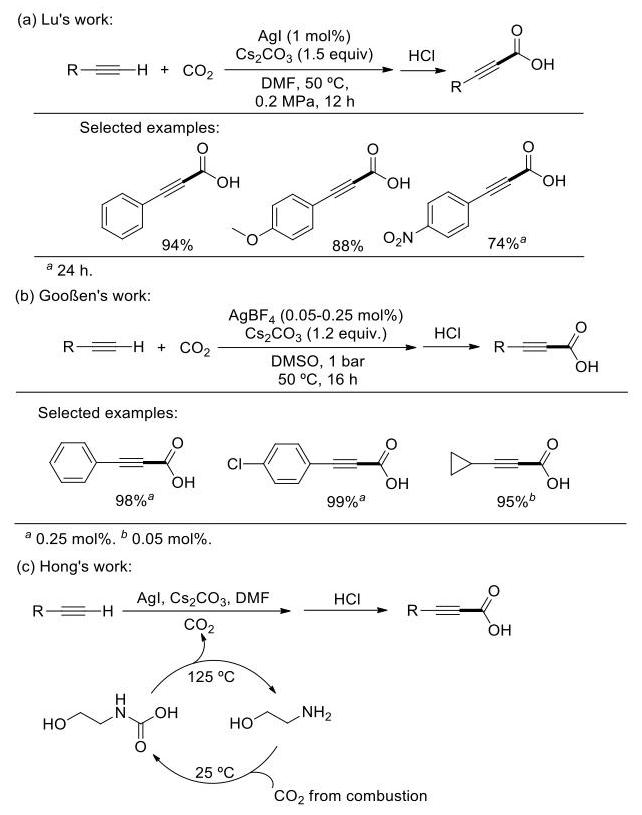
Scheme 13. Ligand-free Ag(I)-catalyzed carboxylation of terminal alkynes[34,48- 49]
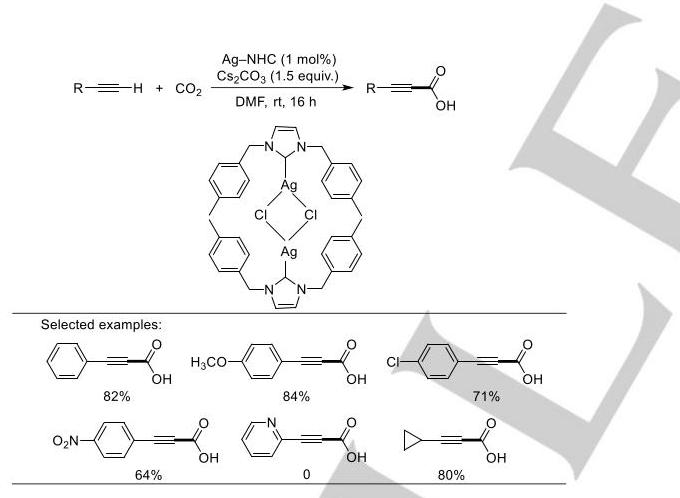
Scheme 14. Ag-NHC complex-catalyzed carboxylation of terminal alkynes[51]
In 2015, Ma, Cheng and coworkers[55] reported the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by Ag@ MIL-101, silver nanoparticles supported by metal-organic frameworks (MOFs) MIL-101. The dual-functional catalytic material can achieve ${\mathrm{{CO}}}_{2}$ capture and high-efficiency conversion at the same time at ${50}^{ \circ }\mathrm{C}$ under 1 atm of ${\mathrm{{CO}}}_{2}$ . As a heterogeneous catalyst, Ag@MIL-101 can be easily separated from the reaction solution by centrifugation. This catalyst can be reused at least five times without significant reduction in catalytic activity. They believe the larger pore size of the MOF carrier favors the diffusion of metal precursor ${\mathrm{{AgNO}}}_{3}$ and the deposition of $\mathrm{{Ag}}$ nanoparticles. In addition, the MOF carrier also plays an important role in the efficient performance of the catalytic reaction: it is an excellent material for the adsorption of ${\mathrm{{CO}}}_{2}$ and can increase local ${\mathrm{{CO}}}_{2}$ concentration in the catalytic center; the framework and pores can be used as a “microreactor” to provide a favorable reaction environment; MOF structure can stabilize the overall catalytic material. Islam, Bhaumik and coworkers[56] reported the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by $\mathrm{{Ag}}$ nanoparticles supported over a bivalent cobalt-based MOF carrier. In 2017, Ma and coworkers[57] utilized Ag@MIL-100(Fe) and Ag@UIO-66(Zr) materials for the capture and transformation of ${\mathrm{{CO}}}_{2}$ into alkynyl carboxylic acids. The influence on the catalytic activity of such Ag@MOF systems were compared and discussed systematically. They supposed that acid-base properties of the MOF influence the catalytic activity of Ag@MOF system significantly. Stronger Lewis acidity favors adsorption of aromatic substrates and activation of the C≡C group, while stronger Lewis basicity is beneficial for the capture of ${\mathrm{{CO}}}_{2}$ and the loading of AgNPs. There are many other Ag-catalyzed carboxylation of terminal alkynes developed in recent years. ${}^{\left\lbrack {58} - {62}\right\rbrack }$
2.2.3. Other metal-catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$
In 2015, Zhao, Yao and coworkers[63] reported the direct carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by rare-earth metal complexes (Scheme 15). The structures of the three rare-earth metal complexes were confirmed by X-ray crystal diffraction. Among the three rare-earth metal complexes, Nd-based catalyst D showed the best activity. Phenylacetylene substrates with either an electron-donating group or an electron-withdrawing group afford the product in high yields, while aliphatic alkynes fail to deliver the expected product. The mechanism was proposed as similar to that of monovalent copper or silver salt catalysts.
REVIEW
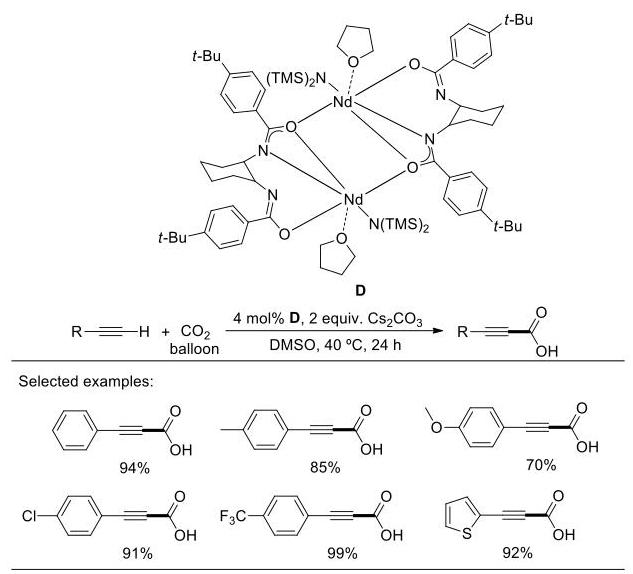
Scheme 15. Complex D-catalyzed carboxylation of terminal alkynes[63]
In 2016, Trivedi, Rath and coworkers ${}^{\left\lbrack {64}\right\rbrack }$ reported the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ catalyzed by MIL-101 supported bimetallic Pd-Cu nanocrystals. In 2018, Ma, Cheng and coworkers[65] investigated the triply bonded dimolybdenum compound ${\mathrm{{Mo}}}_{2}{\left( {\mathrm{O}}^{\prime }\mathrm{{Bu}}\right) }_{6}$ as a homogeneous catalyst for the carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ (Scheme 16). For the carboxylation of diethynylbenzenes, the asymmetric compounds were selectively obtained. Notably, this is the first report of the application of a metal-metal bond-containing species in the catalytic conversion of ${\mathrm{{CO}}}_{2}$ .
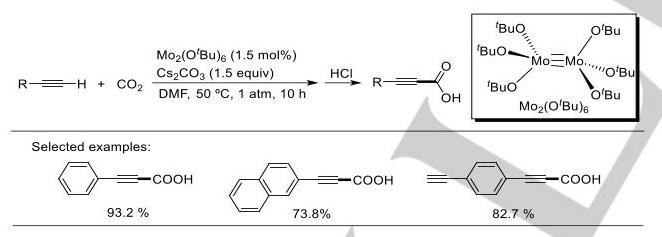
Scheme 16. ${\mathrm{{Mo}}}_{2}{\left( {\mathrm{O}}^{t}\mathrm{{Bu}}\right) }_{6}$ -catalyzed carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {65}\right\rbrack }$
2.2.4. Transition-metal-free carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$
In 2001, Duñach and coworkers ${}^{\left\lbrack {66}\right\rbrack }$ employed electrochemical methods to convert terminal alkynes and ${\mathrm{{CO}}}_{2}$ to alkynyl carboxylic acids. For α,ω-diynes, this method can selectively give monocarboxylated products.
In 2011, Zhang and coworkers[67] reported the direct carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ promoted by ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ (Scheme 17). They found that the reaction temperature and pressure have significant effect on the reaction outcome: high temperature may cause decarboxylation side reactions; ${\mathrm{{CO}}}_{2}$ pressure is positively correlated with reaction rate. Both aromatic and aliphatic alkynes can undergo the carboxylation reaction smoothly. In 2013, Jang, Lee and coworkers[68] reported the direct carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ mediated by 1,5,7-Triazabicyclo [4.4.0] dec-5- ene (TBD). When acetylene is used as the substrate, succinic acid salt can be obtained by catalytic hydrogenation with Pd/C after the carboxylation. Succinic acid can be used to produce some biodegradable polymers (such as polybutylene succinate). Similar to ethylene carbonate, supercritical carbon dioxide is also a green reaction medium. In 2014, Suo, Han and coworkers[69] found that 1, 8-diazabicyclo [5.4.0] undec-7-ene (DBU) can promote the reaction in supercritical carbon dioxide medium (Scheme 18). In addition to facilitating deprotonation of the terminal alkyne, DBU may also play a role in capturing ${\mathrm{{CO}}}_{2}$ . They also reported the carboxylation of terminal alkynes catalyzed by ${\mathrm{{Cul}}}^{\left\lbrack {69}\right\rbrack }$ or ${\mathrm{{AgOAc}}}^{\left\lbrack {70}\right\rbrack }$ in supercritical carbon dioxide medium (Scheme 18). The Liu group has also developed a CsF-promoted carboxylation of aryl(hetoaryl) terminal alkynes with ${\mathrm{{CO}}}_{2}$ (1 atm) ${}^{\left\lbrack {58}\right\rbrack }$ and sequential silylation and carboxylation strategy. ${}^{\left\lbrack {71}\right\rbrack }$
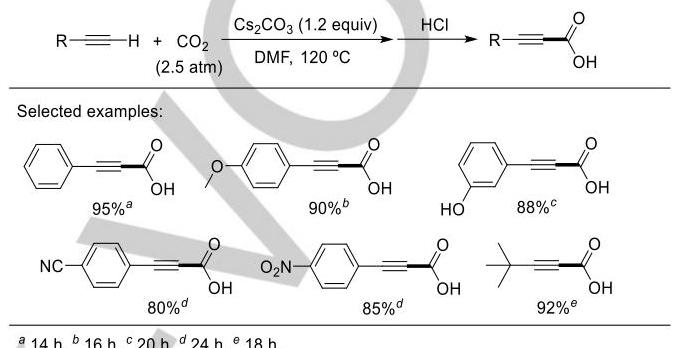
Scheme 17. ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ -promoted carboxylation of terminal alkynes ${}^{\left\lbrack {67}\right\rbrack }$
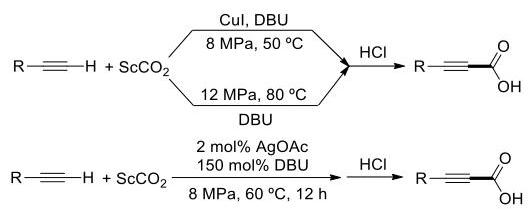
Scheme 18. Carboxylation of terminal alkynes in supercritical ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {69} - {70}\right\rbrack }$
2.3. Application of carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ in the multicomponent synthesis of heterocycles
Compared with traditional multistep synthesis, multicomponent reactions avoid the separation and purification of intermediates, which has the advantages of significantly shortening reaction time and reducing waste generation. In recent years, considerable efforts have been made toward multicomponent reactions of ${\mathrm{{CO}}}_{2}$ .
REVIEW
In 2008, Anastas and coworkers[40] reported Agl-catalyzed multicomponent reactions of phenylacetylene, propargylic bromide and ${\mathrm{{CO}}}_{2}$ . The 1,6-diyine produced by the carboxylative coupling reaction can afford an aryl naphthalene lactone via a [2+2+2] cyclization reaction (Scheme 19). In 2010, further improvements have been made to this reaction ${}^{\left\lbrack {72}\right\rbrack }$ and this approach has also been employed in the synthesis of six arylnaphthalene lactone natural products. ${}^{\left\lbrack {73}\right\rbrack }$ In 2014, Müller and coworkers[74] reported the carboxylation-methylation-Michael addition-cyclocondensation of terminal alkynes catalyzed by $\mathrm{{Cu}}{\left( {\mathrm{{PPh}}}_{3}\right) }_{3}{\mathrm{{NO}}}_{3}$ (Scheme 20). The multicomponent reaction can produce pyrimidones or 3- hydroxypyrazoles without isolation and purification of the intermediate propiolate. Recently, Xu, Gong and coworkers reported a similar reaction for 4H-quinolizin-4-one skeleton in the presence of EtBr. ${}^{\left\lbrack {75}\right\rbrack }$ In 2015, He and coworkers ${}^{\left\lbrack {76}\right\rbrack }$ reported CuCl/PPh3-catalyzed carboxylation-cyclization of terminal alkynes (Scheme 21). ${\mathrm{{PPh}}}_{3}$ is not only the ligand for the carboxylation step but also the catalyst for the second cyclization step. In 2016, Müller and coworkers[77] merged the copper-catalyzed carboxylation-propargylation of terminal alkynes with azido-alkyne cycloaddition to obtain a functionalized 1,2,3-triazole. At the same time, they also found that the above four-component reaction can be further integrated with a Michael reaction to develop a five-component reaction (Scheme 22). In 2016, Liang, Pan and coworkers ${}^{\left\lbrack {78}\right\rbrack }$ reported a copper-catalyzed carboxylative coupling/cycloaddition of terminal alkynes, 1,2-dibromoethane, ${\mathrm{{CO}}}_{2}$ and sodium azide similar to Müller’s reaction (Scheme 23).
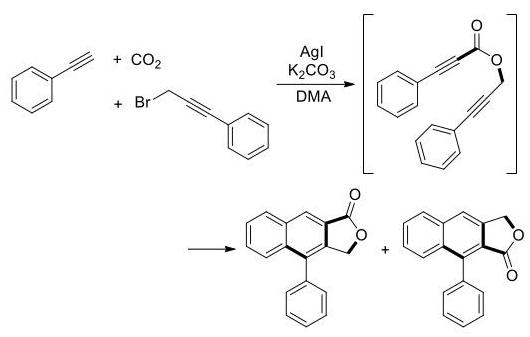
Scheme 19. Synthesis of arylnaphthalene lactones using the Agl-catalyzed muticomponent coupling reaction ${}^{\left\lbrack {40}\right\rbrack }$
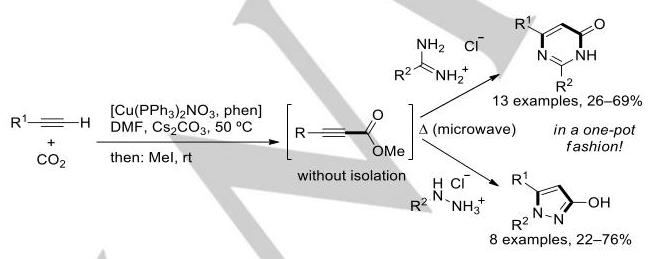
Scheme 20. One-pot synthesis of pyrimidones and 3-hydroxypyrazoles initiated by $\mathrm{{Cu}}\left( \mathrm{I}\right)$ -catalyzed carboxylation of terminal alkynes ${}^{\left\lbrack {74}\right\rbrack }$
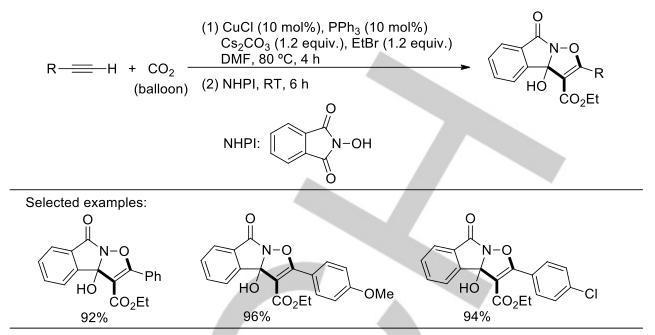
Scheme 21. Copper(I)/PPh3-catalyzed tandem carboxylation/annulation of terminal alkynes ${}^{\left\lbrack {76}\right\rbrack }$
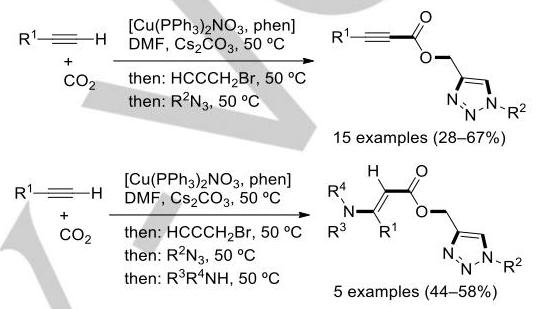
Scheme 22. Copper(I)-catalyzed sequential alkyne carboxylation-propargylation-azide cycloaddition and related five-component reaction[77]
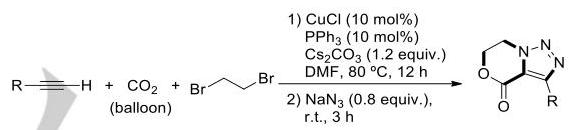
Scheme 23. Copper-catalyzed carboxylative coupling/cycloaddition of terminal alkynes,1,2-dibromoethane, ${\mathrm{{CO}}}_{2}$ and ${\mathrm{{NaN}}}_{3}{}^{\left\lbrack {78}\right\rbrack }$ In 2016, Liang, Pan and coworkers ${}^{\left\lbrack {79}\right\rbrack }$ reported a multicomponent heterocycle synthesis reaction of terminal alkynes, α-haloketones, and ${\mathrm{{CO}}}_{2}$ based on capture and utilization of ${\mathrm{{CO}}}_{2}$ (Scheme 24). Aqueous morpholine solution can be used as an adsorbent to directly capture ${\mathrm{{CO}}}_{2}$ from air. They believe this reaction goes through a copper-catalyzed carboxylative coupling and a base-mediated acyl shift/cyclization/decarbonylation sequence.
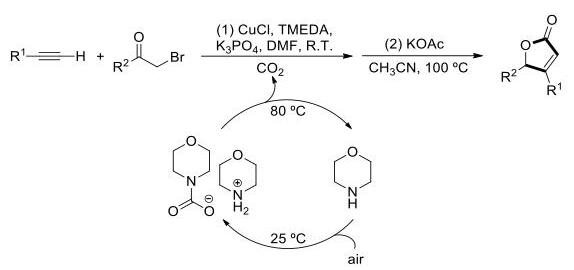
Scheme 24. Muticomponent reaction of terminal alkynes, α-haloketones and ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {79}\right\rbrack }$
REVIEW
2.4. Application of carboxylation of terminal alkynes with ${\mathrm{{CO}}}_{2}$ in the hydrothiolation of alkynes
Transition metal-catalyzed hydrothiolation of alkynes is an important method for the synthesis of vinyl sulfides. This reaction faces challenges of regioselectivity and stereoselectivity. In 2012, Ying, Zhang and coworkers ${}^{\left\lbrack {80}\right\rbrack }$ found that the stereoselectivity of the reaction can be modulated by changing the atmosphere under which the reaction proceeds (Scheme 25). Z-vinyl sulfides can be stereoselectively obtained under ${\mathrm{{CO}}}_{2}$ atmosphere, while $E$ -vinyl sulfides are mainly obtained under argon or air. A small amount of water can speed up the reaction. A possible reaction mechanism was proposed: in the ${\mathrm{{CO}}}_{2}$ atmosphere, the terminal alkynes can be converted into propiolate, and then a stereoselective decarboxylative hydrothiolation reaction occurs. Finally, the Z-vinyl sulfide is selectively obtained.
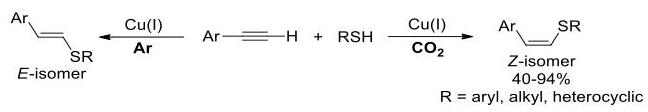
Scheme 25. Stereoselective hydrothiolation of terminal alkynes[80]
3. Carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$
3.1. Carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds of (hetero)arenes with ${\mathrm{{CO}}}_{2}$
3.1.1. Chelation-free carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds of (hetero)arenes with ${\mathrm{{CO}}}_{2}$
3.1.1.1. Transition-metal-catalyzed chelation-free carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds of (hetero)arenes with ${\mathrm{{CO}}}_{2}$
In 1984, Fujiwara and coworkers ${}^{\left\lbrack {81}\right\rbrack }$ reported a divalent palladium-catalyzed direct carboxylation of aromatic compounds (benzene, anisole, furan, thiophene) with ${\mathrm{{CO}}}_{2}$ . The pressure of ${\mathrm{{CO}}}_{2}$ can be one atmosphere, but the catalyst has a low turnover number (TON) (for benzene, TON = ca. 1.3). In 2014, based on their previous work ${}^{\left\lbrack {82}\right\rbrack }$ , Iwasawa and coworkers reported the Rh(I)- catalyzed direct carboxylation of $\mathrm{C} - \mathrm{H}$ bonds of simple arenes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {83}\right\rbrack }$ (Scheme 26). The use of bulky bidentate phosphine ligands and addition of 1,1,3,3-tetramethylurea (TMU) as a co-solvent may serve to stabilize the coordinatively unsaturated Rh(I) species. Both electron-rich and electron-deficient arenes can participate in this reaction (for benzene, TON = 37). Steric hindrance has a great influence on the regioselectivity of this reaction. For benzofuran and 1-methylindole, the reaction occurs selectively at the C-2 position. The reaction mechanism they proposed is similar to their previous work. ${}^{\left\lbrack {82}\right\rbrack }$ In 2017, Iwasawa and coworkers ${}^{\left\lbrack {84}\right\rbrack }$ studied the mechanism of rhodium-catalyzed carboxylation of simple aromatic compounds with ${\mathrm{{CO}}}_{2}$ . 14- Electron complexes, RhMe(dcype) and RhPh(dcype), participated in the $\mathrm{C} - \mathrm{H}$ bond activation step and the carboxylation step, respectively. Interestingly, the undesired carboxylation of
RhMe(dcype) to give acetic acid was found to be much faster than the desired C-H bond activation reaction under stoichiometric conditions; however, the C-H bond activation reaction could occur under catalytic conditions. In 2012, Hölscher, Leitner and coworkers ${}^{\left\lbrack {85}\right\rbrack }$ studied the direct carboxylation of $\mathrm{C} - \mathrm{H}$ bonds of unactivated arenes (such as benzene) with ${\mathrm{{CO}}}_{2}$ by DFT calculations. The study has shown that some ruthenium pincer complexes can catalyze the reaction theoretically, and the predicted catalyst turnover frequency (TOF) can be up to ${10}^{5} - {10}^{7}{\mathrm{\;h}}^{-1}$ . In 2014, they ${}^{\left\lbrack {86}\right\rbrack }$ conducted further research on the effect of spatial configuration of the ruthenium pincer complexes on catalysis by DFT calculations.
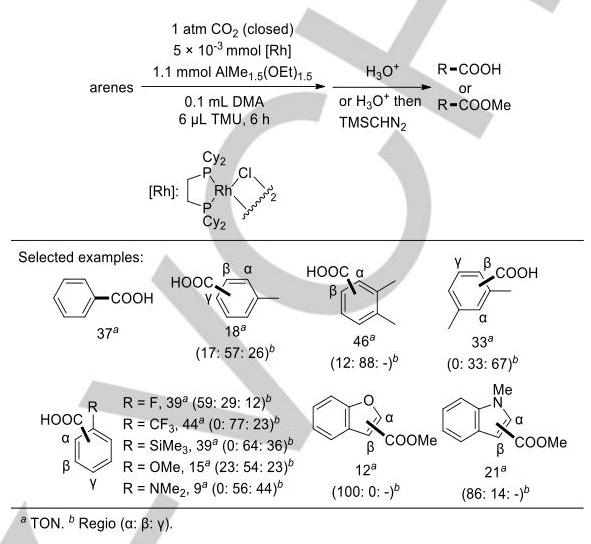
Scheme 26. Rh(I)-catalyzed direct carboxylation of C-H bond of simple arenes ${}^{\left\lbrack {83}\right\rbrack }$
In 2010, Nolan and coworkers[87] reported the carboxylation of electron-deficient heterocycles and aromatic rings catalyzed by Au(I)-NHC complex [(IPr)AuOH] (Scheme 27). The reaction occurs at the most acidic C-H bond regioselectively. For the carboxylation of weakly acidic C-H bonds such as N-methylimidazole and $N$ -methylbenzimidazole, it is necessary to use a more basic catalyst [(ItBu)AuOH]. Catalyst recovery can be achieved by simple extraction. The authors isolated and characterized the reaction intermediates (Scheme 28) and applied the intermediates to catalytic carboxylation. Thus, a possible reaction mechanism was proposed. Notably, in 2011, based on their experimental and theoretical calculations, Ahlquist, Wendt and coworkers[88] questioned the mechanism of [(IPr)AuOH]-catalyzed carboxylation of electron-deficient (hetero)aromatics proposed by Nolan and coworkers. In 2012, Geng and coworkers ${}^{\left\lbrack {89}\right\rbrack }$ conducted theoretical studies on the mechanism of oxazole with ${\mathrm{{CO}}}_{2}$ catalyzed by $\mathrm{{Au}}\left( \mathrm{I}\right) - \mathrm{{NHC}}$ complexes. In 2010, Hou ${}^{\left\lbrack {90}\right\rbrack }$ and Nolan ${}^{\left\lbrack {91}\right\rbrack }$ individually reported similar carboxylations of electron-deficient heterocycles or aromatic rings catalyzed by $\mathrm{{Cu}}\left( \mathrm{I}\right) - \mathrm{{NHC}}$ complex. Hou and coworkers[90] used [Cu(IPr)Cl] as catalyst and 1.1 equivalents of potassium $t$ -butoxide as the base (Scheme 29a). The substrate scope covers various substituted benzoxazoles. The yields of less acidic N-methylbenzimidazoles and 1,3,4-oxadiazoles are lower, while benzothiazole, 4-phenyloxazole and benzofuran give only trace amounts of carboxylated products. Similar to Nolan and coworkers’ [(IPr)AuOH] catalytic system[87], Hou and coworkers also isolated and characterized key intermediates for their reactions (including X-ray crystal diffraction) and proved that these key intermediates can catalyze the carboxylation of benzoxazoles. The reaction mechanism they proposed is similar to that of Nolan and coworkers’ [(IPr)AuOH] catalytic system[ ${}^{87}$ ]. Ariafard, Yates and coworkers[ ${}^{92}$ ] studied the mechanism of C-H carboxylation of electron-deficient heteroarenes catalyzed by Cu(I)-NHC complexes by DFT calculations. They proposed a reaction mechanism different from that of Hou and coworkers: it is energetically more favorable for a metal-stabilized carbene intermediate react with ${\mathrm{{CO}}}_{2}$ to give carboxylated products (Scheme 30). The [Cu(IPr)OH]-CsOH system reported by Nolan and coworkers ${}^{\left\lbrack {91}\right\rbrack }$ can be applied to the carboxylation of benzoxazole, benzothiazole, oxazole, 1,2,4-trifluorobenzene and 1,2,4,5-tetrafluorobenzene (Scheme 29b). For 1,2,4,5- tetrafluorobenzene, its monocarboxylation and dicarboxylation can be modulated by controlling the amount of base. In 2010, considering that benzoxazole have similar acidity with terminal alkynes, Gooßen and coworkers [41] applied their copper-complex catalytic system to the carboxylation of benzoxazole. In 2012, Fukuzawa and coworkers ${}^{\left\lbrack {93}\right\rbrack }$ reported the carboxylation of electron-deficient benzoxazoles and benzothiazoles catalyzed by 1,2,3-triazole-derived Cu(I)-NHC complexes [(TPr)CuCl] (Scheme 31). Compared with the [Cu(IPr)Cl] catalyst used by Hou and coworkers, [(TPr)CuCl] has higher catalytic activity. In 2017, Hong and coworkers[94] developed diethylene glycol-functionalized imidazo[1,5-a]pyridin-3-ylidenes (DEG-ImPy) bifunctional NHC ligands. DEG-ImPy Cu(I) complexes can efficiently catalyze the direct $\mathrm{C} - \mathrm{H}$ carboxylation of benzoxazole with ${\mathrm{{CO}}}_{2}$ , giving higher isolated yields than those with the (IPr)CuCl catalyst (Scheme 32).
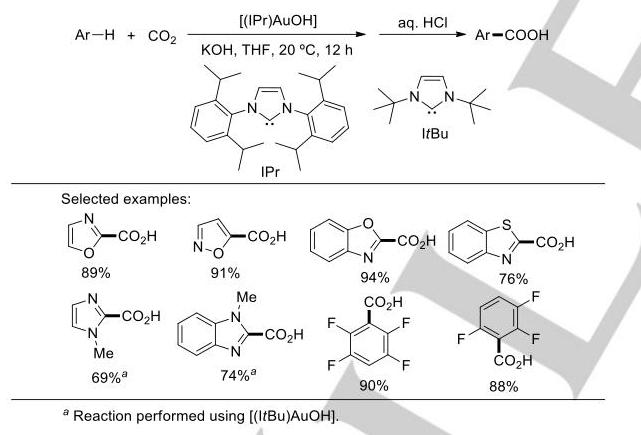
Scheme 27. [(IPr)AuOH]-catalyzed carboxylation of C-H bond of electron-deficient (hetero)arenes ${}^{\left\lbrack {87}\right\rbrack }$
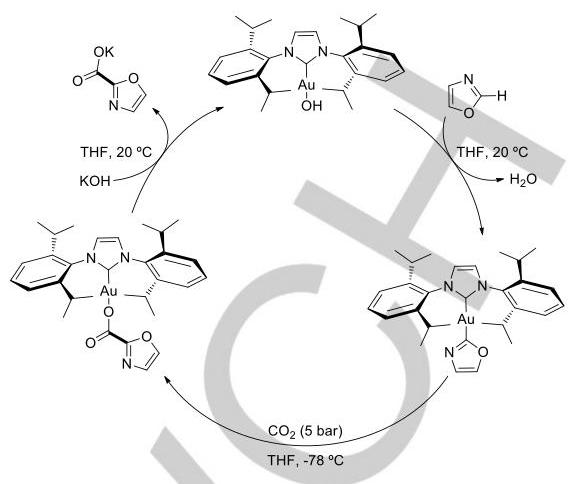
Scheme 28. Possible intermediates involved in the [(IPr)AuOH]-catalyzed carboxylation of $\mathrm{C} - \mathrm{H}$ bond of oxazole ${}^{\left\lbrack {87}\right\rbrack }$
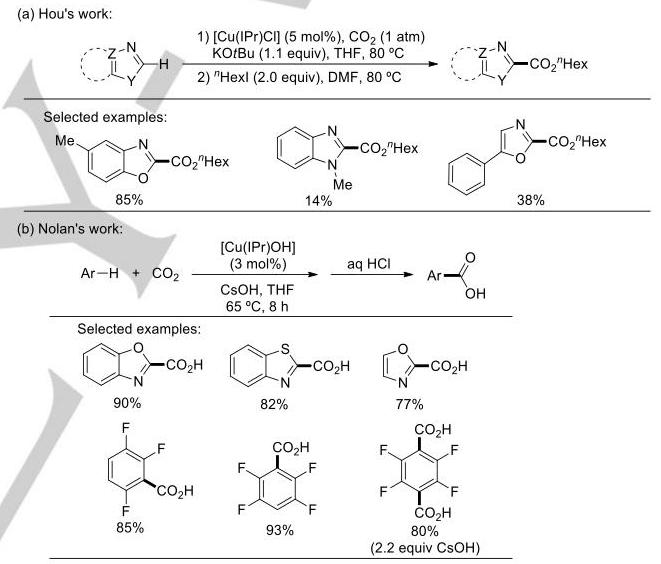
Scheme 29. [Cu(IPr)Cl]- or [Cu(IPr)OH]-catalyzed carboxylation of C-H bond of electron-deficient (hetero)arenes[90-91]
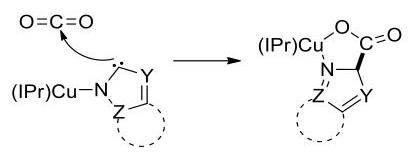
Scheme 30. Reaction of the metal-stabilized carbene intermediate with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {92}\right\rbrack }$
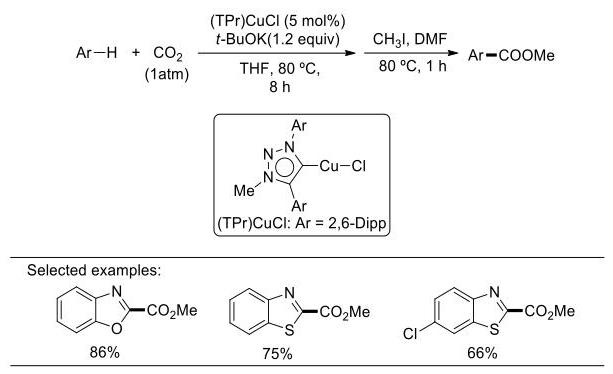
Scheme 31. [(TPr)CuCl]-catalyzed carboxylation of $\mathrm{C} - \mathrm{H}$ bond of benzoxazole and benzothiazole derivatives ${}^{\left\lbrack {93}\right\rbrack }$
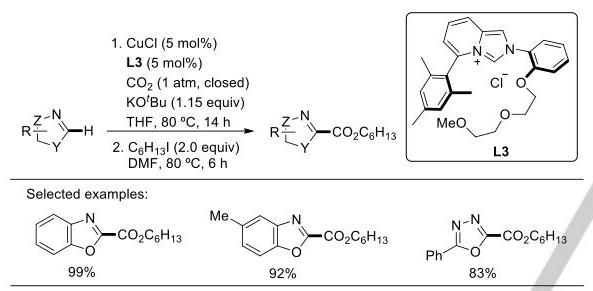
Scheme 32. The direct $\mathrm{C} - \mathrm{H}$ carboxylation of benzoxazole with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {94}\right\rbrack }$
Catalyst recovery is a problem that limits the wide application of homogeneous catalysis. In 2013, Li and coworkers[95] reported the carboxylation of benzoxazole derivatives catalyzed by a pH-responsive Cu(I)-NHC complex E (Scheme 33). Unlike the [Cu(IPr)Cl] catalyst[90] used by Hou and coworkers, a basic tertiary amine functionality was introduced into Li and coworkers’ catalyst. This type of catalyst is soluble under basic carboxylation conditions. However, when HCI diethyl ether is added after the reaction, the protonated salt of the catalyst will precipitate from the solution. The catalyst can be separated from product by simple centrifugation. And when new substrates and bases are added to the recovered catalyst, the protonated catalyst can be regenerated in the form of a neutral homogeneous catalyst. Moreover, the catalyst can be used for four times without significant drop in catalytic activity. This catalyst can also be used to catalyze the carboxylation of organoboronic esters, while a similar Ag(I)-NHC complex catalyst can catalyze the carboxylation of terminal alkynes.
In 2018, Ma, Cheng and coworkers[65] reported the carboxylation of benzothiazoles and benzoxazoles with ${\mathrm{{CO}}}_{2}$ catalyzed by ${\mathrm{{Mo}}}_{2}{\left( {\mathrm{O}}^{t}\mathrm{{Bu}}\right) }_{6}$ (Scheme 34).
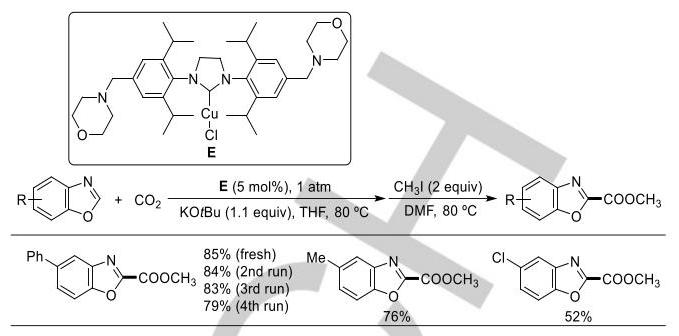
Scheme 33. Complex E-catalyzed carboxylation of benzoxazole derivatives ${}^{\left\lbrack {95}\right\rbrack }$
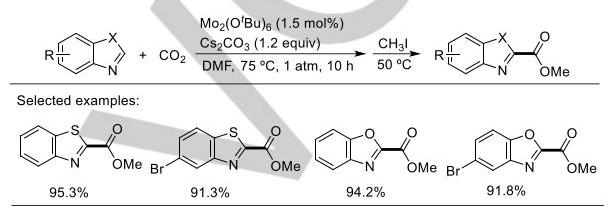
Scheme 34. ${\mathrm{{Mo}}}_{2}{\left( {\mathrm{O}}^{\prime }\mathrm{{Bu}}\right) }_{6}$ -catalyzed carboxylation of benzothiazoles and benzoxazoles with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {65}\right\rbrack }$
3.1.1.2. Transition-metal-free and chelation-free carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds of (hetero)arenes with ${\mathrm{{CO}}}_{2}$
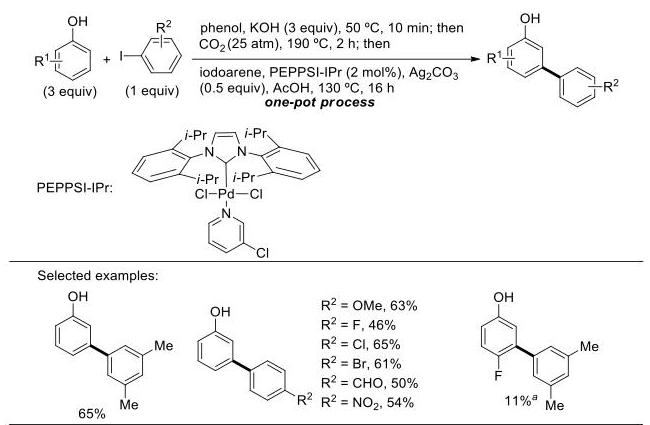
The Kolbe-Schmitt reaction ${}^{\left\lbrack {96}\right\rbrack }$ is one of the most important carboxylation reactions, which refers to the direct ortho C-H carboxylation of phenoxide with ${\mathrm{{CO}}}_{2}$ . This reaction is employed for the industrial synthesis of salicylic acid. In 2014, Larrosa and coworkers[97] reported a “one-pot” direct meta-selective arylation reaction of phenol via a traceless directing group relay strategy (Scheme 35). They first exploited the o/p-directing nature of phenol to react with ${\mathrm{{CO}}}_{2}$ to generate ortho- or para-substituted carboxylic acids (Kolbe-Schmitt reaction), followed by ${\mathrm{{CO}}}_{2}\mathrm{H}$ group-directed ortho-C-H arylation reaction and decarboxylation of carboxylic acids, and finally a phenol meta-arylated product was produced. In this process, ${\mathrm{{CO}}}_{2}$ acts as a traceless directing group. Notably, regardless of the regioselectivity of the carboxylation reaction (ortho or para), the meta-aryl phenol products can be finally obtained. The Kolbe-Schmitt reaction generally requires harsh reaction conditions: high ${\mathrm{{CO}}}_{2}$ pressure (20-100 atm) and high temperature (130-280 °C). In 2016, Larrosa and coworkers ${}^{\left\lbrack {98}\right\rbrack }$ first reported a Kolbe-Schmitt reaction that can be performed at atmospheric pressure (Scheme 36). Using 2,4,6-trimethylphenol as a recyclable additive is a key to success of this reaction. This method avoids separation of phenoxides and has a broader range of substrates. They believe that 2,4,6-trimethylphenol in the reaction may facilitate ${\mathrm{{CO}}}_{2}$ capture. In addition, enzymatic Kolbe-Schmitt reactions have also been reported ${}^{\left\lbrack {99} - {104}\right\rbrack }$ . For example, in 2012, Faber and coworkers ${}^{\left\lbrack {100}\right\rbrack }$ found that different types of enzymes can modulate the regioselectivity of $p$ -hydroxystyrene carboxylation (Scheme 37). In 2016, Zhu and coworkers [104] found that addition of quaternary ammonium salt to the reaction system can shift the equilibrium of enzymatic Kolbe-Schmitt reaction towards carboxylation. The precipitation capacity of the quaternary ammonium salt of the carboxylation product is the driving force for reaction equilibrium shift. In 1995, Kudo and coworkers ${}^{\left\lbrack {105}\right\rbrack }$ reported the carboxylation of alkali-metal 2-naphthoate with high-pressure ${\mathrm{{CO}}}_{2}\left( {{400}\mathrm{{atm}}}\right)$ promoted by molten alkali-metal carbonate and formate. In 2010, Hu and coworkers ${}^{\left\lbrack {106}\right\rbrack }$ found that ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ can facilitate $\mathrm{C} - \mathrm{H}$ bond carboxylation of some aromatic heterocycles such as benzothiazole, benzoxazole, 5-phenyl oxazole, and 2-aryl-1,3,4- oxadiazole (Scheme 38) without use of any transition metal catalysts. They proposed the following reaction mechanism: first, the most acidic C-H bond in the aromatic heterocycle is cleaved with ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ , and the resulting anion reacts with ${\mathrm{{CO}}}_{2}$ to form a new C-C bond (Scheme 38). In 2016, Kanan and coworkers[107] reported the direct carboxylation of the C-H bond of 2-furoic acid promoted by carbonate (Scheme 39). 2-Furoic acid can be easily obtained from lignocellulose. The furan-2,5-dicarboxylic acid produced by carboxylation of 2-furoic acid has wide applications. For example, it can be used to synthesize polyethylene furandicarboxylate (PEF), a substitute for petroleum-derived polyethylene terephthalate (PET). It is believed that ${\mathrm{{CO}}}_{3}{}^{2 - }$ in the molten salt of ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ or ${\mathrm{K}}_{2}{\mathrm{{CO}}}_{3}$ can deprotonate furan-2- carboxylate, while the high concentration of alkali metal cations can stabilize the conjugate base by ion pairing. If ${\mathrm{{CO}}}_{3}{}^{2\text{ - }}$ -promoted C-H carboxylation reaction is combined with protonation of the carboxylate salt and regeneration of ${\mathrm{{CO}}}_{3}{}^{2 - }$ , then the net reaction would be the transformation of $\mathrm{C} - \mathrm{H}$ bond and ${\mathrm{{CO}}}_{2}$ into $\mathrm{C} - {\mathrm{{CO}}}_{2}\mathrm{H}$ . In addition, ${\mathrm{{CO}}}_{2}$ -promoted esterification can convert carboxylates into esters and regenerate ${\mathrm{{CO}}}_{3}{}^{2 - }$ (Scheme 40). The authors also examined C-H carboxylation of caesium 2-thiophene carboxylate, caesium benzoate and benzene with ${\mathrm{{CO}}}_{2}$ . Recently, Fenner and Ackermann ${}^{\left\lbrack {108}\right\rbrack }$ reported a heterocyclic C-H bond carboxylation reaction similar to that of Hu and coworkers ${}^{\left\lbrack {106}\right\rbrack }$ (Scheme 41). The main difference is that Fenner and Ackermann ${}^{\left\lbrack {108}\right\rbrack }$ used potassium tert-butoxide as the base to make carboxylation conditions milder (lower reaction temperature, one atmosphere of ${\mathrm{{CO}}}_{2}$ ). In 2002, Hattori, Miyano and coworkers ${}^{\left\lbrack {109}\right\rbrack }$ reported the direct carboxylation of naphthalene, anthracene and phenanthrene with ${\mathrm{{CO}}}_{2}$ promoted by two equivalents of ${\mathrm{{AlBr}}}_{3}$ . In 2002, Olah, Prakash and coworkers ${}^{\left\lbrack {110}\right\rbrack }$ found that the ${\mathrm{{Al}}}_{2}{\mathrm{{Cl}}}_{6}/\mathrm{{Al}}$ system promotes direct carboxylation of benzene or substituted benzene with ${\mathrm{{CO}}}_{2}$ (Scheme 42). For mesitylene, the reaction can be carried out at 20 °C. For some alkylbenzenes (such as ethylbenzene, tert-butylbenzene, and p-xylene), side reaction pathways such as isomerization or disproportionation lead to a decrease in product yield. Addition of metallic aluminum can significantly increase the yield and selectivity of the reaction. They believe the role of aluminum may be to react with HCI generated in the carboxylation reaction to suppress other side reactions, and at the same time to generate part of aluminum chloride in situ. Based on experimental and theoretical calculations, they proposed a reaction mechanism. Aluminum chloride can electrophilically activate ${\mathrm{{CO}}}_{2}$ , and then electrophilic substitution on the aromatic ring occurs. Hattori and coworkers have done a great deal of work on the electrophilic carboxylation of aromatic rings with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {111} - {113}\right\rbrack }$ . For example, they found that using ${\mathrm{{AlBr}}}_{3}$ and ${\mathrm{R}}_{3}\mathrm{{SiCl}}$ together can significantly promote direct carboxylation of aromatics ${}^{\left\lbrack {112}\right\rbrack }$ . Also, this system solves the disproportionation problem often occuring in the carboxylation of ethylbenzene and $t$ -butylbenzene ${}^{\left\lbrack {110}\right\rbrack }$ . In 2012, Müller [114] and Hattori [113] independently reported the carboxylation of aromatics with ${\mathrm{{CO}}}_{2}$ mediated by silyl cation. In 2009, Munshi and coworkers ${}^{\left\lbrack {115}\right\rbrack }$ found that pre-reacting ${\mathrm{{CO}}}_{2}$ with Lewis acid for a period before adding toluene can promote the carboxylation of toluene. This phenomenon is consistent with the mechanism proposed by Olah, Prakash and coworkers[110] In 2010, Munshi and coworkers[116] achieved a catalytic cycle of ${\mathrm{{Al}}}_{2}{\mathrm{{Cl}}}_{6}$ used in the carboxylation of toluene through rational selection of fluorine-containing solvents and bases. In 2012, Munshi and coworkers ${}^{\left\lbrack {117}\right\rbrack }$ further studied the kinetics of carboxylation of toluene. In 2014, Gu and Cheng ${}^{\left\lbrack {118}\right\rbrack }$ reported the carboxylation of toluene promoted by aluminum-supported aluminum chloride. In 2017, Cheng and coworkers[119] further showed that the addition of silica compounds can promote ${\mathrm{{AlCl}}}_{3}$ catalytic activity in carboxylation of toluene. In 2016, Ma and coworkers[120] made a theoretical investigation of boron-doped lithium clusters, ${\mathrm{{BLi}}}_{\mathrm{n}}\left( {n = 3 - 6}\right)$ , on the carboxylation of simple aromatic compounds with ${\mathrm{{CO}}}_{2}$ . In 2009, Hattori and coworkers ${}^{\left\lbrack {121}\right\rbrack }$ reported the carboxylation of 1-substituted indoles and pyrrole promoted by equimolar amounts of dimethylaluminum chloride. In 2016, they ${}^{\left\lbrack {122}\right\rbrack }$ further extended the substrate scope for this reaction (Scheme 43) and proposed a possible reaction mechanism. For 1-unsubstituted indole, the yield was poor. In 2012, the ${y}^{\left\lbrack {123}\right\rbrack }$ reported similar carboxylation of thiophene and benzothiophene assisted by equimolar amounts of ethylaluminum dichloride. In 2012, Kobayashi and coworkers[124] reported the carboxylation of unprotected indole derivatives with ${\mathrm{{CO}}}_{2}$ mediated by lithium tert-butoxide (Scheme 44). The key of this reaction is the use of a large excess of lithium tert-butoxide as base to inhibit decarboxylative side reaction. Their reaction mechanism is as follows. First, lithium tert-butoxide captures the proton of $\mathrm{N} - \mathrm{H}$ , and the resulting anion reacts with ${\mathrm{{CO}}}_{2}$ to form $\mathrm{N} -$ centered or C-3 carboxylates. Since the carboxylation reaction is reversible, C-3 carboxylates are generated at high temperatures. Notably, the free NH of unprotected indole derivatives plays an important role in the reaction. 1-Methyl indole cannot undergo this carboxylation under the same conditions, which demonstrates that deprotonation by lithium tert-butoxide is a key step in this reaction. In 2015, they ${}^{\left\lbrack {125}\right\rbrack }$ extended the reaction to the carboxylation of pyrrole derivatives with ${\mathrm{{CO}}}_{2}$ . In the same year, Skrydstrup and coworkers[ ${}^{126}$ ] reported the cyclization of 2-alkynyl indole using a catalytic amount of organic base TBD and five equivalents of ${\mathrm{{CO}}}_{2}$ (Scheme 45). Cyclization of aryl-, heteroaryl-, aliphatic-, alkenyl-substituted 2-alkynyl indole, and 2-terminal 2- alkynyl indole are all viable. They believed the reaction went through the following steps. First, TBD reacts with ${\mathrm{{CO}}}_{2}$ to generate an addition product. Then the C-3 position of the indole attacks the addition product or directly attacks ${\mathrm{{CO}}}_{2}$ to generate a new C-C bond. Next, a rearomatized intermediate is obtained after deprotonation. This intermediate undergoes C-O bond formation to give the final product, while regenerating the TBD catalyst (Scheme 46). In 2017, Wong and coworkers[127] performed DFT calculations for this reaction. They proposed a bifunctional activation of indole and ${\mathrm{{CO}}}_{2}$ simultaneously by the bicyclic guanidine catalyst. In 2016, Ma and coworkers[128] conducted a theoretical study of the $\mathrm{C} - \mathrm{H}$ bond insertion reaction of pyrrole or $N$ -methylpyrrole with short-lived ${\mathrm{{CO}}}_{2}$ . In 2016, Xi and coworkers ${}^{\left\lbrack {129}\right\rbrack }$ reported the lactamization reaction of o-arylaniline with ${\mathrm{{CO}}}_{2}$ mediated by MeOTf and TBD (Scheme 47). They propose that isocyanate may be the active intermediate in this metal-free reaction.
${}^{a}$ Yield determined by ${}^{1}\mathrm{H}$ NMR analysis using an internal standard.
Scheme 35. The one-pot direct meta-selective arylation of phenols based on a traceless directing group relay strategy ${}^{\left\lbrack {97}\right\rbrack }$
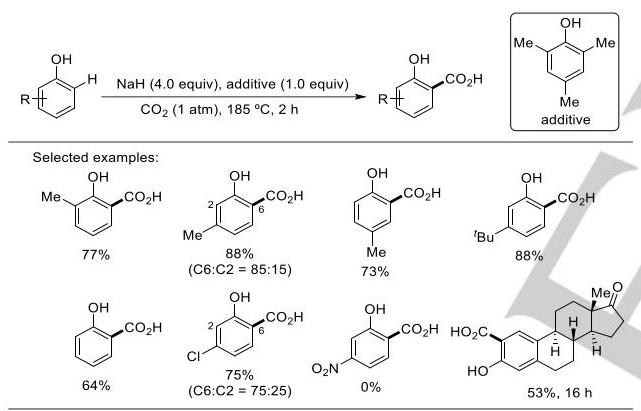
Scheme 36. Kolbe-Schmitt carboxylation at atmospheric pressure ${}^{\left\lbrack {98}\right\rbrack }$
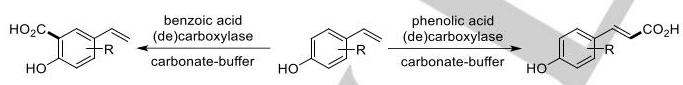
Scheme 37. Enzymatic regioselective carboxylation of hydroxystyrene derivatives ${}^{\left\lbrack {100}\right\rbrack }$
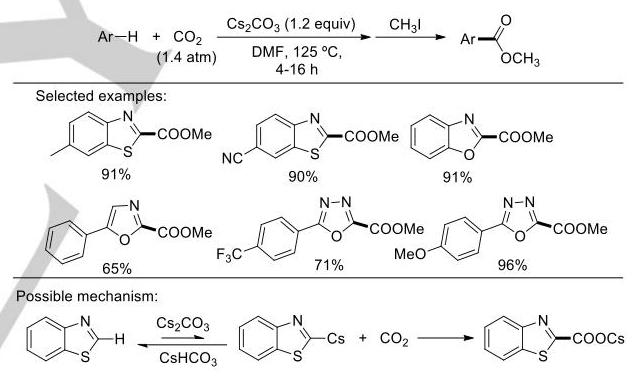
Scheme 38. ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ -promoted carboxylation of $\mathrm{C} - \mathrm{H}$ bond of electron-deficient aromatic heterocycles[106]
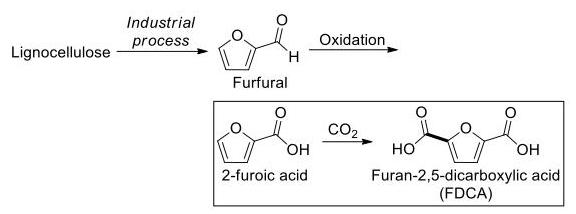
Scheme 39. Synthesis of furan-2,5-dicarboxylic acid from lignocellulose using direct carboxylation of 2-furoic acid[107]
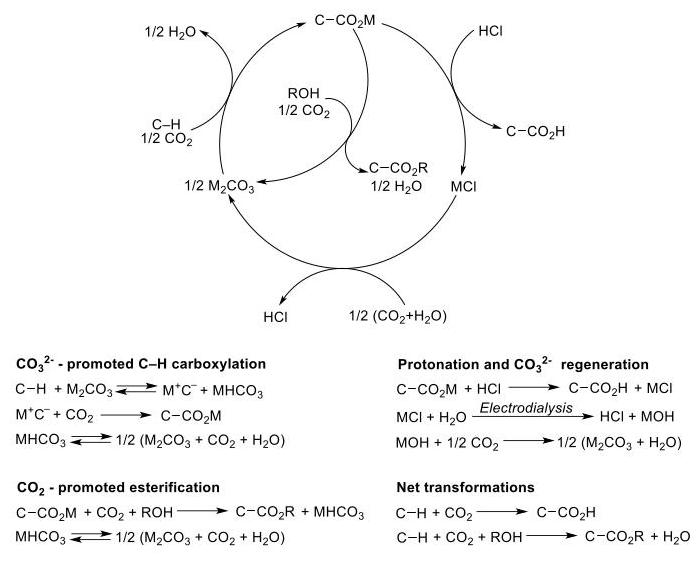
Scheme 40. ${\mathrm{{CO}}}_{2}$ utilization cycle ${}^{\left\lbrack {107}\right\rbrack }$
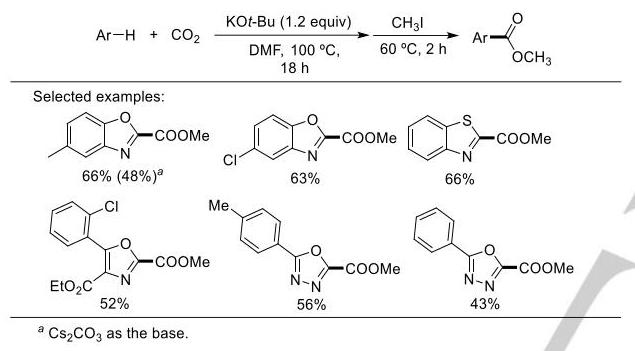
Scheme 41. KOt-Bu-promoted carboxylation of C-H bond of electron-deficient aromatic heterocycles[108]
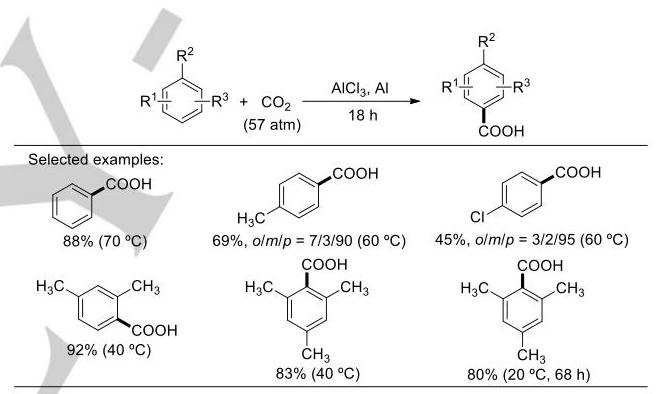
Scheme 42. ${\mathrm{{AICl}}}_{3}/\mathrm{{Al}}$ -promoted carboxylation of benzene and substituted aromatics[110]
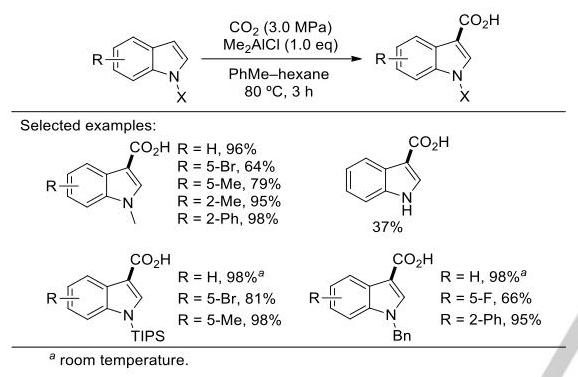
Scheme 43. Me2AICI-mediated carboxylation of indoles ${}^{\left\lbrack {122}\right\rbrack }$
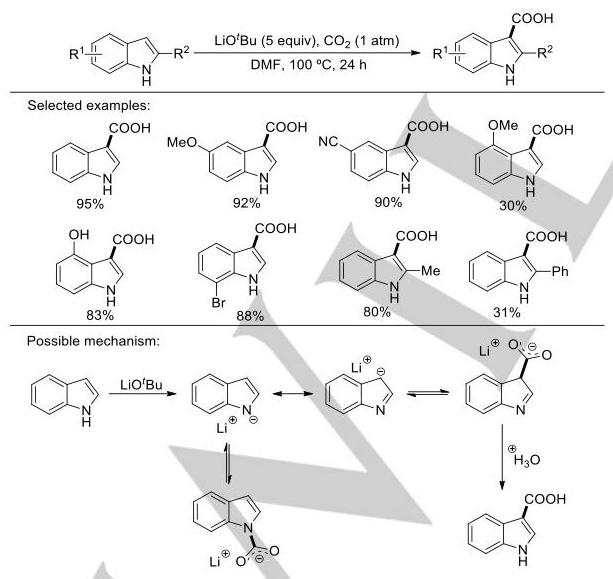
Scheme 44. LiO’Bu-mediated carboxylation of unprotected indole derivatives[124]
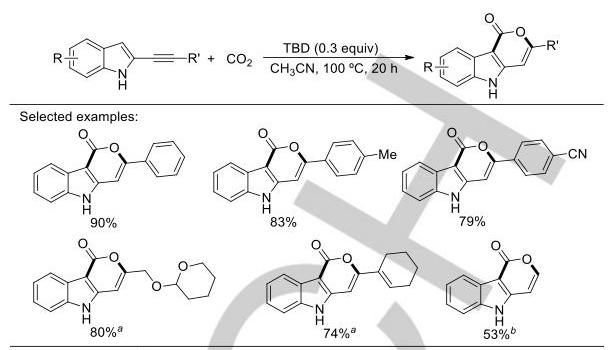
${}^{a}$ TBD: 1.0 equiv. ${}^{b}$ TBD: 3.0 equiv,70 ${}^{ \circ }\mathrm{C}$ .
Scheme 45. TBD-catalyzed/promoted cyclization of 2-alkynyl indoles with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {126}\right\rbrack }$
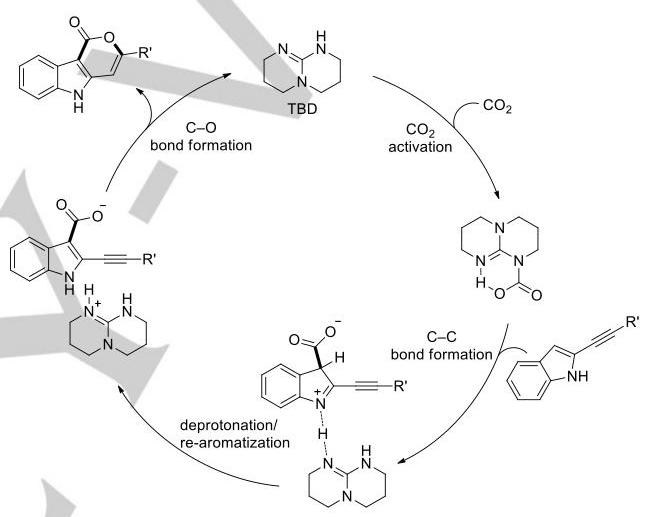
Scheme 46. Possible mechanism for TBD-catalyzed/promoted cyclization of 2- alkynyl indoles with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {126}\right\rbrack }$
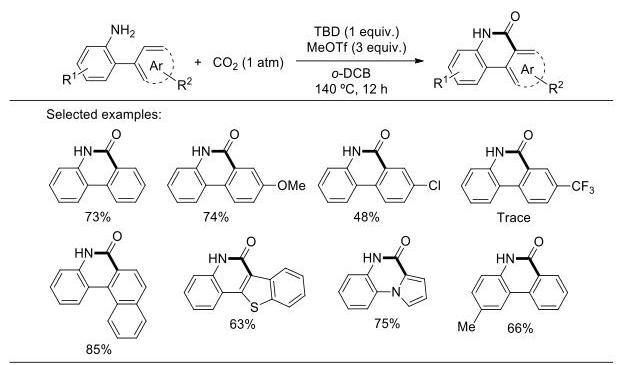
Scheme 47. MeOTf- and TBD-mediated lactamization of ortho-arylanilines with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {129}\right\rbrack }$
In 2017, Yu, Zhi and coworkers[130] developed a novel lactonization of heteroaryl and alkenyl $\mathrm{C} - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ to synthesize important coumarin derivatives under transition-metal-free conditions (Scheme 48).
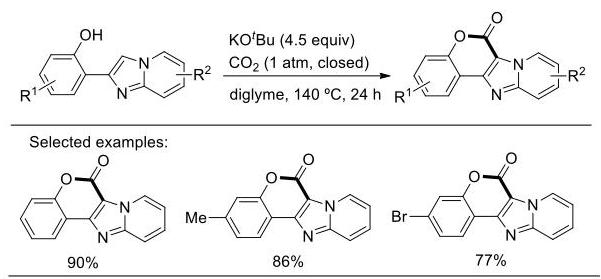
Scheme 48. The lactonization of heteroaryl C-H bonds with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {130}\right\rbrack }$
3.1.2. Chelation-assisted carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds of (hetero)arenes with ${\mathrm{{CO}}}_{2}$
In 2011, Iwasawa and coworkers ${}^{\left\lbrack {82}\right\rbrack }$ reported the Rh(I)-catalyzed chelation-assisted direct carboxylation of aryl $\mathrm{C} - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ (Scheme 49). They found that 2-arylpyridines and 1- arylpyrazoles could undergo C-H bond activation to produce the corresponding carboxylated products in the presence of Rh(I) catalyst and two equivalents of ${\mathrm{{AIMe}}}_{2}\left( \mathrm{{OMe}}\right)$ . The choice of phosphine ligand is critical for the reaction: more sterically hindered monodentate phosphine ligands such as $\mathrm{P}{\left( \mathrm{{mes}}\right) }_{3}$ and ${\mathrm{{PCy}}}_{3}$ perform better, and bidentate phosphine ligands are detrimental to the reaction. The reaction mechanism is proposed as follows: first, Rh(I)CI reacts with methylaluminum reagent to generate methyl rhodium (I); then, chelation-assisted aryl C-H bond activation (oxidative addition) generates aryl trivalent rhodium, and reductive elimination of methane produces the key monovalent aryl rhodium species; this species undergoes nucleophilic addition to ${\mathrm{{CO}}}_{2}$ and then transmetalation with the methylaluminum reagent to obtain aluminium carboxylate, while methyl rhodium (I) is regenerated (Scheme 49). In 2017, Lu and coworkers[131] investigated the mechanism and effects of ligands and reagents in Rh-catalyzed C(sp ${}^{2}$ )-H bond carboxylation with ${\mathrm{{CO}}}_{2}$ through DFT calculations. Based on DFT calculations, a similar mechanism was proposed (Scheme 50). In the catalytic cycle, ${\mathrm{{CO}}}_{2}$ insertion is the rate-determining step. In addition to its role as a methylation reagent, ${\mathrm{{AlMe}}}_{2}\left( \mathrm{{OMe}}\right)$ can also serve as a Lewis acid to promote ${\mathrm{{CO}}}_{2}$ insertion. The origin of the promoting effect of $\mathrm{P}{\left( \text{ mes }\right) }_{3}$ on the reactivity is derived from the stabilizing effect due to the favorable agostic interaction of Rh with the ortho-methyl group of P(mes)3. In 2016, Ma and coworkers[132] successfully inserted ${\mathrm{{CO}}}_{2}$ into aryl $\mathrm{C} - \mathrm{H}$ bonds on the backbone of metal-organic framework UiO-67(dcppy) to generate free carboxylate groups (Scheme 51). Their catalytic system and the proposed reaction mechanism are similar to those of Iwasawa and coworkers[82]. UiO-67(dcppy)-COOH obtained by this reaction can be used as a heterogeneous Brønsted acid catalyst to catalyze the methanolysis of epoxides. In 2014, Mita and Sato and coworkers[133] found that 1- arylpyrazole derivatives can sequentially undergo ruthenium-catalyzed $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ silylation and fluoride-mediated carboxylation (Scheme 52). They believe the pyrazole group can play two roles
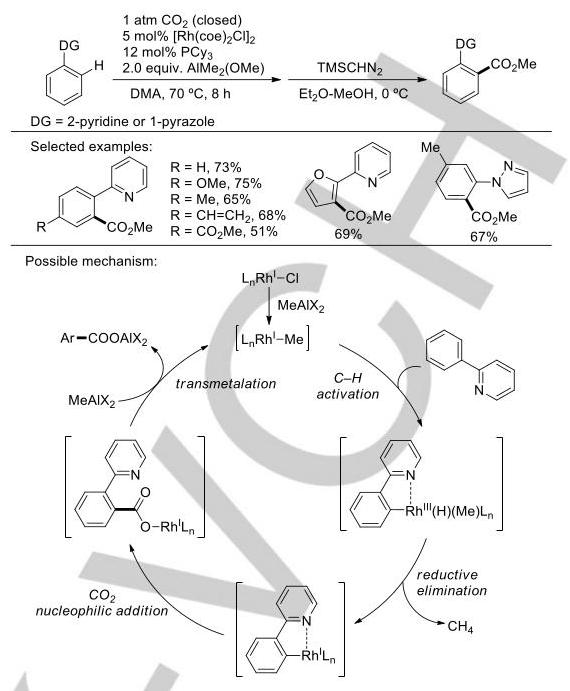
Scheme 49. Rh(I)-catalyzed direct carboxylation of aryl C-H bond via chelation-assisted C-H bond activation[82]
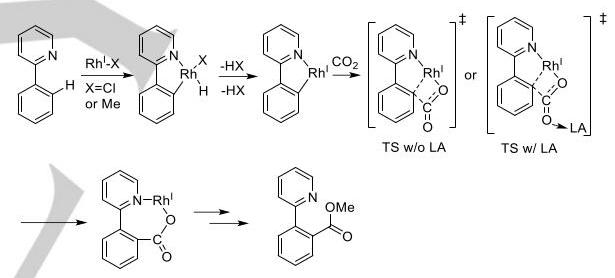
Scheme 50. Proposed mechanism of the Rh-catalysed C-H bond carboxylation based on DFT calculations[131]
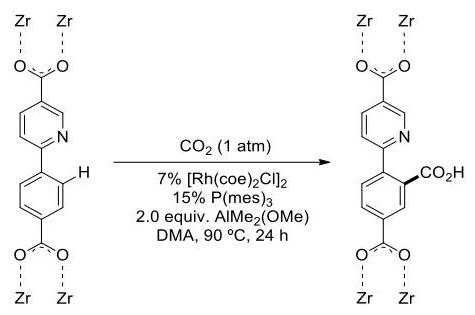
Scheme 51. Insertion of ${\mathrm{{CO}}}_{2}$ into aryl $\mathrm{C} - \mathrm{H}$ bonds of the UiO-67(dcppy) backbone ${}^{\left\lbrack {132}\right\rbrack }$
REVIEW
- promoting ortho silylation via formation of a five-membered ruthenacycle intermediate and promoting carboxylation by an electron-withdrawing pyrazole nitrogen atom. In 2015, Hou and coworkers ${}^{\left\lbrack {134}\right\rbrack }$ reported a formal C-H bond carboxylation of aromatic rings containing directing groups or heterocycles catalyzed by a Cu(I)-NHC complex (Scheme 53). The reaction is carried out in two steps: Firstly, the aromatic ring is deprotonated with iBu ${}_{3}$ Al(TMP)Li (TMP=2,2,6,6- tetramethylpiperidide) to form the corresponding arylaluminum intermediate which is then catalyzed by Cu(I)-NHC complex to give the corresponding carboxylated products. This reaction has high regioselectivity, broad substrate scope and good tolerance of functional groups. For heterocycles such as benzofuran, benzothiophene, and N-Boc indole, the reaction occurs selectively at the C-2 position.
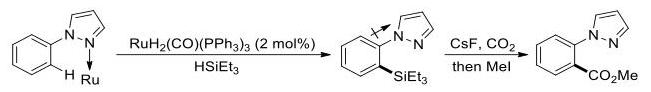
Scheme 52. Sequential C-H bond silylation and carboxylation of 1-arylpyrazole derivatives[133]
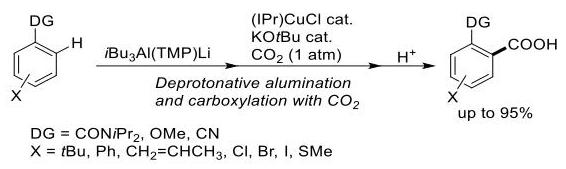
Scheme 53. $\mathrm{C} - \mathrm{H}$ carboxylation of aromatic compounds by deprotonative alumination and carboxylation ${}^{\left\lbrack {134}\right\rbrack }$
In 2018, Li and coworkers ${}^{\left\lbrack {135}\right\rbrack }$ reported a ligand-enabled site-selective carboxylation of 2-arylphenols under atmospheric pressure of ${\mathrm{{CO}}}_{2}$ through a ${\mathrm{{Rh}}}_{2}{\left( \mathrm{{OAc}}\right) }_{4}$ -catalysed and chelation-assisted C-H bond activation (Scheme 54). Remarkably, the reaction occurs selectively on the less nucleophilic phenyl group via promotion by a phosphine ligand, which overrides the site selectivity dictated by the well-known Kolbe-Schmitt type reaction. Later, they have successfully applied this Rh" catalysis approach to other similar types of substrate. ${}^{\left\lbrack {136} - {137}\right\rbrack }$ In 2018, Yu and coworkers[138] reported the first Pd(TFA)2- catalyzed benzamide-directed carbonylation of unactivated aryl $\mathrm{C - H}$ bonds with atmospheric ${\mathrm{{CO}}}_{2}$ to afford a series of phthalimides (Scheme 55a). The mechanistic investigations suggested that the palladacycle might be an active intermediate, which further reacts with ${\mathrm{{CO}}}_{2}$ to generate the desired product. In another work, ${}^{\left\lbrack {139}\right\rbrack }$ with benzylic alcohol as a directing group, they realized Pd(TFA)2-catalyzed lactonization of unactivated aryl C-H bonds with ${\mathrm{{CO}}}_{2}$ to afford phthalides (Scheme 55b). Remarkably, the DFT calculations indicated that lactonization might involve insertion of two ${\mathrm{{CO}}}_{2}$ into the $\mathrm{O} - \mathrm{{Pd}}$ bond of the palladacycle, which is interesting and novel from previous reports.
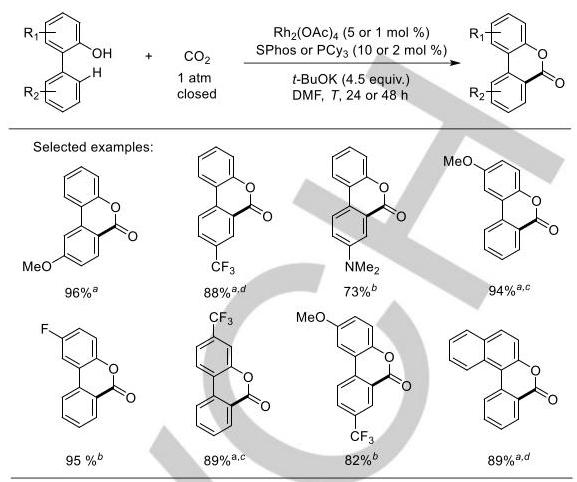
${}^{a}{24}\mathrm{\;h},{90}^{ \circ }\mathrm{C},{\mathrm{{Rh}}}_{2}{\left( \mathrm{{OAc}}\right) }_{4}\left( {5\mathrm{\;{mol}}\% }\right)$ , SPhos $\left( {{10}\mathrm{\;{mol}}\% }\right)$ . ${}^{c}{100}^{ \circ }\mathrm{C}$ . ${}^{d}{120}^{ \circ }\mathrm{C}$ . ${}^{b}{48}\mathrm{\;h},{100}^{ \circ }\mathrm{C},{\mathrm{{Rh}}}_{2}{\left( \mathrm{{OAc}}\right) }_{4}\left( {1\mathrm{\;{mol}}\% }\right) ,{\mathrm{{PCy}}}_{3}\left( {2\mathrm{\;{mol}}\% }\right)$ .
${}^{a}{24}\mathrm{\;h},{90}^{ \circ }\mathrm{C},{\mathrm{{Rh}}}_{2}{\left( \mathrm{{OAc}}\right) }_{4}\left( {5\mathrm{\;{mol}}\% }\right)$,SPhos$\left( {{10}\mathrm{\;{mol}}\% }\right)$。${}^{c}{100}^{ \circ }\mathrm{C}$。${}^{d}{120}^{ \circ }\mathrm{C}$。${}^{b}{48}\mathrm{\;h},{100}^{ \circ }\mathrm{C},{\mathrm{{Rh}}}_{2}{\left( \mathrm{{OAc}}\right) }_{4}\left( {1\mathrm{\;{mol}}\% }\right) ,{\mathrm{{PCy}}}_{3}\left( {2\mathrm{\;{mol}}\% }\right)$。
Scheme 54. Ligand-enabled site-selectivity in a versatile rhodium(II)-catalyzed aryl $\mathrm{C} - \mathrm{H}$ carboxylation of aromatic compounds by deprotonative
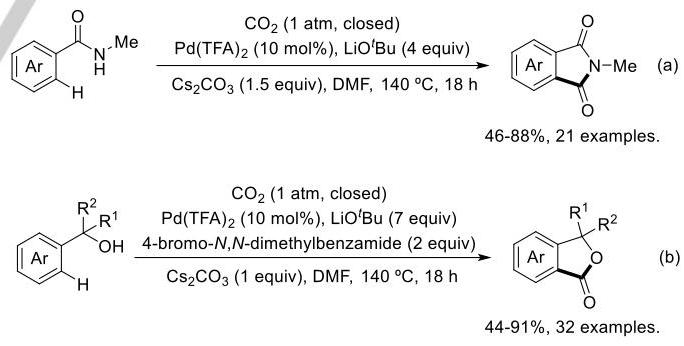
Scheme 55. Pd-catalyzed directed C-H bond carboxylation with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {138} - {139}\right\rbrack }$
3.2. Carboxylation of vinylic $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$
In 2013, Iwasawa and coworkers[140] first reported a Pd(II)- catalyzed direct carboxylation (lactonization) reaction of alkenyl C-H bonds with ${\mathrm{{CO}}}_{2}$ (Scheme 56a). Their reaction mechanism is proposed as follows. First, one molecule of 2-hydroxystyrene is chelated with $\mathrm{{Pd}}{\left( \mathrm{{OAc}}\right) }_{2}$ to activate the vinylic $\mathrm{C} - \mathrm{H}$ bond, and then reacts with the cesium salt of a second molecule of 2- hydroxystyrene to produce a hexacyclic alkenyl-palladium intermediate. The next step is a reversible nucleophilic carboxylation of this intermediate with ${\mathrm{{CO}}}_{2}$ to give a palladium carboxylate intermediate. The palladium carboxylate intermediate reacts with another molecule of 2-hydroxystyrene and ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ to give the coumarin product, while the cyclometalated intermediate is regenerated (Scheme 57). Through stoichiometric experiments they isolated the cyclometalated intermediates, whose structures were confirmed by X-ray crystal diffraction. Interestingly, a transition-metal-free version of this reaction was reported by Yu, Zhi and coworkers in 2017 (Scheme 56b).[130] The reaction conditions are much harsher for the transition-metal-free version compared with the Pd-catalyzed version.
In 2016, Yu and coworkers ${}^{\left\lbrack {141}\right\rbrack }$ developed a transition-metal-free lactamization of alkenyl and heteroaryl $\mathrm{C} - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ (Scheme 58). A series of 2-quinolinone and polyheterocycles were synthesized. Using excessive amounts of sodium tert-butoxide as base and high temperature are key to the efficient transformation. Isocyanate is proposed as the active intermediate in this reaction. In 2016, based on previous work[112-113, 122-123], Tanaka, Hattori and coworkers reported a EtAICl2/2,6-disubstituted pyridine-mediated carboxylation of a-aryl alkenes and trisubstituted alkenes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {142}\right\rbrack }$ (Scheme 59). In terms of practical yield, this is the first reaction in which a vinyl hydrogen atom is replaced by a carboxyl group. The choice of a base with suitable steric hindrance and basicity is crucial to this reaction. Notably, the base can be used as a catalyst in this reaction but the amount of ${\mathrm{{EtAICl}}}_{2}$ must be stoichiometric. In 2018, Yu and coworkers ${}^{\left\lbrack {143}\right\rbrack }$ reported a novel synthesis of 1,3- oxazin-6-ones from enamides and ${\mathrm{{CO}}}_{2}$ through C-H carboxylation and one-pot cyclization (Scheme 60). Nucleophilic attack on ${\mathrm{{CO}}}_{2}$ by the electron-rich alkene is demonstrated for this reaction. The direct synthesis of acrylates from ${\mathrm{{CO}}}_{2}$ and ethylene is an ideal and economical way, but this reaction is incredibly challenging. In industry, sodium acrylate is a monomer used for the synthesis of sodium polyacrylate, a versatile polymeric material. In 2012, Limbach and coworkers ${}^{\left\lbrack {144}\right\rbrack }$ first realized a nickel-catalyzed synthesis of sodium acrylate from ${\mathrm{{CO}}}_{2}$ and ethylene (Scheme 61). The turnover number (TON) of the catalyst can exceed 10. Critical to success was the separation and characterization of several key organonickel intermediates of elementary reactions in the catalytic cycle as well as combination of experimental data and theoretical calculations. Since the base sodium tert-butoxide can react with ${\mathrm{{CO}}}_{2}$ to produce relatively stable carbonic acid half esters, the catalytic cycle must be divided into two parts, a ${\mathrm{{CO}}}_{2}$ -rich regime and a ${\mathrm{{CO}}}_{2}$ -poor regime. This shortcoming has greatly limited the practicality of this method. In 2014, Vogt and coworkers[ ${}^{145}$ ] used a nickel catalyst system to further increase catalytic TON to 21 (Scheme 62). Compared with the work of Limbach and coworkers, Vogt and coworkers’ method is easier to operate and avoids repeated replacement of reactant gases. Believed by Vogt and coworkers, the challenge of this reaction is that the nickelalactone intermediate do not readily undergo $\beta$ -hydride elimination due to conformation of the five-membered ring. They found that adding “hard cation soft anion” salt Lil to the reaction system can effectively promote decomposition of the nickelalactone intermediate to produce lithium acrylate (Scheme 62). The role of Lil in the reaction has also been supported by DFT calculations. In 2014, Limbach and coworkers[146] made further improvements to their previously reported system. ${}^{\left\lbrack {144}\right\rbrack }$ Through screen of ligands, solvents, bases, etc., they succeeded in realizing nickel-catalyzed carboxylation of a series of alkenes (including ethylene, styrene and 1,3- butadiene) ${}^{\left\lbrack {146}\right\rbrack }$ (Scheme 63). The choice of base is crucial for the reaction: a suitable base not only has strong basicity so that the nickelalactone intermediate can be deprotonated to release the product, but also has weaker nucleophilicity to avoid reacting directly with ${\mathrm{{CO}}}_{2}$ . Sodium 2-fluorophenoxide is the most suitable base in their selection. For ethylene, the catalyst TON can reach 107. Except for 2-vinylpyridine which gives a branched $\alpha ,\beta$ - unsaturated carboxylic acid salt, other alkenes are capable of selectively producing linear products. In terms of stereoselectivity, this reaction has high (E)-selectivity. Terminal, unactivated alkenes or internal ones cannot participate in this reaction. In 2015, Limbach and coworkers[147] reported the first palladium-catalyzed carboxylation of ethylene and conjugated dienes (Scheme 64a). Their proposed reaction mechanism is similar to the nickel-catalyzed reaction mechanism ${}^{\left\lbrack {146}\right\rbrack }$ . In 2015, Schaub and coworkers ${}^{\left\lbrack {148}\right\rbrack }$ improved the catalyst system of Limbach and coworkers: by using the $\left\lbrack {\mathrm{{Pd}}{\left( {\mathrm{{PPh}}}_{3}\right) }_{4}}\right\rbrack /$ dcpe system, sodium 2,6-di-tert-butyl-4-methylphenolate as base, catalyst TON can reach a level comparable to that of Limbach and coworkers’ nickel catalyst system (TON = 106) while avoiding use of zinc; in addition, the less toxic anisole is used as solvent (Scheme 64b). They also proposed a new concept that combines product separation, catalyst recovery and base regeneration. Schaub and coworkers[149] also studied the elementary reactions involved in nickel-catalyzed carboxylation of some simple alkenes. Recently, Schaub and coworkers[150] found that using [Pd(PPh3)4]/dcpe system and sodium tert-butoxide as base, ethylene, 1,3- butadiene and non-activated terminal alkenes or internal alkenes can undergo carboxylation at ${145}^{ \circ }\mathrm{C}$ (Scheme 64c). Sodium tert-butoxide is more basic than sodium 2-fluorophenoxide and applicable to a broader range of substrates for this system. Notably, the decomposition temperature of carbonate esters given by the reaction of sodium tert-butoxide and ${\mathrm{{CO}}}_{2}$ is ${60}^{ \circ }\mathrm{C}$ . Later, by using amide solvents, they have further improved their Pd catalysis system to achieve turnover numbers greater than 500.[^151] In 2017, Cummins and coworkers developed a series of novel cis-macrocyclic diphosphines ligands, and found these ligands show promising catalytic activity in Ni- ${}^{\left\lbrack {152}\right\rbrack }$ and Co-catalyzed ${}^{\left\lbrack {153}\right\rbrack }{\mathrm{{CO}}}_{2}$ /ethylene coupling reaction.
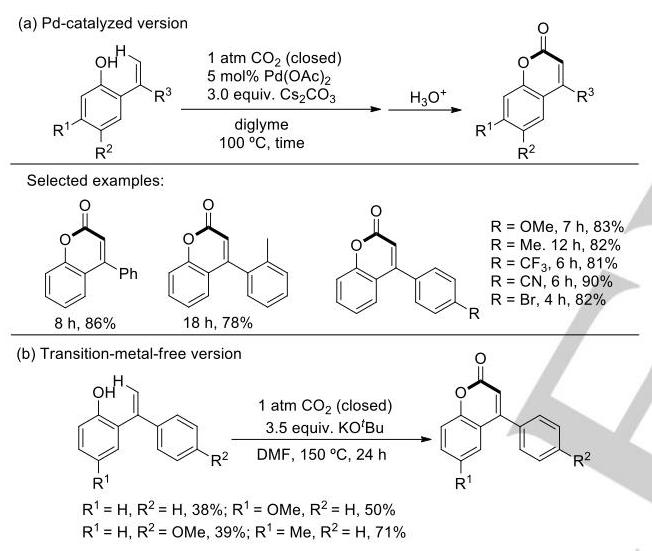
Scheme 56. Pd-catalyzed direct carboxylation of alkenyl C-H bonds of 2- hydroxystyrenes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {140}\right\rbrack }$ and its transition-metal-free version ${}^{\left\lbrack {130}\right\rbrack }$
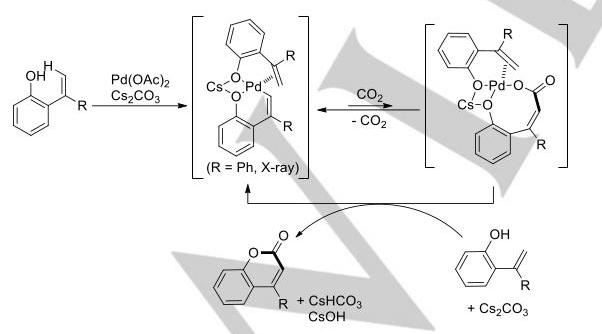
Scheme 57. Possible mechanism for Pd-catalyzed direct carboxylation of alkenyl C-H bonds of 2-hydroxystyrenes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {140}\right\rbrack }$
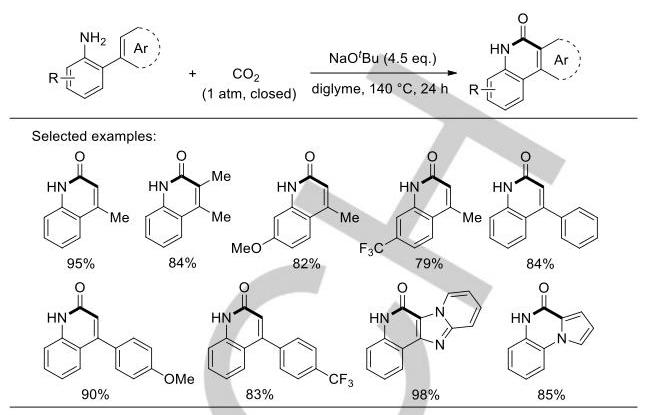
Scheme 58. ${\mathrm{{NaO}}}^{t}\mathrm{{Bu}}$ -promoted lactamization of 2-alkenylanilines or 2- heteroarylanilines with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {141}\right\rbrack }$
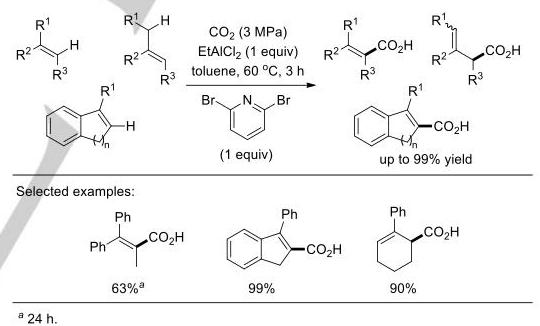
Scheme 59. EtAlCl2/2,6-dibromopyridine-mediated carboxylation of alkenes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {142}\right\rbrack }$
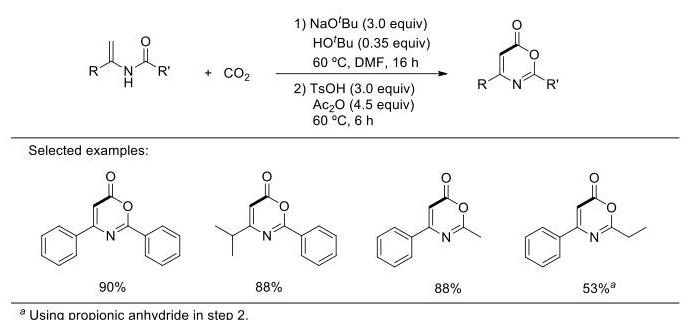
Scheme 60. C-H carboxylation of enamides with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {143}\right\rbrack }$
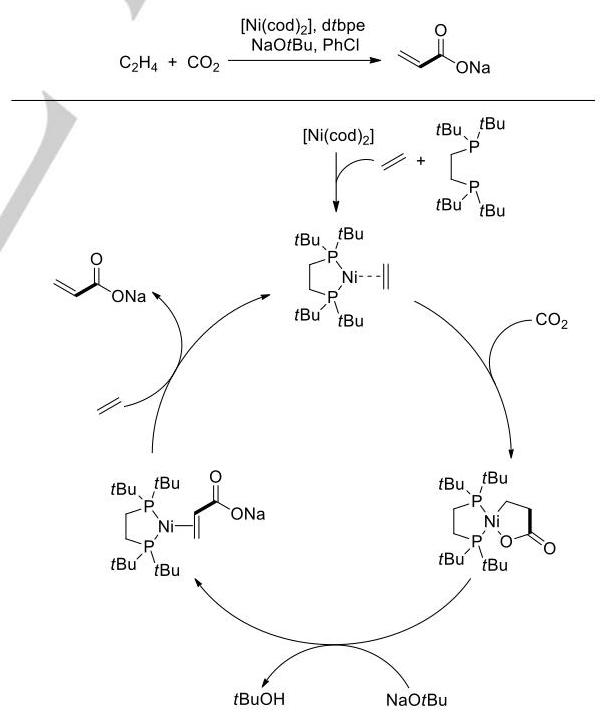
Scheme 61. Possible catalytic cycle for Ni-catalyzed formation of sodium acrylate from ${\mathrm{{CO}}}_{2}$ , ethylene, and base ${}^{\left\lbrack {144}\right\rbrack }$
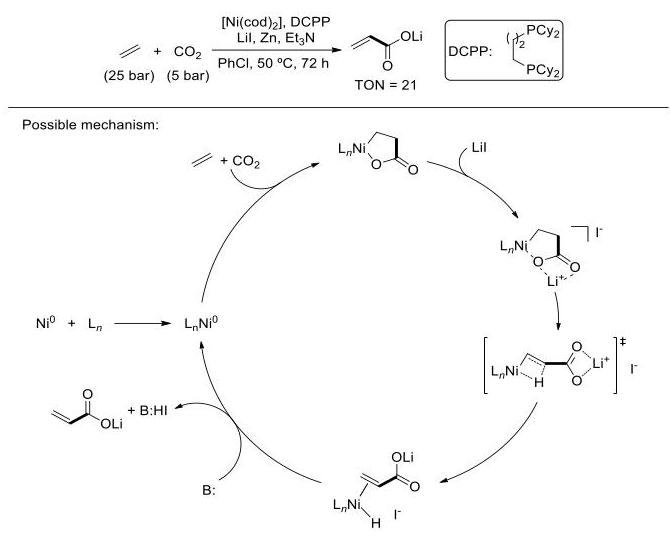
Scheme 62. Ni-catalyzed formation of lithium acrylate ${}^{\left\lbrack {145}\right\rbrack }$
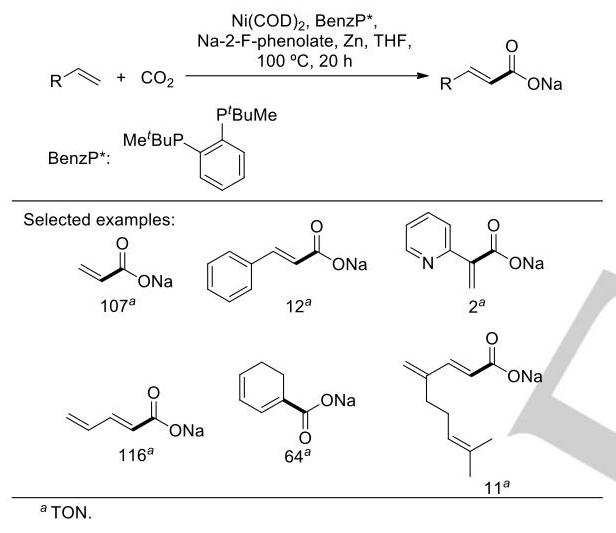
Scheme 63. Ni-catalyzed direct carboxylation of olefins with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {146}\right\rbrack }$
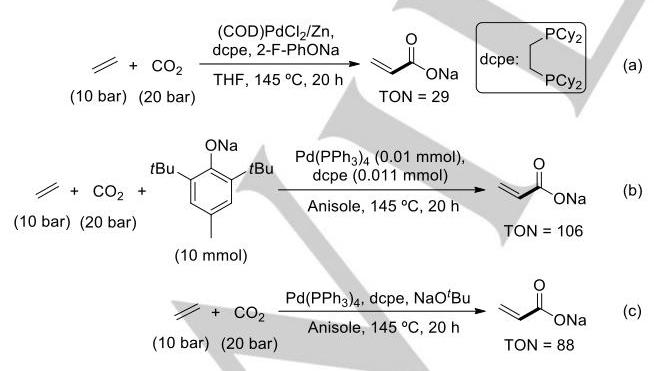
Scheme 64. Pd-catalyzed formation of sodium acrylate ${}^{\left\lbrack {147} - {148},{150}\right\rbrack }$
${\mathrm{{CO}}}_{2}$ In 2017, Mo and coworkers ${}^{\left\lbrack {154}\right\rbrack }$ reported ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ -promoted carboxylation of the diazo group ipso- $\mathrm{C} - \mathrm{H}$ bond with ${\mathrm{{CO}}}_{2}$ under very mild reaction conditions (Scheme 65). With intrinsic nucleophilicity, diazo group has been found to react with various electrophiles, such as acyl chloride, aldehyde, ketone, imine and boron compounds. This is the first report of utilizing ${\mathrm{{CO}}}_{2}$ in diazo addition reaction. The carboxylation exhibits high reactivity and is amenable to subsequent diversification. The resulting α-diazo carboxylate can be readily converted to esters and amides in a one-pot fashion. In addition to stable $\alpha$ -diazo carbonyl compounds, labile TMSCHN2 is also a competent substrate for the desired transformation. This method provide a modular synthesis of a series of unsymmetrical 1,3-diester/keto/amide diazo compounds in moderate to excellent yields (up to 99%) with good functional group compatibility. In 2018, Lee and coworkers[155] reported an interesting umpolung reactivity of aromatic aldehydes with ${\mathrm{{CO}}}_{2}$ (Scheme 66). By using KCN and Ti(OiPr)4, they achieved direct carboxylation reaction of aromatic aldehydes with ${\mathrm{{CO}}}_{2}$ (1 atm) to afford $\alpha$ -keto acids products under mild reaction conditions. This method has been expanded to the synthesis of α-amino acids derivatives, illustrating the chemical synthesis of essential bioactive molecules from ${\mathrm{{CO}}}_{2}$ .
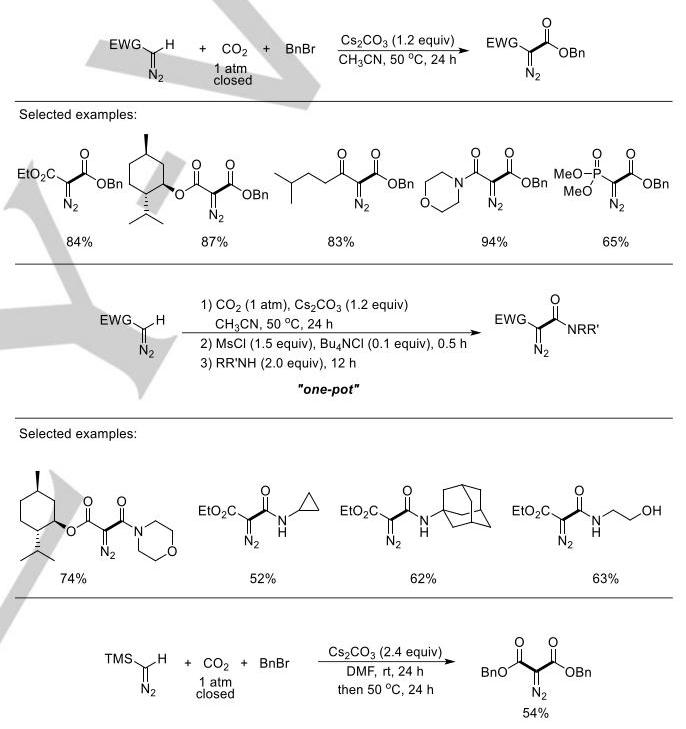
Scheme 65. ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ -promoted carboxylation of the diazo group ipso-C-H bond with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {154}\right\rbrack }$
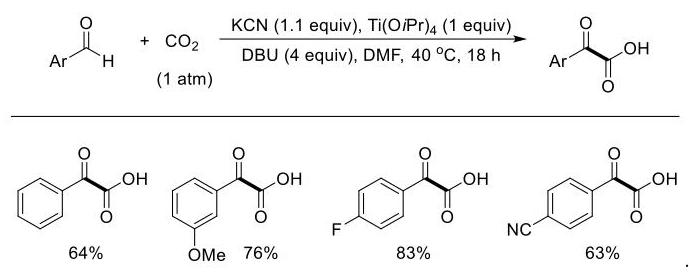
Scheme 66. Carboxylation reaction of aromatic aldehydes with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {155}\right\rbrack }$
4. Carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$
A large body of work ${}^{\left\lbrack {156} - {175}\right\rbrack }$ has been done on the carboxylation of carbonyl $\alpha - \mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bond with ${\mathrm{{CO}}}_{2}$ . For example, in 2006, Shoji Hara and coworkers[174] reported an electrochemical carboxylation of aliphatic ketones with ${\mathrm{{CO}}}_{2}$ to furnish $\beta$ - ketocarboxylic acids in moderate to good yields (Scheme 67). It is believed that ${\mathrm{{CO}}}_{2}$ is reduced to the corresponding radical anion under the reaction conditions; this radical anion species can facilitate enolate formation. In 2014, Macyk and coworkers reported a photocatalytic carboxylation of acetylacetone with CO2 by using ZnS-based photocatalysts. ${}^{\left\lbrack {175}\right\rbrack }$ In 2008, Jessop and coworkers ${}^{\left\lbrack {171}\right\rbrack }$ combined the carboxylation of ketones promoted by DBU with asymmetric hydrogenation of $\beta$ -ketocarboxylic acids catalyzed by ${\mathrm{{RuCl}}}_{2}\left\lbrack {\left( \mathrm{S}\right) - \mathrm{{BINAP}}}\right\rbrack$ for the asymmetric synthesis of $\beta$ - hydroxycarboxylic acids (Scheme 68). In 2011, Beckman and Munshi ${}^{\left\lbrack {172}\right\rbrack }$ reported the carboxylation of ketones catalyzed by a reversible ${\mathrm{{CO}}}_{2}$ carrier (RCC) (Scheme 69). RCC is obtained by reacting ${\mathrm{{CO}}}_{2}$ with DBU which is supported on methylhydrosiloxane dimethyl siloxane copolymer. The extent of carboxylation reaction might be related to the percentage of enol tautomer of the substrate. The catalyst can be conveniently recycled by filtration. This RCC strategy holds potential for low pressure ${\mathrm{{CO}}}_{2}$ utilization in chemical transformations.
Normally, β-ketocarboxylic acids are thermodynamic unstable due to high tendency towards decarboxylation. Thus, in situ conversion of β-ketocarboxylic acids into stable compounds by tandem reactions becomes a very viable strategy. ${}^{\left\lbrack {176} - {182}\right\rbrack }$ In 2012, Yamada and coworkers[177] reported a silver-catalyzed carboxylative cyclization of alkynyl-substituted ketones (Scheme 70a). The reaction mechanism is proposed as follows: in the presence of base, the substrate generates an enolate which reacts with ${\mathrm{{CO}}}_{2}$ to form a $\beta$ -ketocarboxylate; and the silver catalyst activates C≡C triple bond to capture the $\beta$ - ketocarboxylate, followed by formation of the lactone product. For ketones having two different protons $\alpha$ and ${\alpha }^{\prime }$ to the carbonyl group, the reaction can selectively generate y-lactones. The geometry of the $C = C$ double bond in the product was confirmed to be of Z-geometry by X-ray crystal diffraction and NOE experiments. Based on the above work, in 2013, Yamada and coworkers[178] reported a silver-catalyzed carboxylative cyclization of o-alkynyl acetophenone (Scheme 70b). In this reaction, the silver catalyst activated C=C triple bond captures the carbonyl group of the $\beta$ -ketocarboxylic acids intermediate to furnish the dihydroisobenzofuran framework. In 2014, Zhang and Lu and coworkers[179] reported a reaction (Scheme 70c) similar to Yamada and coworkers’ work. In 2017, Yamada and coworkers[182] reported the silver-catalyzed carboxylation of conjugated ynones with ${\mathrm{{CO}}}_{2}$ to furnish highly functionalized tetronic acids (Scheme 70d). Notably, in 2016, Zhang and coworkers[181] reported a copper-catalyzed double carboxylation of o-alkynyl acetophenone (Scheme 71). This reaction undergoes carboxylation, intramolecular cyclization and a second carboxylation. Substrates with alkyl-substituted alkynyl group have higher selectivity for double carboxylation than substrates with aryl-substituted alkynyl group. In 2015, Zhang and coworkers ${}^{\left\lbrack {180}\right\rbrack }$ reported the carboxylative cyclization of o-hydroxy-and o-acetamidoacetophenone promoted by DBU (Scheme 72). In the reaction of o-acetamidoacetophenone, the acyl group migrated from the nitrogen atom to the carbon atom.
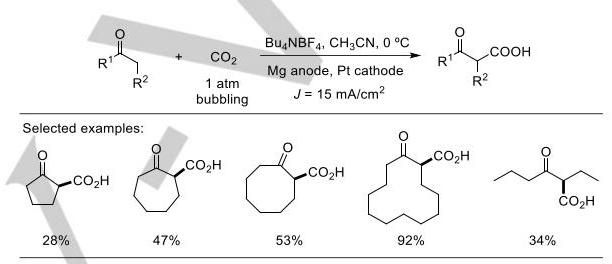
Scheme 67. Electrochemical carboxylation of aliphatic ketones with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {174}\right\rbrack }$
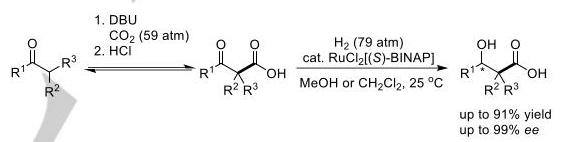
Scheme 68. Carboxylation of ketones followed by asymmetric
hydrogenation [171]
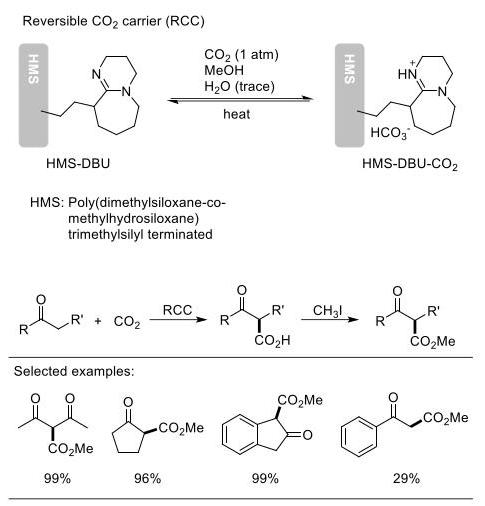
Scheme 69. Reversible ${\mathrm{{CO}}}_{2}$ carrier (RCC)-catalyzed carboxylation of ketones[172]
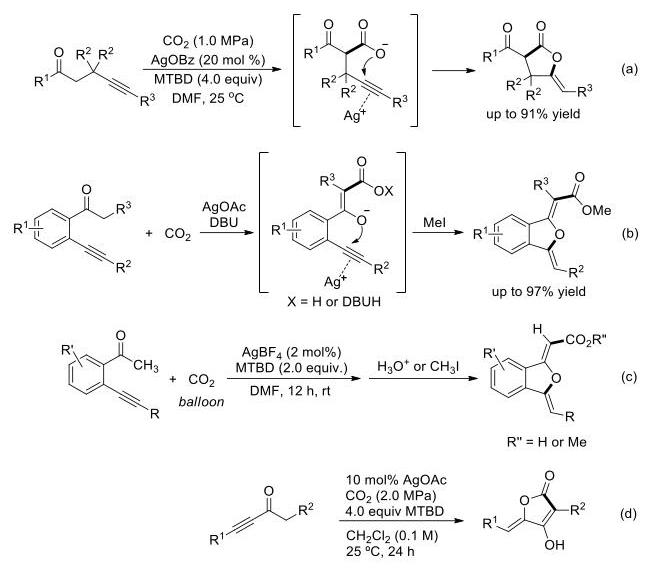
Scheme 70. Silver-catalyzed carboxylative cyclization of ketones containing an alkyne group ${}^{\left\lbrack {177} - {179},{182}\right\rbrack }$
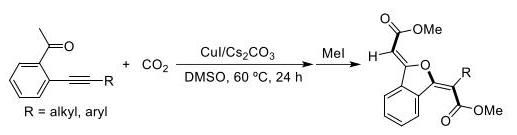
Scheme 71. Copper-catalyzed double carboxylation of o-alkynyl acetophenone ${}^{\left\lbrack {181}\right\rbrack }$
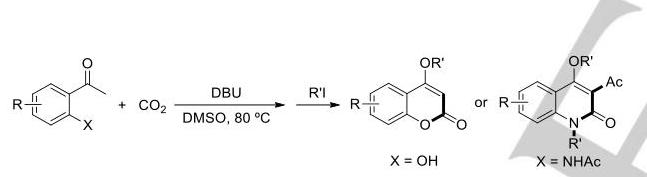
Scheme 72. DBU-promoted carboxylative cyclization of o-hydroxy- and o-acetamidoacetophenone ${}^{\left\lbrack {180}\right\rbrack }$
Reports on carboxylation of carbonyl $\mathrm{y} - \mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bond with ${\mathrm{{CO}}}_{2}$ are relatively rare. In 1999, Yoshida and coworkers[183] reported the carboxylative cyclization of 4-methyl-3-penten-2-one with ${\mathrm{{CO}}}_{2}$ promoted by ethylzinc carbamate. In 2016, Zhang and coworkers ${}^{\left\lbrack {184}\right\rbrack }$ reported the carboxylative cyclization of various substituted 1-propenone with ${\mathrm{{CO}}}_{2}$ promoted by $\mathrm{{CsF}}$ or ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ (Scheme 73). This reaction may have undergone a tandem carbonyl $\gamma - \mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bond carboxylation and base-promoted cyclization. This transition-metal-free reaction was carried out in dilute DMSO to inhibit substrate dimerization. Using ${}^{18}\mathrm{O}$ -labeled carbon dioxide, they found only one oxygen atom in the a-pyrone product came from carbon dioxide (Scheme 74). This proves that the cyclization reaction may have experienced an intramolecular attack of the enolate oxygen on the carbonyl moiety in the resulting carboxylate group. In 2012, Mita and Sato and coworkers[185] reported a formal carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ . The sequential protocol consists of two reactions: an ${\left\lbrack \mathrm{{Ir}}\left( \mathrm{{cod}}\right) \mathrm{{Cl}}\right\rbrack }_{2}$ - or ${\mathrm{{Ru}}}_{3}{\left( \mathrm{{CO}}\right) }_{12}$ - catalyzed silylation of benzylic $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds which is directed by a nitrogen atom-containing directing group (quinoline, quinoxaline, pyridine, etc.), and a fluoride-mediated carboxylation (Scheme 75). The $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{{Si}}$ bond can undergo fluoride-promoted carboxylation while the $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{{Si}}$ bond does not. This advantage provides convenience for further manipulation of the carboxylated products. In 2013, Mita and Sato and coworkers[ ${}^{186}$ ] expanded the substrate scope of this reaction. They found that ${\left\lbrack \mathrm{{Ir}}\left( \operatorname{cod}\right) \mathrm{{OMe}}\right\rbrack }_{2}$ or ${\left\lbrack \mathrm{{Rh}}\left( \mathrm{{cod}}\right) \mathrm{{Cl}}\right\rbrack }_{2}$ can catalyze the silylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds adjacent to nitrogen or oxygen atoms that is directed by a nitrogen-containing directing group (pyridine, isoquinoline, quinoline, etc.). With the participation of fluoride ions, the resulting silylated product can be converted into an α-amino acid derivative. In 2017, Jamison and coworkers[ ${}^{187}$ reported photoredox α- carboxylation of amines with ${\mathrm{{CO}}}_{2}$ for continuous flow synthesis of $\alpha$ -amino acids (Scheme 76). The single-electron reduction of ${\mathrm{{CO}}}_{2}$ is the key in this reaction. In 2015, Murakami and coworkers[188] reported a light-driven carboxylation of $o$ -alkylphenyl ketones with ${\mathrm{{CO}}}_{2}$ (Scheme 77). The reaction mechanism is proposed as follows: the substrate absorbs a photon to produce the excited state; the oxygen atom of the excited carbonyl group abstracts a hydrogen of the ortho-methyl group to form 1,4-biradical species which is then converted into o-quinodimethane. The (E)-isomer can undergo a [4+2] cycloaddition with ${\mathrm{{CO}}}_{2}$ , and finally ring-opening to form a carboxylic acid product while the $\left( Z\right)$ -isomer will undergo 1,5- hydrogen shift to regenerate the starting material. The reaction can be promoted by ultraviolet light or sunlight. Recently, Su and coworkers[189] studied the mechanism of this reaction through theoretical calculations. In 1995, Kudo and coworkers ${}^{\left\lbrack {190}\right\rbrack }$ reported the ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ -promoted carboxylation of allylic C-H bonds of caesium 3-crotonate with ${\mathrm{{CO}}}_{2}$ . In 2016, Murakami and coworkers ${}^{\left\lbrack {191}\right\rbrack }$ reported the carboxylation of allylic C-H bonds of simple alkene with ${\mathrm{{CO}}}_{2}$ promoted by light, ketone and copper complex systems (Scheme 78). They proposed a mechanism as follows: the ketone and alkene react under light excitation to form tertiary homoallyl alcohol intermediates; the tertiary homoallyl alcohol reacts with copper tert-butoxide generated in situ by copper complexes and potassium tert-butoxide. The resulting copper alkoxide undergoes C-C bond cleavage via β-carbon elimination to furnish the ketone and allylcopper intermediate; nucleophilic addition of the allylcopper intermediate to ${\mathrm{{CO}}}_{2}$ forms the copper carboxylate, which undergoes ligand exchange with potassium tert-butoxide to produce the carboxylate product and regenerate copper tert-butoxide. The above mechanism was supported by several control experiments. Mita, Sato and coworkers[192] reported a cobalt-catalyzed allylic $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ carboxylation with ${\mathrm{{CO}}}_{2}$ to regioselectively afford linear 3-butenoic acid motifs with good functional group tolerance (Scheme 79). The formation of a low-valent allyl-Co species is the key in this reaction. Hou and coworkers[193] reported copper-catalysed allylic C-H carboxylation of allyl aryl ethers with ${\mathrm{{CO}}}_{2}$ to afford 2-aryloxy butenoates (Scheme 80). The reaction consists of deprotonative allylic C-H bond alumination of allyl aryl ethers with an aluminum ate base $\left( {i{\mathrm{{Bu}}}_{3}\mathrm{{Al}}\left( \mathrm{{TMP}}\right) \mathrm{{Li}}}\right)$ and subsequent carboxylation of the resulting allylaluminum species with ${\mathrm{{CO}}}_{2}$ catalysed by an NHC-copper complex. Notably, they also found that DBU can catalyze the isomerization of 2-aryloxy-3-butenoate products to (Z)-2- aryloxy-2-butenoate isomers. $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bond activation and ${\mathrm{{CO}}}_{2}$ carboxylation promoted by transition metal complexes have received growing attention in recent years[194-203]. In 2008, Langer and Oro and coworkers[199] reported a reaction in which ${\mathrm{{CO}}}_{2}$ was formally inserted into the ligand C-H bond of an iridium complex (Scheme 81). In 2010, they[200] published further research on this reaction. In 2012, Annibale and Song[201] also reported similar reactions (Scheme 82). They proposed the following mechanism: a backbone carbon of diazafluorenide makes nucleophilic attack on ${\mathrm{{CO}}}_{2}$ , and then proton migration occurs. Different from the work of Langer and Oro and coworkers, the reaction of Annibale and Song is reversible. Braunstein and coworkers ${}^{\left\lbrack {195},{198}\right\rbrack }$ have reported similar reversible carboxylation reactions. In 2013, on the basis of the complexes of 4,5-diazafluorenide ligand and divalent ruthenium used in previous work, Song and coworkers[202] investigated the reactivity of the actor ligand toward ${\mathrm{{CO}}}_{2}$ with trivalent rhodium, monovalent rhodium and monovalent copper as spectator metal centers. They proposed the role of the spectator metal center probably was to modulate strength of the resulting C-C bond and adjust acidity of the C-H bond. Interestingly, they also first found that metal-free compounds were also capable of reversible tandem ${\mathrm{{CO}}}_{2}$ and $\mathrm{C} - \mathrm{H}$ activation ${}^{\left\lbrack {202} - {203}\right\rbrack }$ (Scheme 83). Similar to ${\mathrm{{CO}}}_{2}$ , methane is also a cheap, available and quite inert C1 source. $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ carboxylation of methane can be used to synthesize acetic acid which is widely used in industry. ${}^{\left\lbrack {204} - {229}\right\rbrack }$ In 2013, Wang and coworkers [223] investigated the reaction mechanism of the conversion of methane and ${\mathrm{{CO}}}_{2}$ to acetic acid catalyzed by zinc-modified H-ZSM-5 zeolite (Zn/H-ZSM-5). They first demonstrated through solid-state NMR experiments that $\mathrm{{Zn}}/\mathrm{H} - \mathrm{{ZSM}} - 5$ can catalyze the conversion of methane and ${\mathrm{{CO}}}_{2}$ to acetic acid at a temperature range of 523-773 K. The possible catalytic mechanism of the bifunctional catalyst was revealed by solid-state ${}^{13}\mathrm{C}$ and ${}^{1}\mathrm{H}$ MAS NMR: methane is activated as zinc methyl species and Brønsted protons, while ${\mathrm{{CO}}}_{2}$ is activated as surface carbonate ion species; ${\mathrm{{CO}}}_{2}$ is inserted into the $\mathrm{{Zn}} - \mathrm{C}$ bond of zinc methyl species to generate a key surface zinc acetate species which then abstracts the proton of Brønsted acid sites to release the acetic acid product. In 2014, Basset and coworkers[224] reported low temperature activation of methane over silica supported zinc-exchanged phosphotungstic acid (Zn-HPW/SiO2) (Scheme 84). Zn-HPW/SiO ${}_{2}$ can activate methane to form zinc methyl species at room temperature, while at higher temperature methane reacts with ${\mathrm{O}}_{2}$ to give methanol or reacts with ${\mathrm{{CO}}}_{2}$ to give acetic acid. Recently, Zhu and Ge and coworkers[225] reported a theoretical study of Zn-doped ceria-catalyzed C-C coupling reaction of ${\mathrm{{CO}}}_{2}$ and methane. The results show that the Zn dopant stabilizes the methyl group through formation of a Zn-C bond, thereby inhibiting subsequent dehydrogenation of methane. ${\mathrm{{CO}}}_{2}$ can be inserted into the $\mathrm{{Zn}} - \mathrm{C}$ bond in an activated bent configuration. The transition state of this insertion reaction is in the form of a three-centered Zn-C-C arrangement, with an activation energy barrier of ${0.51}\mathrm{{eV}}$ . The acetate ion generated by C-C coupling can bind to the surface proton generated by dissociation of methane and then desorbs in the form of free acetic acid. They have also examined other possible dopants such as AI, Ga, Cd, In, and Ni, and found a positive correlation between the activation barrier of C-C coupling and the electronegativity of dopant. In addition, carboxylation reactions of other simple alkanes with ${\mathrm{{CO}}}_{2}$ have also been reported. ${}^{\left\lbrack {230} - {231}\right\rbrack }$
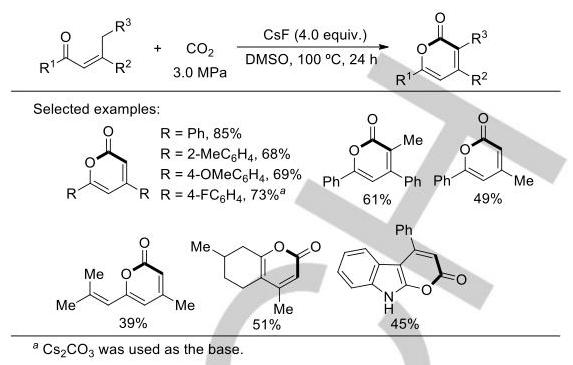
Scheme 73. Carboxylative cyclization of substituted 1-propenyl ketones with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {184}\right\rbrack }$
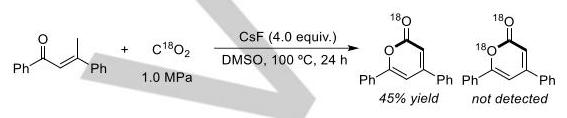
Scheme 74. Carboxylative cyclization of 1-propenyl ketone with ${\mathrm{C}}^{18}{\mathrm{O}}_{2}{}^{\left\lbrack {184}\right\rbrack }$
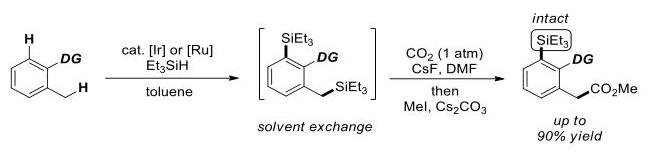
Scheme 75. Ir/Ru-catalyzed C-H silylation and fluoride-mediated carboxylation ${}^{\left\lbrack {185}\right\rbrack }$
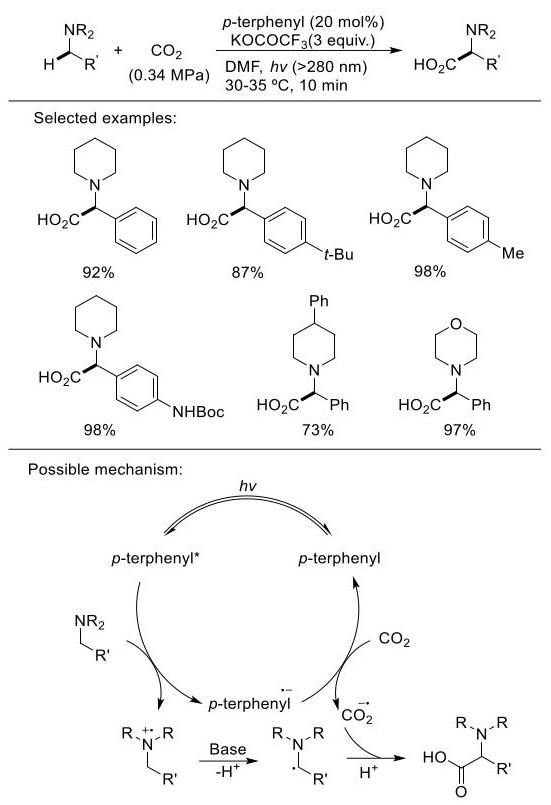
Scheme 76. The photoredox $\alpha$ -carboxylation of amines with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {187}\right\rbrack }$
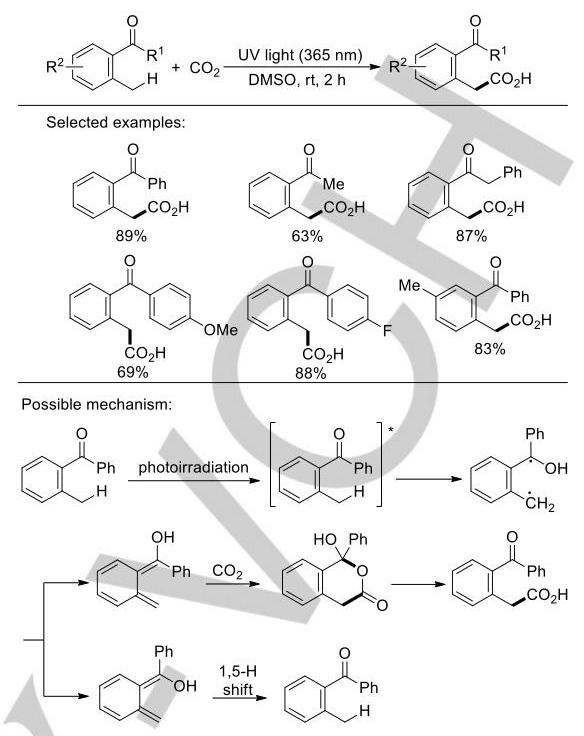
Scheme 77. Light-driven carboxylation of o-alkylphenyl ketones with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {188}\right\rbrack }$
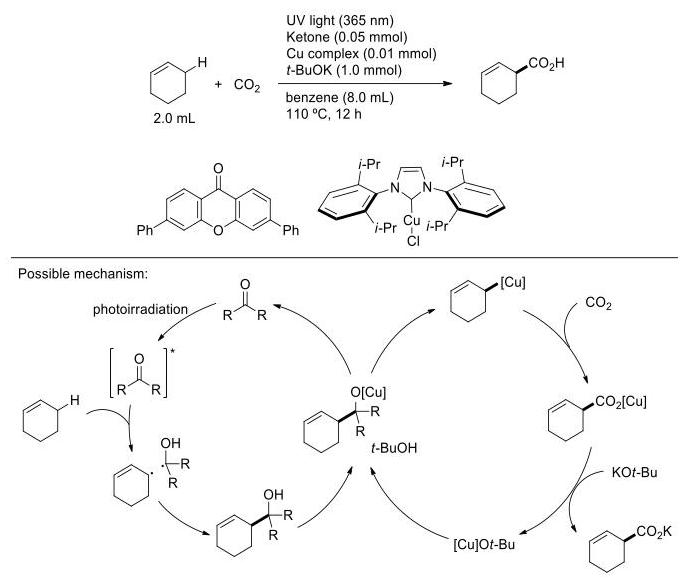
Scheme 78. Carboxylation of cyclohexene with ${\mathrm{{CO}}}_{2}$ in the presence of a ketone and a copper complex under UV irradiation [191]
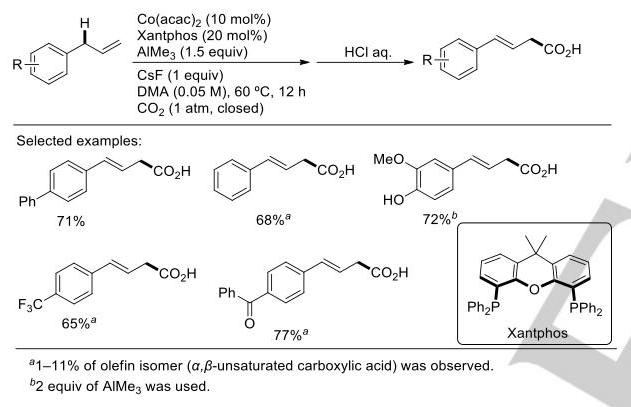
Scheme 79. Cobalt-catalyzed allylic $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ carboxylation with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {192}\right\rbrack }$
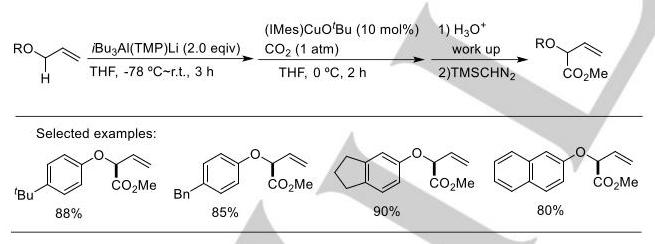
Scheme 80. Copper-catalysed allylic C-H carboxylation of allyl aryl ethers with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {193}\right\rbrack }$
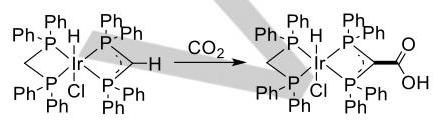
Scheme 81. Formal insertion of ${\mathrm{{CO}}}_{2}$ into a $\mathrm{C} - \mathrm{H}$ bond of ligand in an Ir complex ${}^{\left\lbrack {199}\right\rbrack }$
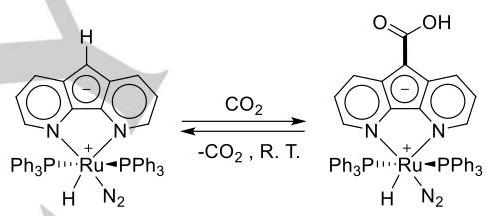
Scheme 82. Reversible carboxylation of a Ru complex ${}^{\left\lbrack {201}\right\rbrack }$
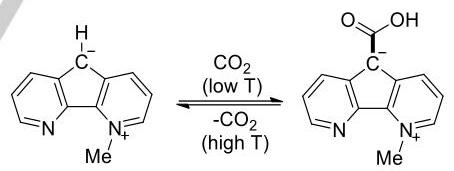
Scheme 83. Reversible metal-free tandem ${\mathrm{{CO}}}_{2}$ and $\mathrm{C} - \mathrm{H}$ activation ${}^{\left\lbrack {202}\right\rbrack }$
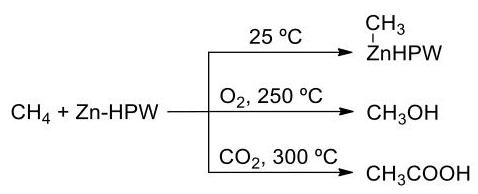
Scheme 84. Activation of methane by Zn-HPW[224]
In 2017, Martin and coworkers[232] reported a remote carboxylation of halogenated aliphatic hydrocarbons with ${\mathrm{{CO}}}_{2}$ (Scheme 85). Neither purification nor isolation of the intermediate halogenated compounds was necessary for this regioconvergent carboxylation of alkanes with ${\mathrm{{CO}}}_{2}$ to obtain single-regioisomer carboxylic acids. Tunable and controllable displacement of the Ni catalyst through a saturated hydrocarbon side-chain enables the switchable site-selective carboxylation at remote $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ sites.
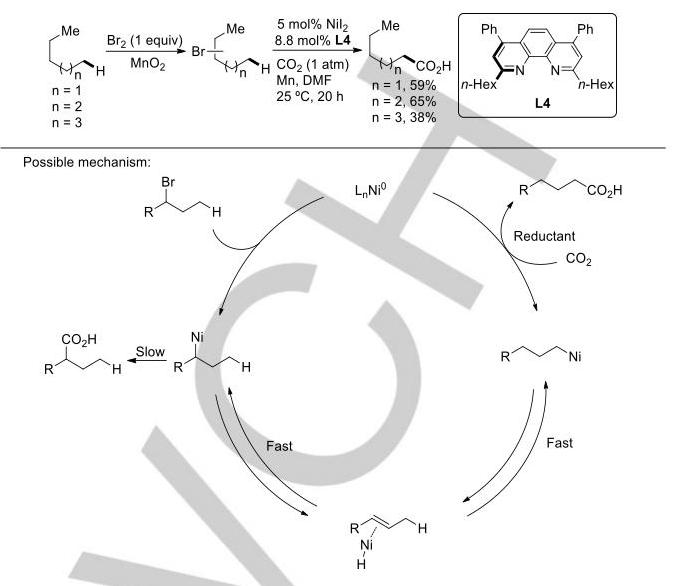
Scheme 85. Remote carboxylation of halogenated aliphatic hydrocarbons with ${\mathrm{{CO}}}_{2}{}^{\left\lbrack {232}\right\rbrack }$
5. Conclusions and Outlook
The carboxylation of $\mathrm{C} - \mathrm{H}$ bonds with ${\mathrm{{CO}}}_{2}$ merges activation of chemically inert $\mathrm{C} - \mathrm{H}$ bonds with thermodynamically stable ${\mathrm{{CO}}}_{2}$ and thus has great research value. This type of reaction has developed rapidly in recent years and has made many important advances. Its advantages are as follows: 1) Using ${\mathrm{{CO}}}_{2}$ as $\mathrm{C}1$ source avoids the use of toxic CO or phosgene; 2) C-H bonds are ubiquitous in organic molecules; 3) It conforms with the requirements of green chemistry in terms of atom economy and step economy; 4) It is easy to be incorporated into tandem or multicomponent reactions; 5) Many reactions have good industrial application prospects, such as the syntheses of furan-2,5- dicarboxylic acid and sodium acrylate. In fact, the Kolbe-Schmitt reaction, a classical $\mathrm{C} - \mathrm{H}$ bond carboxylation with ${\mathrm{{CO}}}_{2}$ , has been an industrial method for producing salicylic acid. At present, there are several problems in this field: the reaction conditions are relatively harsh in many cases, for example, requiring high temperature or high-pressure ${\mathrm{{CO}}}_{2}$ ; there is still much room for expansion in the scope of substrates, especially the carboxylation of $\mathrm{C}\left( {\mathrm{{sp}}}^{3}\right) - \mathrm{H}$ bonds; many reactions still require stoichiometric additives or metal exchange reagents; some carboxylation catalysts are not sufficiently efficient; some reactions are unsatisfactory in terms of chemoselectivity, regioselectivity, and stereoselectivity; the mechanism of some reactions is not clear; green reaction media is still rarely used; the cost and recovery of catalysts remains a problem, etc. In terms of reaction mechanism, the dual activation strategy should be more promising than the single activation of $\mathrm{C} - \mathrm{H}$ bonds or ${\mathrm{{CO}}}_{2}$ . Inexpensive metal catalysis, transition-metal-free, electrochemical and light-driven methods deserve more attention. Combining theoretical calculations with experimental research will certainly promote the growth of this field. More effective integration of ${\mathrm{{CO}}}_{2}$ capture and chemical transformation is still a major challenge in this field.
REVIEW
Acknowledgements
The project is supported by the Natural Science Foundation of China (Grants 21502003 and 21772003). We also thank the “1000-Youth Talents Plan” and Peking University for start-up funds. This work was also partially funded by the Peking University carbon capture, utilization and storage project supported by BHP Billiton.
The project is supported by the Natural Science Foundation of China (Grants 21502003 and 21772003). We also thank the “1000-Youth Talents Plan” and Peking University for start-up funds. This work was also partially funded by the Peking University carbon capture, utilization and storage project supported by BHP Billiton.
Keywords: Carbon dioxide $\cdot \mathrm{C} - \mathrm{H}$ bond activation $\cdot$
Carboxylation $\cdot$ Carboxylic acid $\cdot$ Catalysis
[1] T. Sakakura, J.-C. Choi, H. Yasuda, Chem. Rev. 2007, 107, 2365.
[2] M. Aresta, A. Dibenedetto, A. Angelini, Chem. Rev. 2014, 114, 1709.
[3] J. Luo, I. Larrosa, ChemSusChem 2017, 10, 3317.
[4] W. Zhang, N. Zhang, C. Guo, X. Lü, Chinese. J. Org. Chem. 2017, 37, 1309.
[5] J. Artz, T. E. Müller, K. Thenert, J. Kleinekorte, R. Meys, A. Sternberg, A. Bardow, W. Leitner, Chem. Rev. 2018, 118, 434.
[6] T. Andreu, J.-H. Francisco, B. Marino, M. Toni, M. Ruben, Angew. Chem. Int. Ed. 2018, 0. doi: 10.1002/anie.201803186
[7] W. Zhang, X. LÜ, Chin. J. Catal. 2012, 33, 745.
[8] A. H. Liu, B. Yu, L. N. He, Greenhouse Gas Sci. Technol. 2015, 5, 17.
[9] B. Yu, L. N. He, ChemSusChem 2015, 8, 52.
[10] S. Wang, G. Du, C. Xi, Org. Biomol. Chem. 2016, 14, 3666.
[11] X.-F. Wu, F. Zheng, Top. Curr. Chem. 2016, 375, 4.
[12] G. Yuan, C. Qi, W. Wu, H. Jiang, Current Opinion in Green and Sustainable Chemistry 2017, 3, 22.
[13] Y.-G. Chen, X.-T. Xu, K. Zhang, Y.-Q. Li, L.-P. Zhang, P. Fang, T.-S. Mei, Synthesis 2018, 02, 35.
[14] A. Correa, R. Martín, Angew. Chem. Int. Ed. 2009, 48, 6201.
[15] Y. Zhang, S. N. Riduan, Angew. Chem. Int. Ed. 2011, 50, 6210.
[16] S. Zhang, X.-D. Li, L.-N. He, Acta Chim. Sinica. 2016, 17.
[17] N. Kuhl, M. N. Hopkinson, J. Wencel - Delord, F. Glorius, Angew. Chem. Int. Ed. 2012, 51, 10236.
[18] G. Yan, X. Wu, M. Yang, Org. Biomol. Chem. 2013, 11, 5558.
[19] C. Cheng, J. F. Hartwig, Chem. Rev. 2015, 115, 8946.
[20] Z. Huang, H. N. Lim, F. Mo, M. C. Young, G. Dong, Chem. Soc. Rev. 2015, 44, 7764.
[21] C. Liu, J. Yuan, M. Gao, S. Tang, W. Li, R. Shi, A. Lei, Chem. Rev. 2015, 115, 12138.
[22] W. R. Gutekunst, P. S. Baran, Chem. Soc. Rev. 2011, 40, 1976.
[23] J. Yamaguchi, A. D. Yamaguchi, K. Itami, Angew. Chem. Int. Ed. 2012, 51, 8960.
[24] K. Okamoto, J. Zhang, J. B. Housekeeper, S. R. Marder, C. K. Luscombe, Macromolecules. 2013, 46, 8059.
[25] I. Tommasi, Catalysts 2017, 7, 380.
[26] Y. Fukue, S. Oi, Y. Inoue, J. Chem. Soc. Chem. Commun. 1994, 2091.
[27] S. Oi, Y. Fukue, K. Nemoto, Y. Inoue, Macromolecules. 1996, 29, 2694.
[28] W.-Z. Zhang, W.-J. Li, X. Zhang, H. Zhou, X.-B. Lu, Org. Lett. 2010, 12, 4748.
[29] R. Yuan, Z. Lin, ACS Catal. 2014, 4, 4466.
[30] K. Inamoto, N. Asano, K. Kobayashi, M. Yonemoto, Y. Kondo, Org. Biomol. Chem. 2012, 10, 1514.
[31] J.-N. Xie, B. Yu, Z.-H. Zhou, H.-C. Fu, N. Wang, L.-N. He, Tetrahedron Lett. 2015, 56, 7059.
[32] B. Yu, Z.-F. Diao, C.-X. Guo, C.-L. Zhong, L.-N. He, Y.- N. Zhao, Q.-W. Song, A.-H. Liu, J.-Q. Wang, Green Chem. 2013, 15, 2401.
[33] B. Yu, J.-N. Xie, C.-L. Zhong, W. Li, L.-N. He, ACS Catal. 2015, 5, 3940.
[34] X. Zhang, W.-Z. Zhang, X. Ren, L.-L. Zhang, X.-B. Lu, Org. Lett. 2011, 13, 2402.
[35] X. Zhang, W.-Z. Zhang, L.-L. Shi, C. Zhu, J.-L. Jiang, X.- B. Lu, Tetrahedron 2012, 68, 9085.
36] L. Zhang, W. Zhang, L. Shi, X. Ren, X. Lu, Chin. J. Catal. 2013, 34, 1179.
[37] C.-X. Guo, B. Yu, J.-N. Xie, L.-N. He, Green Chem. 2015, 17, 474.
[38] Z.-Z. Zhang, R.-J. Mi, F.-J. Guo, J. Sun, M.-D. Zhou, X.- C. Fang, Journal of Saudi Chemical Society 2017, 21, 685.
[39] F.-J. Guo, Z.-Z. Zhang, J.-Y. Wang, J. Sun, X.-C. Fang, M.-D. Zhou, Tetrahedron 2017, 73, 900.
[40] N. Eghbali, J. Eddy, P. T. Anastas, J. Org. Chem. 2008, 73, 6932.
L. J. Gooßen, N. Rodríguez, F. Manjolinho, P. P. Lange, Adv. Synth. Catal. 2010, 352, 2913.
[42] D. Yu, Y. Zhang, P. Natl. Acad. Sci. Usa. 2010, 107, 20184.
[43] L. Yang, Y. Yuan, H. Wang, N. Zhang, S. Hong, RSC Adv. 2014, 4, 32457.
[44] M. Trivedi, G. Singh, A. Kumar, N. P. Rath, Dalton Trans. 2015, 44, 20874.
[45] G. N. Bondarenko, E. G. Dvurechenskaya, E. S. Magommedov, I. P. Beletskaya, Catal. Lett. 2017, 147, 2570.
[46] T. Wendling, E. Risto, T. Krause, L. J. Gooßen, Chem. Eur. J. 2018, 24, 6019.
[47] C. Liu, Y. Luo, W. Zhang, J. Qu, X. Lu, Organometallics. 2014, 33, 2984.
[48] M. Arndt, E. Risto, T. Krause, L. J. Gooßen, ChemCatChem 2012, 4, 484.
[49] S. H. Kim, K. H. Kim, S. H. Hong, Angew. Chem. Int. Ed. 2014, 53, 771.
[50] J. Jover, F. Maseras, J. Org. Chem. 2014, 79, 11981.
[51] S. Li, J. Sun, Z. Zhang, R. Xie, X. Fang, M. Zhou, Dalton Trans. 2016, 45, 10577.
[52] D. Yu, M. X. Tan, Y. Zhang, Adv. Synth. Catal. 2012, 354, 969.
[53] D. Yu, F. Zhou, D. S. W. Lim, H. Su, Y. Zhang, ChemSusChem 2017, 10, 836.
[54] E. D. Finashina, L. M. Kustov, O. P. Tkachenko, V. G. Krasovskiy, E. I. Formenova, I. P. Beletskaya, Russ. Chem. B+. 2014, 63, 2652.
[55] X. H. Liu, J. G. Ma, Z. Niu, G. M. Yang, P. Cheng, Angew. Chem. Int. Ed. 2015, 54, 988.
[56] R. A. Molla, K. Ghosh, B. Banerjee, M. A. Iqubal, S. K. Kundu, S. M. Islam, A. Bhaumik, J. Colloid Interf. Sci. 2016, 477, 220.
REVIEW
[57] N.-N. Zhu, X.-H. Liu, T. Li, J.-G. Ma, P. Cheng, G.-M. Yang, Inorg. Chem. 2017, 56, 3414.
[58] B. Yu, P. Yang, X. Gao, Z. Z. Yang, Y. F. Zhao, H. Y. Zhang, Z. M. Liu, New J. Chem. 2017, 41, 9250.
[59] Y. Yuan, C. Chen, C. Zeng, B. Mousavi, S. Chaemchuen, F. Verpoort, ChemCatChem 2017, 9, 882.
[60] Z. Wu, L. Sun, Q. Liu, X. Yang, X. Ye, Y. Hu, Y. Huang, Green Chem. 2017, 19, 2080.
[61] H. Díaz Velázquez, Z.-X. Wu, M. Vandichel, F. Verpoort, Catal. Lett. 2017, 147, 463.
[62] Q.-Q. Dang, C.-Y. Liu, X.-M. Wang, X.-M. Zhang, ACS Appl. Mater. Interfaces 2018, 10, 27972.
[63] H. Cheng, B. Zhao, Y. Yao, C. Lu, Green Chem. 2015, 17, 1675.
[64] M. Trivedi, Bhaskaran, A. Kumar, G. Singh, A. Kumar, N. P. Rath, New J. Chem. 2016, 40, 3109.
[65] J.-H. Chen, C.-H. Deng, S. Fang, J.-G. Ma, P. Cheng, Green Chem. 2018, 20, 989.
[66] F. Köster, E. Dinjus, E. Duñach, Eur. J. Org. Chem. 2001, 2001, 2507.
[67] D. Yu, Y. Zhang, Green Chem. 2011, 13, 1275.
[68] X. Wang, Y. N. Lim, C. Lee, H. Y. Jang, B. Y. Lee, Eur. J. Org. Chem. 2013, 2013, 1867.
[69] F.-W. Li, Q.-L. Suo, H.-L. Hong, N. Zhu, Y.-Q. Wang, L.- M. Han, Tetrahedron Lett. 2014, 55, 3878.
[70] F.-W. Li, Q.-L. Suo, H.-L. Hong, N. Zhu, Y.-Q. Wang, L.- M. Han, Chinese. J. Org. Chem. 2015, 34, 2172.
[71] Y. Bo, Y. Peng, G. Xiang, Y. Zhenzhen, Z. Yanfei, Z. Hongye, L. Zhimin, Sci. China Chem. 2018, 61, 449.
[72] P. Foley, N. Eghbali, P. T. Anastas, Green Chem. 2010, 12, 888.
[73] P. Foley, N. Eghbali, P. T. Anastas, J. Nat. Prod. 2010, 73, 811.
[74] E. Schreiner, S. Braun, C. Kwasnitschka, W. Frank, T. J. J. Müller, Adv. Synth. Catal. 2014, 356, 3135.
[75] C.-C. Dong, J.-F. Xiang, L.-J. Xu, H.-Y. Gong, J. Org. Chem. 2018, 83, 9561.
[76] J.-N. Xie, B. Yu, C.-X. Guo, L.-N. He, Green Chem. 2015, 17, 4061.
[77] E. Schreiner, T. Wilcke, T. J. J. Müller, Synlett 2016, 27, 379.
[78] F.-S. Wu, W. Tong, Y. Liang, H.-S. Wang, Q.-H. Teng, Y.-M. Pan, RSC Adv. 2016, 6, 63855.
[79] F.-H. Cui, S.-X. Su, Y.-l. Xu, Y. Liang, H.-s. Wang, Y.- M. Pan, Org. Chem. Front. 2016, 3, 1304.
[80] S. Nurhanna Riduan, J. Y. Ying, Y. Zhang, Org. Lett. 2012, 14, 1780.
[81] H. Sugimoto, I. Kawata, H. Taniguchi, Y. Fujiwara, J. Organomet. Chem. 1984, 266, c44.
[82] H. Mizuno, J. Takaya, N. Iwasawa, J. Am. Chem. Soc. 2011, 133, 1251.
[83] T. Suga, H. Mizuno, J. Takaya, N. Iwasawa, Chem. Commun. 2014, 50, 14360.
[84] T. Suga, T. Saitou, J. Takaya, N. Iwasawa, Chem. Sci. 2017, 8, 1454.
[85] A. Uhe, M. Hölscher, W. Leitner, Chem. Eur. J. 2012, 18, 170.
[86] S. D. Stoychev, C. M. Conifer, A. Uhe, M. Holscher, W. Leitner, Dalton Trans. 2014, 43, 11180.
87] I. I. F. Boogaerts, S. P. Nolan, J. Am. Chem. Soc. 2010, 132,8858.
8] M. T. Johnson, J. Marthinus Janse van Rensburg, M. Axelsson, M. S. G. Ahlquist, O. F. Wendt, Chem. Sci. 2011, 2, 2373.
[89] X. Zhang, Z. Geng, Y. Wang, X. Hou, D. Wang, J. Mol. Catal. A-Chem. 2012, 363-364, 31.
[90] L. Zhang, J. Cheng, T. Ohishi, Z. Hou, Angew. Chem. Int. Ed. 2010, 49, 8670.
[91] I. I. F. Boogaerts, G. C. Fortman, M. R. L. Furst, C. S. J. Cazin, S. P. Nolan, Angew. Chem. Int. Ed. 2010, 49, 8674.
[92] A. Ariafard, F. Zarkoob, H. Batebi, R. Stranger, B. F. Yates, Organometallics. 2011, 30, 6218.
[93] H. Inomata, K. Ogata, S.-i. Fukuzawa, Z. Hou, Org. Lett. 2012, 14, 3986.
[94] D.-A. Park, J. Y. Ryu, J. Lee, S. Hong, RSC Adv. 2017, 7, 52496.
[95] W. Wang, G. Zhang, R. Lang, C. Xia, F. Li, Green Chem. 2013, 15, 635.
[96] A. S. Lindsey, H. Jeskey, Chem. Rev. 1957, 57, 583.
[97] J. Luo, S. Preciado, I. Larrosa, J. Am. Chem. Soc. 2014, 136, 4109.
[98] J. Luo, S. Preciado, P. Xie, I. Larrosa, Chem. Eur. J. 2016, 22, 6798.
[99] K. Kirimura, H. Gunji, R. Wakayama, T. Hattori, Y. Ishii, Biochem. Bioph. Res. Co 2010, 394, 279.
[100] C. Wuensch, S. M. Glueck, J. Gross, D. Koszelewski, M. Schober, K. Faber, Org. Lett. 2012, 14, 1974.
[101] C. Wuensch, N. Schmidt, J. Gross, B. Grischek, S. M. Glueck, K. Faber, J. Biotechnol. 2013, 168, 264.
[102] C. Wuensch, J. Gross, G. Steinkellner, A. Lyskowski, K. Gruber, S. M. Glueck, K. Faber, RSC Adv. 2014, 4, 9673.
[103] C. Wuensch, T. Pavkov - Keller, G. Steinkellner, J. Gross, M. Fuchs, A. Hromic, A. Lyskowski, K. Fauland, K. Gruber, S. M. Glueck, K. Faber, Adv. Synth. Catal. 2015, 357, 1909.
[104] J. Ren, P. Yao, S. Yu, W. Dong, Q. Chen, J. Feng, Q. Wu, D. Zhu, ACS Catal. 2016, 6, 564.
[105] K. Kudo, M. Shima, Y. Kume, F. Ikoma, S. Mori, N. Sugita, J. Jpn. Petrol. Inst. 1995, 38, 40.
[106] O. Vechorkin, N. Hirt, X. Hu, Org. Lett. 2010, 12, 3567.
[107] A. Banerjee, G. R. Dick, T. Yoshino, M. W. Kanan, Nature 2016, 531, 215.
[108] S. Fenner, L. Ackermann, Green Chem. 2016, 18, 3804.
[109] Y. Suzuki, T. Hattori, T. Okuzawa, S. Miyano, Chem. Lett. 2002, 31, 102.
[110] G. A. Olah, B. Török, J. P. Joschek, I. Bucsi, P. M. Esteves, G. Rasul, G. K. Surya Prakash, J. Am. Chem. Soc. 2002, 124, 11379.
[111] K. Nemoto, H. Yoshida, Y. Suzuki, N. Morohashi, T. Hattori, Chem. Lett. 2006, 35, 820.
[112] K. Nemoto, H. Yoshida, N. Egusa, N. Morohashi, T. Hattori, J. Org. Chem. 2010, 75, 7855.
[113] M. Konno, M. Chiba, K. Nemoto, T. Hattori, Chem. Lett. 2012, 41, 913.
[114] A. Schäfer, W. Saak, D. Haase, T. Müller, Angew. Chem. Int. Ed. 2012, 51, 2981.
[115] P. Munshi, E. J. Beckman, Ind. Eng. Chem. Res. 2009, 48, 1059.
[116] P. Munshi, E. J. Beckman, S. Padmanabhan, Ind. Eng. Chem. Res. 2010, 49, 6678.
[117] A. N. Sarve, P. A. Ganeshpure, P. Munshi, Ind. Eng. Chem. Res. 2012, 51, 5174.
[118] M. Gu, Z. Cheng, Ind. Eng. Chem. Res. 2014, 53, 9992.
REVIEW
[119] X. Zhang, Z. Cheng, Aiche J. 2017, 63, 185.
[120] J.-F. Ma, F. Ma, Z.-J. Zhou, Y.-T. Liu, RSC Adv. 2016, 6, 84042.
[121] K. Nemoto, S. Onozawa, N. Egusa, N. Morohashi, T. Hattori, Tetrahedron Lett. 2009, 50, 4512.
[122] K. Nemoto, S. Tanaka, M. Konno, S. Onozawa, M. Chiba, Y. Tanaka, Y. Sasaki, R. Okubo, T. Hattori, Tetrahedron 2016, 72, 734.
[123] K. Nemoto, S. Onozawa, M. Konno, N. Morohashi, T. Hattori, Bull. Chem. Soc. Jpn. 2012, 85, 369.
[124] W.-J. Yoo, M. G. Capdevila, X. Du, S. Kobayashi, Org. Lett. 2012, 14, 5326.
[125] W. J. Yoo, T. V. Q. Nguyen, M. G. Capdevila, S. Kobayashi, Heterocycles 2015, 90, 1196.
[126] Z. Xin, C. Lescot, S. D. Friis, K. Daasbjerg, T. Skrydstrup, Angew. Chem. Int. Ed. 2015, 54, 6862.
[127] C. W. Kee, K. Q. E. Peh, M. W. Wong, Chem. Asian J. 2017, 12, 1780.
[128] F. Ma, T. Miao, Z.-J. Zhou, H.-L. Xu, Chem. Phys. Lett. 2016, 647, 46.
[129] S. Wang, P. Shao, G. Du, C. Xi, J. Org. Chem. 2016, 81, 6672.
[130] Z. Zhang, T. Ju, M. Miao, J.-L. Han, Y.-H. Zhang, X.-Y. Zhu, J.-H. Ye, D.-G. Yu, Y.-G. Zhi, Org. Lett. 2017, 19, 396.
[131] X. Lv, L. Zhang, B. Sun, Z. Li, Y.-B. Wu, G. Lu, Catal. Sci. Technol. 2017, 7, 3539.
[132] W. Y. Gao, H. Wu, K. Leng, Y. Sun, S. Ma, Angew. Chem. Int. Ed. 2016, 55, 5472.
[133] T. Mita, H. Tanaka, K. Michigami, Y. Sato, Synlett 2014, 25, 1291.
[134] A. Ueno, M. Takimoto, W. N. O Wylie, M. Nishiura, T. Ikariya, Z. Hou, Chem. Asian J. 2015, 10, 1010.
[135] L. Fu, S. Li, Z. Cai, Y. Ding, X.-Q. Guo, L.-P. Zhou, D. Yuan, Q.-F. Sun, G. Li, Nature Catalysis 2018, 1, 469.
[136] Z. Cai, S. Li, Y. Gao, G. Li, Adv. Synth. Catal. 2018, 360, 4005.
[137] R. Huang, S. Li, L. Fu, G. Li, Asian Journal of Organic Chemistry 2018, 7, 1376.
[138] L. Song, G.-M. Cao, W.-J. Zhou, J.-H. Ye, Z. Zhang, X.- Y. Tian, J. Li, D.-G. Yu, Org. Chem. Front. 2018, 5, 2086.
[139] L. Song, L. Zhu, Z. Zhang, J.-H. Ye, S.-S. Yan, J.-L. Han, Z.-B. Yin, Y. Lan, D.-G. Yu, Org. Lett. 2018, 20, 3776.
[140] K. Sasano, J. Takaya, N. Iwasawa, J. Am. Chem. Soc. 2013, 135, 10954.
[141] Z. Zhang, L. L. Liao, S. S. Yan, L. Wang, Y. Q. He, J. H. Ye, J. Li, Y. G. Zhi, D. G. Yu, Angew. Chem. Int. Ed. 2016, 55, 7068.
[142] S. Tanaka, K. Watanabe, Y. Tanaka, T. Hattori, Org. Lett. 2016, 18, 2576.
[143] Z. Zhang, C.-J. Zhu, M. Miao, J.-L. Han, T. Ju, L. Song, J.-H. Ye, J. Li, D.-G. Yu, Chinese. J. Chem. 2018, 36, 430.
[144] M. L. Lejkowski, R. Lindner, T. Kageyama, G. É. Bódizs, P. N. Plessow, I. B. Müller, A. Schäfer, F. Rominger, P. Hofmann, C. Futter, S. A. Schunk, M. Limbach, Chem. Eur. J. 2012, 18, 14017.
[145] C. Hendriksen, E. A. Pidko, G. Yang, B. Schäffner, D. Vogt, Chem. Eur. J. 2014, 20, 12037.
[146] N. Huguet, I. Jevtovikj, A. Gordillo, M. L. Lejkowski, R. Lindner, M. Bru, A. Y. Khalimon, F. Rominger, S. A. Schunk, P. Hofmann, M. Limbach, Chem. Eur. J. 2014, 20, 16858.
[147] S. C. E. Stieber, N. Huguet, T. Kageyama, I. Jevtovikj, P. Ariyananda, A. Gordillo, S. A. Schunk, F. Rominger, P. Hofmann, M. Limbach, Chem. Commun. 2015, 51, 10907.
S. Manzini, N. Huguet, O. Trapp, T. Schaub, Eur. J. Org. Chem. 2015, 2015, 7122.
[149] I. Jevtovikj, S. Manzini, M. Hanauer, F. Rominger, T. Schaub, Dalton Trans. 2015, 44, 11083.
[150] S. Manzini, N. Huguet, O. Trapp, R. A. Paciello, T. Schaub, Catal. Today 2017, 281, 379.
[151] S. Manzini, A. Cadu, A.-C. Schmidt, N. Huguet, O. Trapp, R. Paciello, T. Schaub, ChemCatChem 2017, 9, 2269.
[152] I. Knopf, D. Tofan, D. Beetstra, A. Al-Nezari, K. Al-Bahily, C. C. Cummins, Chem. Sci. 2017, 8, 1463.
[153] I. Knopf, M.-A. Courtemanche, C. C. Cummins, Organometallics. 2017, 36, 4834.
[154] Q. Liu, M. Li, R. Xiong, F. Mo, Org. Lett. 2017, 19, 6756.
[155] M. Juhl, J.-W. Lee, Angew. Chem. Int. Ed. 2018, 57, 12318.
G. Bottaccio, G. P. Chiusoli, Chem. Commun. (London) 1966, 618a.
[157] E. J. Corey, R. H. K. Chen, J. Org. Chem. 1973, 38, 4086.
[158] Y. Otsuji, M. Arakawa, N. Matsumura, E. Haruki, Chem. Lett. 1973, 2, 1193.
[159] T. Ito, Y. Takami, Chem. Lett. 1974, 3, 1035.
[160] K. Chiba, T. Akama, K. Sakakibara, K. Horie, Chem. Lett. 1978, 7, 1387.
[161] H. Sakurai, A. Shirahata, A. Hosomi, Tetrahedron Lett. 1980, 21, 1967.
[162] K. Chiba, K. Sakakibara, A. Ota, Nippon Kagaku Kaishi 1983, 1983, 118.
[163] R. E. Tirpak, R. S. Olsen, M. W. Rathke, J. Org. Chem. 1985, 50, 4877.
[164] H. Hogeveen, W. M. P. B. Menge, Tetrahedron Lett. 1986, 27, 2767.
[165] H. Mori, Bull. Chem. Soc. Jpn. 1988, 61, 435.
[166] K. Tanaka, R. Wakita, T. Tanaka, J. Am. Chem. Soc. 1989, 111, 2428.
[167] Y. Hirai, T. Aida, S. Inoue, J. Am. Chem. Soc. 1989, 111, 3062.
[168] K. Chiba, H. Tagaya, M. Karasu, T. Ono, M. Kugiya, Chem. Lett. 1990, 19, 39.
K. Chiba, H. Tagaya, S. Miura, M. Karasu, Chem. Lett. 1992, 21, 923.
H. Abe, S. Inoue, J. Chem. Soc. Chem. Commun. 1994, 1197.
[171] B. J. Flowers, R. Gautreau - Service, P. G. Jessop, Adv. Synth. Catal. 2008, 350, 2947.
[172] E. J. Beckman, P. Munshi, Green Chem. 2011, 13, 376.
[173] B. R. Van Ausdall, N. F. Poth, V. A. Kincaid, A. M. Arif, J. Louie, J. Org. Chem. 2011, 76, 8413.
[174] H. Senboku, Y. Yamauchi, T. Fukuhara, S. Hara, Electrochemistry. 2006, 74, 612.
[175] T. Baran, A. Dibenedetto, M. Aresta, K. Kruczala, W. Macyk, ChemPlusChem 2014, 79, 708.
[176] P. D. Re, E. Sandri, Chemische Berichte 1960, 93, 1085.
[177] S. Kikuchi, K. Sekine, T. Ishida, T. Yamada, Angew. Chem. Int. Ed. 2012, 51, 6989.
[178] K. Sekine, A. Takayanagi, S. Kikuchi, T. Yamada, Chem. Commun. 2013, 49, 11320.
[179] W.-Z. Zhang, L.-L. Shi, C. Liu, X.-T. Yang, Y.-B. Wang, Y. Luo, X.-B. Lu, Org. Chem. Front. 2014, 1, 275.
REVIEW
[180] W. Z. Zhang, S. Liu, X. B. Lu, Beilstein J. Org. Chem. 2015, 11, 906.
[181] W.-Z. Zhang, M.-W. Yang, X.-T. Yang, L.-L. Shi, H.-B. Wang, X.-B. Lu, Org. Chem. Front. 2016, 3, 217.
[182] Y. Sadamitsu, K. Komatsuki, K. Saito, T. Yamada, Org. Lett. 2017, 19, 3191.
[183] M. Zhou, Y. Yoshida, S. Ishii, H. Noguchi, Synthetic. Commun. 1999, 29, 2241.
[184] W.-Z. Zhang, M.-W. Yang, X.-B. Lu, Green Chem. 2016, 18, 4181.
[185] T. Mita, K. Michigami, Y. Sato, Org. Lett. 2012, 14, 3462.
[186] T. Mita, K. Michigami, Y. Sato, Chem. Asian J. 2013, 8, 2970.
[187] H. Seo, M. H. Katcher, T. F. Jamison, Nat. Chem. 2016, 9, 453.
[188] Y. Masuda, N. Ishida, M. Murakami, J. Am. Chem. Soc. 2015, 137, 14063.
[189] S.-H. Su, M.-D. Su, RSC Adv. 2016, 6, 50825.
[190] K. Kudo, F. Ikoma, S. Mori, N. Sugita, J. Jpn. Petrol. Inst. 1995, 38, 48.
[191] N. Ishida, Y. Masuda, S. Uemoto, M. Murakami, Chem. Eur. J. 2016, 22, 6524.
[192] K. Michigami, T. Mita, Y. Sato, J. Am. Chem. Soc. 2017, 139, 6094.
[193] A. Ueno, M. Takimoto, Z. Hou, Org. Biomol. Chem. 2017, 15, 2370.
[194] A. D. English, T. Herskovitz, J. Am. Chem. Soc. 1977, 99, 1648.
[195] P. Braunstein, D. Matt, Y. Dusausoy, J. Fischer, A. Mitschler, L. Ricard, J. Am. Chem. Soc. 1981, 103, 5115.
[196] A. Behr, E. Herdtweck, W. A. Herrmann, W. Keim, W. Kipshagen, J. Chem. Soc. Chem. Commun. 1986, 1262.
[197] A. Behr, E. Herdtweck, W. A. Herrmann, W. Keim, W. Kipshagen, Organometallics. 1987, 6, 2307.
[198] P. Braunstein, D. Matt, D. Nobel, J. Am. Chem. Soc. 1988, 110, 3207.
[199] J. Langer, M. J. Fabra, P. Garcia-Orduna, F. J. Lahoz, L. A. Oro, Chem. Commun. 2008, 4822.
[200] J. Langer, M. J. Fabra, P. Garcia-Orduna, F. J. Lahoz, H. Gorls, L. A. Oro, M. Westerhausen, Dalton Trans. 2010, 39, 7813.
[201] V. T. Annibale, D. Song, Chem. Commun. 2012, 48, 5416.
[202] V. T. Annibale, D. A. Dalessandro, D. Song, J. Am. Chem. Soc. 2013, 135, 16175.
[203] Y. Yang, M. Xu, D. Song, Chem. Commun. 2015, 51, 11293.
[204] M. Kira, T. Miyazawa, S.-y. Koshihara, Y. Segawa, H. Sakurai, Chem. Lett. 1995, 24, 3.
[205] Y. Fujiwara, Y. Taniguchi, K. Takaki, M. Kurioka, T. Jintoku, T. Kitamura, in Stud. Surf. Sci. Catal., Vol. 107 (Eds.: M. de Pontes, R. L. Espinoza, C. P. Nicolaides, J. H. Scholtz, M. S. Scurrell), Elsevier, 1997, pp. 275.
[206] Y. Taniguchi, T. Hayashida, T. Kitamura, Y. Fujiwara, in Stud. Surf. Sci. Catal., Vol. 114 (Eds.: T. Inui, M. Anpo, K. Izui, S. Yanagida, T. Yamaguchi), Elsevier, 1998, pp. 439.
[2
[2
[2
[2
[2
‘] W. Huang, K. C. Xie, J. P. Wang, Z. H. Gao, L. H. Yin, Q. M. Zhu, J. Catal. 2001, 201, 100.
[208] C.-j. Liu, Y. Li, Y.-p. Zhang, Y. Wang, J. Zou, B. Eliasson, B. Xue, Chem. Lett. 2001, 30, 1304.
[209] E. M. Wilcox, M. R. Gogate, J. J. Spivey, G. W. Roberts, in Stud. Surf. Sci. Catal., Vol. 136 (Eds.: E. Iglesia, J. J. Spivey, T. H. Fleisch), Elsevier, 2001, pp. 259.
[210] E. M. Wilcox, G. W. Roberts, J. J. Spivey, Applied Catalysis A: General 2002, 226, 317.
[211] Y. Li, C.-J. Liu, B. Eliasson, Y. Wang, Energ. Fuel. 2002, 16, 864.
[212] E. M. Wilcox, G. W. Roberts, J. J. Spivey, Catal. Today 2003, 88, 83.
[213] Y.-p. Zhang, Y. Li, Y. Wang, C.-j. Liu, B. Eliasson, Fuel Process. Technol 2003, 83, 101.
[214] M. Zerella, S. Mukhopadhyay, A. T. Bell, Org. Lett. 2003, 5, 3193.
[215] S. Mukhopadhyay, M. Zerella, A. T. Bell, in Stud. Surf. Sci. Catal., Vol. 147 (Eds.: X. Bao, Y. Xu), Elsevier, 2004, pp. 523.
[216] Y.-H. Ding, W. Huang, Y.-G. Wang, Fuel Process. Technol 2007, 88, 319.
[217] J. J. Spivey, E. M. Wilcox, G. W. Roberts, Catal. Commun. 2008, 9, 685.
[218] H. Sellers, R. J. Spiteri, M. Perrone, J. Phys. Chem. C 2009, 113, 2340.
[219] W. Huang, W. Z. Sun, F. Li, Aiche J. 2009, 56, 1279.
[220] V. Havran, M. P. Duduković, C. S. Lo, Ind. Eng. Chem. Res. 2011, 50, 7089.
[221] R. Zhang, L. Song, H. Liu, B. Wang, Appl. Catal. A-Gen. 2012, 443-444, 50.
[222] W. Panjan, J. Sirijaraensre, C. Warakulwit, P. Pantu, J. Limtrakul, Phys. Chem. Chem. Phys. 2012, 14, 16588.
[223] J.-F. Wu, S.-M. Yu, W. D. Wang, Y.-X. Fan, S. Bai, C.- W. Zhang, Q. Gao, J. Huang, W. Wang, J. Am. Chem. Soc. 2013, 135, 13567.
U. Patil, Y. Saih, E. Abou-Hamad, A. Hamieh, J. D. A. Pelletier, J. M. Basset, Chem. Commun. 2014, 50, 12348.
Y. Zhao, C. Cui, J. Han, H. Wang, X. Zhu, Q. Ge, J. Am. Chem. Soc. 2016, 138, 10191.
[226] B. D. Montejo-Valencia, Y. J. Pagán-Torres, M. M. Martínez-Iñesta, M. C. Curet-Arana, ACS Catal. 2017, 7, 6719.
[227] A. M. Rabie, M. A. Betiha, S.-E. Park, Appl. Catal. B-Environ 2017, 215, 50.
[228] R. Shavi, J. Ko, A. Cho, J. W. Han, J. G. Seo, Applied Catalysis B: Environmental 2018, 229, 237.
[229] M. Dutra, M. Schmal, R. Guardani, Catal. Lett. 2018, 148, 3413.
[230] W. Sangthong, M. Probst, J. Limtrakul, Chemphyschem 2014, 15, 514.
[231] C.-C. Chen, C.-H. Yeh, C.-C. Chang, J.-J. Ho, Ind. Eng. Chem. Res. 2015, 54, 1539.
[232] F. Juliá-Hernández, T. Moragas, J. Cornella, R. Martin, Nature 2017, 545, 84.