Direct Carboxylation of the Diazo Group ipso-C(sp2)–H bond with Carbon Dioxide: Access to Unsymmetrical Diazomalonates and Derivatives
中文 / Bilingual · English version
Qianyi Liu, Man Li, Rui Xiong, and Fanyang Mo*®
Department of Energy and Resources Engineering, College of Engineering, Peking University, Beijing 100871, China
3 Supporting Information
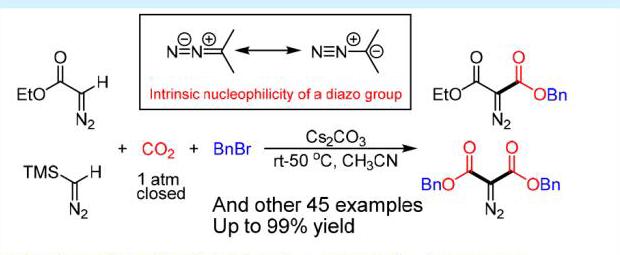
ABSTRACT: The direct carboxylation of the ipso- $\mathrm{C}\left( {\mathrm{{sp}}}^{2}\right) - \mathrm{H}$ bond of a diazo compound with carbon dioxide under mild reaction conditions is described. This method is transition-metal-free, uses a weak base, and proceeds at ambient temperature under atmospheric pressure in carbon dioxide. The carboxylation exhibits high reactivity and is amenable to subsequent diversification. A series of unsymmetrical 1,3-diester/keto/amide diazo compounds are obtained with moderate to excellent yields (up to 99%) with good functional group compatibility.
Non-transitional metal catalysis - Atmospheric pressure
Normal temperature
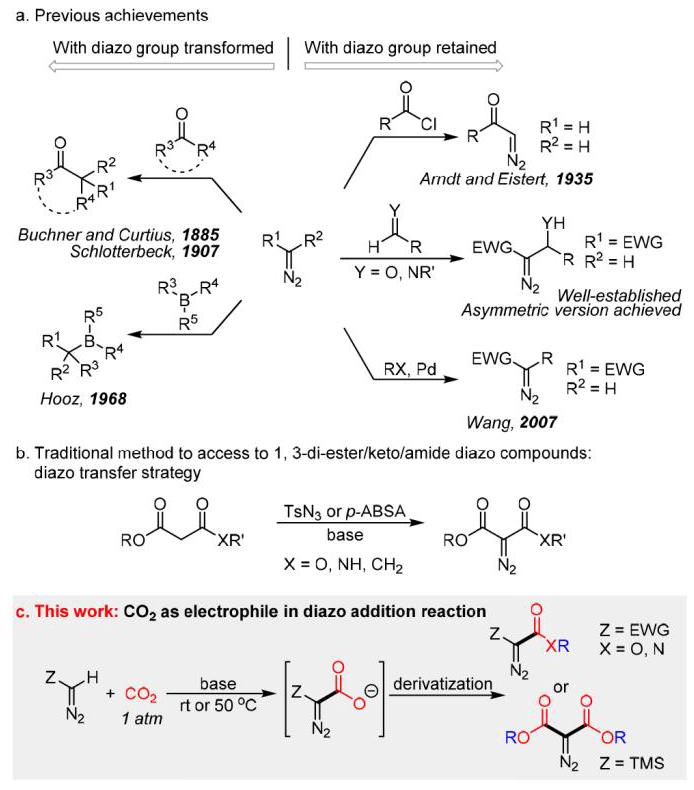
Nucleophilic addition reactions of diazo compounds with various electrophiles have been known for more than a century. ${}^{1}$ Many of these reactions have been developed into useful methods for organic synthesis. For example, the Buchner-Curtius-Schlotterbeck reaction combines aldehydes or ketones with aliphatic diazoalkanes to form homologated ketones. ${}^{2}$ In 1935, Arndt and Eistert reported the nucleophilic addition of diazomethane to acyl chlorides to afford $\alpha$ -diazo ketones. ${}^{3}$ Diazo compounds are generally sensitive to acids but are relatively stable under basic conditions. Base-promoted 1,2- or 1,4-additions to carbonyl compounds with the diazo group retained, as well as their asymmetric version, have been well documented in past decades. ${}^{1,4}$ Besides carbonyl compounds, organoboron reagents have been found to react with diazo compounds under mild conditions. ${}^{5}$ This type of reaction was pioneered by Hooz in ${1968}^{6}$ and further developed by Wang ${}^{7}$ and Barluenga. ${}^{8}$ A palladium-catalyzed cross-coupling reaction using diazo compounds as the nucleophilic partner was developed by Wang in 2007. ${}^{9}$
Looking through this history (Scheme 1a), the most common and available electrophile, carbon dioxide $\left( {\mathrm{{CO}}}_{2}\right)$ , surprisingly has not been used in this addition reaction. ${\mathrm{{CO}}}_{2}$ , a greenhouse gas, has attracted much attention as an inexpensive, nontoxic, nonflammable, renewable, and abundant C1 building block for organic synthesis. ${}^{10}$ The carboxylation of $\mathrm{C} - \mathrm{H}$ bonds using ${\mathrm{{CO}}}_{2}$ is emerging as a straightforward and atom-economic way to synthesize carboxylic acids and their derivatives. ${}^{11}$
We present here an addition reaction of the diazo compound ipso-C-H bond to ${\mathrm{{CO}}}_{2}$ under very mild reaction conditions with the diazo group retained. The resulting $\alpha$ -diazo carboxylate intermediates can be readily converted to esters and amides in a one-pot method (Scheme 1c). From a synthetic perspective, this method is particularly suitable for accessing unsymmetrical diazomalonates and their derivatives (vide infra). While these products can also be prepared via direct diazo transfer onto activated methylenes (Scheme 1b), the unsymmetrical malonate diester starting materials are not readily available and often require special techniques to prepare. ${}^{12}$ In contrast, our method provides modular synthesis of unsymmetrical diazomalonates.
Scheme 1. Diazo Group as Nucleophile
Received: November 17, 2017
Published: December 6, 2017
An electron-withdrawing group at the $\alpha$ position stabilizes the diazo group, which makes the diazo compounds easy to handle but also makes the ipso-C-H bond more acidic. The literature contains many reports of transition-metal-free, base-promoted additions of acidic $\mathrm{C} - \mathrm{H}$ bonds to ${\mathrm{{CO}}}_{2} \cdot {}^{13}$ The carboxylation of several representative types of acidic $\mathrm{C} - \mathrm{H}$ bonds, such as at the $\alpha$ position of a carbonyl compound ${}^{{13a} - 1}$ and at certain sites on an aromatic heterocycle, ${}^{{13j}, m, n}$ has been achieved using various bases such as DBU and alkali carbonates. Thus, we envisioned that a judicious choice of base would be critical to promoting the desired reactivity of diazo compounds with carbon dioxide.
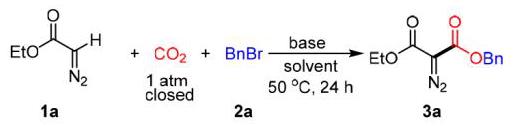
We initiated our studies by investigating the carboxylation of ethyl diazoacetate (EDA) 1a with 1 atm of ${\mathrm{{CO}}}_{2}$ in the presence of a base and an additional electrophile (benzyl bromide 2a in this case) to trap the newly formed carboxylate anion. A variety of bases were examined with 3a obtained with 84% isolated yield using ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ as the base (Table 1, entry 5). The use of an organic base, DBU, was unsuccessful (entry 6). Other solvents, such as dichloromethane, 1,4-dioxane, THF, or DMSO, proved to be less efficient (entries 7-10), while DMF exhibited a positive solvent effect that afforded a 79% yield of 3a (entry 11). Since a stoichiometric amount of carbonate salt is used as the base, the source of the carboxyl group must be accurately identified. To this end, a control experiment was conducted in a ${\mathrm{N}}_{2}$ atmosphere (i.e., in the absence of ${\mathrm{{CO}}}_{2}$ , entry 12). Not surprisingly, this reaction gave only trace amounts of product, indicating that ${\mathrm{{CO}}}_{2}$ is the main source of the carboxyl group for this carboxylation reaction.
Table 1. Optimization of the Reaction Conditions ${}^{a}$
| entry | base | solvent | yield ${}^{b}\left( \% \right)$ |
| 1 | ${\mathrm{K}}_{2}{\mathrm{{CO}}}_{3}$ | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | 9 |
| 2 | ${\mathrm{{Li}}}_{2}{\mathrm{{CO}}}_{3}$ | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | nd |
| 3 | ${\mathrm{{KO}}}^{t}\mathrm{{Bu}}$ | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | trace |
| 4 | CsF | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | 17 |
| 5 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | 84 |
| 6 | DBU | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | trace |
| 7 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | DCM | trace |
| 8 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | 1,4-dioxane | trace |
| 9 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | THF | trace |
| 10 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | DMSO | 30 |
| 11 | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | DMF | 79 |
| ${12}^{c}$ | ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ | ${\mathrm{{CH}}}_{3}\mathrm{{CN}}$ | trace |
${}^{a}$ Reaction conditions: 1a ( ${0.5}\mathrm{{mmol}}$ ), ${\mathrm{{CO}}}_{2}$ (1 atm, closed, ${50}\mathrm{\;{mL}}$ Schlenk flask, about 2 mmol, 4 equiv based on the ideal gas law at 20 ${}^{ \circ }\mathrm{C}$ ), 2a (0.75 mmol, 1.5 equiv), base (0.6 mmol, 1.2 equiv), solvent (2 $\mathrm{{mL}}),{24}\mathrm{\;h}$ , nd, not detected. ${}^{b}$ Isolated yields were reported. ${}^{c}{\mathrm{\;N}}_{2}$ atmosphere.
These optimal reaction conditions were then used for substrate scope studies with the results shown in Table 2. Substituted benzyl bromides, allyl bromide, and $\alpha$ -bromo carbonyl compounds were all good electrophiles for esterification of the carboxylate, which was formed in situ by the addition of the diazo group to ${\mathrm{{CO}}}_{2}$ , furnishing high yields of the products (>80% in most cases). Other common alkyl bromides were less reactive. Thus, alkyl iodides were employed and/or the solvent was switched to DMF (3l, 3n, and 3q) to enhance the reactivity of the esterification step. The chemo-selectivity of this transformation was further demonstrated by the compatibility of ketone and aldehyde functional groups in cases $3\mathbf{g}$ and $3\mathbf{h}$ . $\alpha$ -Diazo alkyl ketones also worked well and produced diazo $\beta$ -keto esters with $> {80}\%$ yields (3r and 3s), while an $\alpha$ -diazo aryl ketone gave a relatively low yield (3t, 45%). This carboxylation-esterification strategy was also applied to the synthesis of $\alpha$ -diazo amides (3u-x) and $\alpha$ - diazo phosphonate (3y) with 65-94% yields.
Table 2. Carboxylation-Esterification of Diazo Compounds ${}^{a}$
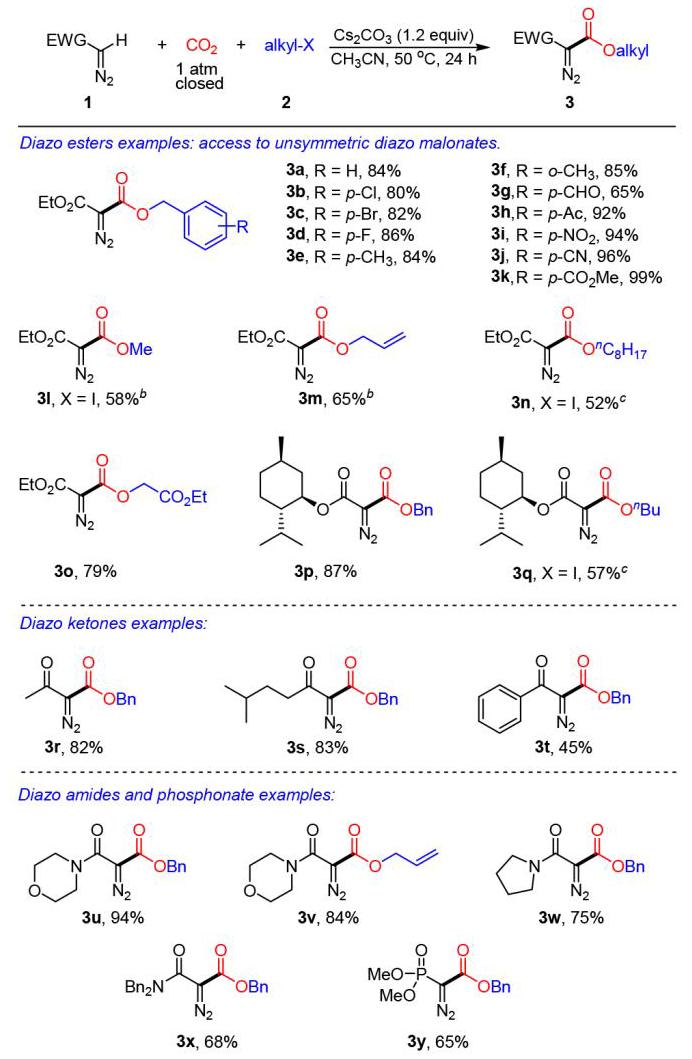
${}^{a}$ Reaction conditions: $1\left( {{0.5}\mathrm{{mmol}}}\right) ,{\mathrm{{CO}}}_{2}$ (1 atm, closed, ${50}\mathrm{\;{mL}}$ Schlenk flask, about 2 mmol, 4 equiv based on the ideal gas law at 20 ${}^{ \circ }\mathrm{C}),2\left( {{0.75}\mathrm{{mmol}},{1.5}\mathrm{{equiv}}}\right) ,{\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}\left( {{0.6}\mathrm{{mmol}},{1.2}\mathrm{{equiv}}}\right)$ , acetonitrile $\left( {2\mathrm{\;{mL}}}\right) ,{50}^{ \circ }\mathrm{C},{24}\mathrm{\;h}$ . Bromides were used unless specified otherwise. Isolated yields were given. ${}^{b}{80}{}^{ \circ }\mathrm{C}$ . ${}^{c}\mathrm{{DMF}}$ is used as solvent. For more details, see the Supporting Information.
The utility of this methodology was further investigated by studying the carboxylation-amidation of $\alpha$ -diazo compounds. The scope is summarized in Table 3. Carboxylation of the diazo ipso- $\mathrm{C} - \mathrm{H}$ bond with ${\mathrm{{CO}}}_{2}$ generated the $\alpha$ -diazo carboxylate intermediate, which reacted with methylsulfonyl chloride to form a mixed anhydride. Finally, an amine was introduced to deliver the final amide products. This protocol gave a range of primary and secondary amines with the amide products obtained with moderate to good yields (50-74%).
Table 3. Carboxylation-Amidation of Diazo Compounds ${}^{a}$
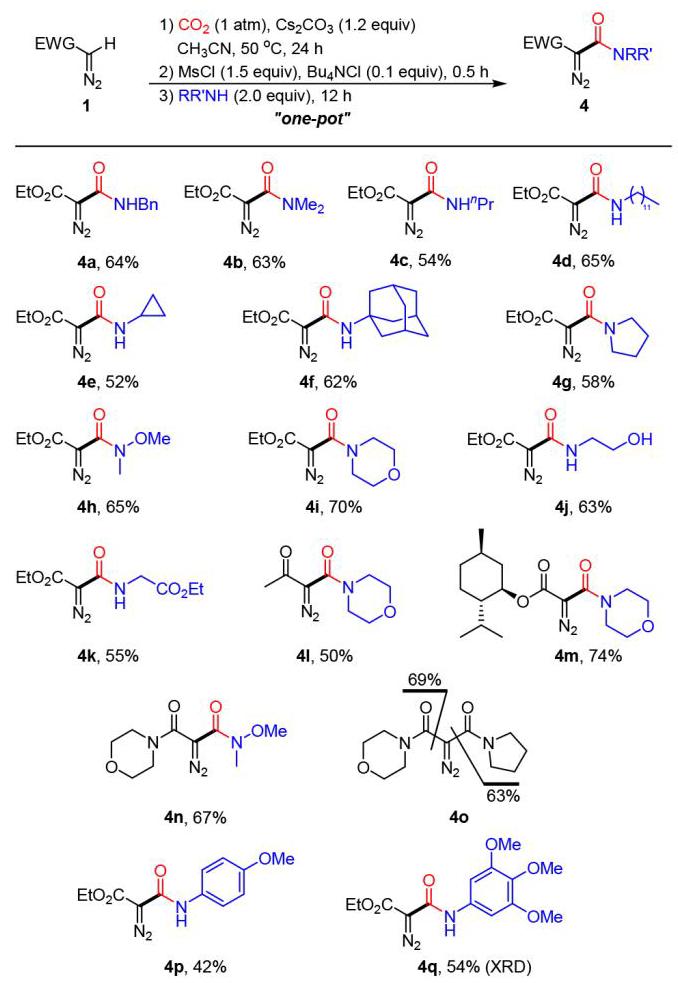
${}^{a}$ Reaction conditions: $1\left( {{0.5}\mathrm{{mmol}}}\right) ,{\mathrm{{CO}}}_{2}(1\mathrm{\;{atm}}$ , closed, ${50}\mathrm{\;{mL}}$ Schlenk flask, about 2 mmol, 4 equiv based on the ideal gas law at 20 ${}^{ \circ }\mathrm{C}),{\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ (0.6 mmol,1.2 equiv), acetonitrile (2 mL), ${50}^{ \circ }\mathrm{C},{24}\mathrm{\;h}$ . Then MsCl (0.75 mmol,1.5 equiv), ${\mathrm{{Bu}}}_{4}\mathrm{{NCl}}$ (0.05 mmol,0.1 equiv), ${50}{}^{ \circ }\mathrm{C},{0.5}\mathrm{\;h}$ . Amine (1 mmol,2 equiv), ${50}{}^{ \circ }\mathrm{C},{12}\mathrm{\;h}$ . Isolated yields were reported.
In addition to stable diazo compounds with an electron-withdrawing group at the $\alpha$ position of the diazo group, labile ${\mathrm{{TMSCHN}}}_{2}$ is also a competent substrate for the desired transformation. This can be attributed to the mild reaction conditions and the high intrinsic reactivity of the diazo $\mathrm{C} - \mathrm{H}$ addition to ${\mathrm{{CO}}}_{2}$ . Thus, increasing the stoichiometry of the base and the halide results in the carboxylation-esterification of ${\mathrm{{TMSCHN}}}_{2}$ proceeding on both sides of the diazo group to furnish symmetrical diazomalonates 5 with moderate to good yields (Table 4, 5a-e, 38-68%).
The deprotonation of monosubstituted diazocarbonyls and subsequent $\mathrm{C} - \mathrm{C}$ bond formation by nucleophilic addition to the electrophiles is well documented. ${}^{1,{14}}$ Accordingly, we propose that our reaction is also an example of deprotona-tion-nucleophilic addition. We also performed a number of mechanistic experiments to support the deprotonation-addition process with the details shown in the Supporting Information.
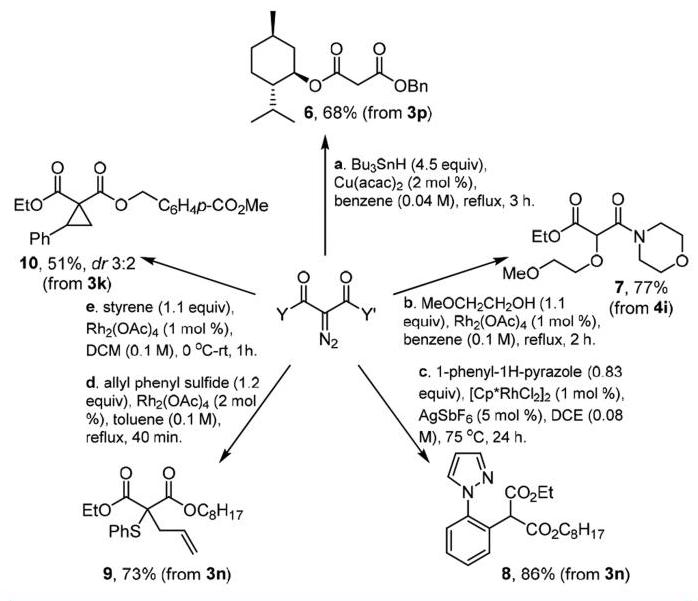
To demonstrate the utility of these unsymmetrical diazo malonates/ketones/amides, several 3 and 4 products were subjected to typical transformations via metal-carbene chemistry. For example, reduction to the methylene group (6), ${}^{15}\mathrm{O} - \mathrm{H}$ insertion (7), ${}^{16}$ formal C-H insertion (8), ${}^{17}$ the sulfonium ylide $\left\lbrack {2,3}\right\rbrack$ -sigmatropic rearrangement (9), ${}^{18}$ and cyclopropanation ${\left( {10}\right) }^{19}$ can be efficiently performed in a single step from the corresponding diazo starting materials (Scheme 2).
Table 4. Carboxylation-Esterification of TMSCHN ${}_{2}{}^{a}$
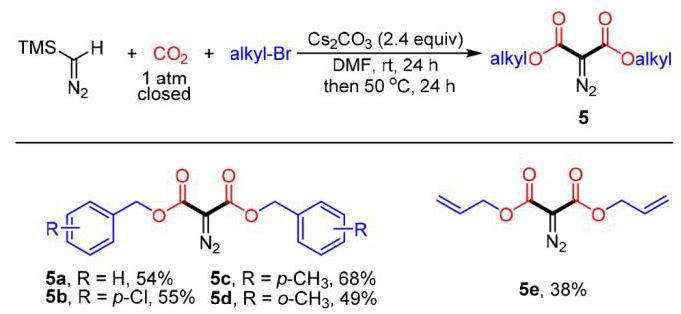
${}^{a}$ Reaction conditions: TMSCHN ${}_{2}$ (0.5 mmol,0.25 mL of $2\mathrm{M}$ in hexane), ${\mathrm{{CO}}}_{2}$ (1 atm, closed, ${100}\mathrm{\;{mL}}$ Schlenk tube, about $4\mathrm{{mmol}},8$ equiv based on the ideal gas law at ${20}^{ \circ }\mathrm{C}),{\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ (1.2 mmol,2.4 equiv), alkyl bromide (3.0 mmol,6 equiv), DMF (2 mL), rt, ${24}\mathrm{\;h}$ , then ${50}^{ \circ }\mathrm{C},{24}\mathrm{\;h}$ . Isolated yields were reported.
Scheme 2. Derivative Reactions of Unsymmetrical 1,3- Dicarbonyl Diazo Compounds
In summary, we have demonstrated the first use of ${\mathrm{{CO}}}_{2}$ as an electrophile for the diazo addition reaction. The reaction harnesses the intrinsic nucleophilicity of the diazo group, so no strong base, no transition metal, and no harsh conditions are needed. The resulting $\alpha$ -diazo carboxylate can be readily converted to esters and amides in a one-pot method. This method provides a new retrosynthetic disconnection for chemists in the preparation of unsymmetrical 1,3-dicarbonyl 2-diazo compounds and their derivatives.
ASSOCIATED CONTENT
G Supporting Information
The Supporting Information is available free of charge on the ACS Publications website at DOI: 10.1021/acs.or-glett.7b03573.
Experiemental procedures, X-ray data for 4q, and NMR spectra (PDF)
Organic Letters
Accession Codes
CCDC 1558263 contains the supplementary crystallographic data for this paper. These data can be obtained free of charge via www.ccdc.cam.ac.uk/data_request/cif, or by contacting The Cambridge Crystallographic Data Centre, 12 Union Road, Cambridge CB2 1EZ, UK; fax: +44 1223 336033.
AUTHOR INFORMATION
Corresponding Author
ORCID ®
Fanyang Mo: 0000-0002-4140-3020
Notes
The authors declare no competing financial interest.
ACKNOWLEDGMENTS
The project was supported by the Natural Science Foundation of China (Grant Nos. 21502003 and 21772003). We also thank the “1000-Youth Talents Plan” and Peking University for startup funds.
The project was supported by the Natural Science Foundation of China (Grant Nos. 21502003 and 21772003). We also thank the “1000-Youth Talents Plan” and Peking University for startup funds.
REFERENCES
(1) Zhang, Y.; Wang, J. Chem. Commun. 2009, 5350.
(2) (a) Buchner, E.; Curtius, T. Ber. Dtsch. Chem. Ges. 1885, 18, 2371. (b) Schlotterbeck, F. Ber. Dtsch. Chem. Ges. 1907, 40, 1826. (c) Gutsche, C. D. Org. React. 1954, 8, 364. (d) Candeias, N. R.; Paterna, R.; Gois, P. M. P. Chem. Rev. 2016, 116, 2937.
(3) Arndt, F.; Eistert, B. Ber. Dtsch. Chem. Ges. B 1935, 68, 200.
(4) (a) Ye, T.; McKervey, M. A. Chem. Rev. 1994, 94, 1091. (b) Doyle, M. P.; McKervey, M. A.; Ye, T. Modern Catalytic Methods for Organic Synthesis with Diazo Compounds: From Cyclopropanes to Ylides; John Wiley & Sons: New York, 1998. (c) Zhang, Z.; Wang, J. Tetrahedron 2008, 64, 6577. (d) Ford, A.; Miel, H.; Ring, A.; Slattery, C. N.; Maguire, A. R.; McKervey, M. A. Chem. Rev. 2015, 115, 9981. (e) Qiu, D.; Qiu, M.; Ma, R.; Zhang, Y.; Wang, J. Huaxue Xuebao 2016, 74, 472.
(5) Li, H.; Zhang, Y.; Wang, J. Synthesis 2013, 45, 3090.
(6) (a) Hooz, J.; Linke, S. J. Am. Chem. Soc. 1968, 90, 5936. (b) Hooz, J.; Linke, S. J. Am. Chem. Soc. 1968, 90, 6891.
(7) (a) Peng, C.; Zhang, W.; Yan, G.; Wang, J. Org. Lett. 2009, 11, 1667. (b) Li, H.; Wang, L.; Zhang, Y.; Wang, J. Angew. Chem., Int. Ed. 2012, 51, 2943. (c) Li, H.; Shangguan, X.; Zhang, Z.; Huang, S.; Zhang, Y.; Wang, J. Org. Lett. 2014, 16, 448.
(8) Barluenga, J.; Tomás-Gamasa, M.; Aznar, F.; Valdés, C. Nat. Chem. 2009, 1, 494.
(9) Peng, C.; Cheng, J.; Wang, J. J. Am. Chem. Soc. 2007, 129, 8708. (10) (a) Aresta, M.; Dibenedetto, A. Dalton Trans. 2007, 2975. (b) Sakakura, T.; Choi, J.-C.; Yasuda, H. Chem. Rev. 2007, 107, 2365. (c) Mikkelsen, M.; Jorgensen, M.; Krebs, F. C. Energy Environ. Sci. 2010, 3, 43. (d) Cokoja, M.; Bruckmeier, C.; Rieger, B.; Herrmann, W. A.; Kühn, F. E. Angew. Chem., Int. Ed. 2011, 50, 8510. (e) Huang, K.; Sun, C.-L.; Shi, Z.-J. Chem. Soc. Rev. 2011, 40, 2435. (f) Omae, I. Coord. Chem. Rev. 2012, 256, 1384. (g) Tsuji, Y.; Fujihara, T. Chem. Commun. 2012, 48, 9956. (h) Yang, Z.-Z.; He, L.-N.; Gao, J.; Liu, A.- H.; Yu, B. Energy Environ. Sci. 2012, 5, 6602. (i) Yu, B.; Diao, Z.-F.; Guo, C.-X.; He, L.-N. J. CO2 Util. 2013, 1, 60. (j) Aresta, M.; Dibenedetto, A.; Angelini, A. Chem. Rev. 2014, 114, 1709. (k) Maeda, C.; Miyazaki, Y.; Ema, T. Catal. Sci. Technol. 2014, 4, 1482. (l) Yeung, C. S.; Dong, V. M. Top. Catal. 2014, 57, 1342. (m) Fiorani, G.; Guo, W.; Kleij, A. W. Green Chem. 2015, 17, 1375. (n) Liu, Q.; Wu, L.; Jackstell, R.; Beller, M. Nat. Commun. 2015, 6, 5933. (o) Yu, B.; He, L.-N. ChemSusChem 2015, 8, 52. (p) Börjesson, M.; Moragas, T.; Gallego, D.; Martin, R. ACS Catal. 2016, 6, 6739.
(11) (a) Zhang, W.; Lu, X. Chin. J. Catal. 2012, 33, 745. (b) Cai, X.; Xie, B. Synthesis 2013, 45, 3305. (c) Liu, A.-H.; Yu, B.; He, L.-N. Greenhouse Gases: Sci. Technol. 2015, 5, 17. (d) Yu, D.; Teong, S. P.; Zhang, Y. Coord. Chem. Rev. 2015, 293-294, 279. (e) Sekine, K.; Yamada, T. Chem. Soc. Rev. 2016, 45, 4524. (f) Wang, S.; Du, G.; Xi, C. Org. Biomol. Chem. 2016, 14, 3666. (g) Wu, X.-F.; Zheng, F. Top. Curr. Chem. 2017, 375, 4.
(12) Niwayama, S.; Cho, H. Chem. Pharm. Bull. 2009, 57, 508.
(13) (a) Bottaccio, G.; Chiusoli, G. P. Chem. Commun. (London) 1966, 618a. (b) Haruki, E.; Arakawa, M.; Matsumura, N.; Otsuji, Y.; Imoto, E. Chem. Lett. 1974, 3, 427. (c) Sakurai, H.; Shirahata, A.; Hosomi, A. Tetrahedron Lett. 1980, 21, 1967. (d) Mori, H.; Satake, Y. Chem. Pharm. Bull. 1985, 33, 3469. (e) Tirpak, R. E.; Olsen, R. S.; Rathke, M. W. J. Org. Chem. 1985, 50, 4877. (f) Hisakazu, M. Bull. Chem. Soc. Jpn. 1988, 61, 435. (g) Chiba, K.; Tagaya, H.; Miura, S.; Karasu, M. Chem. Lett. 1992, 21, 923. (h) Abe, H.; Inoue, S. J. Chem. Soc., Chem. Commun. 1994, 1197. (i) Flowers, B. J.; Gautreau-Service, R.; Jessop, P. G. Adv. Synth. Catal. 2008, 350, 2947. (j) Vechorkin, O.; Hirt, N.; Hu, X. Org. Lett. 2010, 12, 3567. (k) Dingyi, Y.; Yugen, Z. Green Chem. 2011, 13, 1275. (1) Wang, X.; Lim, Y. N.; Lee, C.; Jang, H.-Y.; Lee, B. Y. Eur. J. Org. Chem. 2013, 2013, 1867. (m) Banerjee, A.; Dick, G. R.; Yoshino, T.; Kanan, M. W. Nature 2016, 531, 215. (n) Fenner, S.; Ackermann, L. Green Chem. 2016, 18, 3804. (o) Zhang, W.-Z.; Yang, M.-W.; Lu, X.-B. Green Chem. 2016, 18, 4181.
(14) (a) Aggarwal, V. K.; Sheldon, C. G.; Macdonald, G. J.; Martin, W. P. J. Am. Chem. Soc. 2002, 124, 10300. (b) Jiang, N.; Wang, J. Tetrahedron Lett. 2002, 43, 1285. (c) Matsuya, Y.; Ohsawa, N.; Nemoto, H. J. Am. Chem. Soc. 2006, 128, 13072. (d) Xiao, F.; Liu, Y.; Wang, J. Tetrahedron Lett. 2007, 48, 1147. (e) Draghici, C.; Brewer, M. J. Am. Chem. Soc. 2008, 130, 3766.
(15) Tan, Z.; Qu, Z.; Chen, B.; Wang, J. Tetrahedron 2000, 56, 7457. (16) Davis, O. A.; Bull, J. A. Angew. Chem., Int. Ed. 2014, 53, 14230.
(17) Yu, X.; Yu, S.; Xiao, J.; Wan, B.; Li, X. J. Org. Chem. 2013, 78, 5444.
(18) Takano, S.; Tomita, S.; Takahashi, M.; Ogasawara, K. Chem. Lett. 1987, 16, 1569.
(19) De Simone, F.; Saget, T.; Benfatti, F.; Almeida, S.; Waser, J. Chem. - Eur. J. 2011, 17, 14527.