Transition metal-catalyzed ketone-directed or mediated C–H functionalization
中文 / Bilingual · English version
Zhongxing Huang, Hee Nam Lim, Fanyang Mo, Michael C. Young and Guangbin Dong*
Transition metal-catalyzed C-H functionalization has evolved into a prominent and indispensable tool in organic synthesis. While nitrogen, phosphorus and sulfur-based functional groups (FGs) are widely employed as effective directing groups (DGs) to control the site-selectivity of C-H activation, the use of common FGs (e.g. ketone, alcohol and amine) as DGs has been continuously pursued. Ketones are an especially attractive choice of DGs and substrates due to their prevalence in various molecules and versatile reactivity as synthetic intermediates. Over the last two decades, transition metal-catalyzed C-H functionalization that is directed or mediated by ketones has experienced vigorous growth. This review summarizes these advancements into three major categories: use of ketone carbonyls as DGs, direct $\beta$ -functionalization, and $\alpha$ -alkylation/alkenylation with unactivated olefins and alkynes. Each of these subsections is discussed from the perspective of strategic design and reaction discovery.
CrossMark
CrossMark
Eclick for updates
Cite this: DOI: 10.1039/c5cs00272a
Received 31st March 2015
DOI: 10.1039/c5cs00272a
DOI: 10.1039/c5cs00272a
1. Introduction
Transition metal-catalyzed C-H functionalization has emerged as an increasingly important tool for organic synthesis; however, control of site-selectivity remains a challenging and ongoing task. ${}^{1}$ Among all of the approaches developed hitherto, the use of a directing group (DG) represents a versatile and reliable strategy to govern which C-H bond will be functionalized. ${}^{2}$ Through coordination of a DG to a transition metal, the proximity effect can often override the inherent steric/electronic preferences and ultimately dictate the site-selectivity of $\mathrm{C} - \mathrm{H}$ bond activation. Consequently, nitrogen-based functional groups (FGs) and hetero-cycles, as well as soft elements (e.g. sulfur and phosphorus), are widely employed as DGs due to their excellent coordination ability with transition metals. In contrast, the use of a more common FG as DG, such as regular ketones is much rarer due to their relatively low coordinating ability. Undoubtedly the C-H functionalization reactions involving these common FGs as DGs or substrates are highly attractive. For example, ketone moieties are widely found in various bioactive molecules and functional materials; ${}^{3}$ they can also be readily transformed into a diverse range of other FGs, and are thus often employed as versatile synthetic intermediates. For this reason simple ketone-directed or mediated C-H functionali-zation is of high synthetic value.
Conventional ketone-functionalization methods take advantage of the intrinsic electrophilicity of the carbonyl and acidity of the $\alpha - \mathrm{C} - \mathrm{H}$ bonds (Scheme 1A). Carbonyl attack by various nucleophiles allows functionalization at the ipso carbon, whereas deprotonation of the $\alpha$ -hydrogen or enolization with a strong (Lewis) acid permits substitution at the $\alpha$ -positions by reaction with various electrophiles. Alternatively, recent advancements in transition-metal chemistry enable novel ketone-based $\mathrm{C} - \mathrm{H}$ bond transformations that do not occur under traditional conditions. This field has taken off since Murai’s pioneering work on ketone-directed Ru-catalyzed C-H/ olefin coupling reactions in 1993 (vide infra). ${}^{4}$ To date, a number of innovative approaches and methods have been developed.
This review focuses on the simple-ketone-directed or mediated C-H functionalization reactions that are catalyzed by transition metals, covered through January of 2015 (Scheme 1B). The content is organized into three parts: use of ketone carbonyl as the DG, direct $\beta$ -functionalization, and $\alpha$ -alkylation/alkenylation with unactivated olefins and alkynes. It is noteworthy that transition-metal enolate-mediated cross coupling reactions, e.g. Buchwald-Hartwig-Miura $\alpha$ -arylation, ${}^{5}$ and Lewis acid-mediated enolate chemistry with regular electrophiles ${}^{6}$ will not be included. Reactions with related ketone derivatives, such as silyl enol ethers, oximes, $\beta$ -keto esters and $\alpha ,\beta$ -unsaturated ketones, will also not be discussed. While not intended to be comprehensive, this review summarizes major developments from the perspective of strategic design and reaction discovery.
2. Ketone as the directing group
Based on the reaction type, this section is divided into three subsections: carbon-carbon (C-C) bond formation via addition to alkenes and alkynes, C-H arylation, and carbon-heteroatom (C-X) bond formation. While a large number of transformations have been reported using ketone as the directing group, it should be noted that the substrate scopes of these reactions are limited to ${\mathrm{{sp}}}^{2}$ C-H bonds in aromatic, and in some cases, vinyl ketones. Applications of the DG-based strategy to other types of ketones, such as alkyl ketones ( $\beta - {\mathrm{{sp}}}^{3}\mathrm{C} - \mathrm{H}$ bonds, vide infra), are largely underdeveloped, likely due to the weak coordinating ability of ketones.
Department of Chemistry, University of Texas at Austin, Austin, Texas 78712, USA.
Over the past two decades, the DG-based strategy has been established as one prominent approach for C-H functionaliza-tion. ${}^{2}$ One common mode of reactivity consists of coordination of the DG to a transition metal before C-H cleavage and subsequent formation of stable metallacycles (Scheme 2A). To date, a diverse range of DGs are available, among which ketones are often considered weakly coordinating due to their relatively low Lewis basicity. As demonstrated by the ${\mathrm{{BF}}}_{3}$ affinity scale, ketones’ coordinating capability is considerably lower than other directing groups, including pyridines, sulfoxides and amides (Scheme 2B). ${}^{7}$ However, from a chronological viewpoint, ketones are actually one of the earliest DGs employed in transition metal-catalyzed C-H functionalization.
2.1 Coupling with alkenes and alkynes
In 1993, Murai and coworkers reported the first Ru-catalyzed ortho-alkylation reaction of aromatic ketones with olefins, also known as the hydroarylation of olefins (Scheme 3). ${}^{4}$ The ortho-C-H bond of the ketone was selectively coupled with the olefin to afford the alkylation product in up to quantitative yield with ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ as the catalyst in toluene at ${135}{}^{ \circ }\mathrm{C}$ . Detailed studies of the substrate scope revealed that a large collection of olefins can participate in this reaction. ${}^{8}$ Vinylsilanes proved to be superior substrates that gave complete linear selectivity. In the case of styrene, a 6 : 1 ratio between linear and branched products was obtained. Despite the good reactivity with terminal olefins, internal, isomerizable terminal, and acrylate-type olefins gave poor yields or no reaction. Regarding the scope of ketones, aromatic ketones bearing various alkyl, aryl, or hetero-aryl substituents were all alkylated smoothly under the reaction conditions. It is interesting to note that while $\alpha$ -tetralone afforded the product in quantitative yield, 1-indanone showed no reactivity. As explained by the authors, the loss of reactivity with 1-indanone was attributed to the difficulty of forming the five-membered ring metallacycle due to the strained benzo-fused structure.
The proposed catalytic cycle, supported by both mechanistic studies ${}^{9 - {12}}$ and DFT calculations, ${}^{13}$ is initiated by the generation of a $\mathrm{{Ru}}\left( 0\right)$ species through the reduction of ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ with olefin substrates (Scheme 4). The ketone moiety coordinates to
Hee Nam Lim (Far right): Dr Hee Nam Lim received his BSc degree in 2006 and MSc in 2008 from the Department of Chemical Engineering at Hanyang University, Seoul, Korea, under the supervision of Professor Kee-Jung Lee. In 2013, He obtained his PhD degree in Organic Chemistry from SUNY Stony Brook, under the supervision of Professor Kathlyn A. Parker. He is currently a postdoc fellow in Professor Guangbin Dong’s group at the University of Texas at Austin. His research is focused on the $\alpha - C - H$ functionaliza-tion of ketones.
Zhongxing Huang (Center), Hee Nam Lim (Far right), Fanyang Mo (2nd from right), Michael C. Young (2nd from left) and Guangbin Dong (Far left)
Zhongxing Huang (Center): Zhongxing Huang received his BSc degree in Chemistry from Peking University in 2012 under the supervision of Professor Jianbo Wang and Yan Zhang. He is currently a PhD student in Professor Guangbin Dong’s group at the University of Texas at Austin. His research is focused on transition-metal-catalyzed C-H functionalization.
Fanyang Mo (2nd from right): Dr Fanyang Mo obtained his MSc degree in Applied Chemistry in 2006 from Beijing Institute of Technology under the supervision of Professor Zhiming Zhou, and then obtained his PhD degree in Organic Chemistry in 2010 from Peking University under the supervision of Professor Jianbo Wang. After spending 10 months at the Scripps Research Institute under the supervision of Professor Qinghai Zhang, he joined Professor Guangbin Dong’s group at the University of Texas at Austin in 2011. In 2015, he joined the department of engineering at Peking University as an assistant professor.
Michael C. Young (2nd from left): Michael Young received his BSc and MSc degrees in Chemistry under the tutelage of Prof. Brian D. Dinkelmeyer studying hydrogen-bonding self-assembly at Western Carolina University. He went on to pursue PhD work at the University of California - Riverside under Prof. Richard J. Hooley studying the self-assembly of metal-organic polyhedra and their applications towards organocatalysis. He is currently engaged in postdoctoral work in Prof. Guangbin Dong’s group, investigating supramolecular strategies for directed $\mathrm{C} - \mathrm{H}$ functionalization based on aldehyde and ketone functional groups.
Guangbin Dong (Far left): Guangbin Dong received his BSc degree from Peking University and completed his PhD degree in Chemistry from Stanford University with Professor Barry M. Trost where he was a Larry Yung Stanford Graduate Fellow. In 2009, he began research with Prof. Robert H. Grubbs at California Institute of Technology as a Camille and Henry Dreyfus Environmental Chemistry Fellow. In 2011, he joined the Department of Chemistry and Biochemistry at the University of Texas at Austin as an assistant professor and a CPRIT Scholar. His research interests lie at the development of powerful chemical tools for addressing questions of biological importance. and delivers the $\mathrm{{Ru}}\left( 0\right)$ within proximity of the ortho-C-H bond, followed by oxidative addition to generate ruthenium hydride 3. Subsequent migratory insertion of olefin into the Ru-H bond proceeds in a highly regio- and chemoselective manner to give intermediate 5, which after reductive elimination gives the alkylation product and restores the active $\mathrm{{Ru}}\left( 0\right)$ catalyst. The reductive elimination of complex 5 proved to be the rate determining step of the catalytic cycle. While the reaction was run at ${135}^{ \circ }\mathrm{C}$ , the oxidative addition of $\mathrm{{Ru}}\left( 0\right)$ to the aryl $\mathrm{C} - \mathrm{H}$ bond was found to proceed even at ${50}^{ \circ }\mathrm{C}$ .
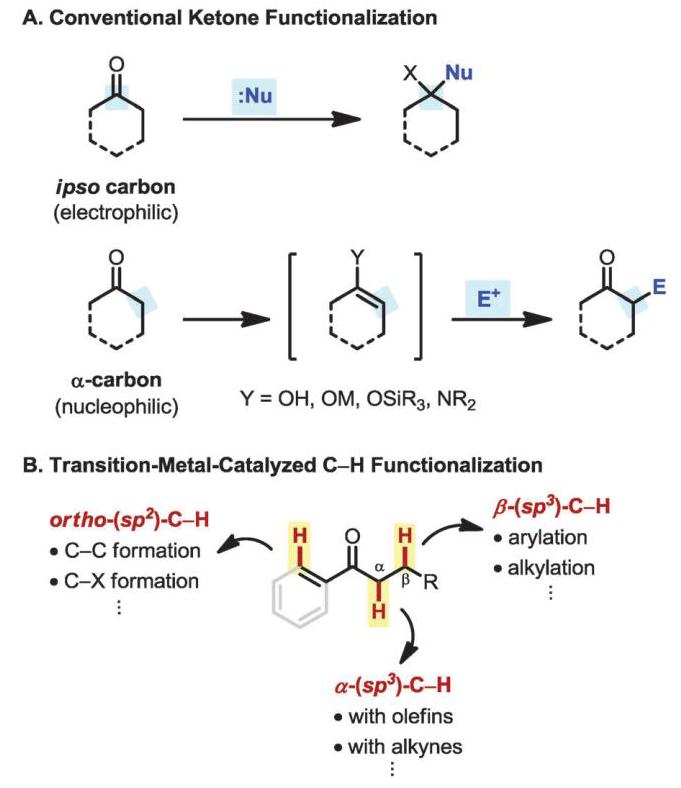
Scheme 1 Functionalization of ketones.
A. Ketone-Directed C-H Functionalization
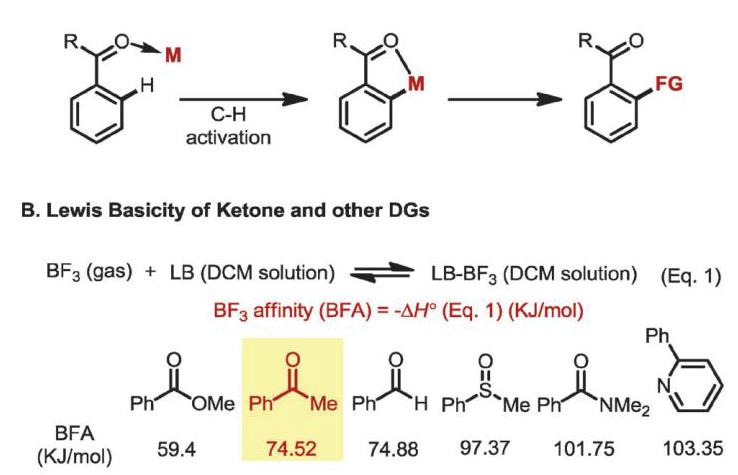
Scheme $2\mathrm{C} - \mathrm{H}$ functionalization directed by ketones.
A more reactive precatalyst was discovered during the mechanistic studies by Kakiuchi, Murai, and coworkers. ${}^{14}$ The spectroscopic evidence supported the generation of a Rh-hydride complex (6) from the reaction of ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ and trimethylvinylsilane (Scheme 5, eqn (1)). This species, once used as the catalyst, could enable the ortho-alkylation to proceed even at room temperature (eqn (2)). When the alkylation was run at ${120}{}^{ \circ }\mathrm{C}$ , complex 6 achieved a turnover number (TON) close to 1000 (eqn (3)).
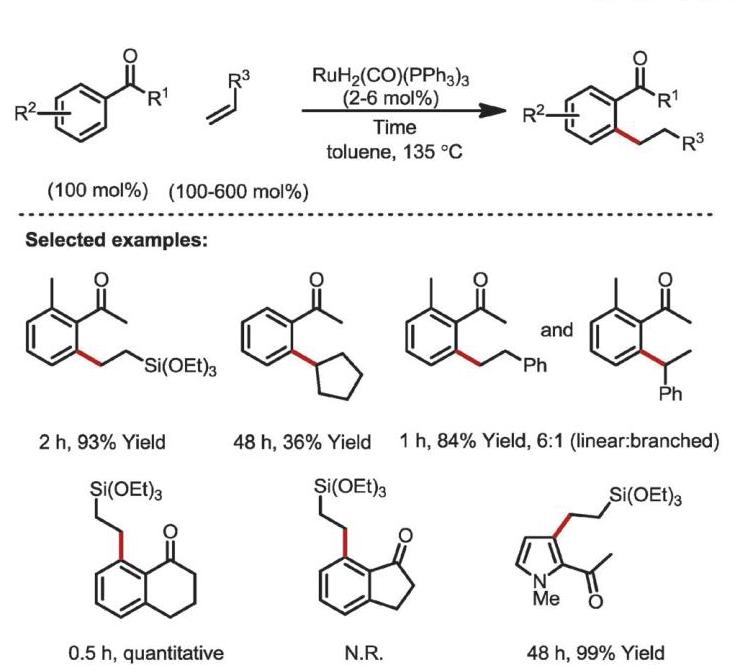
Scheme 3 Ruthenium-catalyzed ortho-alkylation of aromatic ketones.
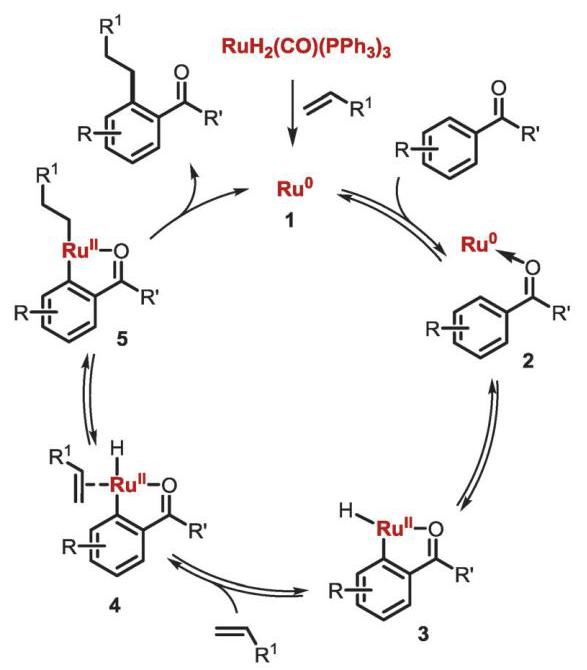
Scheme 4 Proposed catalytic cycle for the ruthenium-catalyzed ortho-alkylation of aromatic ketones.
Other groups have also developed improved catalysts for this transformation. Using ${\mathrm{{RuH}}}_{2}{\left( {\mathrm{H}}_{2}\right) }_{2}{\left( {\mathrm{{PCy}}}_{3}\right) }_{2}$ (7) as the catalyst, the Chaudret ${}^{{15},{16}}$ and Leitner ${}^{{17},{18}}$ groups reported ortho-alkylation with ethylene that proceeded at room temperature (Scheme 6). A variant ruthenium complex with two tricyclopentylphosphine (PCyp3) ligands was also synthesized, characterized, and demonstrated to possess outstanding catalytic activity towards the same ethylation reaction. ${}^{19}$ Compared to Murai’s original precatalyst ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ , these modified ruthenium species have two dihydrogen and two tricycloalkylphosphine ligands. The lability of dihydrogen and strongly electron-donating tricyclohexylphosphine ligand were proposed to generate an active and electron-rich $\mathrm{{Ru}}\left( 0\right)$ catalyst promptly. However, coupling with olefins other than ethylene was not demonstrated using these modified catalysts.
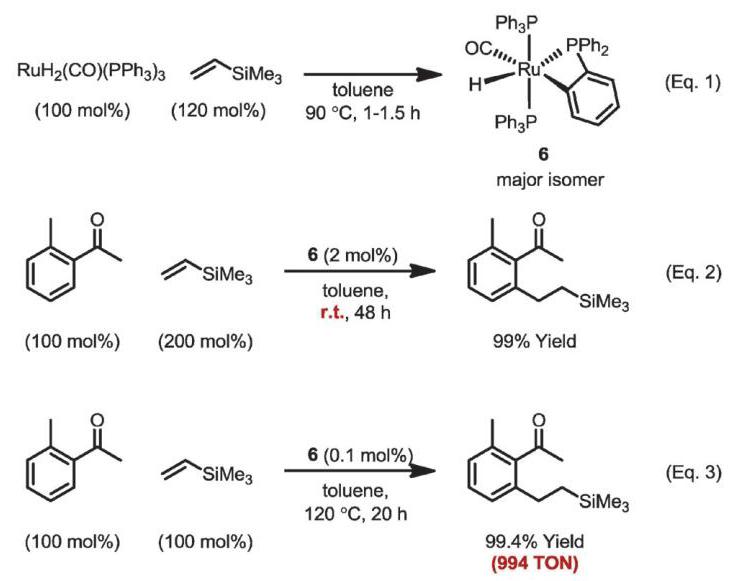
Scheme 5 Formation of active catalyst of ruthemium-catalyzed ortho-alkylation.
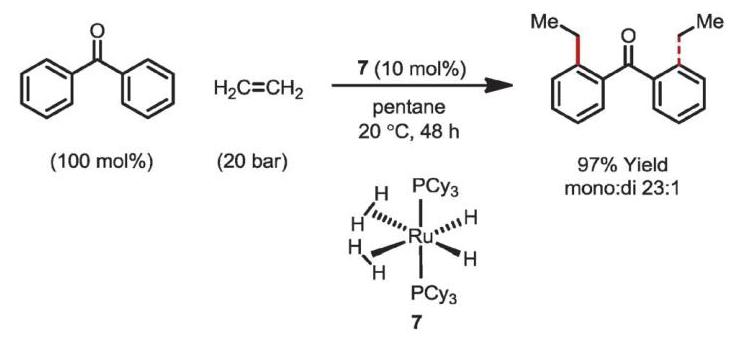
Scheme 6 RuH ${}_{2}{\left( {\mathrm{H}}_{2}\right) }_{2}{\left( {\mathrm{{PCy}}}_{3}\right) }_{2}$ -catalyzed ortho-ethylation with ethylene.
Another important contribution for the catalyst modification came from the Darses group. ${}^{{20} - {23}}$ They developed several protocols for the ortho-alkylation of aromatic ketones using a combination of less expensive Ru salts, such as ${\left\lbrack {\mathrm{{RuCl}}}_{2}\left( p\text{ -cymene }\right) \right\rbrack }_{2}$ and ${\mathrm{{RuCl}}}_{3}$ , along with triarylphosphine ligands and formate salts (Scheme 7). The formate salts were proposed to reduce the Ru(II) or $\mathrm{{Ru}}\left( \mathrm{{III}}\right)$ catalyst in situ to generate the active $\mathrm{{Ru}}\left( 0\right)$ catalyst. This strategy turned out to be highly efficient and the TON could reach more than 1000 (eqn (4)). Besides the lower cost of the precatalysts, the addition of phosphine ligands as a separate reagent presents another advantage over the original catalytic system using ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ . This is because the ruthenium dihydride complexes are often nontrivial to synthesize and sensitive to air and moisture, thus the addition of ligand as a separate reagent makes it easier to evaluate the ligand effect. The authors discovered that the ligand played a vital role in controlling the regioselectivity. For example, the coupling between 1-acetonaphthone and styrene with dicyclohexyl(phenyl)- phosphine significantly enhanced linear selectivity compared to triphenylphosphine (eqn (5)). ${}^{20}$
Besides aromatic C-H bonds, the olefinic C-H bond can also participate in the Ru-catalyzed alkylation with alkenes (Scheme 8). Trost ${}^{24}$ and Kakiuchi/Murai ${}^{{25},{26}}$ independently demonstrated that both linear (eqn (6)) and cyclic vinyl ketones (eqn (7)) are suitable substrates for the alkylation reaction, although $E/Z$ isomerisation from linear enone substrates was observed in some cases. ${}^{24}$ With regard to the alkene scope, vinyl silanes and styrene-type olefins proved to be superior substrates for the coupling. Alkyl olefins and acrylate derivatives also participated in the reaction, although a large excess was required to obtain good yields.
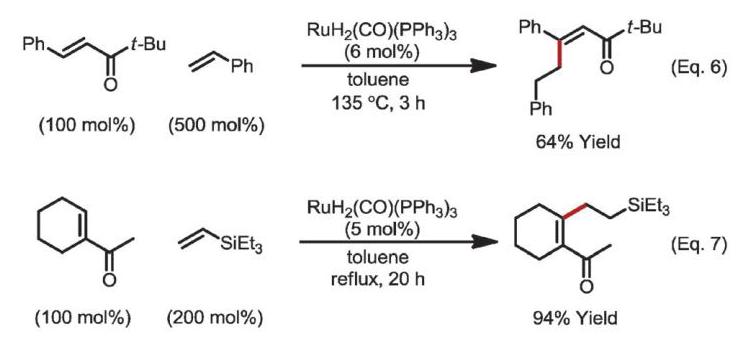
Scheme 8 Ketone-directed ortho-alkylation of olefinic C-H bonds.
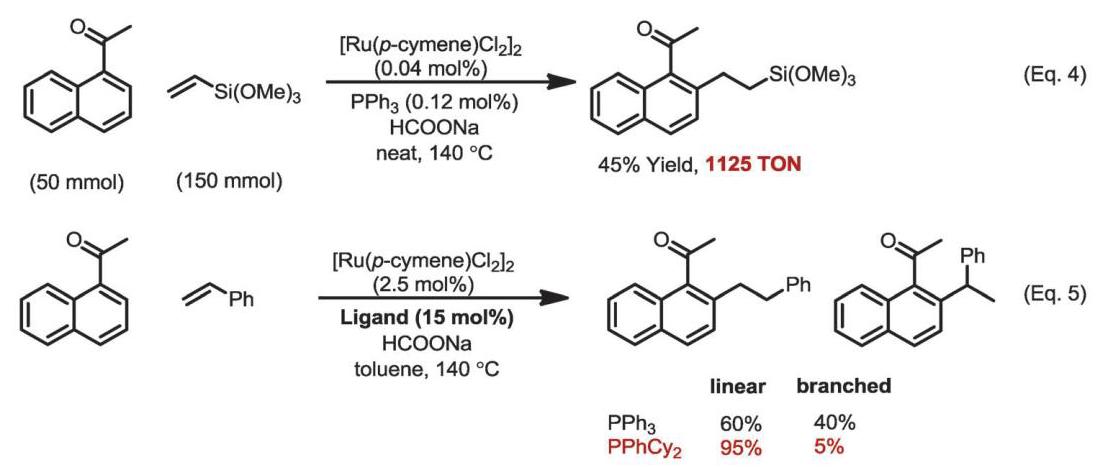
Scheme 7 Ketone-directed ortho-alkylation using Ru(II) precursors.
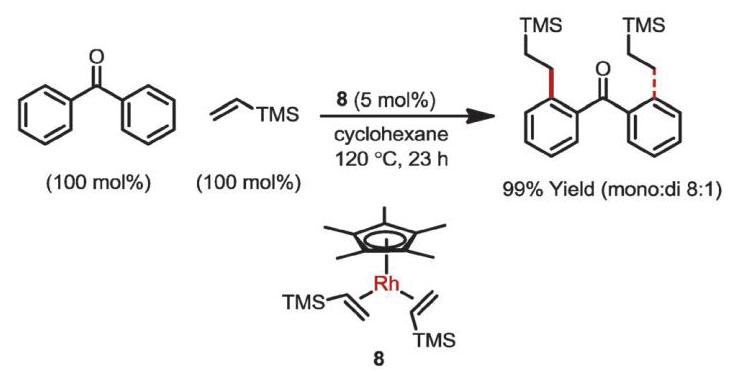
Scheme 9 Rhodium-catalyzed ortho-alkylation of aromatic ketones with olefins.
In addition to ruthenium catalysts, ortho-alkylation of aromatic ketones with olefins has also been reported using other transition-metal catalysts. The rhodium bis-olefin complex 8 was used by Brookhart and coworkers for the hydroarylation of olefins (Scheme 9). ${}^{27}$ A distinct feature of this rhodium catalysis is the non-site-selective oxidative addition of rhodium into aromatic C-H bonds before the migratory insertion. Chelation of the ketone moiety was proposed to lower the barrier of reductive elimination, which is the rate-determining step, thus affording the ortho-selective alkylation product. Besides the distinct mechanism, this rhodium-catalyzed alkylation can accommodate isomerizable olefins (e.g. 1-pentene) that were hard to react under Murai’s ruthenium catalysis.
In 2002, Jun and coworkers reported a benzylamine-assisted ortho-alkylation of aromatic ketones catalyzed by Wilkinson’s catalyst (Scheme 10). ${}^{28}$ A substoichiometric amount (50 mol%) of benzylamine acts as a co-catalyst through transient condensation with the substrate, which converts ketones to stronger coordinating ketimines. After the rhodium-catalyzed ortho-alkylation, the resulting alkylated ketimine would be hydrolyzed to release the product and regenerate the benzylamine (eqn (8)). A similar strategy was later applied to the $\beta$ -alkylation of 4-phenyl-3-buten-2-one (9) (eqn (9)). ${}^{29}$ In this case, a secondary amine was employed to form a transient enamine with 9, followed by directed olefinic C-H activation with a rhodium catalyst. It is interesting to note that after the alkylation reaction, an olefin migration was proposed to take place to give $\beta ,\gamma$ -enone 10 as the major product. Although the substrate scopes of these amine-assisted alkylations were only briefly investigated, the reaction conditions were compatible with a variety of olefins, including vinyl silanes, styrene-type olefins and even isomerizable $\alpha$ -olefins, which represent a key advantage over the Ru-catalyzed ortho-alkylation reactions.
An iridium-catalyzed hydroarylation of olefins was reported by Shibata and coworkers in 2008 (Scheme 11). ${}^{30}$ In their study, the cationic iridium complex $\left\lbrack {\operatorname{Ir}{\left( \operatorname{cod}\right) }_{2}}\right\rbrack$ BARF was used as the catalyst to promote ortho-alkylation of aromatic ketones with styrene-type olefins, giving the linear products as the major isomers (eqn (10)). It is noteworthy that when the chiral bidentate phosphine ligand (R)-MeO-BIPHEP (12) was used, the coupling between norbornene and 11 afforded the product with moderate enantioselectivity (eqn (11)).
With Murai’s protocols, formation of linear alkylation products is favored over branched ones. In 2014, Bower and coworkers reported an alternative iridium-catalyzed system that is highly selective for branched products (Scheme 12). ${}^{31}$ The key for the inversion of the selectivity is the use of an electron-deficient bis-phosphine ligand (13), which can presumably accelerate the reductive elimination. Under their reaction conditions, various aromatic ketones were coupled with styrenes giving better than 20:1 branched/linear selectivity.
An important variation of Murai’s ketone-directed alkylation reaction is the oxidative ortho-olefination of aromatic ketones with alkenes. While Murai’s reaction is viewed as a redox-neutral addition of a $\mathrm{C} - \mathrm{H}$ bond to an olefin, the olefination reaction can be considered as a formal dehydrogenative coupling, also known as the oxidative Heck reaction. ${}^{32}$ It is noteworthy to mention that an olefination product was also observed by Murai and coworkers in the ${\mathrm{{Ru}}}_{3}{\left( \mathrm{{CO}}\right) }_{12}$ -catalyzed addition of $\mathrm{C} - \mathrm{H}$ bonds to olefins, although the dehydrogenative coupling was directed by imidates instead of ketones. ${}^{33}$ In 2011, Glorius and coworkers reported a rhodium-catalyzed C-H olefination reaction directed by ketones (Scheme 13). ${}^{34}$ When styrene-type olefins were coupled with acetophenone derivatives, 1,2-disubstituted trans-alkenes were generated as the sole regioisomer. This selectivity represents a distinct feature from Murai’s ruthenium catalysis, where a mixture of branched and linear alkylation products were produced with styrene-type olefins. In addition, acrylate-type olefins, previously unreactive under Murai’s conditions, afforded the alkenylation products in good yields with this rhodium catalysis.
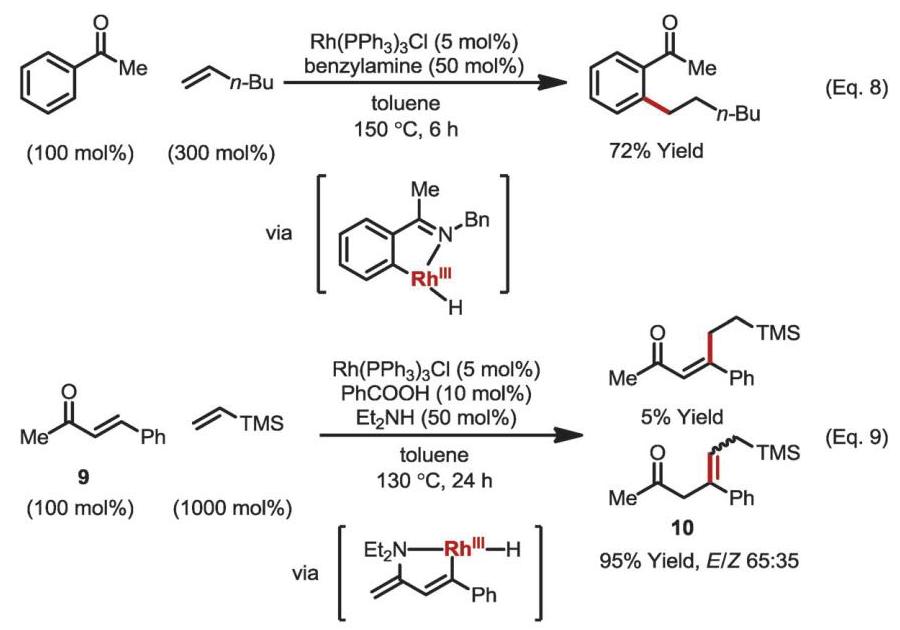
Scheme 10 Amine-assisted ortho-alkylation of aromatic ketones and enones.
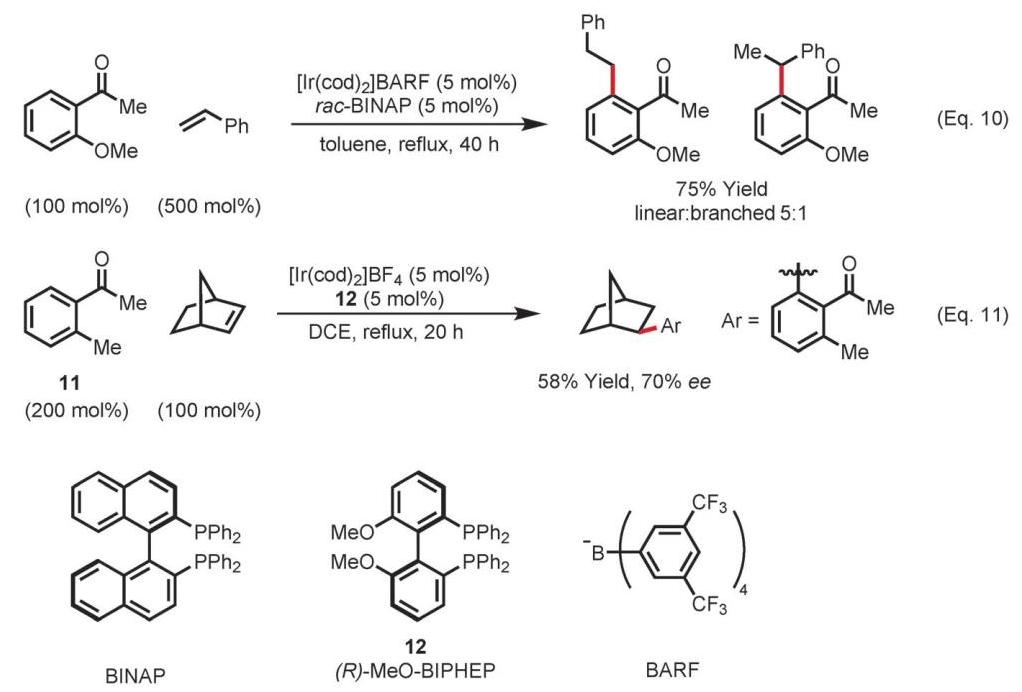
Scheme 11 Iridium-catalyzed ortho-alkylation of aromatic ketones with olefins.
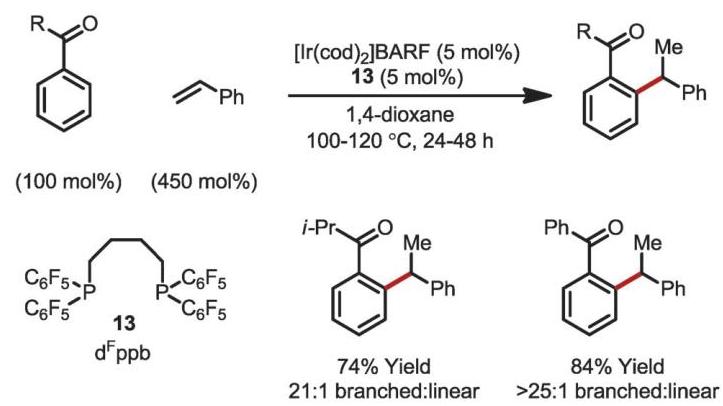
Scheme 12 Branched-selective ortho-alkylation of aromatic ketones with olefins.
The use of a cationic rhodium catalyst, in conjunction with a stoichiometric $\mathrm{{Cu}}\left( \mathrm{{II}}\right)$ oxidant, triggered a different reaction pathway from Murai’s original alkylation reaction (Scheme 14). The activation of the ortho-C-H bond was proposed to proceed through a concerted metalation-deprotonation (CMD) pathway with the $\mathrm{{Rh}}\left( \mathrm{{III}}\right)$ catalyst. Unlike the oxidative addition pathway observed with $\mathrm{{Ru}}\left( 0\right)$ , the CMD mechanism yielded a $\mathrm{{Rh}}\left( \mathrm{{III}}\right)$ -aryl complex and an acid instead of a metal-hydride species. Subsequent migratory insertion of the olefin into the Rh(III)- aryl bond and $\beta$ -hydride elimination afforded the olefination product. The $\mathrm{{Cu}}\left( \mathrm{{II}}\right)$ oxidant was proposed to be responsible for regenerating the active $\mathrm{{Rh}}\left( \mathrm{{III}}\right)$ catalyst. An analogous Ru(III)-catalyzed ortho-alkenylation with olefins was reported by Jeganmohan and coworkers. ${}^{35}$
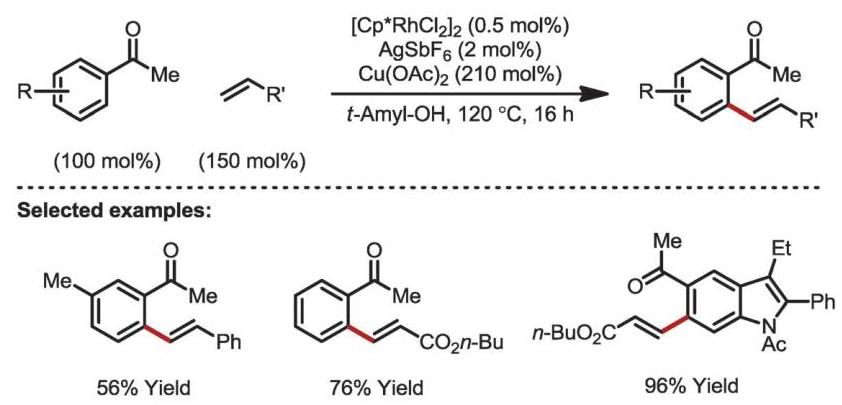
Scheme 13 Rhodium-catalyzed ortho-olefination of aromatic ketones with olefins.
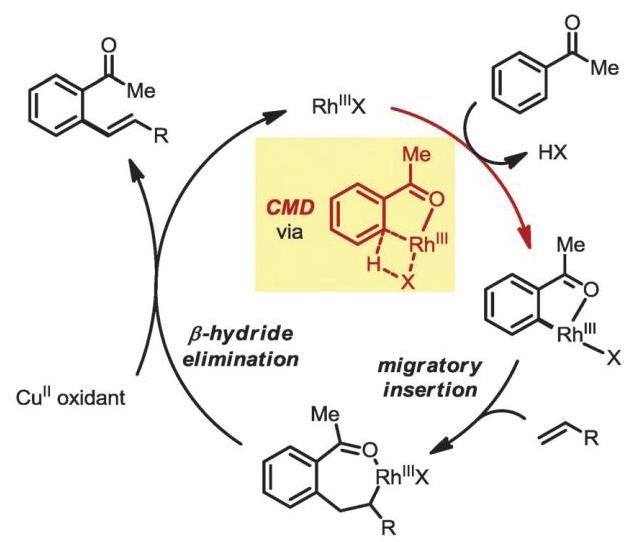
Scheme 14 Catalytic cycle of rhodium-catalyzed ortho-olefination.
When a cationic rhodium catalyst $\left\lbrack {{\mathrm{{Cp}}}^{ * }\mathrm{{Rh}}{\left( {\mathrm{{CH}}}_{3}\mathrm{{CN}}\right) }_{3}}\right\rbrack {\left( {\mathrm{{SbF}}}_{6}\right) }_{2}$ was used with stoichiometric silver acetate, Li and coworkers discovered aromatic ketones and enones underwent a C-H alkylation/aldol condensation sequence instead of an oxidative olefination reaction (Scheme 15). ${}^{36}$ The cascade reaction was proposed to proceed through 14, a similar intermediate as in Glorius’s olefination reaction. However, instead of $\beta$ -hydride elimination, intermediate 14 would be protonated under the reaction conditions to give 15, followed by silver-mediated aldol condensation to give an indene as the product. The possibility of direct aldol condensation with intermediate 14 cannot be excluded. A small amount of water was demonstrated to be beneficial to the cascade reaction, probably by facilitating the protonation step.
Although less studied than coupling with olefins, ketone-directed addition of the ortho-C-H bonds to alkynes also proves to be efficient. Murai and coworkers demonstrated that both aromatic ${}^{37}$ and olefinic ${}^{38}\mathrm{C} - \mathrm{H}$ bonds can react with alkynes to give the alkenylation products using the same ruthenium catalyst (Scheme 16). While internal alkynes gave good yields, terminal alkynes failed to provide access to the desired products, presumably due to competing side reactions, e.g. dimerization. When unsymmetrical ethylphenylacetylene was submitted to the reaction, all four possible stereo- and regioisomers (16) were isolated. However, reactions with trimethylsilyl-substituted acetylenes all gave 1-trimethylsilyl-2-aryl alkenes as the exclusive regioisomers. Besides ruthenium catalysis, Shibata and coworkers also reported the same transformation with a cationic iridium catalyst, although only diphenylacetylene and trimethylsilyl-phenylacetylene were employed. ${}^{30}$
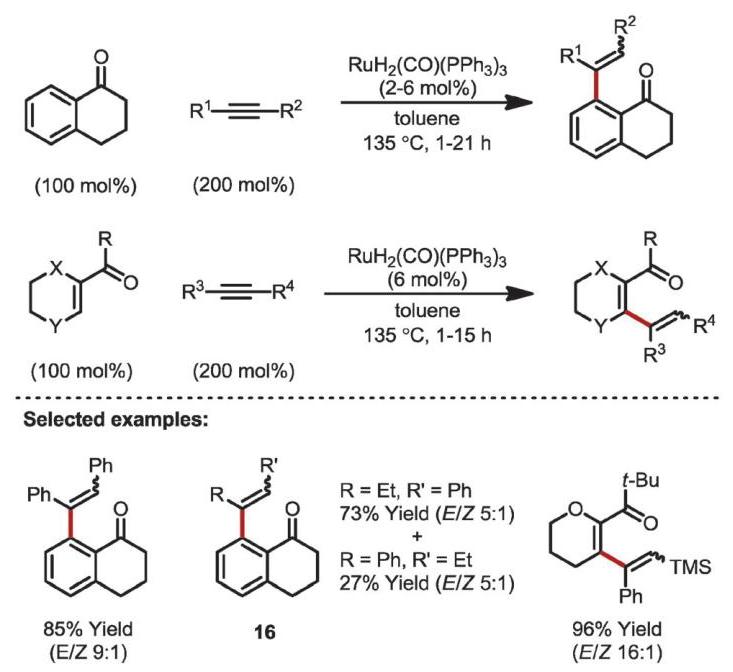
Scheme 16 Ruthenium-catalyzed ortho-alkenylation of aromatic ketones or enones with alkynes.
In 2011, Glorius and coworkers reported that the coupling between aromatic ketones and alkynes can undergo a different pathway, leading to indenol and fulvene derivatives when a cationic Rh(III) catalyst and stoichiometric Cu(II) salt are used (Scheme 17). ${}^{39}$ Similar to the coupling with olefins (vide supra, Scheme 14), ortho-C-H activation with Rh(111) proceeds through a CMD mechanism and the following migratory insertion of alkyne gives vinyl-Rh(III) species 20. Subsequently, an intramolecular addition of vinyl group to the ketone carbonyl yields the intermediate 21, which upon protonation gives the indenol (17) as the final product. However, if a methyl or methylene group is present at the $\alpha$ -position of the ketone $\left( {\mathrm{R}}^{1}\right)$ or the $\gamma$ -position of the alkyne $\left( {\mathrm{R}}^{3}\right)$ , dehydration takes place affording the fulvene derivatives (18 and 19, respectively). Similar transformations have also been reported by Cheng, ${}^{40}$ Jeganmohan, ${}^{41}$ and Shibata ${}^{42}$ using rhodium, ruthenium, and iridium catalysts respectively.
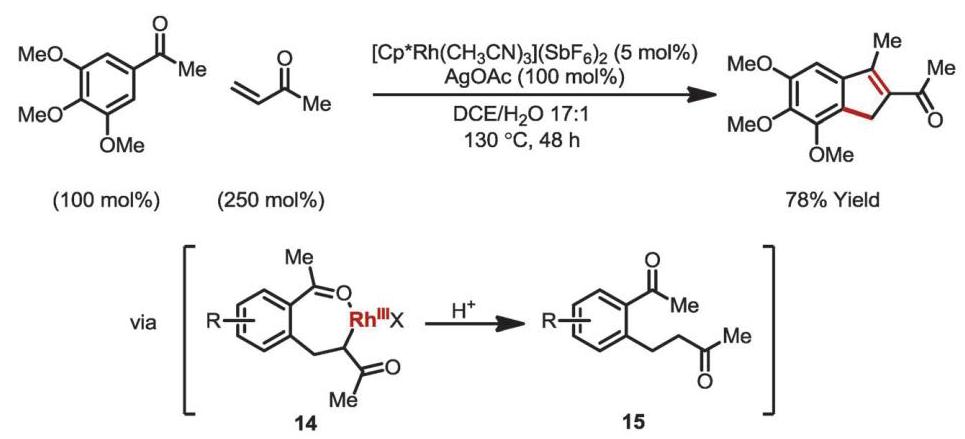
Scheme 15 Rhodium-catalyzed ortho-alkylation/aldol condensation cascade.
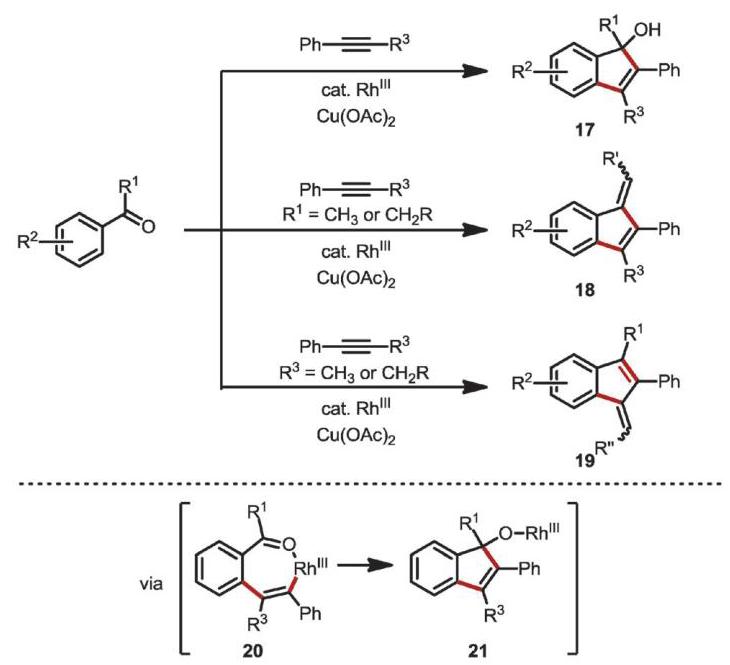
Scheme 17 Rhodium-catalyzed fulvene and indenol synthesis.
Another interesting tandem reaction that involves ketone-directed C-H bond activation is the rhodium-catalyzed cyclization of aromatic ketones and diynes (Scheme 18). In 2007, Shibata and coworkers reported the formation of cyclic 1,3-diene product 23 from the coupling of diynes and aromatic ketones catalyzed by $\left\lbrack {\mathrm{{Rh}}\left( \mathrm{{BIPHEP}}\right) }\right\rbrack {\mathrm{{BF}}}_{4}$ (eqn (12)). ${}^{43}$ When trans-chalcone was used as the substrate, monocyclic trienone 24 was generated (eqn (13)). The tandem process was proposed to initiate via the ketone-directed ortho-C-H activation with the rhodium catalyst. Intermolecular hydro-rhodation of one of the alkyne moieties and subsequent intramolecular carbo-rhodation of the other would give the six-membered rhodacycle 26. The final reductive elimination generates the product and releases the active catalyst.
When an enyne was employed as the substrate, the metal-lated intermediate added to the alkyne first to give the monocyclic product 27 (Scheme 19). The product descended from the initial addition to alkene and subsequent carbo-rhodation of alkyne (28) was not observed. When a chiral rhodium complex was used as the catalyst, good enantioselectivity was obtained. Similar tandem reactions with enynes and diynes as the coupling partners have also been published by Tanaka and coworkers using cationic rhodium catalysts. ${}^{{44},{45}}$
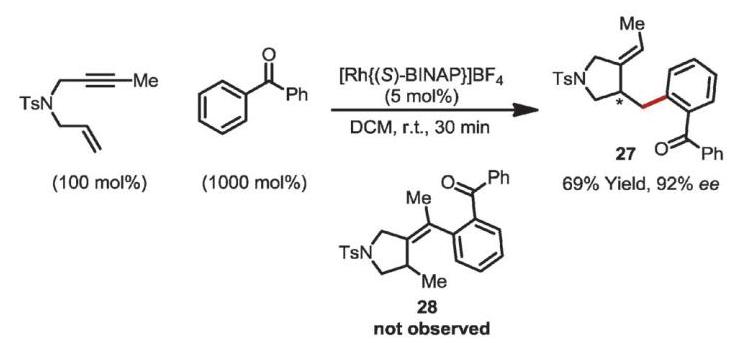
Scheme 19 Rhodium-catalyzed cyclization of enynes and aromatic ketones.
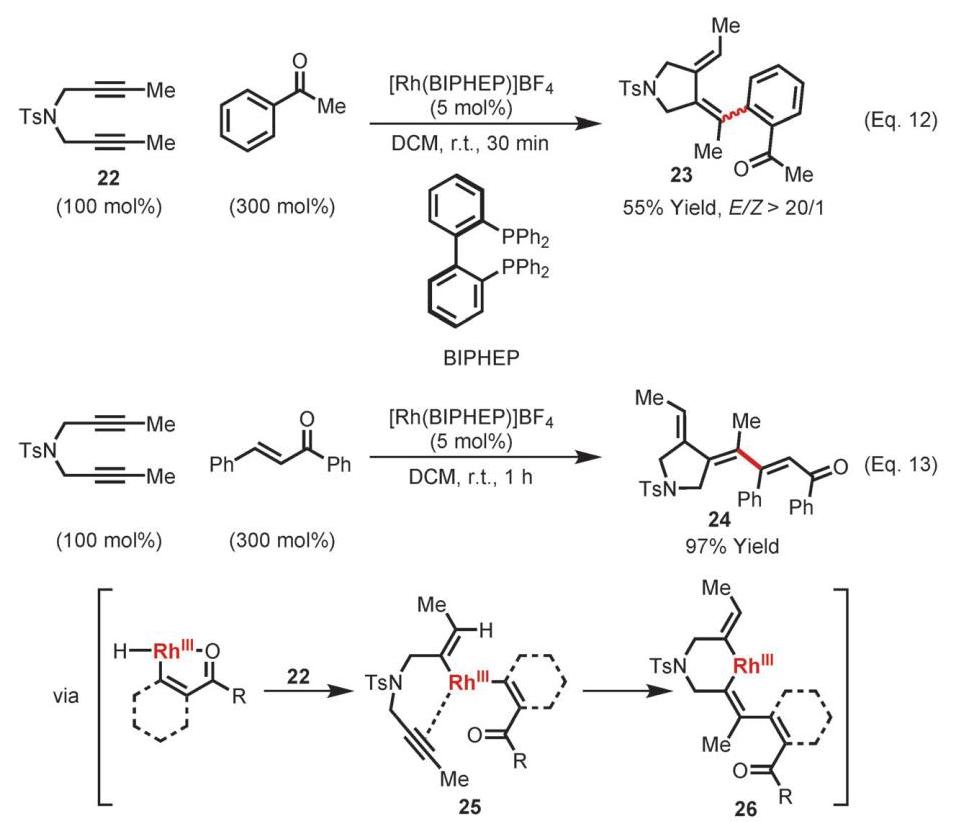
Scheme 18 Rhodium-catalyzed cyclization of diynes and aromatic ketones or chalcones.
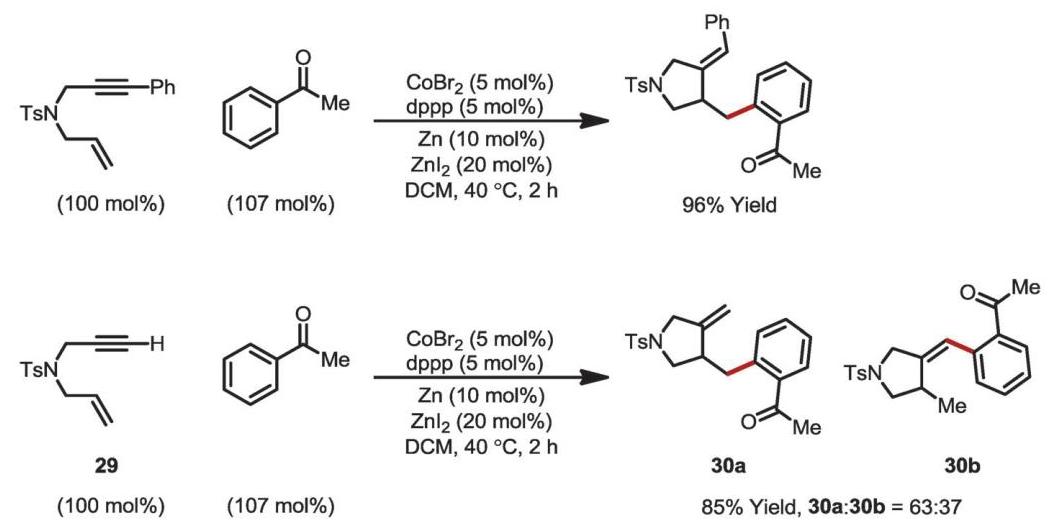
Scheme 20 Cobalt-catalyzed cyclization of enynes and aromatic ketones.
Recently, Cheng and coworkers reported a similar hydroar-ylative cyclization of enynes and aromatic ketones with a cobalt catalyst (Scheme 20). ${}^{46}$ In their case, ${\mathrm{{CoBr}}}_{2}/1,3$ -bis(diphenyl-phosphino)propane (dppp) was used as the precatalyst and the combination of zinc and zinc iodide was proposed to reduce the cobalt salt to the active $\mathrm{{Co}}\left( \mathrm{I}\right)$ catalyst. A large collection of functionalized pyrrolidines, dihydrofurans, and cyclopentanes were accessed as single regioisomers in good yields from simple aromatic ketones and enynes. While examples with terminal alkynes were not shown with rhodium catalysts (vide supra, Scheme 19), enyne 29 proceeded to give the cyclization product 30 as a mixture of regioisomers under the cobalt catalysis.
Although this review focuses on reaction discovery and strategy design, ketone-directed C-H bond addition to alkenes and alkynes has found many synthetic applications. For example, ketone-directed hydroarylation reactions were employed by Woodgate and coworkers in the synthesis of various aromatic diterpenoids (Scheme 21). In their study, the C14 C-H bond of diterpenoid 31 was selectively alkylated in quantitative yield with triethoxyvinylsilane using ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ as the catalyst. ${}^{47}$ The authors also demonstrated the resulting alkyl-alkoxysilane moiety could serve as a handle to access a diverse range of analogues. ${}^{48}$
Due to its high efficiency, the ketone-directed hydroarylation reaction has also found applications in the field of polymer chemistry. Shortly after Murai’s original discovery, Weber and coworkers applied the ruthenium-catalyzed ortho-alkylation of ketones to the copolymerization of aromatic ketones and diene derivatives (Scheme 22A). Copolymers with various backbones and functional groups, such as amines and acetals, were synthesized in high efficiency using ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ as the catalyst. ${}^{{49} - {53}}$ The Weber group also reported the modification of poly-(vinyl-methylsiloxane) (PVMS) using the ruthenium-catalyzed hydroaryl-ation reaction (Scheme 22B). The ortho-C-H bond of the ketone in 9-acetylphenanthrene added to the pendant vinyl groups in PVMS with anti-Markovnikov selectivity and afforded the modified polymer in an excellent yield. ${}^{54}$
2.2 Arylation
In 2003, Kakiuchi and coworkers successfully extended ruthenium-catalyzed ortho-functionalization to the arylation of aromatic ketones (Scheme 23). ${}^{55}$ With ${\mathrm{{RuH}}}_{2}\left( \mathrm{{CO}}\right) {\left( {\mathrm{{PPh}}}_{3}\right) }_{3}$ as the catalyst, a wide panel of aromatic ketones were arylated in good yields with arylboronates as the arene source. To ensure good yields, two equivalents of ketone substrate were required.
Murai’s original ruthenium-catalyzed addition of C-H bonds across olefins is a redox-neutral process requiring no external oxidant or reductant. However, in the case of arylation with aryl-boronates, additional reductants are necessary to trap the hydride generated after the oxidative addition. This is supported by the authors’ observation that an equal amount of benzyl alcohol was isolated as a byproduct after workup (eqn (14)). ${}^{56}$ In the proposed catalytic cycle, addition of Ru(π)-hydride 32 to another ketone substrate generates Ru(π)-alkoxide 33 (Scheme 24). Subsequent transmetalation between 33 and arylboronate transfers the aryl group to the $\mathrm{{Ru}}\left( \mathrm{{II}}\right)$ and meanwhile generates a stoichiometric amount of boronic ester, which is hydrolyzed during the workup to give the benzyl alcohol. Reductive elimination of complex 34 would give the arylation product and restore the active $\mathrm{{Ru}}\left( 0\right)$ catalyst.
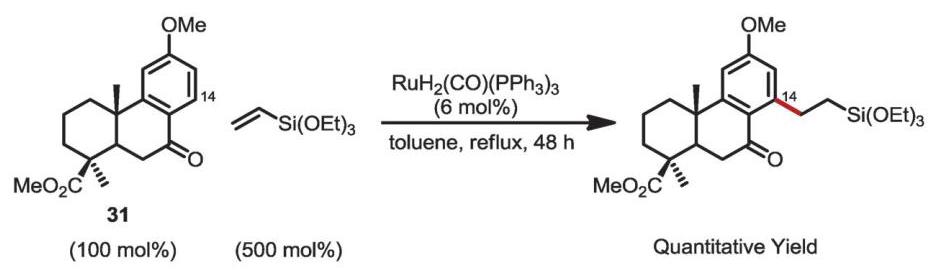
Scheme 21 Ruthenium-catalyzed alkylation of aromatic diterpenoid.
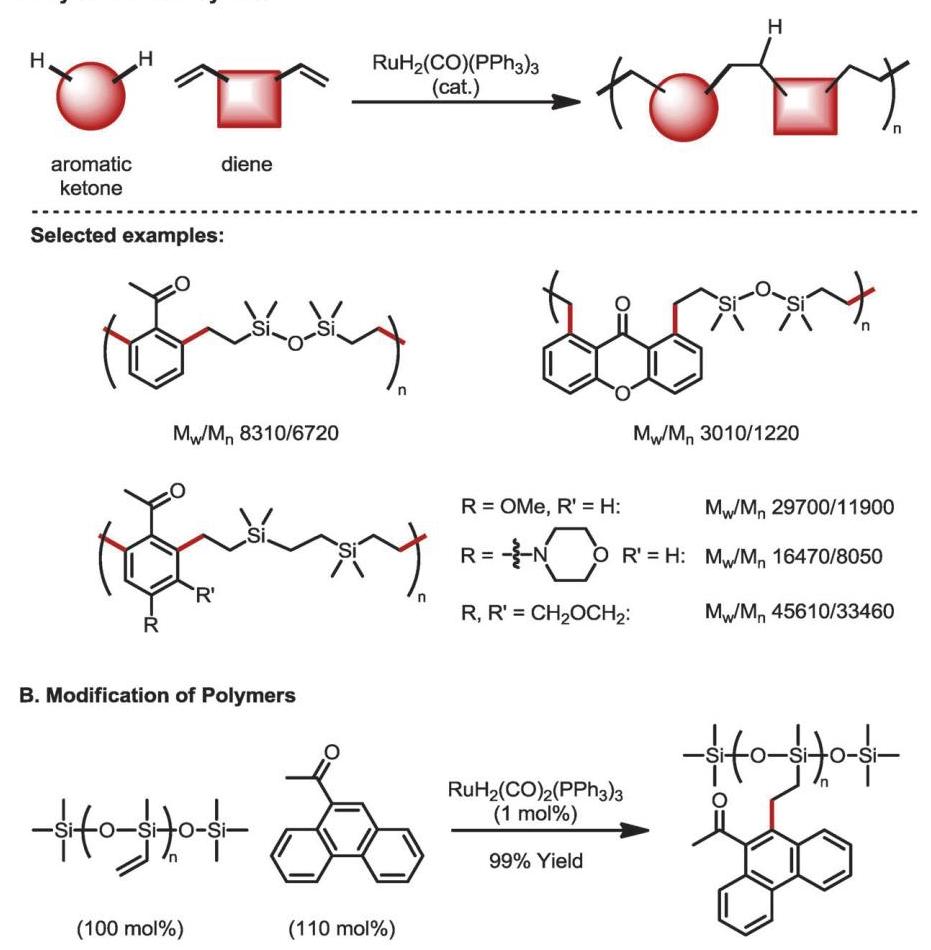
Scheme 22 Ruthenium-catalyzed synthesis and modification of polymers.
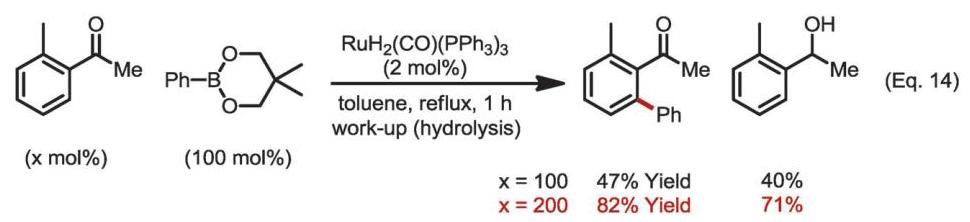
Scheme 23 Ruthenium-catalyzed ortho-arylation of aromatic ketones with arylboronates.
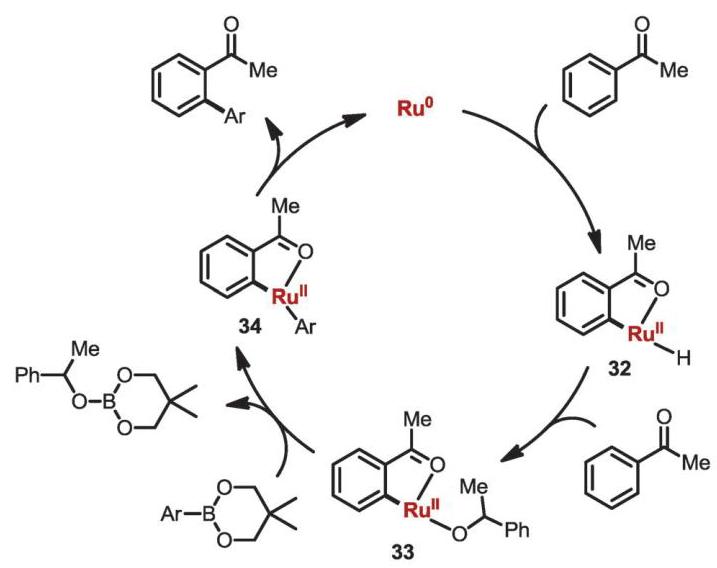
Scheme 24 Proposed catalytic cycle of ruthenium-catalyzed ortho-arylation.
The issue of using excess ketone substrates was later resolved by employing pinacolone as the solvent (Scheme 25, eqn (15)). The efficiency of the arylation reaction can be sustained with aromatic ketones as the limiting reagent, and excess pinacolone serves as the hydride scavenger instead of the aryl ketone substrate. The coupling with alkenylboronates was also reported under similar reaction conditions by the same group (eqn (16)). ${}^{{57},{58}}$ This new olefination reaction can be considered as a complementary approach to the previous C-H addition to alkynes (vide supra, Scheme 16), since products like 35 cannot be accessed directly from coupling with alkynes.
On the other hand, the ortho-arylation of aromatic ketones using aryl halides as the coupling partner was achieved using palladium catalysis. The ortho-arylation of aromatic ketones with aryl bromides was first discovered by Miura and coworkers (Scheme 26). ${}^{59}$ In the presence of a palladium catalyst and a stoichiometric base, benzylphenyl ketone reacted with excess bromobenzene to give poly-arylated ketone 36 as the major product (eqn (17)). Instead of a ketone-directed pathway, the authors proposed an enolate-directed palladation assisted by ${\mathrm{{Cs}}}_{2}{\mathrm{{CO}}}_{3}$ . In a following study by the same group, the ortho- $\mathrm{C} - \mathrm{H}$ bonds of anthrone were also arylated under similar reaction conditions (eqn (18)). ${}^{60}$
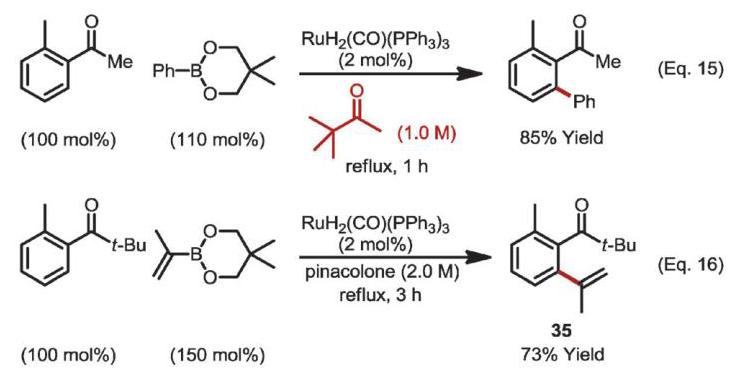
Scheme 25 Ruthenium-catalyzed ortho-arylation using pinacolone as solvent.
In 2010, Cheng and coworkers reported a ketone-directed ortho-arylation with aryl iodides (Scheme 27). ${}^{61}$ Contrary to Miura’s basic conditions, trifluoroacetic acid was used as the solvent for this reaction. In addition, although excess aryl iodides were employed, only mono arylation products were observed for a range of aromatic ketones. However, when electron-neutral or rich aryl iodides were used as the substrates, the yields were largely compromised. It is interesting to note that the use of sec-alkyl aryl ketones triggered a tandem process that gave phenanthrone products (Scheme 28). It was proposed that after the ortho-arylation, a second ketone (or enolate)- directed C-H activation occurs to form a seven-membered palladacycle (41). Subsequent enolate formation, rearrangement, and reductive elimination results in formation of phenan-throne 39.
Besides coupling with pre-functionalized aryl sources (i.e. arylboronates and aryl halides), ketone-directed arylation can also occur through the coupling between two aryl $\mathrm{C} - \mathrm{H}$ bonds. In 2012, the Cheng ${}^{62}$ and Shi ${}^{63}$ groups independently reported the synthesis of fluorenone derivatives from diaryl ketones using palladium acetate as the catalyst and silver oxide as the oxidant (Scheme 29). The proposed pathway begins with ortho-C-H palladation via a CMD mechanism directed by the ketone carbonyl. Due to the weak coordinating ability of the ketone, de-chelation followed by C-C bond rotation can occur, allowing the Pd(II) center to rotate towards the ortho-C-H on the other arene. A second C-H palladation would lead to a six-membered pallacycle (46), which upon reductive elimination would give fluorenone 44 as the product and a $\operatorname{Pd}\left( 0\right)$ species. The silver salt was proposed to oxidize the $\operatorname{Pd}\left( 0\right)$ intermediate and restore the active Pd(π) catalyst.
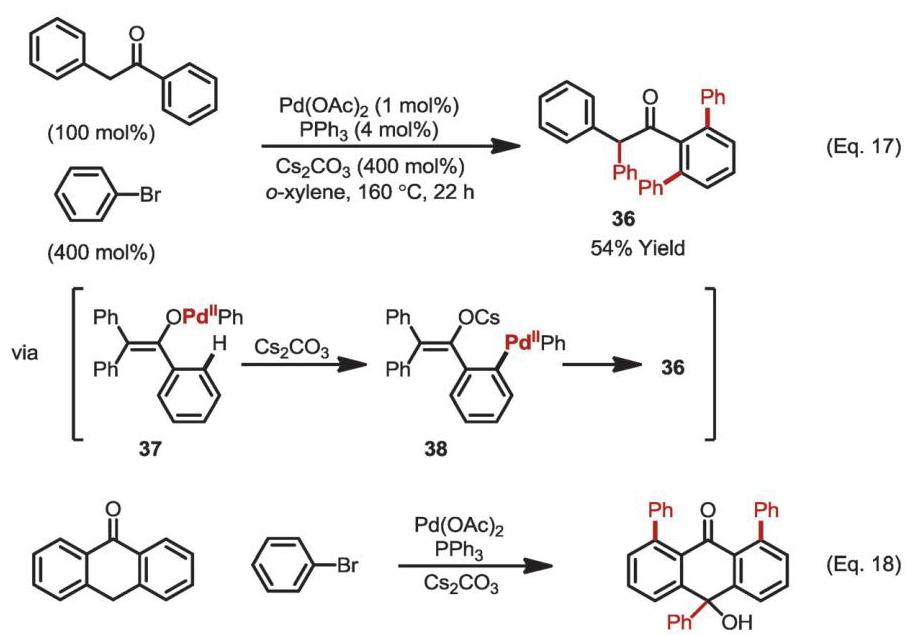
Scheme 26 Palladium-catalyzed multi-arylation of ketones.
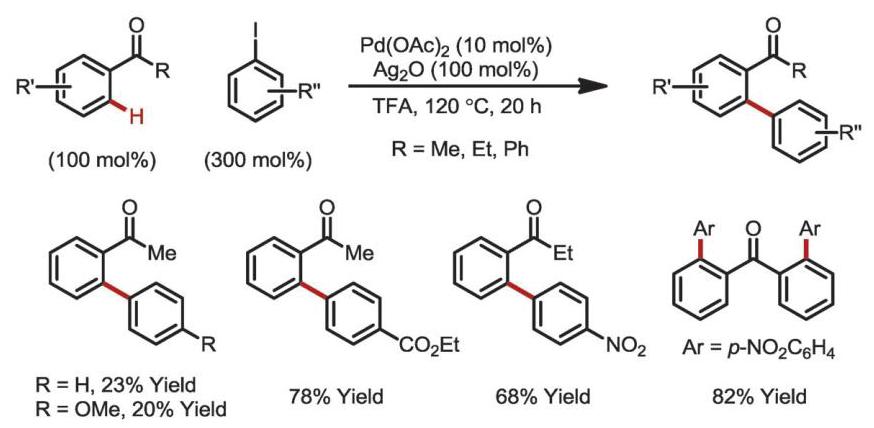
Scheme 27 Palladium-catalyzed ortho-arylation with aryl iodides.
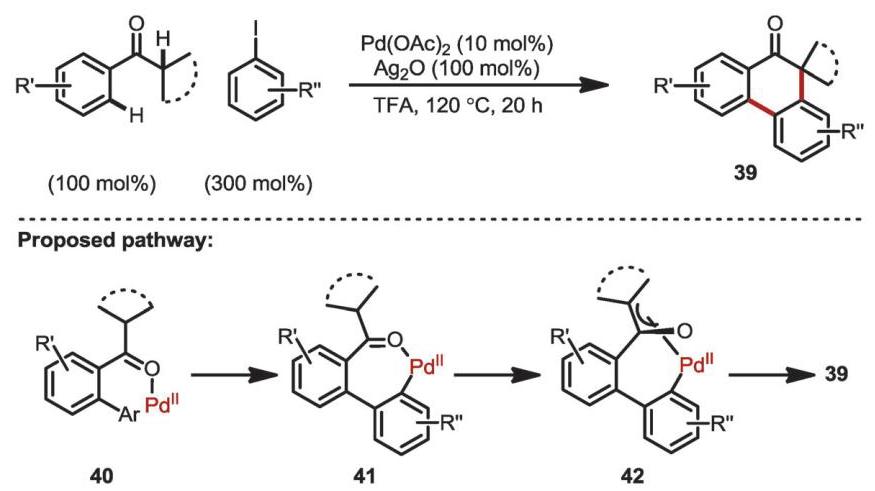
Scheme 28 Palladium-catalyzed ortho-arylation and tandem cyclization.
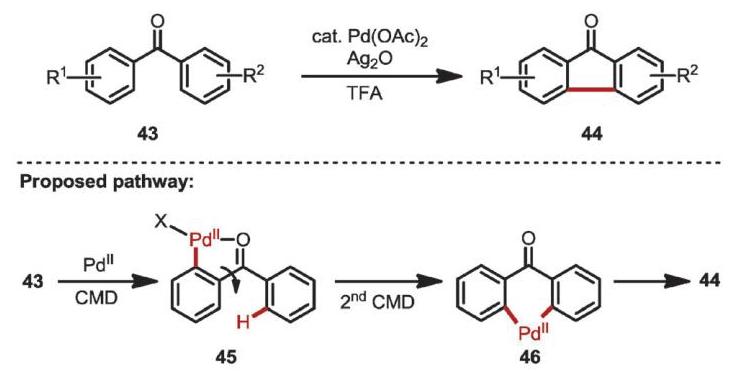
Scheme 29 Palladium-catalyzed fluorenone synthesis via sequential C-H activation.
2.3 Carbon-heteroatom (C-X) bond formation
Compared to C-C bond forming reactions, ketone-directed ortho-C-X (X: heteroatom) bond formation is relatively less studied. Nevertheless, direct and efficient assembly of ortho-heteroatom-substituted ketones via C-H functionalization is of great value to the synthetic community as $o$ -acyl phenols, anilines, and haloarenes are prevalent motifs in bioactive compounds.
The general catalytic cycle for ketone-directed C-X bond formation is illustrated in Scheme 30. The reaction starts with a directed ortho-C-H metalation to give a five-membered metalla-cycle (47), where the oxidation state of the metal remains unchanged. The following oxidation with various oxidants first delivers an anionic heteroatom ligand $\left( \mathrm{X}\right)$ to the metal, and second increases the oxidation state of the metal by two. Depending on the oxidant employed, different heteroatom ligands can be introduced to the metal. The final reductive elimination of the higher-oxidation state metal complex (48) affords the product with a newly formed C-X bond and regenerates the active metal catalyst.
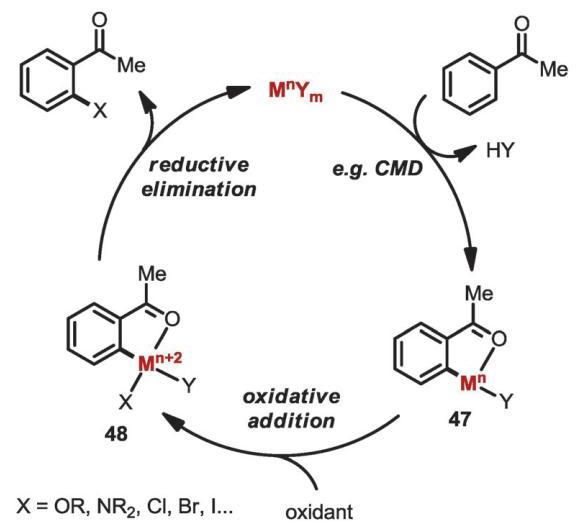
Scheme 30 General mechanism of the ketone-directed ortho-C-X bond formation.
In 2012, the Rao ${}^{64}$ and Dong ${}^{65}$ groups independently reported a palladium-catalyzed ortho-hydroxylation reaction directed by ketones (Scheme 31, eqn (19)). A wide range of ortho-acylphenols were synthesized using palladium acetate or trifluoroacetate as the catalyst in good yields. Both organic and inorganic oxidants, e.g. hypervalent iodine compounds and potassium persulfate, can be used to promote the C-O bond formation. Dong and coworkers further discovered an unusual ortho-carbonylation reaction when running the hydroxylation reaction under CO atmosphere (eqn (20)). ${}^{65}$ While the exact pathway remains unclear, the ethyl-ketal lactone was proposed to come from mixed-anhydride intermediate 50. A similar palladium-catalyzed hydroxylation reaction was later reported by Kwong and coworkers. ${}^{66}$ In addition to palladium catalysis, the ketone-directed hydroxylation reaction can also be catalyzed by ruthenium ${}^{{67},{68}}$ and rhodium ${}^{68}$ under similar conditions (Scheme 32). The study of substrate scopes indicated that, while ruthenium- and rhodium-catalyzed hydroxylation reactions shared similar chemoselectivity (electron-rich arenes are favored), the ruthenium catalysts gave better performance in terms of yields.
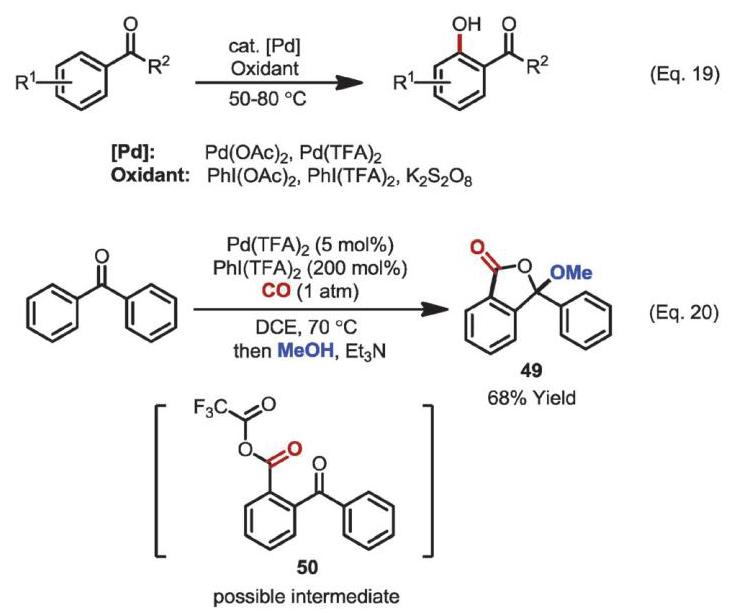
Scheme 31 Palladium-catalyzed ketone-directed ortho-hydroxylation reaction.
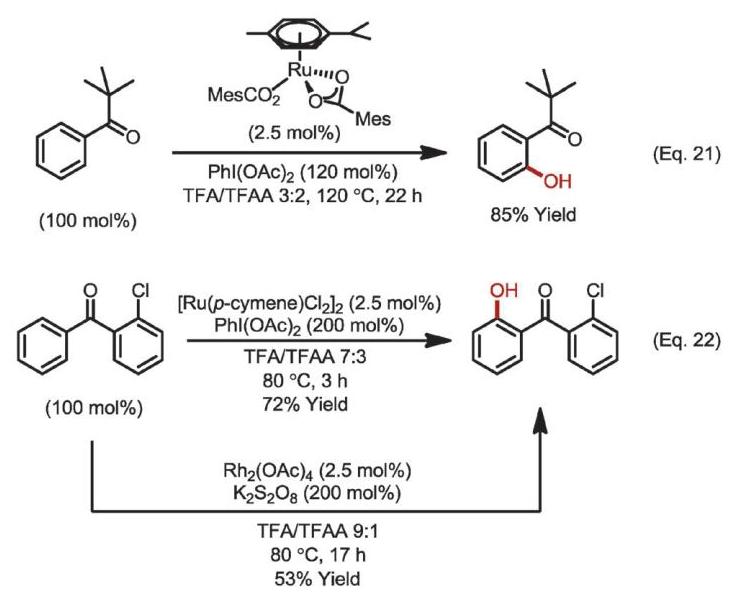
Scheme 32 Ruthenium and rhodium-catalyzed ketone-directed ortho-hydroxylation reaction.
Halogenation is also possible, and $N$ -halo succinimides proved to be efficient oxidants for carbon-halogen bond formation directed by ketones. In 2012, Glorius and coworkers reported rhodium-catalyzed ortho-bromination and iodination reactions with aromatic ketones, using $N$ -bromo and $N$ -iodo-succinimide as the oxidant respectively (Scheme 33). ${}^{69}$ The reaction conditions utilize a cationic rhodium catalyst and stoichiometric pivalic acid or copper acetate, presumably to facilitate the metalation step. The chlorination with $N$ -chlorosuccinimide was reported later by Rao and coworkers using palladium catalysis (Scheme 34). ${}^{70}$ Both triflic acid and potassium persulfate additives play a vital role in sustaining the reactivity. While triflic acid facilitates coordination of the ketone by in situ generation of a more electrophilic palladium catalyst $\mathrm{{Pd}}{\left( \mathrm{{OTf}}\right) }_{2}$ , potassium persulfate is believed to promote the oxidation of $\mathrm{{Pd}}\left( \mathrm{{II}}\right)$ to $\mathrm{{Pd}}\left( \mathrm{{IV}}\right)$ as a co-oxidant.
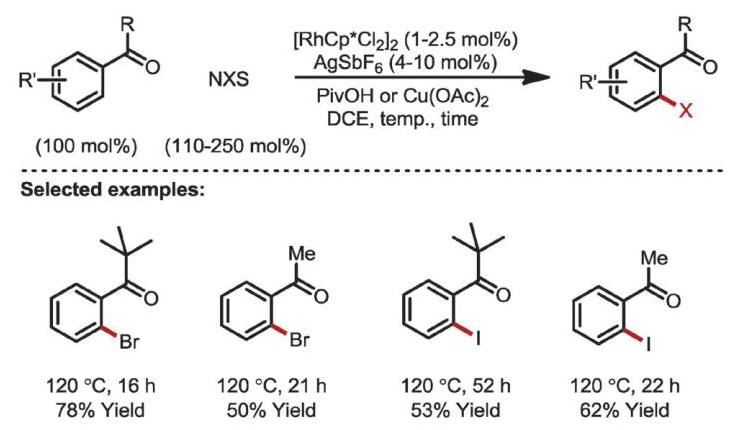
Scheme 33 Palladium-catalyzed ketone-directed ortho-bromination and iodination.
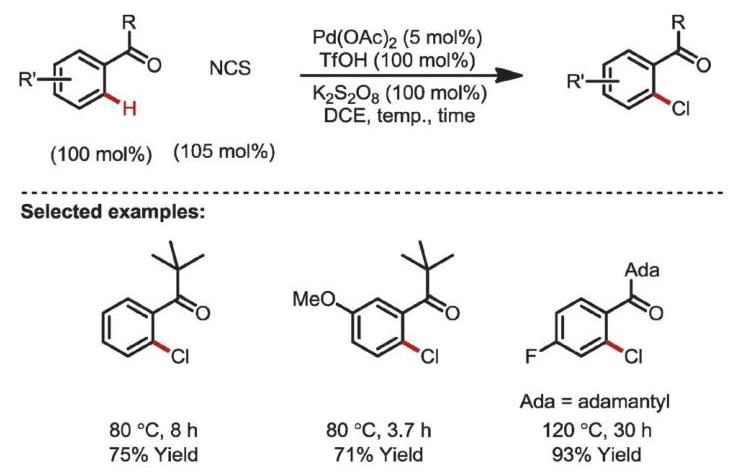
Scheme 34 Palladium-catalyzed ketone-directed ortho-chlorination.
Ketone-directed ortho-amination was first accomplished by Liu and coworkers using sulfonamides or amides as the amine source (Scheme 35). ${}^{71}N$ -Fluoro-2,4,6-trimethylpyridinium triflate 51 or sodium persulfate can be used as the terminal oxidant. Mechanistic studies revealed that during the reaction the sulfonamide competes with the weakly coordinating ketone for complexation with palladium, which results in lower efficiency during the reaction.
The corresponding ortho-amination with sulfonyl azides was independently reported by the Chang, ${}^{72}$ Jiao, ${}^{73}$ and Sahoo ${}^{74}$ groups in 2013 using ${\left\lbrack \mathrm{{Ru}}\left( p\text{ -cymene }\right) {\mathrm{{Cl}}}_{2}\right\rbrack }_{2}$ as the catalyst (Scheme 36). Besides the use of cationic silver salts, additional copper or sodium acetate is indispensable for all three reactions. The silver salt is proposed to generate a more electrophilic ruthenium catalyst allowing better complexation with ketone. The basic acetate ligands are expected to facilitate C-H metalation via a CMD mechanism.
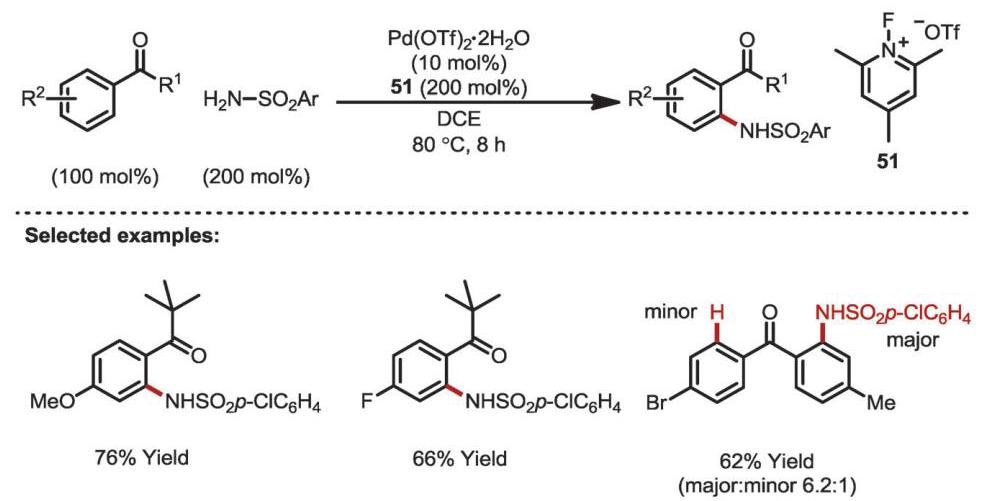
Scheme 35 Palladium-catalyzed ketone-directed ortho-amination using sulfonamides.
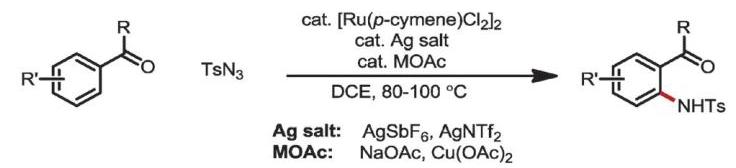
Scheme 36 Palladium-catalyzed ketone-directed ortho-amination using sulfonyl azides.
3. β-C-H functionalization of simple ketones
$\beta$ -Substituted ketones represent an important class of prevalent motifs in bioactive compounds, such as drug candidates, antioxidants, and pesticides. Nevertheless, compared to the $\alpha - \mathrm{C} - \mathrm{H}$ bond, the more distal $\beta - \left( {\mathrm{{sp}}}^{3}\right) \mathrm{C} - \mathrm{H}$ bonds of ketones are usually considered intrinsically unreactive. Conventionally, $\beta$ -functionalized ketones are prepared through the conjugate addition of nucleo-philes to $\alpha ,\beta$ -unsaturated ketones (Scheme 37). ${}^{75}$ As efficient as this protocol is, $\alpha ,\beta$ -unsaturated ketones are often synthesized from the corresponding saturated ketones via dehydrogenation, which takes 1-3 steps and requires stoichiometric oxidants. ${}^{76}$ The direct $\beta$ -C-H functionalization of the carboxylic acid derivatives, e.g. amides and esters, using a DG-based strategy have been extensively studied and reviewed: ${}^{77}$ in contrast, $\beta$ -functionalization of simple ketones is nontrivial to develop due to their higher reactivity but lower coordinating ability. In the past two years, several transition-metal-catalyzed approaches have emerged that enable the direct $\beta - \mathrm{C} - \mathrm{H}$ functionalization of simple ketones.
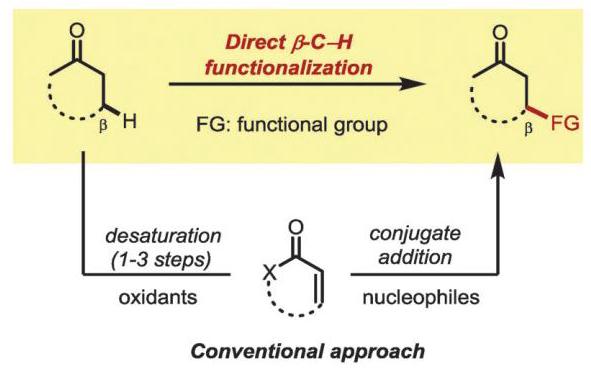
Scheme 37 Direct $\beta$ -C-H functionalization of ketones.
In 2013, MacMillan and coworkers reported a direct $\beta$ -arylation of cyclic ketones with electron-deficient arylnitriles via the combination of photo- and enamine catalysis (Scheme 38). ${}^{78}$ With a photocatalyst, i.e. $\operatorname{Ir}{\left( \mathrm{{ppy}}\right) }_{3}$ , and amine catalyst, i.e. azepane, various cyclic ketones were coupled with 1,4-dicyanobenzene selectively at the $\beta$ -position in the presence of a ${26}\mathrm{\;W}$ light bulb (CFL: compact fluorescent light) (eqn (23)). This transformation is amenable for enantioselective catalysis. When a cinchona-based chiral amine (53) was employed, a moderate ee value was obtained for the $\beta$ -arylation of cyclohexanone (eqn (24))
The proposed mechanism for the $\beta$ -arylation reaction consists of two catalytic cycles mediated by the photo- and amine catalyst, respectively (Scheme 39). In the photocatalytic cycle, the $\operatorname{Ir}{\left( \mathrm{{ppy}}\right) }_{3}$ catalyst is first activated by light to generate an excited *Ir(ppy) ${}_{3}$ . A subsequent single electron transfer (SET) between the electron-deficient 1,4-dicyanobenzene and the activated catalyst reduces the arene to a radical anion (54) and yields an Ir(rv) intermediate 55. In the second cycle, the amine catalyst condenses with the ketone substrate to give an electron-rich enamine intermediate (56). The two catalytic cycles then merge through a single electron transfer process, where the enamine gets oxidized to a nitrogen-centered radical cation (57) and the Ir complex (55) is reduced back to $\operatorname{Ir}{\left( \mathrm{{ppy}}\right) }_{3}$ . The formation of the radical cation increases the acidity of the allylic $\beta$ -C-H bond, which can be deprotonated by a base to give a ${5\pi }$ -electron system that has significant radical character at the $\beta$ -carbon. Coupling between electron-rich $\beta$ -radical 58 and electron-deficient radical anion 54 provides intermediate 59 bearing a new $\beta - \mathrm{C} - \mathrm{C}$ bond. The following re-aromatization and hydrolysis affords the $\beta$ -arylated ketone, releases a cyanide anion, and regenerates the amine catalyst. The use of electron-deficient arylnitriles as the coupling partner is important because the radical anion intermediate 54 generated from the arylnitrile is long living and reluctant to dimerize due to its electron deficiency, which is the key for high chemo and site-selectivity for the $\beta - \mathrm{C} - \mathrm{C}$ bond formation.
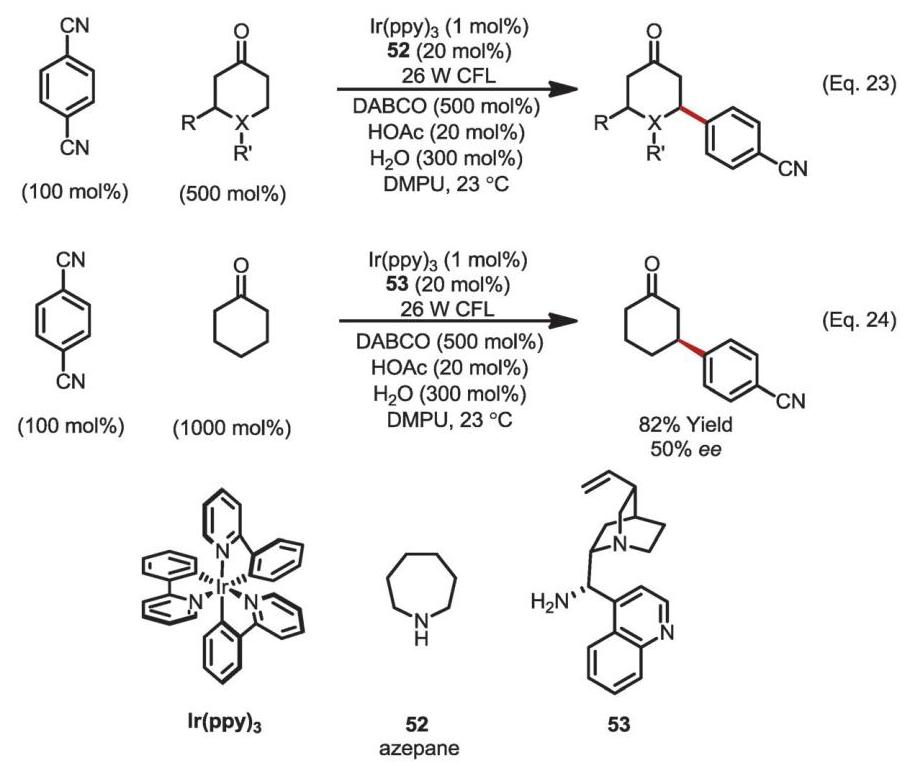
Scheme 38 Direct $\beta$ -arylation of ketones via photoredox organocatalysis.
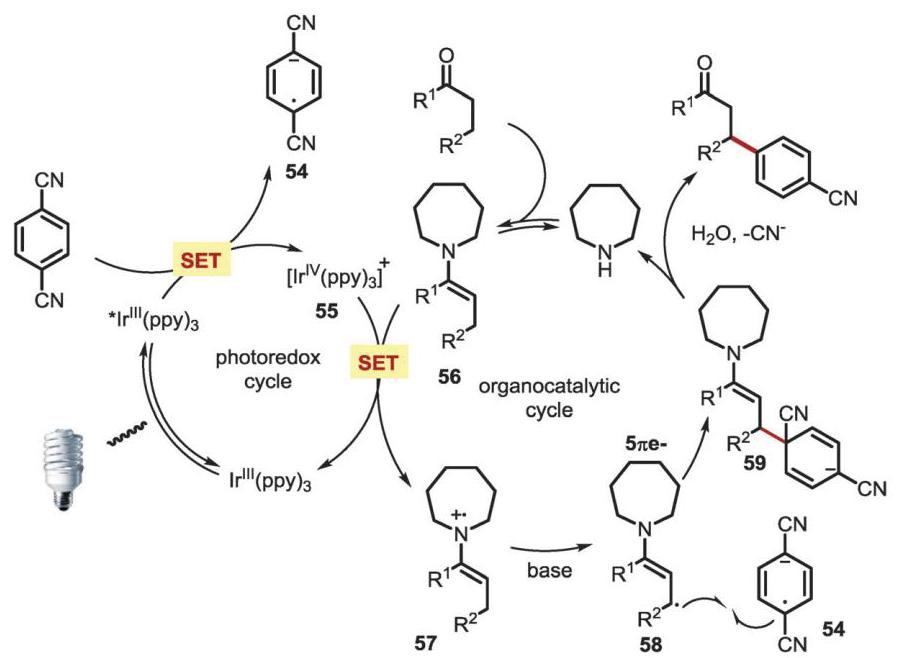
Scheme 39 Proposed catalytic cycle of direct $\beta$ -arylation of ketones.
Later, the same group extended this activation mode to other ketone $\beta - \mathrm{C} - \mathrm{H}$ functionalization reactions. With the use of the same photocatalyst [Ir(ppy) ${}_{3}$ ] and amine catalyst (azepane), cyclic ketones can be coupled with diarylketones to give $\gamma$ -hydroxyketone adducts (Scheme 40, eqn (25)). ${}^{79}$ This transformation can be considered as a formal $\beta$ -aldol reaction. A similar mechanism as the $\beta$ -arylation reaction was proposed by the authors, where the major difference is that a ketyl radical is used as the coupling partner instead of the aryl radical anion. A stoichiometric amount of ${\mathrm{{LiAsF}}}_{6}$ salt was found important to sustain the efficiency of the coupling, presumably due to its capability of prohibiting the dimerization of ketyl radicals. When an alkyl aryl ketone was employed as the substrate, the more reducing photocatalyst Ir(p-OMe-ppy) ${}_{3}$ , a higher loading of the amine catalyst, and an additional light bulb are necessary to promote the transformation (eqn (26)). This can be attributed to a higher reduction potential for the alkyl aryl ketones compared to the diaryl ketones.
Another interesting ketone $\beta$ -functionalization was reported by Fagnoni and coworkers in 2014 using a different transition-metal-catalyzed photoredox approach (Scheme 41). ${}^{80}$ Using tetrabutylammonium decatungstate (TBADT) as the catalyst and a Xe lamp or sunlight as the light source, various cyclo-pentanones can be coupled with electron-deficient alkenes to give $\beta$ -alkylated products (eqn (27)). The light source was proposed to excite the TBADT catalyst to a state containing electronegative oxygen-centered radicals. The activated catalyst can directly abstract a $\beta$ -hydrogen of the cyclopentanone to generate a carbon-centered radical. Subsequent 1,4-addition to a Michael acceptor, followed by hydrogen transfer back from the catalyst, results in the $\beta$ -alkylation product.
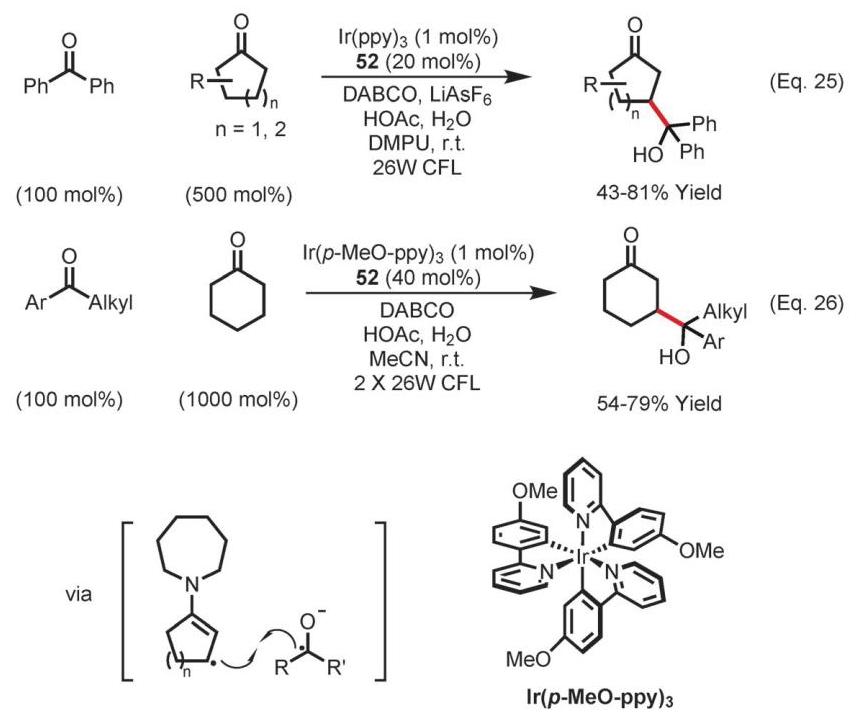
Scheme ${40\beta }$ -Aldol coupling of cyclic ketones with aryl ketones via photoredox catalysis.
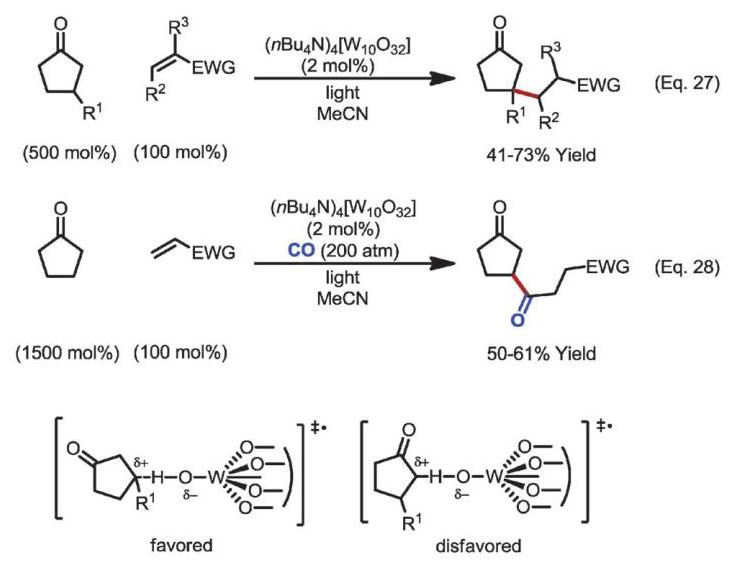
Scheme 41 Photocatalyzed β-alkylation and acylation of cyclopentanones.
The $\beta$ -selectivity of this transformation is determined during the hydrogen abstraction step. When the electronegative oxygen-centered radical approaches a C-H bond, a partial positive charge will build on the carbon atom in the transition state. Because the partial positive charge is destabilized by the neighboring carbonyl group, the abstraction of the $\alpha$ -hydrogen is disfavored, resulting in a high $\beta$ -selectivity. In addition to the alkylation, a $\beta$ -acylation of cyclopentanones was also achieved when the reaction was run under high pressure of carbon monoxide (eqn (28)). Despite the high $\beta$ -selectivity for cyclopentanone, other cyclic ketones, e.g. cyclohexanone, gave a mixture of $\beta$ - and $\gamma$ -C-H functionalization products.
In 2013, the Dong group reported a palladium-catalyzed direct ketone $\beta$ -arylation with aryl halides (Scheme 42). ${}^{81}$ Under the reaction conditions, $\mathrm{{Pd}}{\left( \mathrm{{TFA}}\right) }_{2}/\mathrm{P}{\left( \mathrm{i} - \mathrm{{Pr}}\right) }_{3}$ is used as the pre-catalyst and silver trifluoroacetate is employed as an additive. While the catalytic system is formally similar to those used for $\alpha$ -arylation, the reaction gives complete $\beta$ -position selectivity, which is likely due to the mildly acidic conditions. Aryl iodides with various electronic properties (electron-rich and poor) can participate, and both linear and cyclic ketones with different ring sizes are suitable substrates (eqn (29)). In addition, base- or nucleophile-sensitive functional groups, such as aldehydes, Weinreb amides, and methyl ketones are tolerated. Aryl bromides also react but give a decreased yield (eqn (30)).
While the mechanism remains unclear, this transformation aims to combine palladium-catalyzed ketone dehydrogenation, ${}^{82}\mathrm{C} - \mathrm{X}$ bond activation, and conjugate addition into one reaction vessel. The proposed catalytic cycle is depicted in Scheme 43. It begins with ketone dehydrogenation by a Pd(π) catalyst (60) to give an $\alpha ,\beta$ -unsaturated ketone $/\mathrm{{Pd}}\left( 0\right)$ intermediate (63). Subsequent oxidative addition with an aryl iodide affords an Pd(π)-aryl species (64), which can undergo migratory insertion and protonation of the resulting Pd(π)-enolate 65 to deliver the $\beta$ -arylation product and restore the active Pd(π) catalyst. Alternatively, a two-catalytic-cycle mechanism, involving ketone dehydrogenation and reductive Heck coupling is also possible.
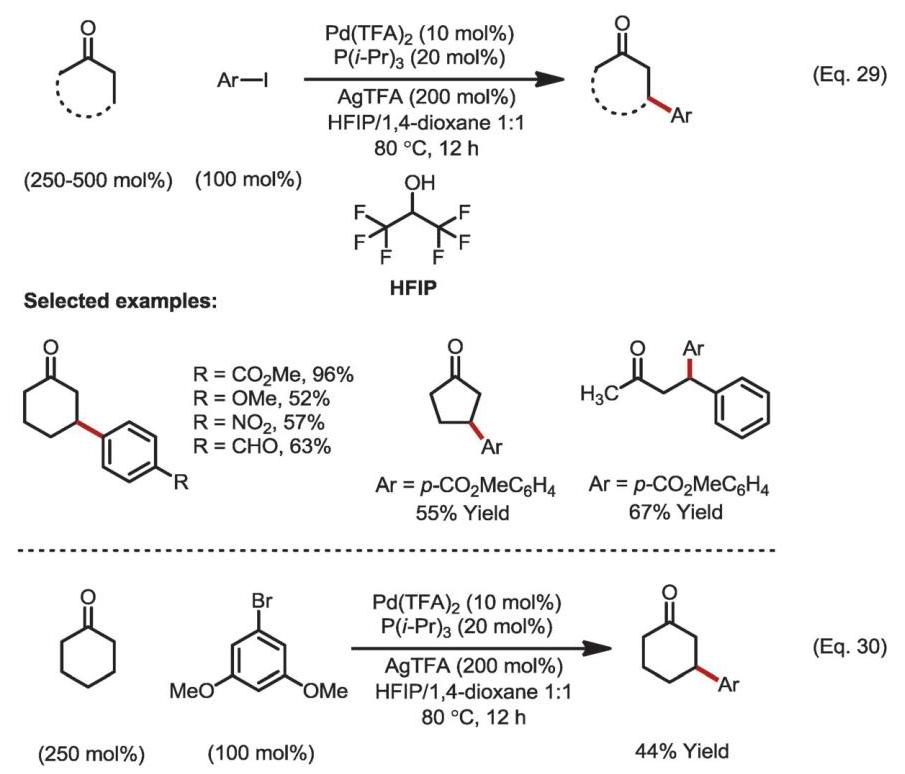
Scheme 42 Palladium-catalyzed direct $\beta$ -arylation of ketones with aryl halides.
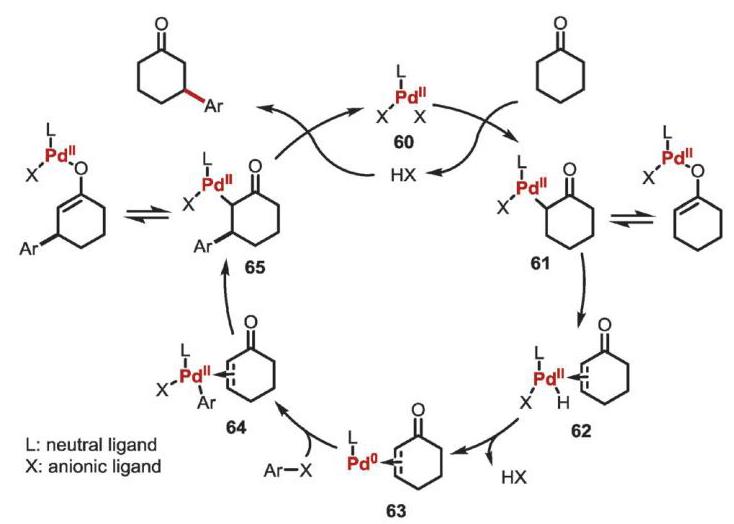
Scheme 43 Proposed strategy of direct $\beta$ -arylation of ketones with aryl halides.
4. Addition of $\alpha - \mathrm{C} - \mathrm{H}$ bonds to unactivated alkenes or alkynes
4.1 Addition to alkenes
Direct addition of the $\alpha - \mathrm{C} - \mathrm{H}$ bond of ketones across unactivated alkenes represents a transformation of significant synthetic value. Traditionally, α-alkylation of ketones is realized through enolate formation and subsequent substitution with an alkylating reagent, e.g. alkyl halides in most cases (Scheme 44A). Efficient as this reaction is, it suffers from several drawbacks, including highly basic conditions, generation of stoichiometric waste, lack of regioselectivity, high cost of the alkylation reagents and over-alkylation. ${}^{83}$ The Stork enamine reaction can also be used to prepare $\alpha$ -alkylated ketones, ${}^{84}$ although the use of highly reactive alkylating reagents (‘hot electrophiles’), as well as the formation of stoichiometric amine salts are inevitable (Scheme 44B).
A. Enolate Alkylation
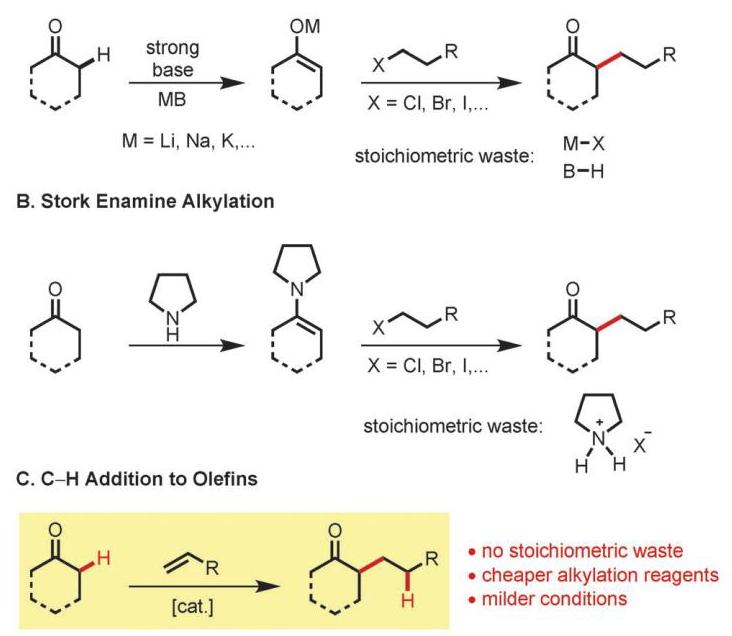
Scheme 44 α-Alkylation of ketones.
Compared to conventional alkylation reagents, such as alkyl halides and Michael acceptors, simple olefins are more accessible and generally less expensive. Moreover, the addition of the $\alpha$ -C-H bond of ketones to olefins ostensibly generates no stoichiometric waste, thus maximizing the atom economy of the alkylation reaction (Scheme 44C). Although the addition of activated methylene compounds (e.g. $\alpha - \mathrm{C} - \mathrm{H}$ bond of 1,3-dicarbonyl compounds) to olefins has been well studied, ${}^{85}$ examples of using less-acidic simple ketones as substrate are limited.
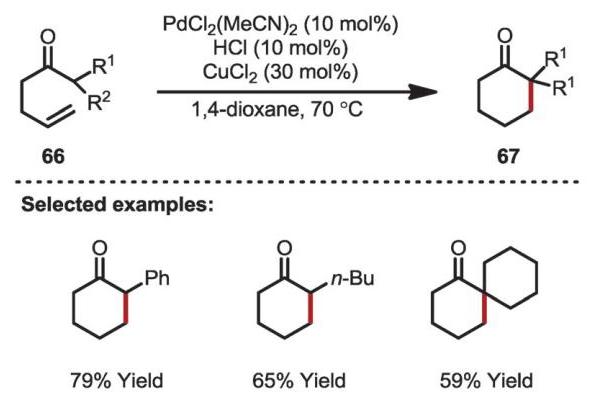
Scheme 45 Palladium-catalyzed intramolecular $\alpha$ -alkylation of ketones with olefins.
In 2003, Widenhoefer and coworkers reported an intramolecular cyclization with a $\gamma ,\sigma$ -enone using palladium catalysis (Scheme 45). ${}^{86}$ A catalytic amount of hydrochloric acid was employed to promote enol formation, and ${\mathrm{{CuCl}}}_{2}$ was used to stabilize the $\mathrm{{Pd}}\left( \mathrm{{II}}\right)$ catalyst. A number of $\gamma ,\delta$ -enones (66) cyclized efficiently in a 6-endo-trig fashion to afford 2-substituted cyclo-hexanones (67). When a tertiary substituent is present on the ketone, a spirocyclic structure can be formed in a similar yield. Based on deuterium-labeling studies, the authors proposed that after intramolecular nucleophilic attack of the enol to the olefin, the resulting Pd(π)-alkyl species (69) would undergo a “chain walk” to place palladium on the $\alpha$ -position through a series of $\beta$ -hydride elimination/re-insertion steps (Scheme 46). Protonation of the Pd(π)-enolate (72) would release the product and regenerate the active palladium catalyst. ${}^{87}$
In 2011, an efficient gold-catalyzed intramolecular $\alpha$ -alkylation with aliphatic olefins was developed by Che and coworkers (Scheme 47). ${}^{88}$ Compared to Widenhoefer’s protocol, the gold-catalyzed method has several distinct features. First, no additional protic acids are employed because the cationic gold is Lewis acidic enough to facilitate enol formation. Second, the 5-exo-trig cyclization to give five-membered rings is favored over the 6-endo-trig pathway. Mechanistically, the alkyl-Au(1) species generated from the enol addition step was proposed to undergo direct protonation to give the alkylation product as opposed to the aforementioned “chain walk” process. No coupling with aromatic olefins was reported with either Widenhoefer or Che’s catalytic system, which can likely be attributed to the acid-lability of aryl olefins.
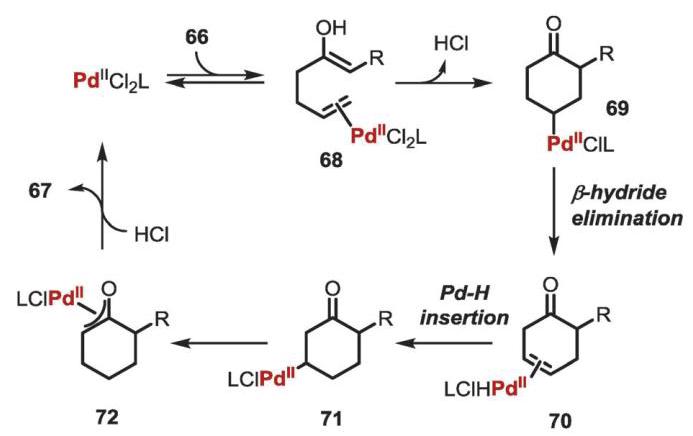
Scheme 46 Proposed pathway of palladium-catalyzed intramolecular $\alpha$ - alkylation.
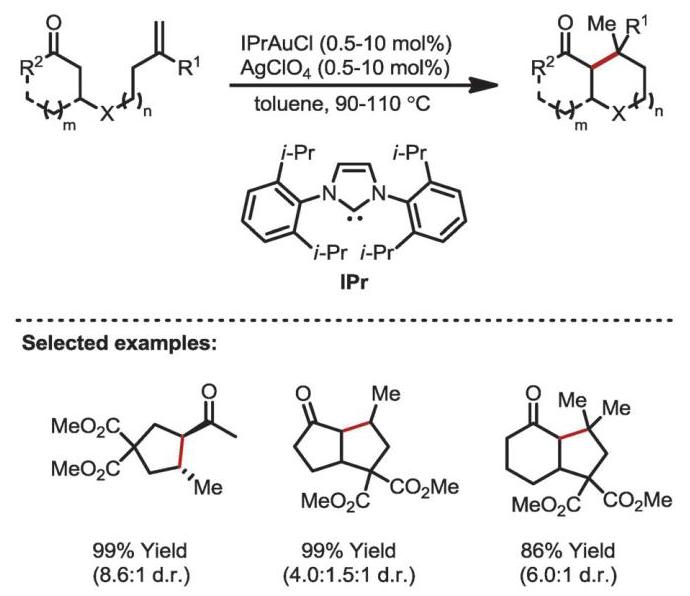
Scheme 47 Gold-catalyzed intramolecular $\alpha$ -alkylation of ketones with olefins.
Besides intramolecular alkylation reactions, several approaches for the more challenging intermolecular $\alpha$ -alkylation of ketones with simple olefins have also emerged. Intermolecular alkylation with aryl olefins, e.g. styrene, is known to be catalyzed by strong bases such as ${\mathrm{{KO}}}^{t}\mathrm{{Bu}}$ in polar solvents such as DMSO. ${}^{{89} - {91}}$ In 2012, Kanai and coworkers reported a copper-catalyzed addition of ketone $\alpha - \mathrm{C} - \mathrm{H}$ bonds to styrene derivatives (Scheme 48). ${}^{92}$ Mesityl cuprate and ${\mathrm{{CuO}}}^{\mathrm{i}}\mathrm{{Pr}}$ were both found to be efficient precatalysts. Ketones with diverse structures can be coupled with styrene derivatives to afford linear alkylation products. While the exact mechanism remains unclear, a dual role of the copper catalyst was proposed: (1) it acts as a strong base to generate a $\mathrm{{Cu}}\left( \mathrm{I}\right)$ -enolate; (2) it serves as a $\pi$ acid to activate the styrene.
An unusual radical approach also proved to be feasible for the addition of $\alpha - \mathrm{C} - \mathrm{H}$ bonds of ketones to aliphatic olefins. ${}^{93}$ In 2000, Ishii and coworkers reported an $\alpha$ -alkylation reaction of ketones with alkenes using a combination of catalytic $\mathrm{{Mn}}{\left( \mathrm{{OAc}}\right) }_{2}$ and $\mathrm{{Co}}{\left( \mathrm{{OAc}}\right) }_{2}$ (Scheme 49, eqn (31)). Both cyclic and linear ketones can couple with 1-octene or isopropenyl acetate to give the alkylation products. Acetic acid was used as solvent, and the reaction was run under a mixed atmosphere of nitrogen and oxygen. In the proposed mechanism, the ketone substrate was first oxidized by in situ generated $\mathrm{{Mn}}{\left( \mathrm{{OAc}}\right) }_{3}$ to give an $\alpha$ -keto radical species (74). Subsequent radical addition to the alkene, followed by hydrogen abstraction from either the solvent or another ketone, would provide the alkylation product. The presence of oxygen was critical, suggesting an oxidative process. Although the cobalt salt is not necessary for product formation, it is believed to assist the re-oxidation of $\mathrm{{Mn}}{\left( \mathrm{{OAc}}\right) }_{2}$ by oxygen. A major drawback of this approach is the use of excess ketone, presumably to sustain the efficiency of the radical generation. When styrene was used as the substrate, an interesting peroxo bicycle was formed (eqn (32)).
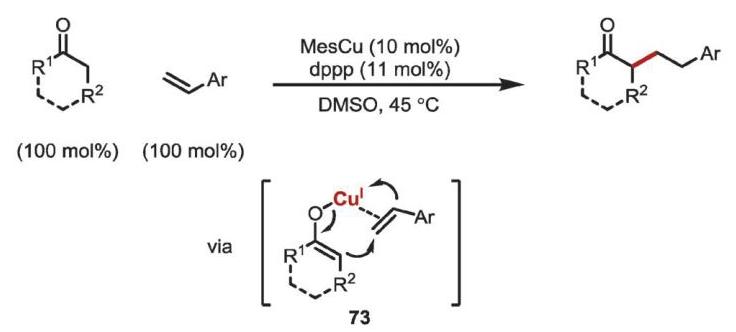
Scheme 48 Copper-catalyzed intermolecular $\alpha$ -alkylation of ketones with olefins.
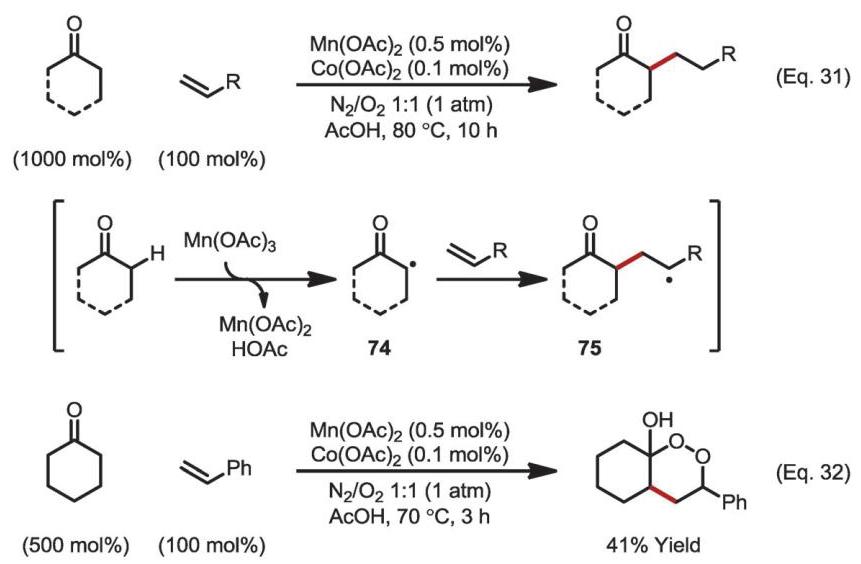
Scheme 49 Manganese-catalyzed intermolecular $\alpha$ -alkylation of ketones with olefins.
In 2014, Mo and Dong reported a Rh(1)-catalyzed $\alpha$ -alkylation of ketones with various alkenes $v$ ia bifunctional catalysis (Scheme 50). ${}^{94}$ The dual activation of the ketone and olefin was achieved through the use of $\mathrm{{Rh}}\left( \mathrm{I}\right) /7$ -azaindoline as the bifunctional catalyst. In the proposed catalytic cycle, the secondary amine motif in 7-azaindoline (81) would first condense with ketone to form enamine 77, which converts a ketone ${\mathrm{{sp}}}^{3}\alpha - \mathrm{C} - \mathrm{H}$ bond to an ${\mathrm{{sp}}}^{2} \; \mathrm{C} - \mathrm{H}$ bond. One benefit of the transient enamine formation is that ${\mathrm{{sp}}}^{2}\mathrm{C} - \mathrm{H}$ bonds are generally easier to be activated than ${\mathrm{{sp}}}^{3}$ bonds both thermodynamically and kinetically. ${}^{95}$ Directed by the pyridine moiety, oxidative addition of $\mathrm{{Rh}}\left( \mathrm{I}\right)$ into the $\alpha - \mathrm{C} - \mathrm{H}$ bond would then yield six-membered rhodacycle 78. Subsequent migratory insertion of the olefin into the Rh-hydride bond, followed by reductive elimination to form the C-C bond and hydrolysis of the enamine, furnished the alkylation product and re-generated the bifunctional catalyst.
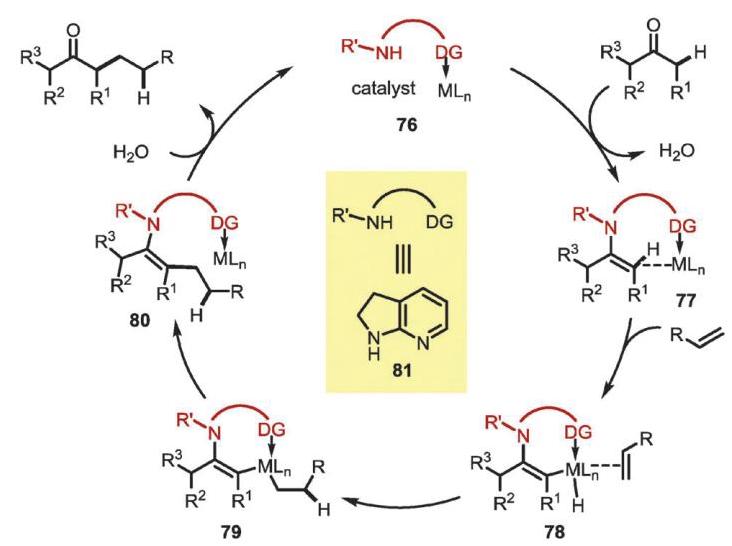
Scheme 50 Proposed strategy of $\alpha$ -alkylation of ketones using bifunctional catalysis.
Based on the proposed strategy, the optimized conditions consist of ${\left\lbrack \mathrm{{Rh}}{\left( \mathrm{{coe}}\right) }_{2}\mathrm{{Cl}}\right\rbrack }_{2}/\mathrm{{bis}}\lbrack 1,3$ -bis(2,4,6-trimethylphenyl)imidazol- 2-ylidene] (IMes) as the precatalyst, and catalytic tosylsulfonic acid to promote enamine formation (Scheme 51). This strategy enabled coupling of both aliphatic and aromatic alkenes giving the alkylation products with linear selectivity. Both cyclic and linear ketones can be coupled with various $\alpha$ -olefins to give the corresponding alkylation products. In addition, due to the pH- and redox-neutral conditions, many sensitive functional groups, such as free alcohols, amines and malonates etc., were found compatible.
4.2 Addition to alkynes
The direct addition of ketone $\alpha - \mathrm{C} - \mathrm{H}$ bonds to alkynes represents a promising approach to access enone derivatives. For example, the intramolecular ketone-alkyne cyclization has been extensively studied and employed to construct cyclic enone structures. ${}^{{96} - {103}}$ Although a large collection of catalysts and reaction conditions have been developed to facilitate this cyclization, the substrate scope is primarily focused on 1,3-dicarbonyl compounds. Nevertheless, $\pi$ -acid-catalyzed ketone-mediated transformations have been recently developed.
In 2010, Davies and coworkers reported a gold-catalyzed intramolecular cycloisomerisation of alkynes and simple ketones (Scheme 52). ${}^{104}$ A number of tethered keto-alkynes could cyclize in the presence of a cationic gold catalyst to give fused or spiro-enone derivatives. While the direct addition of the $\alpha - \mathrm{C} - \mathrm{H}$ bond to the alkyne represents a possible pathway, mechanistic studies indicate an alkyne hydration/aldol condensation pathway seems to be more reasonable. In the proposed reaction pathway, gold-catalyzed alkyne hydration with adventitious water would first generate diketone 82, which upon intramolecular aldol condensation gives the enone product.
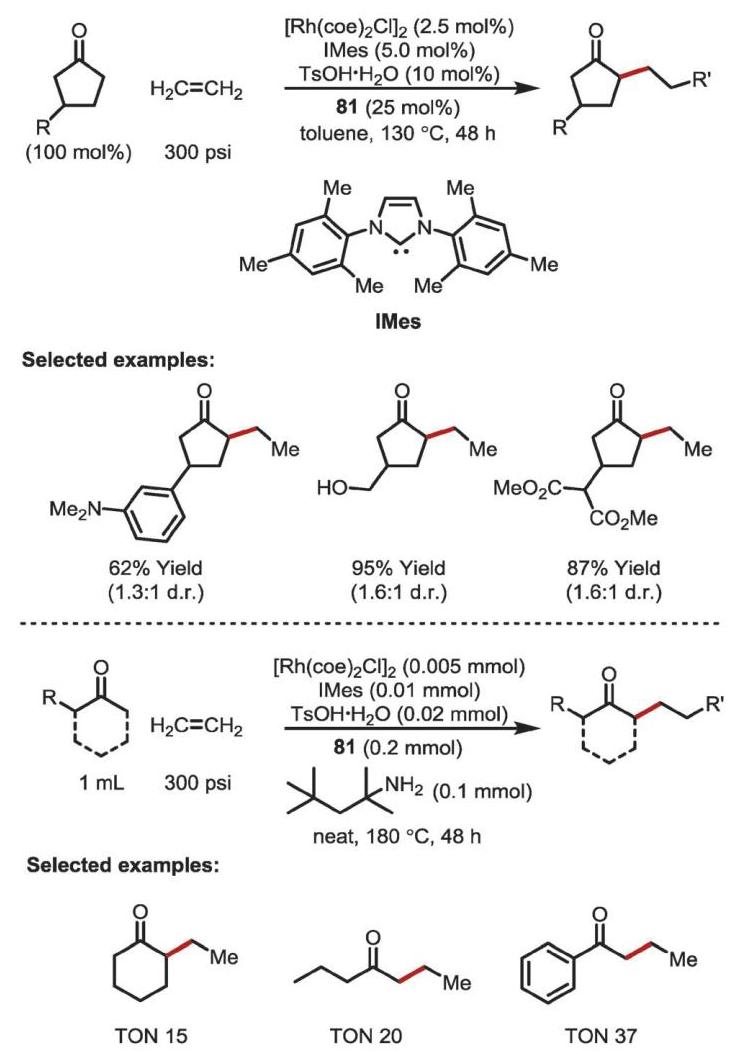
Scheme 51 Rhodium-catalyzed $\alpha$ -alkylation of ketones with bifunctional ligand.
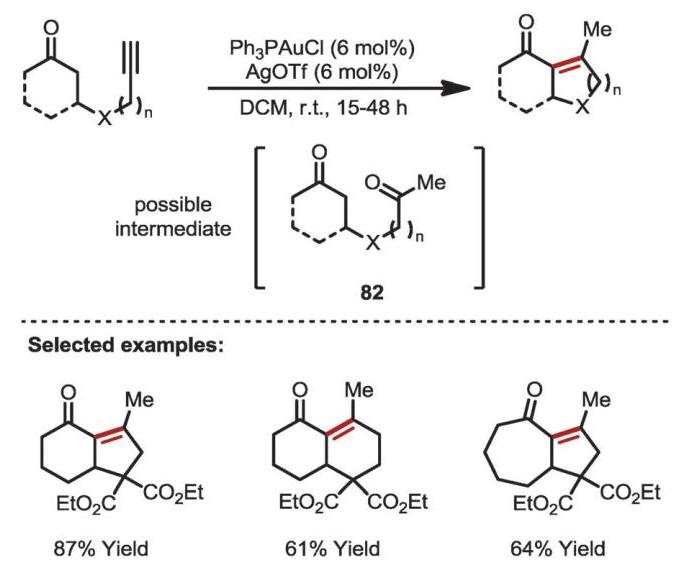
Scheme 52 Gold-catalyzed intramolecular $\alpha$ -alkenylation of ketones with alkynes.
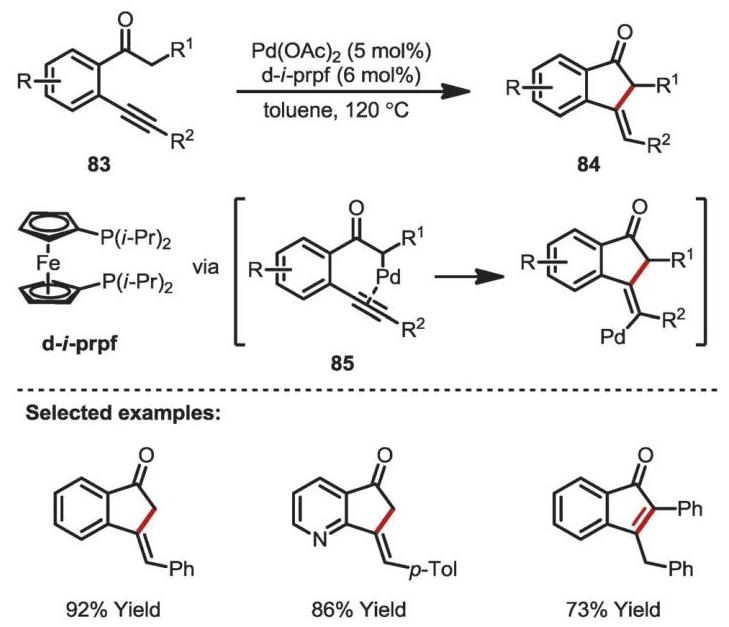
Scheme 53 Palladium-catalyzed intramolecular $\alpha$ -alkenylation of ketones with alkynes.
A palladium-catalyzed cyclization of keto-alkynes was later reported by Gevorgyan and coworkers. ${}^{105}2$ -Alkynyl acetophenone derivatives can readily cyclize to form $\beta$ -alkylidene indanone structures (Scheme 53). Complete stereoselectivity for the $E$ isomers was observed for the cyclization. Interestingly, when the ketone possesses an $\alpha$ -aryl substituent, a conjugated indenone structure was formed instead of the indanone. DFT calculation implied the reaction proceeded through palladium enolate 85, a similar intermediate to that proposed in Kanai’s alkylation reaction with olefins (vide supra, Scheme 48).
The intermolecular addition of ketones to alkynes has limited success using simple ketones as the substrate. No transition metal-catalyzed example has been reported. To date, stoichiometric Lewis acid-mediated addition of silyl enol ethers into terminal alkynes ${}^{{106} - {109}}$ and strong base-promoted addition of potassium enolates into aryl terminal acetylenes ${}^{{110},{111}}$ represent the only known examples.
5. Conclusion
In summary, the transition metal-catalyzed ketone-directed or mediated C-H functionalization has been a rapidly growing field over the past two decades. While the mode of activation varies from case to case, the use of transition metals has driven transformations of ketones beyond the intrinsic reactivity allowing C-H functionalization to occur at the less reactive position or with unactivated coupling partners. Among all the topics covered in this review, the aromatic ortho C-H function-alization using ketone DGs has been most well studied. Future endeavors are expected to demonstrate more applications with these methods and introduce more types of FGs. Although the functionalization of ketone aliphatic C-H bonds using transition-metal catalysis is a relatively new area, an avalanche of novel approaches have sprung up recently. While it is still too early to assess the synthetic potential of these methods, they nonetheless offer new perspectives on how strategy and catalyst design can enable the reactivity and selectivity that are not expected from conventional viewpoints.
Acknowledgements
We thank the University of Texas at Austin and Cancer Prevention and Research Institute of Texas (R1118) for a start-up fund, and thank the Welch Foundation (F-1781) and NSF (CAREER: CHE-1254935) for research grants. G.D. is a Searle Scholar and Sloan fellow.
References
1 For recent reviews on transition metal-catalyzed C-H func-tionalization, see: (a) H. M. L. Davies, J. Mol. Catal. A: Chem., 2002, 189, 125-135; (b) F. Kakiuchi and S. Murai, Acc. Chem. Res., 2002, 35, 826-834; (c) X. Chen, K. M. Engle, D.-H. Wang and J.-Q. Yu, Angew. Chem., Int. Ed., 2009, 48, 5094-5115; (d) L. Ackermann, R. Vicente and A. R. Kapdi, Angew. Chem., Int. Ed., 2009, 48, 9792-9826; (e) R. Jazzar, J. Hitce, A. Renaudat, J. Sofack-Kreutzer and O. Baudoin, Chem. - Eur. J., 2010, 16, 2654-2672; (f) O. Daugulis, Top. Curr. Chem., 2010, 292, 57-84; (g) D. A. Colby, R. G. Bergman and J. A. Ellman, Chem. Rev., 2010, 110, 624-655; (h) I. A. I. Mkhalid, J. H. Barnard, T. B. Marder, J. M. Murphy and J. F. Hartwig, Chem. Rev., 2010, 110, 890-931; (i) O. Baudoin, Chem. Soc. Rev., 2011, 40, 4902-4911; (j) H. Li, B.-J. Li and Z.-J. Shi, Catal. Sci. Technol., 2011, 1, 191-206; (k) H. Lu and X. P. Zhang, Chem. Soc. Rev., 2011, 40, 1899-1909; (l) J. Wencel-Delord, T. Dröge, F. Liu and F. Glorius, Chem. Soc. Rev., 2011, 40, 4740-4761; (m) P. B. Arockiam, C. Bruneau and P. H. Dixneuf, Chem. Rev., 2012, 112, 5879-5918; (n) G. Song, F. Wang and X. Li, Chem. Soc. Rev., 2012, 41, 3651-3678.
2 For recent reviews on directing group strategy, see: (a) T. W. Lyons and M. S. Sanford, Chem. Rev., 2010, 110, 1147-1169; (b) G. Rousseau and B. Breit, Angew. Chem., Int. Ed., 2011, 50, 2450-2494; (c) M. Zhang, Y. Zhang, X. Jie, H. Zhao, G. Li and W. Su, Org. Chem. Front., 2014, 1, 843-895.
3 For selected examples, see: (a) S. R. Mancha, C. M. Regnery, J. R. Dahlke, K. A. Miller and D. J. Blake, Bioorg. Med. Chem. Lett., 2013, 23, 562-564; (b) G. N. Roth, A. Chandra and M. G. Nair, J. Nat. Prod., 1998, 61, 542-545; (c) N. Misawa, R. Nakamura, Y. Kagiyama, H. Ikenaga, K. Furukawa and K. Shindo, Tetrahedron, 2005, 61, 195-204; (d) L. S. Melvin, M. R. Johnson, C. A. Harbert, G. M. Milne and A. Weissman, J. Med. Chem., 1984, 27, 67-71; (e) D. P. G. Hamon, P. J. Hayball, R. A. Massy-Westropp, J. L. Newton and J. G. Tamblyn, Tetrahedron: Asymmetry, 1996, 7, 263-272.
4 S. Murai, F. Kakiuchi, S. Sekine, Y. Tanaka, A. Kamatani, M. Sonoda and N. Chatani, Nature, 1993, 366, 529-531.
5 For recent reviews on $\alpha$ -arylation of ketones, see: (a) C. C. C. Johansson and T. J. Colacot, Angew. Chem., Int. Ed., 2010, 49, 676-707; (b) F. Bellina and R. Rossi, Chem. Rev., 2010, 110, 1082-1146; (c) D. A. Culkin and J. F. Hartwig, Acc. Chem. Res., 2003, 36, 234-245.
6 For selected reviews, see: (a) R. C. Larock, Comprehensive Organic Transformation, Wiley-VCH, New York, 2nd edn, 1999, p. 709; (b) R. Mahrwald, Chem. Rev., 1999, 99, 1095-1120; (c) C. Palomo, M. Oiarbide and J. M. Garcia, Chem. Soc. Rev., 2004, 33, 65-75.
7 C. Laurance and J.-F. Gal, Lewis Basicity and Affinity Scales: Data and Measurement, Wiley-VCH, Weinheim, 2010.
8 F. Kakiuchi, S. Sekine, Y. Tanaka, A. Kamatani, N. Chatani and S. Murai, Bull. Chem. Soc. Jpn., 1995, 68, 62-83.
9 S. Murai, F. Kakiuchi, S. Sekine, Y. Tanaka, A. Kamatani and N. Chatani, Pure Appl. Chem., 1994, 66, 1527-1534.
10 M. Sonoda, F. Kakiuchi, N. Chatani and S. Murai, J. Organomet. Chem., 1995, 504, 151-152.
11 M. Sonoda, F. Kakiuchi, N. Chatani and S. Murai, Bull. Chem. Soc. Jpn., 1997, 70, 3117-3128.
12 K. Hiraki, T. Ishimoto and H. Kawano, Bull. Chem. Soc. Jpn., 2000, 73, 2099-2108.
13 T. Matsubara, N. Koga, D. G. Musaev and K. Morokuma, J. Am. Chem. Soc., 1998, 120, 12692-12693.
14 F. Kakiuchi, T. Kochi, E. Mizushima and S. Murai, J. Am. Chem. Soc., 2010, 132, 17741-17750.
15 Y. Guari, S. Sabo-Etienne and B. Chaudret, J. Am. Chem. Soc., 1998, 120, 4228-4229.
16 Y. Guari, A. Castellanos, S. Sabo-Etienne and B. Chaudret, J. Mol. Catal. A: Chem., 2004, 212, 77-82.
17 S. Busch and W. Leitner, Chem. Commun., 1999, 2305-2306.
18 S. Busch and W. Leitner, Adv. Synth. Catal., 2001, 343, 192-195.
19 M. Grellier, L. Vendier, B. Chaudret, A. Albinati, S. Rizzato, S. Mason and S. Sabo-Etienne, J. Am. Chem. Soc., 2005, 127, 17592-17593.
20 R. Martinez, R. Chevalier, S. Darses and J.-P. Genet, Angew. Chem., Int. Ed., 2006, 45, 8232-8235.
21 R. Martinez, M.-O. Simon, R. Chevalier, C. Pautigny, J.-P. Genet and S. Darses, J. Am. Chem. Soc., 2009, 131, 7887-7895.
22 M.-O. Simon, R. Martinez, J.-P. Genet and S. Darses, J. Org. Chem., 2010, 75, 208-210.
23 M.-O. Simon, J.-P. Genet and S. Darses, Org. Lett., 2010, 12, 3038-3041.
24 B. M. Trost, K. Imi and I. W. Davies, J. Am. Chem. Soc., 1995, 117, 5371-5372.
25 F. Kakiuchi, Y. Tanaka, T. Sato, N. Chatani and S. Murai, Chem. Lett., 1995, 679-680.
26 T. Sato, F. Kakiuchi, N. Chatani and S. Murai, Chem. Lett., 1998, 893-894.
27 C. P. Lenges and M. Brookhart, J. Am. Chem. Soc., 1999, 121, 6616-6623.
28 C.-H. Jun, C. W. Moon, J.-B. Hong, S.-G. Lim, K.-Y. Chung and Y.-H. Kim, Chem. - Eur. J., 2002, 8, 485-492.
29 C.-H. Jun, C. W. Moon, Y.-M. Kim, H. Lee and J. H. Lee, Tetrahedron Lett., 2002, 43, 4233-4236.
30 K. Tsuchikama, M. Kasagawa, Y.-K. Hashimoto, K. Endo and T. Shibata, J. Organomet. Chem., 2008, 693, 3939-3942.
31 G. E. M. Crisenze, N. G. McCreanor and J. F. Bower, J. Am. Chem. Soc., 2014, 136, 10258-10261.
32 For a recent review, see: C. S. Yeung and V. M. Dong, Chem. Rev., 2011, 111, 1215.
33 F. Kakiuchi, T. Sato, M. Yamauchi, N. Chatani and S. Murai, Chem. Lett., 1999, 19-20.
34 F. W. Patureau, T. Besset and F. Glorius, Angew. Chem., Int. Ed., 2011, 50, 1064-1067.
35 K. Padala and M. Jeganmohan, Org. Lett., 2011, 13, 6144-6147.
36 X.-Y. Shi and C.-J. Li, Org. Lett., 2013, 15, 1476-1479.
37 F. Kakiuchi, Y. Yamamoto, N. Chatani and S. Murai, Chem. Lett., 1995, 681-682.
38 F. Kakiuchi, T. Uetsuhara, Y. Tanaka, N. Chatani and S. Murai, J. Mol. Catal. A: Chem., 2002, 182, 511-514.
39 F. W. Patureau, T. Besset, N. Kuhl and F. Glorius, J. Am. Chem. Soc., 2011, 133, 2154-2156.
40 K. Muralirajan, K. Parthasarathy and C.-H. Cheng, Angew. Chem., Int. Ed., 2011, 50, 4169-4172.
41 R. K. Chinnagolla and M. Jeganmohan, Eur. J. Org. Chem., 2012, 417-423.
42 K. Tsuchikama, M. Kasagawa, K. Endo and T. Shibata, Synlett, 2010, 97-100.
43 K. Tsuchikama, Y. Kuwata, Y.-K. Tahara, Y. Yoshinami and T. Shibata, Org. Lett., 2007, 9, 3097-3099.
44 K. Tanaka, Y. Otake, A. Wada, K. Noguchi and M. Hirano, Org. Lett., 2007, 9, 2203-2206.
45 K. Tanaka, Y. Otake, H. Sagae, K. Noguchi and M. Hirano, Angew. Chem., Int. Ed., 2008, 47, 1312-1316.
46 R. Santhoshkumar, S. Mannathan and C.-H. Cheng, Org. Lett., 2014, 16, 4208-4211.
47 P. W. R. Harris and P. D. Woodgate, J. Organomet. Chem., 1997, 530, 211-223.
48 P. W. R. Harris and P. D. Woodgate, Tetrahedron, 2000, 56, 4001-4015.
49 H. Guo and W. P. Weber, Polym. Bull., 1994, 32, 525-528.
50 H. Guo, M. A. Tapsak and W. P. Weber, Polym. Bull., 1994, 34, 49-55.
51 T. M. Londergan, Y. You, M. E. Thompson and W. P. Weber, Macromolecules, 1998, 31, 2784-2788.
52 D. Huang, J. Yang and W. P. Weber, Polym. Bull., 2000, 43, 465-470.
53 H. Guo, M. A. Tapsak and W. P. Weber, Polym. Bull., 1994, 33, 417-423.
54 S. K. Gupta and W. P. Weber, Macromolecules, 2002, 35, 3369-3373.
55 F. Kakiuchi, S. Kan, K. Igi, N. Chatani and S. Murai, J. Am. Chem. Soc., 2003, 125, 1698-1699.
56 F. Kakiuchi, Y. Matsuura, S. Kan and N. Chatani, J. Am. Chem. Soc., 2005, 127, 5936-5945.
57 S. Ueno, N. Chatani and F. Kakiuchi, J. Org. Chem., 2007, 72, 3600-3602.
58 S. Ueno, T. Kochi, N. Chatani and F. Kakiuchi, Org. Lett., 2009, 11, 855-858.
59 T. Satoh, Y. Kametani, Y. Terao, M. Miura and M. Nomura, Tetrahedron Lett., 1999, 40, 5345-5348.
60 Y. Terao, Y. Kametani, H. Wakui, T. Satoh, M. Miura and M. Nomura, Tetrahedron, 2001, 57, 5967-5974.
61 P. Gandeepan, K. Parthasarathy and C.-H. Cheng, J. Am. Chem. Soc., 2010, 132, 8569-8571.
62 P. Gandeepan, C.-H. Hung and C.-H. Cheng, Chem. Commun., 2012, 48, 9379-9381.
63 H. Li, R.-Y. Zhu, W.-J. Shi, K.-H. He and Z.-J. Shi, Org. Lett., 2012, 14, 4850-4853.
64 G. Shan, X. Yang, L. Ma and Y. Rao, Angew. Chem., Int. Ed., 2012, 51, 13070-13074.
65 F. Mo, L. J. Trzepkowski and G. Dong, Angew. Chem., Int. Ed., 2012, 51, 13075-13079.
66 P. Y. Choy and F. Y. Kwong, Org. Lett., 2013, 15, 270-273.
67 V. S. Thirunavukkarasu and L. Ackermann, Org. Lett., 2012, 14, 6206-6209.
68 G. Shan, X. Han, Y. Lin, S. Yu and Y. Rao, Org. Biomol. Chem., 2013, 11, 2318-2322.
69 N. Schröder, J. Wencel-Delord and F. Glorius, J. Am. Chem. Soc., 2012, 134, 8298-8301.
70 X. Sun, G. Shan, Y. Sun and Y. Rao, Angew. Chem., Int. Ed., 2013, 52, 4440-4444.
71 B. Xiao, T.-J. Gong, J. Xu, Z.-J. Liu and L. Liu, J. Am. Chem. Soc., 2011, 133, 1466-1474.
72 J. Kim, J. Kim and S. Chang, Chem. - Eur. J., 2013, 19, 7328-7333.
73 Q.-Z. Zheng, Y.-F. Liang, C. Qin and N. Jiao, Chem. Commun., 2013, 49, 5654-5656.
74 M. Bhanuchandra, M. R. Yadav, R. K. Rit, M. R. Kuram and A. K. Sahoo, Chem. Commun., 2013, 49, 5225-5227.
75 For selected reviews on transition-metal-catalyzed conjugate addition, see: (a) B. E. Rossiter and N. M. Swingle, Chem. Rev., 1992, 92, 771-806; (b) A. Gutnov, Eur. J. Org. Chem., 2008, 4547-4554; (c) T. Hayashi and K. Yamasaki, Chem. Rev., 2003, 103, 2829-2844.
76 (a) D. R. Buckle and I. L. Pinto, in Comprehensive Organic Synthesis, ed. B. M. Trost and I. Fleming, Pergamon Press, Oxford, U.K., 1991, Vol. 7, pp. 119-149; (b) R. C. Larock, Comprehensive Organic Transformations, John Wiley & Sons, New York, 1999, pp. 251-256; (c) J. Muzart, Eur. J. Org. Chem., 2010, 3779-3790.
77 For recent reviews on β-C-H functionalization, see: (a) G. Rouquet and N. Chatani, Angew. Chem., Int. Ed., 2013, 52, 11726-11743; (b) Z. Huang and G. Dong, Tetrahedron Lett., 2014, 55, 5869-5889; (c) G. Qiu and J. Wu, Org. Chem. Front., 2015, 2, 169-178.
78 M. T. Pirnot, D. A. Rankic, D. B. C. Martin and D. W. C. MacMillan, Science, 2013, 339, 1593-1596.
79 F. R. Petronijević, M. Nappi and D. W. C. MacMillan, J. Am. Chem. Soc., 2013, 135, 18323-18326.
80 M. Okada, T. Fukuyama, K. Yamada, I. Ryu, D. Ravelli and M. Fagnoni, Chem. Sci., 2014, 5, 2893-2898.
81 Z. Huang and G. Dong, J. Am. Chem. Soc., 2013, 135, 17747-17750.
82 For recent examples of Pd-catalyzed dehydrogenation of ketones, see: (a) T. Diao and S. S. Stahl, J. Am. Chem. Soc., 2011, 133, 14566-14569; (b) T. Diao, T. J. Wadzinski and S. S. Stahl, Chem. Sci., 2012, 3, 887-891; (c) W. Gao, Z. He, Y. Qian, J. Zhao and Y. Huang, Chem. Sci., 2012, 3, 883-886; (d) T. Diao, D. Pun and S. S. Stahl, J. Am. Chem. Soc., 2013, 135, 8205-8212; (e) D. Pun, T. Diao and S. S. Stahl, J. Am. Chem. Soc., 2013, 135, 8213-8221.
83 (a) M. B. Smith and J. March, March’s Advanced Organic Chemistry, Wiley, New York, 2001; (b) T. Anastas and J. C. Warner, Green Chemistry: Theory and Practice, Oxford Univ. Press, Oxford, 1998; (c) W. F. von Oettingen, The Halogenated Hydrocarbons of Industrial and Toxicological Importance, Elsevier, Amsterdam, New York, 1964.
84 (a) G. Stork, R. Terrell and J. Szmuszkovicz, J. Am. Chem. Soc., 1954, 76, 2029-2030; (b) G. Stork and H. Landesman, J. Am. Chem. Soc., 1956, 78, 5128-5129.
85 For a recent review, see: F. Dénès, A. Pérez-Luna and F. Chemla, Chem. Rev., 2010, 110, 2366-2447.
86 S. Wang, T. Pei, X. Han and R. A. Widenhoefer, Org. Lett., 2003, 5, 2699-2701.
87 X. Han, X. Wang, T. Pei and R. A. Widenhoefer, Chem. - Eur. J., 2004, 10, 6333-6342.
88 Y.-P. Xiao, X.-Y. Liu and C.-M. Che, Angew. Chem., Int. Ed., 2011, 50, 4937-4941.
89 H. Pines, S. V. Kannan and J. Simonik, J. Org. Chem., 1971, 36, 2311-2315.
90 A. L. Rodriguez, T. Bunlaksananusorn and P. Knochel, Org. Lett., 2000, 2, 3285-3287.
91 J.-M. Gaudin and P. Millet, Chem. Commun., 2008, 588-590.
92 S. Majima, Y. Shimizu and M. Kanai, Tetrahedron Lett., 2012, 53, 4381-4384.
93 T. Iwahama, S. Sakaguchi and Y. Ishii, Chem. Commun., 2000, 2317-2318. 94 F. Mo and G. Dong, Science, 2014, 345, 68-72.
95 J. F. Hartwig, Organotransition Metal Chemistry: From Bonding to Catalysis, University Science Books, Sausalito, 2010.
96 J. M. Conia and P. Le Perchec, Synthesis, 1975, 1-19.
97 B. K. Corkey and F. D. Toste, J. Am. Chem. Soc., 2005, 127, 17168-17169.
98 A. Ochida, H. Ito and M. Sawamura, J. Am. Chem. Soc., 2006, 128, 16486-16487.
99 Y. Itoh, H. Tsuji, K.-I. Yamagata, K. Endo, I. Tanaka, M. Nakamura and E. Nakamura, J. Am. Chem. Soc., 2008, 130, 17161-17167.
100 T. Yang, A. Ferrali, F. Sladojevich, L. Campbell and D. J. Dixon, J. Am. Chem. Soc., 2009, 131, 9140-9141.
101 A. Matsuzawa, T. Mashiko, N. Kumagai and M. Shibasaki, Angew. Chem., Int. Ed., 2011, 50, 7616-7619.
102 S. S. K. Boominathan, W.-P. Hu, G. C. Senadi and J.-J. Wang, Adv. Synth. Catal., 2013, 355, 3570-3574.
103 S. Shaw and J. D. White, J. Am. Chem. Soc., 2014, 136, 13578-13581.
104 P. W. Davies and C. Detty-Mambo, Org. Biomol. Chem., 2010, 8, 2918-2922.
105 N. Chernyak, S. I. Gorelsky and V. Gevorgyan, Angew. Chem., Int. Ed., 2011, 50, 2342-2345.
106 M. Yamaguchi, A. Hayashi and M. Hirama, J. Am. Chem. Soc., 1993, 115, 3362-3363.
107 M. Yamaguchi, M. Sehata, A. Hayashi and M. Hirama, J. Chem. Soc., Chem. Commun., 1993, 1708-1709.
108 Y. Hayashi, M. Yamaguchi and M. Hirama, Synlett, 1995, 51-52.
109 M. Yamaguchi, T. Tsukagoshi and M. Arisawa, J. Am. Chem. Soc., 1999, 121, 4074-4075.
110 B. A. Trofimov, E. Y. Schmidt, I. A. Ushakov, N. V. Zorina, E. V. Skital’tseva, N. I. Protsuk and A. I. Mikhaleva, Chem. - Eur. J., 2010, 16, 8516-8521.
111 B. A. Trofimov, E. Y. Schmidt, N. V. Zorina, E. V. Ivanova and I. A. Ushakov, J. Org. Chem., 2012, 77, 6880-6886.